Sulfur isn't complicated once you stop treating it like oxygen's weird cousin
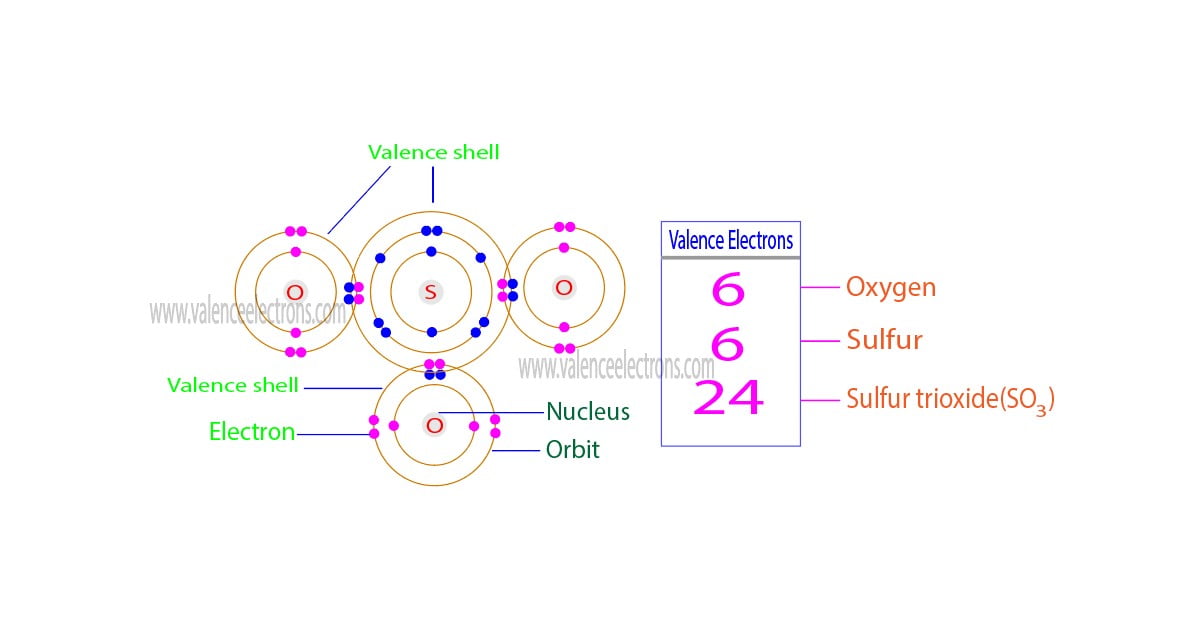
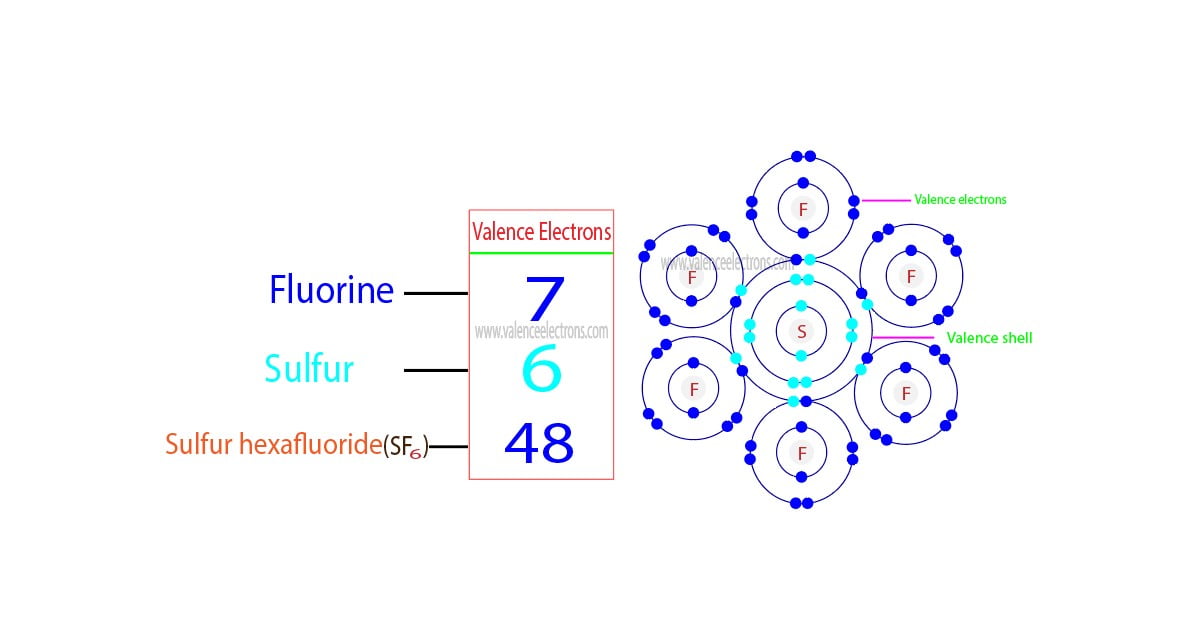
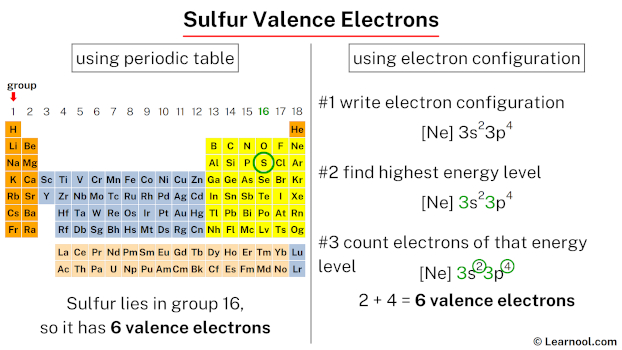
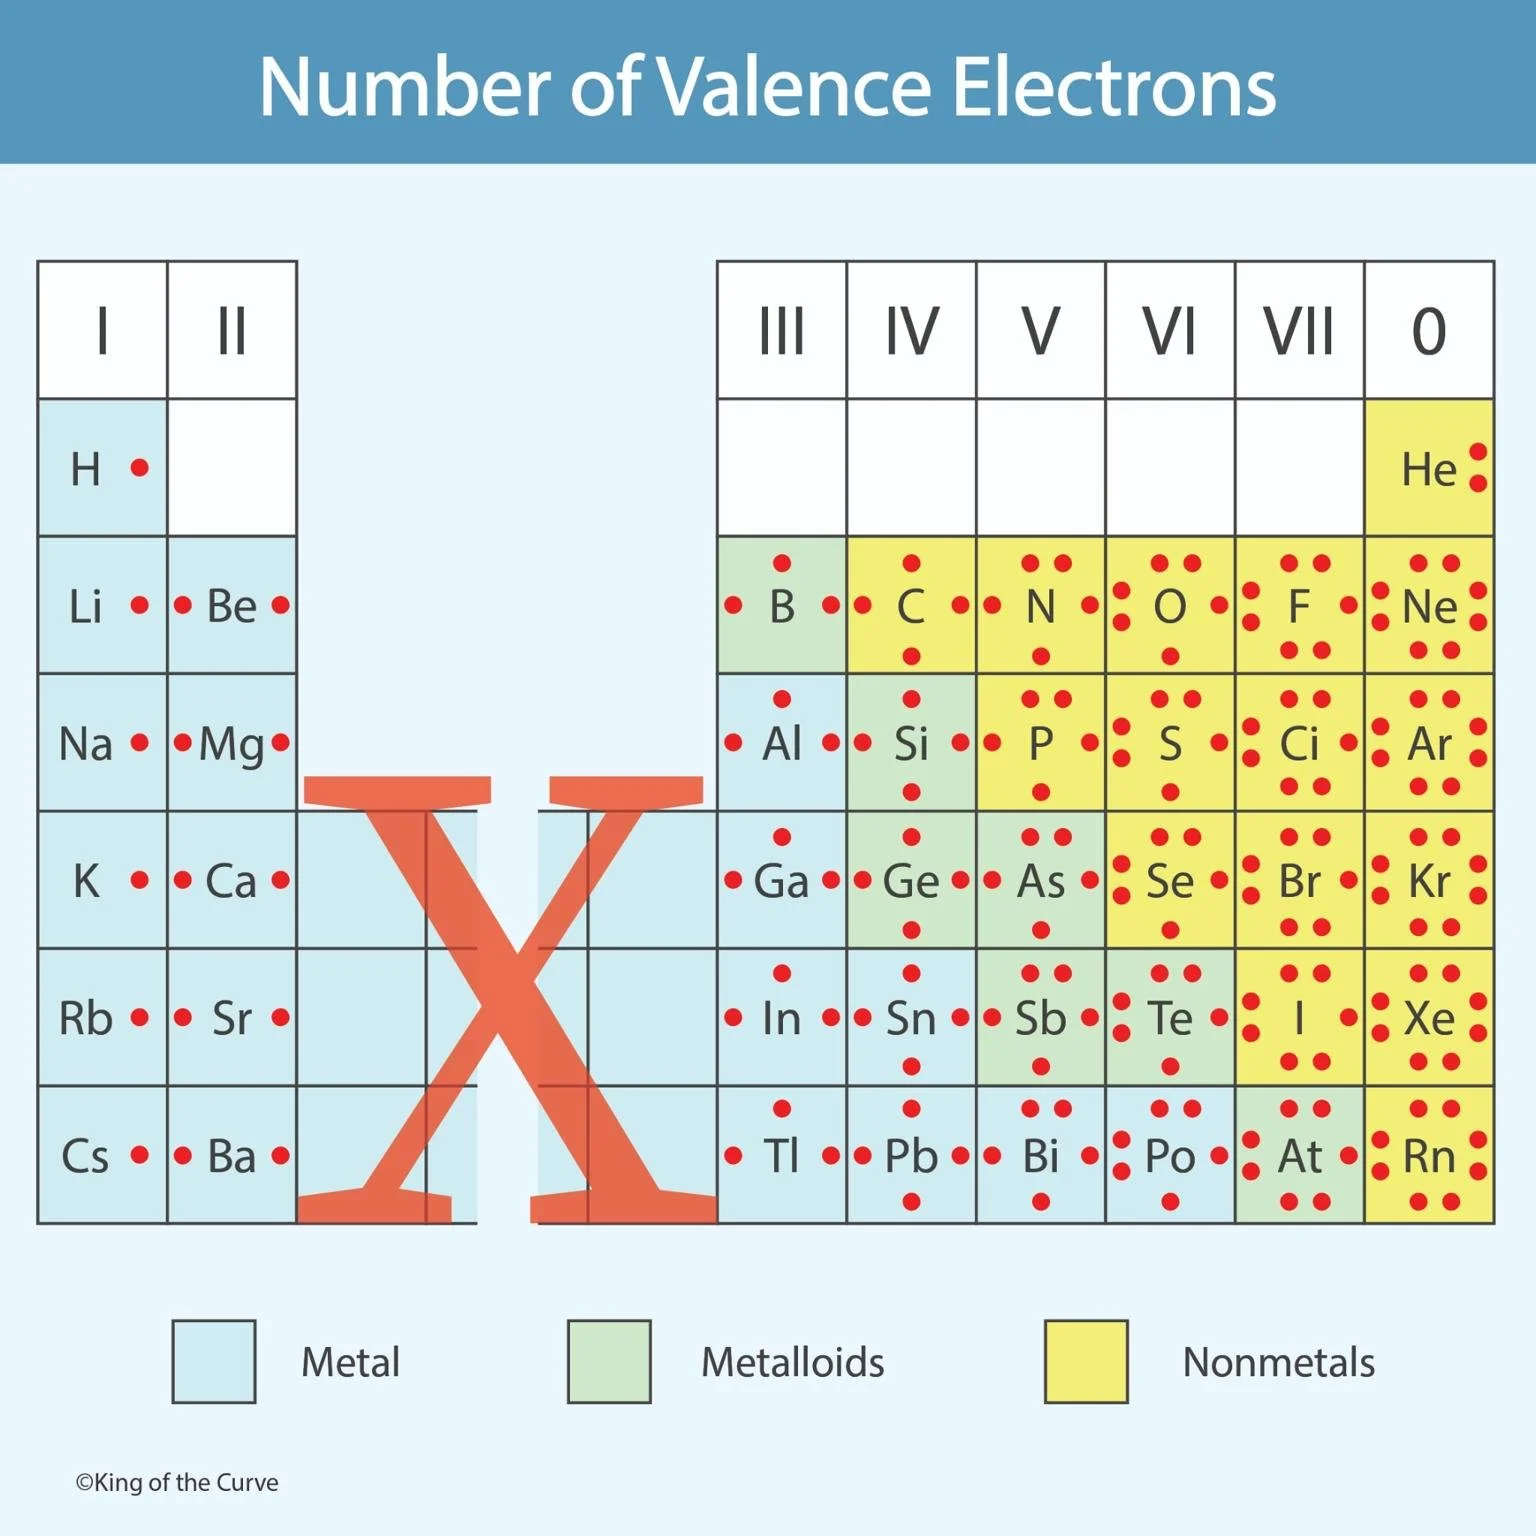
Most people learn sulfur through comparisons. It's in the same group as oxygen, right? Two valence electrons short of a full shell. Same formula, different element. That framing gets you into trouble fast because sulfur does things oxygen simply will not do, and the reason comes down to what happens when you add a third electron shell. Ground state sulfur is [Ne] 3s² 3p. Six valence electrons. Four unpaired slots if you promote one electron from the paired 3p orbital into the empty 3d subshell, which it readily does because that 3d orbitals sits close enough in energy to matter. That expansion capacity is what makes sulfur interesting and what makes it dangerous to model with the same rules you use for carbon or nitrogen. Here's the thing beginners miss: sulfur doesn't just form two bonds like oxygen does. It forms two, four, or six depending on what's around it. SF exists but is unstable. SF is a real compound. SF is commercially produced by the ton. The valence shell can hold twelve electrons in practice, and you'll see compounds where sulfur is formally hypervalent all the time. Think SCl, SO², thiosulfate. Don't try to draw them with simple octet rules and then get frustrated when the math doesn't work. It won't.
I spent about three days once debugging a reaction mechanism where the software kept refusing to protonate a sulfonate intermediate. The program was treating the sulfur center as if it couldn't expand past eight electrons because it was hardcoded with strict octet enforcement. The workaround was switching to a DFT functional that supports diffuse functions and letting the geometry optimizer figure out the coordination number rather than constraining it. Took the calculation from overnight down to about forty minutes on a six-core machine. Let's talk about what sulfur actually does in organic chemistry, since that's where most people encounter it. Thiols, RSH, are more acidic than alcohols. pKa around 10.5 versus 16 for ethanol. The conjugate base,RS, is a better nucleophile than RO in polar aprotic solvents because sulfur is larger and more polarizable. Soft nucleophile, hard electrophile or soft electrophile, depending on what you're targeting. HSAB theory applies cleanly here. Sulfoxides and sulfones come up constantly in synthesis. DMSO isn't just a solvent. It oxidizes primary alcohols to aldehydes with oxalyl chloride in the Swern reaction, and it participates as an active reagent in many other oxidations. The sulfur in DMSO is already in the +4 oxidation state. It can go to +6 in sulfones, and that redox flexibility is why sulfur compounds show up in so many named reactions that have nothing to do with each other on the surface.
The disulfide bond is worth understanding properly. RSSR cleavage and formation is central to biochemistry and to materials chemistry. Reducing agents like DTT or TCEP break disulfides. Iodometry titrates them. In protein folding, the kinetics of disulfide bond formation depend heavily on pH, concentration, and whether the cysteines are sterically accessible. You can have a protein with three disulfide bridges where only one forms correctly under native conditions and the others mispair because the intermediates are trapped in local minima. That's not a theoretical problem. It's the reason refolding experiments fail when you just dilute denatured protein into buffer and wait. Sulfur also does something oxygen doesn't: it forms stable chains. Polymeric sulfur exists. Elemental sulfur at room temperature is mostly S crowns, but heat it above 160°C and the rings open and polymerize into long chains. The melt becomes viscous, then at higher temperatures the chains break again and viscosity drops. You can pour it, quench it, and get amorphous sulfur that slowly re-crystallizes over weeks. This is how you end up with samples that don't match the spec sheet because you melted the wrong grade. On the analytical side, XPS binding energy shifts for sulfur are diagnostic but easy to misassign if you don't account for charge referencing properly. S 2p/ sits around 164 eV for sulfide, 162 eV for thiolate, 168-169 eV for sulfate. If your sample is insulating and you reference to C 1s without correcting for drift, those peaks will shift enough to make a sulfate look like a sulfoxide if you're not careful. I've seen it happen in grant review presentations. Once.
Get the Full Details
Reaction conditions matter more with sulfur than most textbooks imply. Air oxidation of thiols to disulfides is fast enough that you don't always need an oxidant. Just leave a thiol solution open to the bench for a few hours and check the NMR. Dimethyl sulfoxide absorbs water from the atmosphere and slowly degrades to dimethyl sulfide over time, especially if it's been sitting in a non-dried bottle on the shelf for six months. Distillation over calcium hydride fixes that. If you're running a reaction that's sensitive to DMSO degradation products and getting weird byproducts, check your solvent age before you blame your catalyst. Sulfur's redox chemistry is messy in aqueous solution. Thiosulfate disproportionates under acidic conditions. Sulfite gets oxidized by air. Hydrogen sulfide evolves from sulfides when the pH drops below about 7. If you're working with sulfide salts in the hood without checking the pH of your waste stream, you're going to have a bad day and someone else is going to have a worse one. This isn't theoretical. It happened to my group once when a grad student dumped unquenched NaS waste into a mixed acid drain line. The HS release set off the corridor detectors and shut down the building for two hours. Computational chemistry handles sulfur adequately now, but the choice of basis set and functional matters. aug-cc-pVTZ or similar diffuse-augmented sets work for anions and hypervalent species. Standard 6-31G* without diffuse functions will underestimate electron affinity and give poor geometries for sulfates and sulfinates. If you're doing conformational searches on thioethers, remember that the C-S-C angle is wider than C-O-C and the barrier to rotation is lower. The sulfur lone pairs are more diffuse, so n* hyperconjugation stabilizes different conformers than you'd expect from the oxygen analogue.
One practical note about handling elemental sulfur: it's benign compared to most reagents in a sulfur lab. Dust is an irritation, not a catastrophe. The real hazards come from what sulfur makes. HS is lethal at concentrations most people can't smell after the initial warning. SO is a gas that damages lungs and corrodes equipment. Sulfuryl chloride and thionyl chloride are both moisture-sensitive and generate HCl on contact with humid air. Store them under inert atmosphere, use a proper scrubber, and don't assume that because sulfur itself is a cheap yellow powder sitting in a jar on the shelf that the chemistry around it is harmless. The valence shell expansion is the single most important concept for understanding sulfur's behavior. Oxygen can't do it. Sulfur does it regularly. That difference explains why sulfuric acid is a stronger acid than sulfuric acid's oxygen-only analogue would be, why sulfates are tetrahedral and stable, and why sulfur compounds participate in redox chemistry that oxygen compounds simply don't. Once you stop forcing sulfur into oxygen's pattern, the chemistry becomes predictable instead of confusing.