What actually happens inside an enzyme at room temperature
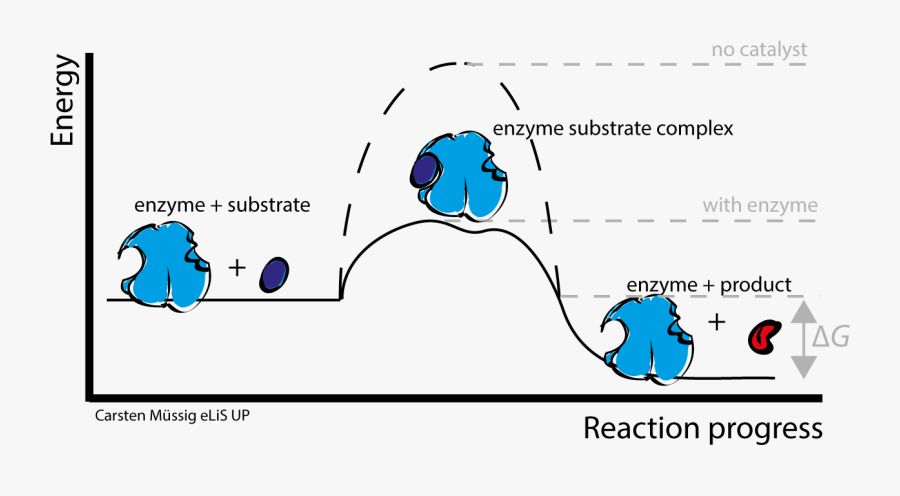
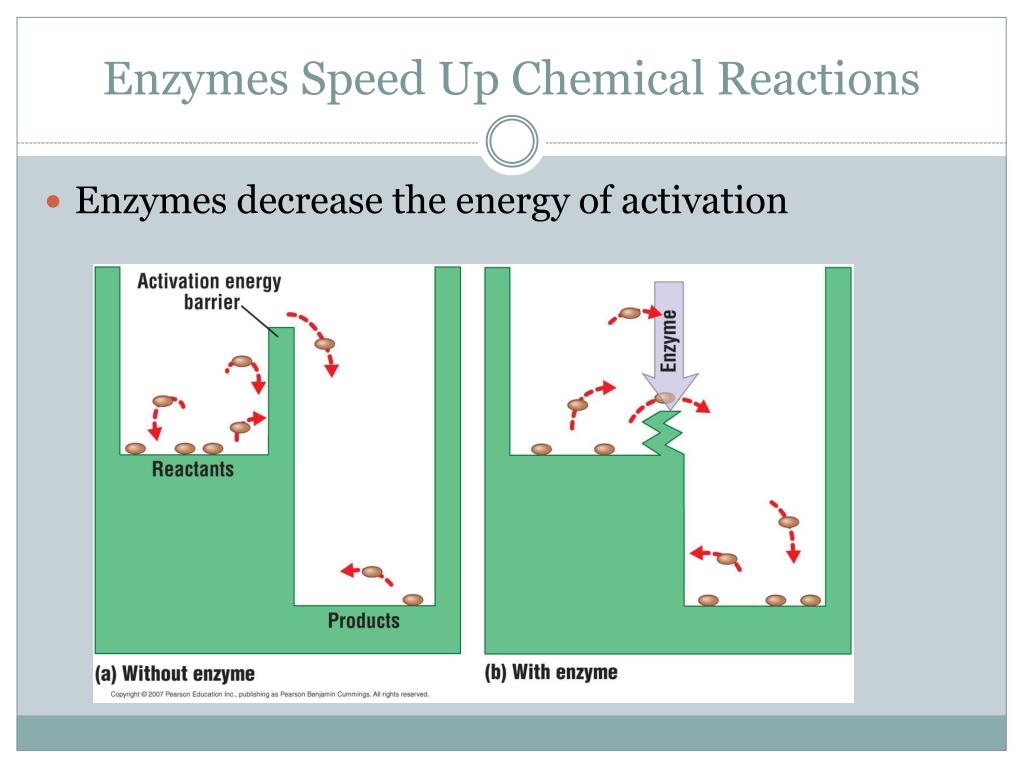
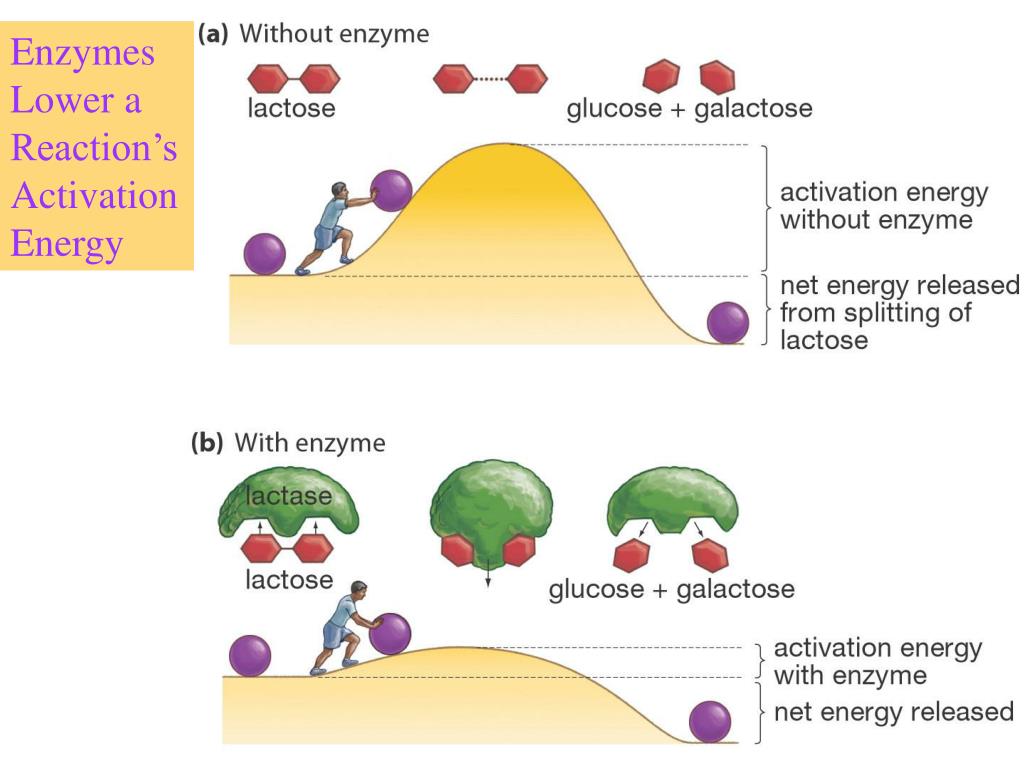
Most people think of enzymes as tiny molecular machines that work like locks and keys. That's the version they teach you in high school biology. The reality is messier, and understanding the actual mechanism matters if you're ever working with real reactions in a lab rather than answering multiple-choice questions. Enzymes are proteins that catalyze chemical reactions by lowering the activation energy required. They don't change the thermodynamics of the reaction. The energy difference between reactants and products stays the same. What changes is the pathway. An uncatalyzed reaction might require crossing an energy barrier of 80 kilojoules per mole. The same reaction with the right enzyme present can happen through a different route where the highest point on the energy landscape is maybe 40 kilojoules per mole. Lower barrier means exponentially more molecules have enough thermal energy to get over it at any given moment. That's the core of it. Not magic. Just physics.
Unlocking The Secrets Of Enzymes How Do They Speed Up Chemical Reactions
The speed-up isn't just one thing. Several mechanisms work in combination, and their relative contribution depends entirely on the specific enzyme and the reaction it catalyzes. I've seen researchers who pick the first explanation they find and run with it as if it applies universally. It doesn't. Proximity and orientation effects is one mechanism. When two substrates need to react, they're floating around in solution colliding randomly. Most collisions are in the wrong orientation and don't lead to product. An enzyme binds both substrates and holds them in a specific geometry that aligns the reactive orbitals. This alone can give rate enhancements of 10 to 100 fold just from holding things still. Not huge compared to some enzymes, but it's real and measurable. Acid-base catalysis is another. Amino acid side chains in the active site can donate or accept protons at positions and times that matter for the reaction. Histidine is the most common player here because its pKa sits right around physiological pH. It can toggle between protonated and deprotonated states without needing extreme conditions. Aspartate and glutamate do similar work at lower pH ranges. Lysine can stabilize negative charges through its positive amine group. These residues aren't sitting there passively. They're positioned so precisely that proton transfer becomes essentially diffusion-limited.
Covalent catalysis involves the enzyme forming a temporary covalent bond with the substrate. Serine proteases are the textbook example. The catalytic serine attacks the peptide bond carbonyl carbon, forming a tetrahedral intermediate and then an acyl-enzyme intermediate. The peptide bond that would normally take thousands of years to hydrolyze in water gets broken in milliseconds. Then water comes in and hydrolyzes the acyl-enzyme to release product and regenerate the free enzyme. The covalent intermediate is higher energy than the starting materials but lower energy than the uncatalyzed transition state. That's the whole point. Metal ion catalysis comes into play when the reaction involves charged species or redox chemistry. Zinc in carbonic anhydrase stabilizes a hydroxide ion and makes it a much better nucleophile. Magnesium in kinases coordinates ATP and positions it correctly while shielding negative charges. Iron-sulfur clusters in oxidoreductases shuttle electrons through proteins. Without the metal, these reactions either wouldn't happen at biologically relevant rates or would require conditions that would denature the protein anyway. Transition state stabilization is probably the most important mechanism and the one people misunderstand the most. The enzyme doesn't bind the substrate tightly. It binds the transition state of the reaction even more tightly. This is the concept that Linus Pauling actually predicted before anyone had structural data to confirm it. The active site is shaped like the transition state geometry, not the ground state geometry. When the substrate enters and starts distorting toward the transition state, the enzyme stabilizes that distorted configuration through electrostatic interactions, hydrogen bonds, and van der Waals contacts. The binding energy released by transition state stabilization is what pays for the catalytic rate enhancement.
Get the Full Details
I learned this the hard way during a project where I was characterizing a mutant form of ketosteroid isomerase. The wild-type enzyme converts 3-oxo-steroids through an enolate intermediate with a rate constant around 10 to the sixth power per second. That's already near diffusion-controlled. I made a point mutation at the active site changing an aspartate to asparagine, expecting maybe a ten-fold reduction in activity. What I got was a thousand-fold drop. The aspartate wasn't doing general base catalysis. It was stabilizing the oxyanion transition state through electrostatic interactions. Changing the charge to neutral removed that stabilization entirely. The lesson was that you can't always predict what a residue does by looking at its chemical properties in isolation. Context matters enormously.
The induced fit model versus conformational selection
For decades the induced fit model reigned supreme. You bind substrate, the enzyme changes shape, and the new conformation is catalytically active. Simple and intuitive. The current thinking is more nuanced. Conformational selection argues that enzymes already exist in an ensemble of different shapes. The substrate simply binds preferentially to the catalytically competent conformation and shifts the equilibrium. Both mechanisms are probably happening simultaneously in most enzymes. The question is which one dominates for a given step in the catalytic cycle. NMR relaxation dispersion experiments have been key here. They can detect minor conformational states that exist at low population but are crucial for catalysis. I've spent a lot of time analyzing data from these experiments and the patterns are sometimes surprising. You'll find a conformer that exists at maybe one percent population in the apo enzyme, and that one percent is the species that does all the chemistry. The other ninety-nine percent is basically spectators. This has practical implications for drug design. If you're screening compounds against the major conformation, you might miss the ones that bind the rare catalytically competent state. Fragment-based screening combined with NMR can catch these interactions that traditional crystallography misses because the binding event itself might stabilize a low-population state.
Catalytic proficiency and why it matters
Enzymes don't just speed up reactions. They speed them up by factors that can exceed 10 to the twelfth power compared to the uncatalyzed reaction. That's what we call catalytic proficiency. The number comes from dividing kcat by the rate constant for the uncatalyzed reaction. For some enzymes, the proficiency is so high that it seems almost impossible. How can a protein achieve rate enhancements this large? The answer involves multiple strategies working together. Ground state destabilization is one. The enzyme might strain the substrate away from its preferred conformation, making it closer to the transition state geometry before any chemistry happens. Desolvation is another. Removing water from the active site creates a low-dielectric environment that strengthens electrostatic interactions and makes charge separation in the transition state less energetically costly. Entropy reduction matters too. By holding substrates in the right orientation, the enzyme eliminates the entropic cost of bringing two molecules together from solution. I worked once with phosphoglucomutase, an enzyme that transfers phosphate groups during glycogen metabolism. The uncatalyzed phosphate transfer in aqueous solution is essentially immeasurable. With the enzyme, the turnover number is around 500 per second. The proficiency calculation gives something on the order of 10 to the eleventh. What's striking is that no single mechanism explains all of that. You need the bound phosphate group acting as a shuttle, the precise positioning of catalytic residues, the exclusion of bulk water, and conformational changes that couple substrate binding to catalysis. Remove any one of these and the rate drops significantly, but the enzyme doesn't go completely dead. They're redundant in a way that's important for robustness.

When enzymes fail and what to do about it
Enzymes are not infinitely robust. They have temperatureoptima, pH optima, and sensitivity to inhibitors that can shut them down completely. Understanding these limitations is as important as understanding how they work when they're functioning normally. Thermal denaturation is the obvious failure mode. Most enzymes lose activity rapidly above forty to fifty degrees Celsius. But some thermophilic enzymes remain active at eighty degrees and above. The difference comes down to protein stability, which is determined by the network of interactions holding the folded structure together. More salt bridges, tighter hydrophobic cores, and shorter surface loops tend to correlate with thermostability. If you're working with an enzyme that's marginally stable, adding glycerol at ten to twenty percent can help. It stabilizes the native state through preferential exclusion. pH sensitivity is another practical concern. Every ionizable group in the active site has a pKa, and the catalytic activity depends on those groups being in the right protonation state. If the pKa values shift due to mutations or environmental changes, the enzyme can become inactive even though the protein structure looks fine. I've seen this happen with histidine residues whose pKa was perturbed by nearby mutations. The enzyme folded correctly and the substrate still bound, but catalysis was severely impaired because the histidine was permanently protonated at physiological pH when it needed to be able to toggle.
Inhibition is the third major concern. Competitive inhibitors bind at the active site and block substrate access. Noncompetitive inhibitors bind elsewhere and alter the enzyme's conformation. Uncompetitive inhibitors only bind to the enzyme-substrate complex. Mixed inhibitors have characteristics of both competitive and noncompetitive binding. Knowing which type you're dealing with matters for designing experiments and interpreting kinetic data. Linearization methods like Lineweaver-Burk plots can be misleading. I recommend using direct nonlinear fitting to the Michaelis-Menten equation whenever possible. The weighting issues with double-reciprocal plots can give you incorrect kinetic parameters, especially at low substrate concentrations where experimental error is largest.
Pre-steady-state kinetics reveals what steady-state hides
Traditional enzyme kinetics measures steady-state conditions where the concentration of the enzyme-substrate complex is constant. The observed rate constant kcat tells you the overall turnover rate but not the individual steps. Pre-steady-state kinetics uses rapid mixing techniques like stopped-flow or quench-flow to observe the reaction during the first milliseconds, before steady state is reached. This can reveal individual rate constants for substrate binding, chemical transformation, and product release. I used pre-steady-state kinetics to characterize a DNA polymerase. The steady-state Km and kcat values suggested reasonable catalytic efficiency. But the pre-steady-state data showed that chemistry was actually faster than product release. The rate-limiting step under steady-state conditions was dissociation of the product, not the chemical transformation. This mattered for understanding fidelity because the kinetic proofreading mechanism depends on the relative rates of correct and incorrect nucleotide incorporation and discrimination happens during the chemistry step, not during product release. If you only look at steady-state kinetics, you might draw the wrong conclusion about where specificity comes from.

Computational approaches to understanding catalysis
Molecular dynamics simulations and quantum mechanics/molecular mechanics calculations have become essential tools for studying enzyme catalysis. You can simulate the motion of every atom in the enzyme and substrate over time and observe conformational changes that experiments can't capture directly. Free energy perturbation calculations can estimate binding affinities and reaction barriers with reasonable accuracy. But these methods have limitations. Force fields aren't perfect. Quantum mechanics calculations are expensive and often limited to small model systems. Sampling remains a fundamental problem. Enzyme motions span timescales from femtoseconds to seconds, and simulations typically cover nanoseconds to microseconds. Enhanced sampling methods help but introduce their own assumptions. I've found that the most reliable approach combines computational results with experimental validation. Use simulations to generate hypotheses, then test them with mutagenesis and kinetic measurements. The computational and experimental data feed each other and constrain the interpretation in ways that neither could achieve alone. Enzyme catalysis remains an active area of research despite centuries of study. New techniques continue to reveal details that challenge old models. The basic principles are well established, but the nuances are still being worked out. If you're approaching this topic, start with the fundamentals and build from there. Don't assume you understand something just because you memorized the textbook explanation. The real mechanisms are more interesting than the simplified versions.