Chair Conformations and Why You Should Care About Axial Bonds
When you're actually working with cyclohexane derivatives in the lab, the difference between a stable compound and a mess that won't crystallize often comes down to which conformation the molecule settles into. I used to draw out chair flips on napkins during grad school and still get tripped up on 1,3-diaxial interactions. It's a practical problem, not a theoretical one.
The Hidden World Of Chair Conformation How Axial Bonds Dictate Molecular Destiny
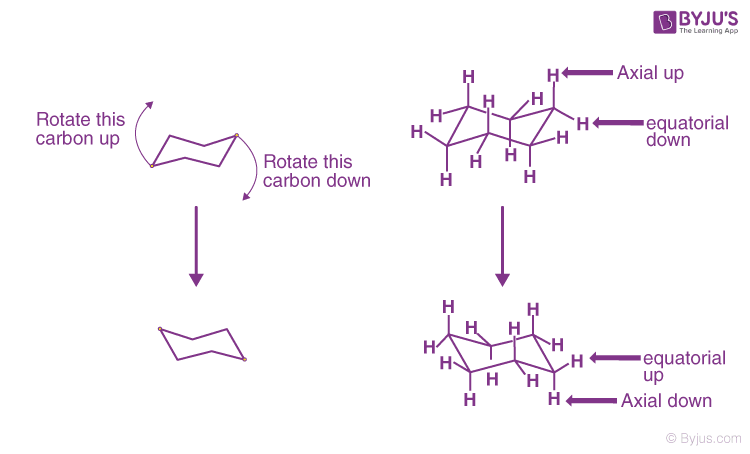
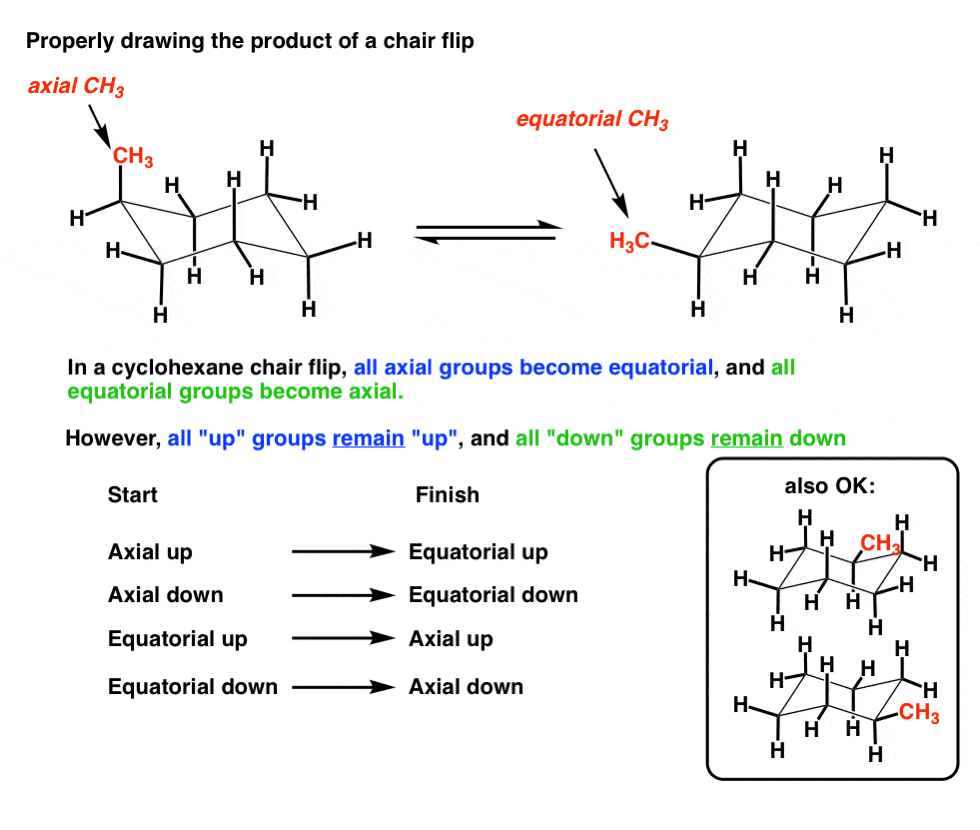
Start with the basics of what actually happens when a cyclohexane ring adopts its chair form. Six carbon atoms arrange themselves so the bond angles stay close to 109.5 degrees, which means you're looking at two types of C-H bonds on each carbon: axial bonds pointing straight up or down relative to the ring plane, and equatorial bonds pointing outward along the equator of the molecule. The axial bonds are the ones that matter most when you're predicting reactivity, because they create steric clashes with other substituents on the same face of the ring.I remember running a synthesis back in my second year where the major product just wouldn't form. I kept drawing the wrong chair conformation in my head. The substrate had a bulky tert-butyl group that should lock the ring in place, but I was underestimating how strongly it would prefer the equatorial position. Once I actually modeled the 1,3-diaxial strain from the axial hydrogens on carbons 3 and 5 bumping into the axial substituent, the whole picture changed. The reaction needed the ring in the less stable chair, and I'd been assuming it would just flip freely. That experiment sat on the bench for two weeks while I figured it out. Sometimes the molecule just doesn't want to do what you need it to do, no matter how clean your mechanism looks on paper. Here's the part most textbooks gloss over: the energy difference between axial and equatorial positions isn't uniform across all substituents. A methyl group costs about 1.74 kcal/mol when forced axial, but a hydroxyl group only costs roughly 0.9 kcal/mol. Hydrogen bonding can partially offset the 1,3-diaxial strain, which means an OH group sitting axially might actually be more stable than you'd expect from steric arguments alone. I've seen people apply A-values blindly to cases where intramolecular H-bonding completely flips the equilibrium. It doesn't always work that way, but you need to check for it. The ring flip itself is fast at room temperature, something like 10^5 flips per second, but bulky groups slow that down considerably. A di-tert-butylcyclohexane won't flip at all under normal conditions, which is useful when you need a conformationally locked substrate for a stereoselective reaction. However, if you're trying to predict the outcome of an elimination or a nucleophilic attack, you have to figure out which chair is actually populated at your reaction temperature, not just which one looks nicer on a test question.
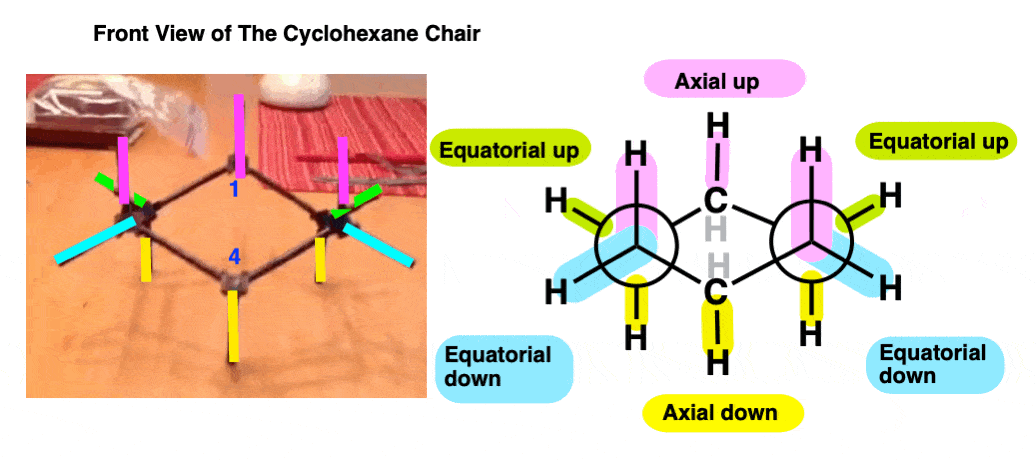
One thing that catches people off guard: the axial bonds on adjacent carbons actually point in opposite directions. Carbon 1 axial points up, carbon 2 axial points down, carbon 3 axial points up again. This is why 1,3-diaxial interactions are the real problem, not 1,2 interactions. The atoms on carbons 1 and 3 are both pointing toward the same face of the ring and sitting right next to each other in space, even though they're separated by one carbon in the connectivity. Visualize it as three vertical lines on the same side, with the middle one sandwiched between the other two. For practical work, when you're drawing these out for a synthesis problem or trying to explain a spectroscopic result, the A-value table is your first reference but not your last word. If you're dealing with a substituted cyclohexanone, the carbonyl changes the geometry slightly, and the axial preference shifts. If you're looking at NMR coupling constants to determine conformation, the diaxial protons on a cyclohexane ring show J values around 10-12 Hz, while axial-equatorial couplings are more like 2-5 Hz. That's usually enough to figure out which protons are axial in your actual compound without having to crystallize it or run a computational model. The main downside to relying purely on chair conformation analysis is that it assumes a static picture. In reality, molecules are vibrating and twisting, and at higher temperatures the population distribution shifts. A substituent that strongly prefers equatorial at room temperature might spend a significant fraction of time axial at 80 or 100 degrees Celsius. If your reaction is running hot, the conformer you'd never consider at ambient temperature could be the reactive species. I learned this the hard way with an elimination reaction that gave a completely different regioisomer when heated, which turned out to proceed through the higher-energy chair where the leaving group happened to be axial and anti-periplanar to the beta hydrogen.
Get the Full Details
If chair conformations aren't giving you enough resolution, there are better tools available now. Molecular mechanics calculations like MMFF94 or UFF can give you approximate energy differences in minutes on a laptop. For anything requiring accuracy beyond rough estimates, DFT at the B3LYP/6-31G* level will give you reliable barrier heights and conformational populations, though it takes longer and needs more setup. The old hand-drawn approach is still the fastest way to get a reasonable answer for simple substituted cyclohexanes, but it breaks down when you start adding heteroatoms, multiple stereocenters, or fused ring systems. The key takeaway is that axial bonds aren't just abstract drawing conventions. They create real steric pressure, influence reaction pathways, and determine which conformer dominates in solution. If you're working with any six-membered ring system and ignoring the chair flip equilibrium, you're probably missing something important about what's actually happening in the flask.