Understanding Enthalpy Changes in Chemical Reactions
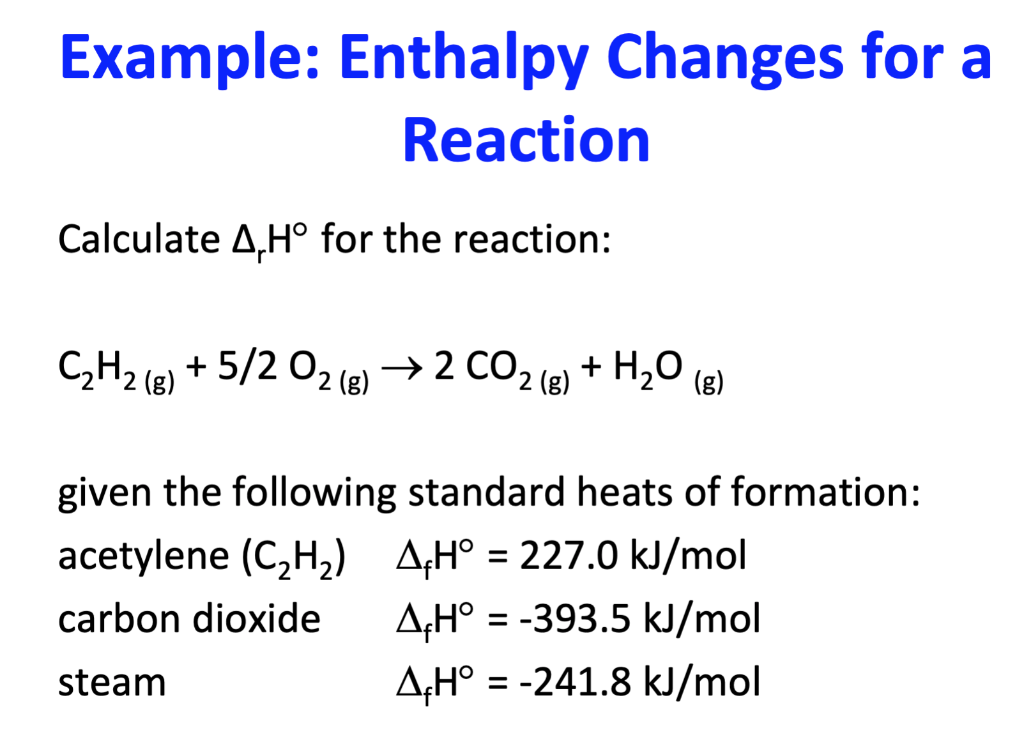
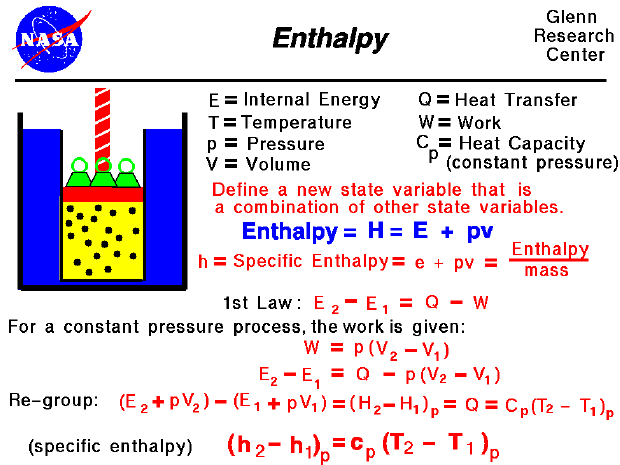
When I started working with thermochemistry back in the early days, most people treated enthalpy like it was some mystical concept reserved for textbook problems. The reality is far more practical. Delta H, the change in enthalpy, tells you whether a reaction releases or absorbs heat under constant pressure. That is the core of it. Getting the sign right matters more than memorizing every equation. This is probably the most natural way to frame the question if you are trying to understand what enthalpy actually does in practice. The enthalpy equation itself is deceptively simple: Delta H equals H products minus H reactants. But the complications start the moment you step away from ideal conditions. I learned this the hard way when running combustion calorimetry on a sample that turned out to have significant water content. My measured Delta H values were consistently 12 percent lower than literature values until I realized the energy was going into vaporizing water rather than just heating the surroundings. The equation you need depends on what data you actually have. Standard enthalpies of formation are the most common route. You look up Delta H f for each compound and apply the same subtraction principle. Formation enthalpies for elements in their standard states are zero by definition. This trips up students regularly because they forget that oxygen gas, hydrogen gas, and solid carbon in graphite form all carry zero values. Once you account for that, the calculation becomes straightforward arithmetic.
Working with Hess Law and Alternative Routes
Hess Law exists precisely because direct measurement is sometimes impossible or impractical. I once needed the enthalpy of formation for nitrogen monoxide. Direct synthesis from N2 and O2 at standard conditions gives negligible product. Instead, I used a cycle involving ammonia oxidation and nitric acid formation, combining three separate reactions with known enthalpies. The math took about ten minutes. The real value was recognizing which intermediate reactions would give me a clean path to the target. Here is a practical workflow I use. First, write out the target equation and identify what you need. Second, find reactions where your target compounds appear as reactants or products. Third, reverse reactions when necessary and remember to flip the sign of Delta H. Fourth, multiply entire equations by coefficients and scale Delta H proportionally. Fifth, add everything together and verify that intermediate species cancel completely. If they do not cancel, you missed a step or reversed something incorrectly.
Bomb Calorimetry and Constant Volume Measurements
Bomb calorimeters measure Delta U, the change in internal energy, not Delta H directly. Converting between the two requires accounting for the work done by gas expansion or compression. The relationship is Delta H equals Delta U plus Delta n gas times R times T. Delta n gas is the change in moles of gaseous species. I usually track this by counting gas moles on each side of the balanced equation. A common pitfall is assuming Delta n gas is zero when no gases appear in the equation. Combustion reactions produce CO2 and H2O, and H2O might be liquid or gas depending on the calorimeter temperature. In bomb calorimetry, water condenses to liquid because the system returns to initial temperature after the reaction. This means your product water count enters as liquid, and you should use the enthalpy of formation for liquid water, not gas. Using the gas value overestimates Delta H by about 44 kilojoules per mole of water formed. Another issue I encounter frequently involves the calibration constant. Bomb calorimeters require determination of the heat capacity of the entire system, including the bomb, water, and stirrer. Standard references like benzoic acid provide known combustion enthalpies for this purpose. If your calibration drifts because of changes in water volume or bomb condition, all subsequent measurements inherit that error. I recalibrate before each batch of samples and record the ambient temperature separately, since small variations affect the final result by a few tenths of a percent.
Get the Full Details

Temperature Dependence and Kirchhoff Equation
Enthalpy changes vary with temperature. The Kirchhoff equation quantifies this dependence: Delta H at T2 equals Delta H at T1 plus the integral of Delta Cp dT from T1 to T2. Delta Cp is the difference in heat capacities between products and reactants. If Delta Cp is approximately constant over your temperature range, the integral simplifies to Delta Cp times the temperature difference. In practice, Delta Cp rarely stays perfectly constant. Heat capacities increase with temperature for most substances, and the variation can be significant over wide ranges. I use tabulated Cp values as polynomial functions of temperature and perform numerical integration when accuracy matters. For rough estimates across narrow temperature windows, assuming constant Delta Cp usually introduces errors under 2 percent, which is acceptable for many applications.
Phase Changes and Latent Heat Contributions
Phase transitions carry their own enthalpy changes that must be included when substances change state during a reaction. Melting, vaporization, and sublimation all have characteristic enthalpies. Fusion enthalpies for most organic compounds fall between 5 and 30 kilojoules per mole. Vaporization enthalpies are typically larger, often 20 to 50 kilojoules per mole. A situation I deal with regularly involves reactions run at temperatures where a reactant or product crosses its phase transition point. Consider a reaction where a solid reactant melts before reacting. You must add the fusion enthalpy to your overall Delta H calculation, even though melting itself is not the chemical transformation. Skipping this step underestimates the total energy requirement. I always check the melting and boiling points of every species against the reaction temperature before committing to a calculation.
Common Mistakes and Practical Shortcuts
Sign errors remain the most frequent problem. When reversing a reaction, you must change the sign of Delta H. When multiplying a reaction by a coefficient, you must scale Delta H by the same factor. Writing the coefficient outside Delta H without multiplying creates incorrect values. I check signs by reading the equation aloud: exothermic reactions get negative Delta H values, endothermic reactions get positive values. If my calculated sign contradicts the physical behavior, I know something is wrong. Unit consistency causes problems more often than people admit. Standard enthalpies are usually reported in kilojoules per mole, but molar quantities depend on how you write the balanced equation. A reaction written with doubled coefficients has doubled Delta H. I verify that my final answer matches the stoichiometry I actually used, not the one I intended to write. Bond enthalpies offer a quick estimation method but come with important caveats. Average bond energies work reasonably well for gas phase reactions involving simple molecules. They become unreliable for condensed phases, reactions involving ionic species, or cases where resonance stabilization differs significantly between reactants and products. I use bond enthalpies for rough checks and prefer formation enthalpies for accurate work.
Real World Applications and Where Calculations Fall Short
Process engineers rely on enthalpy calculations for reactor design, heat exchanger sizing, and safety analysis. Adiabatic flame temperature predictions depend directly on accurate Delta H values. A 5 percent error in combustion enthalpy can shift predicted temperatures by over 100 kelvin, which affects material selection and thermal stress analysis. Biological systems present special challenges. Aqueous solutions, pH dependence, and coupled reactions make simple enthalpy calculations insufficient. I usually supplement thermodynamic data with empirical measurements when working with enzymatic reactions or metabolic pathways. Literature values for biochemical standard states assume pH 7 and specific ionic conditions, which differ from the conventional standard state at pH 0. Sometimes direct measurement beats calculation entirely. If you have access to a calorimeter and the reaction proceeds cleanly, measuring Delta H directly often produces better results than combining multiple literature values. Each combination introduces additional uncertainty from different experimental conditions and reference states. I recommend calculation as a first pass and measurement as confirmation whenever possible.
Computational Tools and Data Sources
NIST Chemistry WebBook provides reliable standard enthalpies of formation for thousands of compounds. The values come from peer-reviewed sources with documented experimental methods. Some entries carry higher uncertainty than others, so I check the confidence ratings before relying on them for critical calculations. Chemical engineering handbooks like Perry's and the CRC Handbook compile extensive thermochemical tables. For reactions under nonstandard conditions, tools like Aspen Plus or HSC Chemistry can perform equilibrium calculations that account for temperature and pressure effects simultaneously. These programs use embedded property databases and save time on iterative calculations, though I still verify key results by hand to catch database inconsistencies. Spreadsheet implementations of Hess Law cycles work well for routine calculations. I set up a template with reaction equations in one column, Delta H values in another, and coefficients that automatically scale the energies. Copying data from reliable sources into this template reduces transcription errors and makes it easy to update values as better measurements become available.