The Reality of Working With Charged Particles
Most people have never actually held an ion. They exist in plasmas, electrolytes, mass spectrometers, and the thin layer between your skin and a balloon you just rubbed on your hair. But they are real, measurable, and absolutely central to almost everything that moves in a battery, passes through a nerve, or gets detected in a lab.
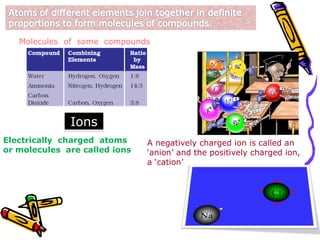
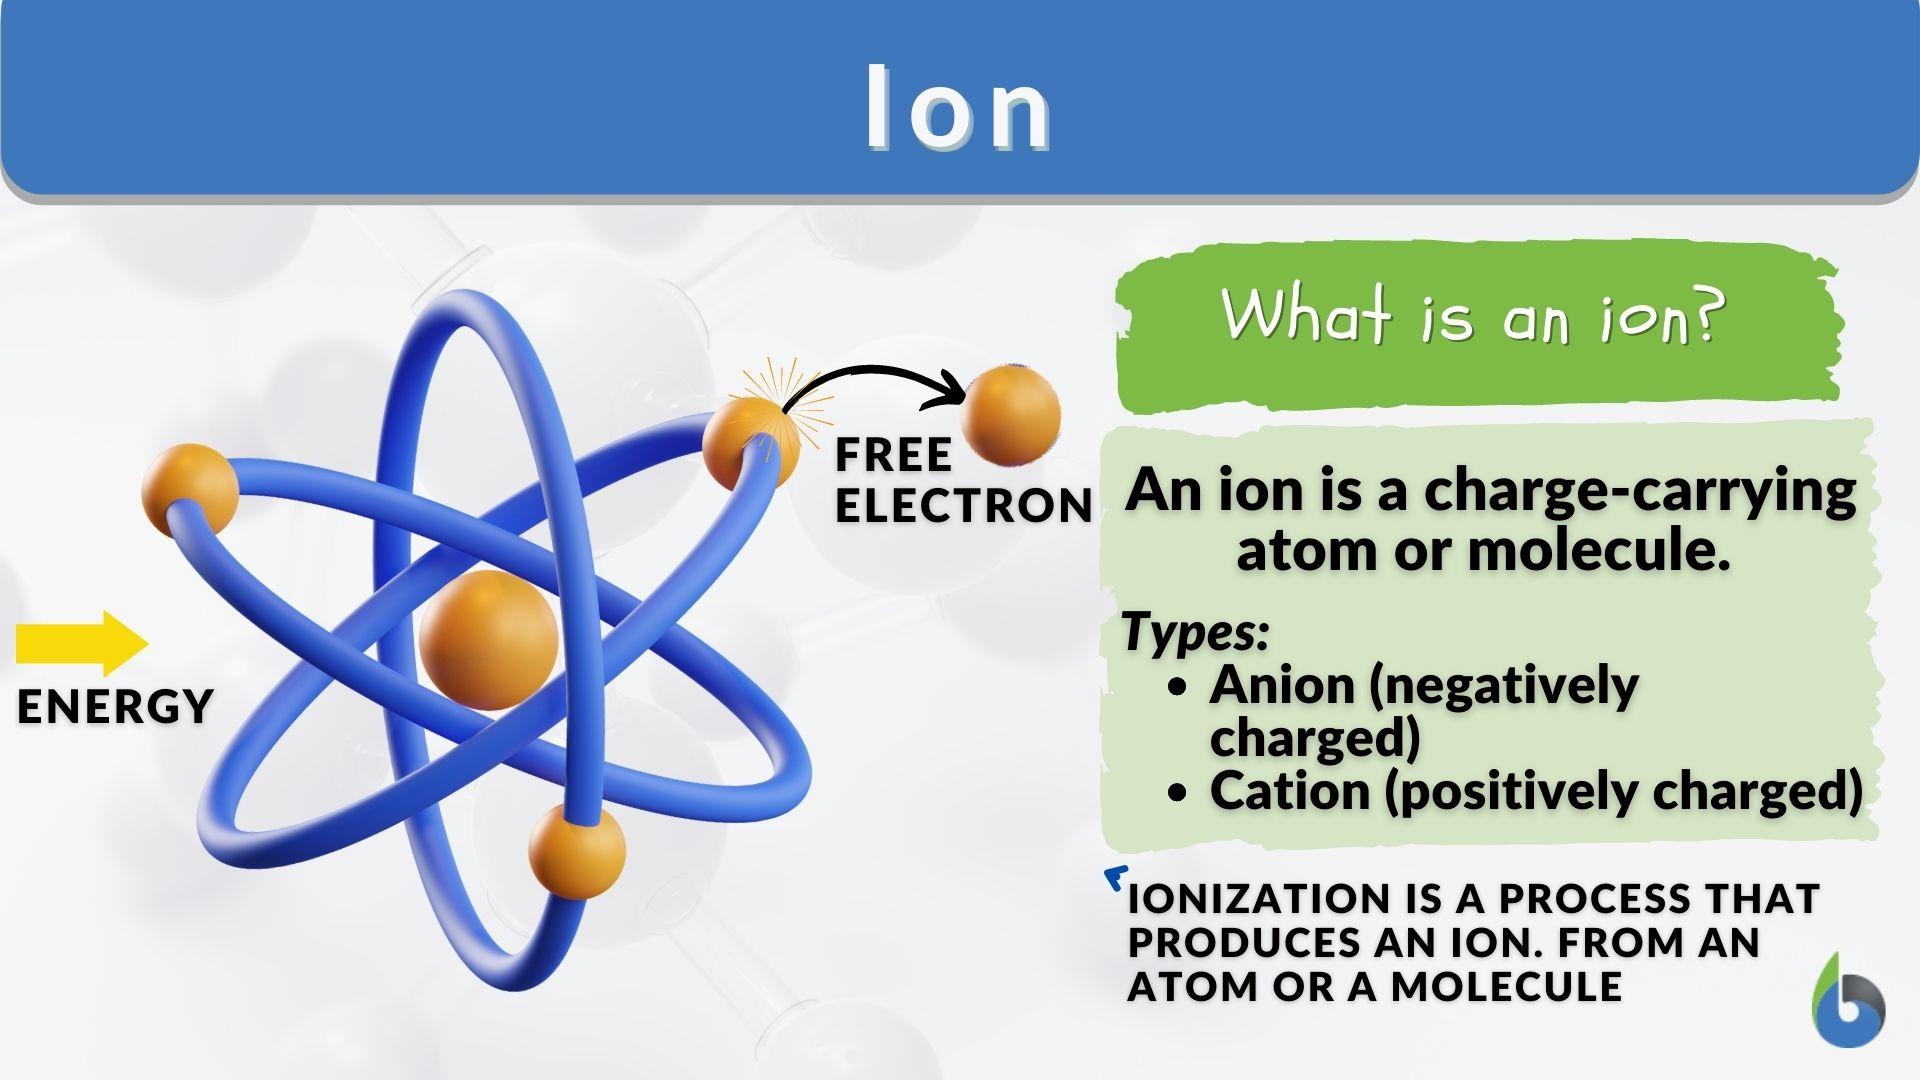
Ion is simply an atom or molecule that has gained or lost one or more electrons, giving it a net electrical charge. A sodium atom loses one electron and becomes Na+. A chloride atom gains one and becomes Cl-. That's it. Everything else is just applying that fact to different systems and watching what happens when those charged particles interact with electric fields, magnetic fields, solvent molecules, or other ions.
What Is An Ion Unlocking The Secrets Of Tiny Electrically Charged Molecules
I used to work in an analytical lab running LC-MS instruments for environmental samples. We were measuring trace pharmaceuticals in wastewater, and the problem wasn't that ions didn't work. The problem was that everything is an ion under the right conditions, and most of them don't want to be detected the way you need them to be.
One specific morning, I spent four hours troubleshooting why my target compound had zero signal. The column was fine. The mobile phase was fine. The standard was fine. It turned out the sample matrix contained high concentrations of phospholipids from the biological material, and those phospholipids were suppressing ionization in the electrospray source. They weren't just competing for charge. They were coating the capillary tip and physically blocking the spray. I couldn't fix it with more sensitive settings. I had to switch to a different sample prep method using solid-phase extraction with a C18 cartridge, then reconstitute in 5% methanol instead of the original aqueous buffer. Signal came back to normal immediately. That's the kind of thing you learn after burning through enough samples.
Here is what actually matters when you are dealing with ions in practice.
How Ions Behave in Real Systems
In solution, ions don't just sit there. They drag solvent molecules along with them, forming solvation shells. In water, small highly charged ions like Mg2+ pull in six water molecules in an octahedral arrangement and hold them tightly. Larger monovalent ions like K+ have looser hydration shells. This affects everything from reaction rates to how fast something moves through a membrane.
In the gas phase, things are completely different. There is no solvent shell. An isolated Na+ ion in a mass spectrometer vacuum is just a bare nucleus surrounded by a few electrons, and its behavior is governed purely by Coulombic attraction and repulsion. This is why mass spectrometry works. You can sort ions by their mass-to-charge ratio using magnetic and electric fields, and the trajectories are predictable to within a fraction of a percent if your instrument is calibrated properly.
When ions move through a medium, their mobility depends on their size, charge, and the viscosity of the medium. In electrophoresis, a charged protein will migrate toward the opposite electrode at a rate determined by its net charge divided by its frictional coefficient. Two proteins with the same molecular weight can separate completely if one has a slightly different surface charge distribution. That's the principle behind SDS-PAGE, except SDS gives everything the same charge density so separation is purely by size.
I have seen people treat ion mobility spectrometry as a magic solution for identifying unknown compounds. It is useful. It is not magic. IMS separates ions based on their collision cross-section with a neutral buffer gas, usually helium or nitrogen. The resolution is typically around 20 to 50, which means you can distinguish between some isomers that have identical mass spectra but different shapes. But if your sample is complex, overlapping drift times will still create ambiguity. I once had a mixture where three isobaric compounds had drift times within 0.3 milliseconds of each other. Without chromatographic separation upstream, they were indistinguishable. You always pair IMS with something else, usually liquid chromatography.
Common Misunderstandings That Cost Time and Money
The first big mistake is assuming ions are inherently unstable. Most ions in solution are perfectly stable. A chloride ion in seawater will sit there indefinitely unless it encounters something it reacts with. In the gas phase, some ions do decompose, but that's because you put energy into them during ionization, not because ions are fundamentally fragile.
The second mistake is thinking that higher charge always means better detection in electrospray. More charge states do increase sensitivity to a point, but they also increase the complexity of the spectrum. A protein that produces a clean +3 to +8 charge state envelope might give you better structural information than a single +2 peak, but deconvoluting that envelope requires proper software and validation. If you are trying to quantify a known compound and you force multiple charge states, you need to integrate all of them consistently across your calibration curve. Mixing charge states between your standards and your samples is a reliable way to get wrong concentrations.
The third mistake is ignoring the counterion. When you dissolve NaCl in water, you get Na+ and Cl-. The sodium doesn't act independently of the chloride in terms of solution chemistry. Activity coefficients, ion pairing, and ionic strength all depend on the complete ionic environment. If you are running electrochemical measurements and your reference electrode drifts, the problem is rarely the ion you are studying. It is usually the supporting electrolyte concentration or contamination from the counterion migrating through the salt bridge.
Practical Tips That Actually Help
If you are preparing ion-containing solutions for analysis, use high-purity water. Standard deionized water from a lab system typically has a resistivity of 18.2 Mohm-cm at 25 degrees Celsius, which corresponds to about 0.055 microsiemens per centimeter of conductivity from residual ions. If you are working at trace levels below one part per billion, that background can be significant. Some labs use sub-boiled distilled water for ultra-trace work, but the main benefit is reducing particulate contamination, not removing dissolved ions, since the distillation process itself doesn't eliminate volatile species like CO2 which forms carbonic acid and contributes carbonate ions.
For mass spectrometry, filter your samples through a 0.22-micrometer membrane before introduction. Particulates don't just clog the LC system. They can scatter the spray plume in ESI and create unpredictable ion suppression effects that vary from run to run. I stopped skipping this step after a batch of dirty samples produced inconsistent results for three days before I realized the filter was the issue.
If you are modeling ion transport in any kind of simulation, don't assume infinite dilution. The Nernst-Planck equation works well at low concentrations, but at higher ionic strengths you need activity corrections. The Debye-Hückel limiting law breaks down above about 0.01 molal, and the extended form helps until roughly 0.1 molal. Beyond that, you are either using empirical parameters or switching to a different model entirely. I've seen people run simulations at 1 molal ionic strength using the limiting law and then wonder why their predicted migration times were off by a factor of three.
For electroplating or battery electrolyte work, temperature control matters more than most people account for. Ion mobility in solution increases roughly 2 to 3 percent per degree Celsius. If your cell temperature fluctuates by five degrees between runs, your measured conductivity or transport number will shift by 10 to 15 percent. That is not instrument noise. That's real physics.
When Ions Simply Don't Work For Your Application
Non-aqueous solvents are a common failure point. If you are trying to dissolve an ionic compound in hexane or toluene, it isn't going to happen regardless of how much you stir or heat it. The dielectric constant of hexane is about 1.9 compared to 78.4 for water. Ions need a high dielectric medium to separate from their counterions and remain solvated. Some ionic liquids can dissolve in lower dielectric solvents, but they tend to form ion pairs and aggregates rather than free ions, which changes their behavior entirely.
Gas-phase ion chemistry also has hard limits. In a standard quadrupole mass spectrometer operating at 10^-6 torr, the mean free path is long enough that ions travel in straight lines between collisions with residual gas molecules. But if you introduce too much sample or the vacuum degraded, collisions become frequent enough that ion optics no longer work predictably. You'll see peak broadening, loss of resolution, and unstable transmission. There is no software fix for a bad vacuum. You replace the pump oil, check for leaks, and run a tune standard to confirm performance.
Another scenario where ion-based methods fail completely is when your analyte cannot be ionized efficiently under any practical condition. Some fully fluorinated compounds, certain polymers, and highly symmetric hydrocarbons produce virtually no signal in electrospray or atmospheric pressure chemical ionization. You can try harder ionization methods like electron impact, but then you risk fragmenting the molecule before you can detect it. Matrix-assisted laser desorption ionization helps for large biomolecules, but it introduces its own set of problems with background noise and reproducibility. Sometimes the right answer is a non-ion-based technique like GC with flame ionization detection, which responds to carbon-hydrogen bonds rather than charged species.
The bottom line is that ions are straightforward conceptually and messy in practice. Understanding the basics takes a weekend. Understanding how they behave in your particular setup takes months of running controls, checking for interferences, and learning which assumptions are actually valid for your samples.