What Alkyl Groups Actually Are and How They Show Up In Real Reactions
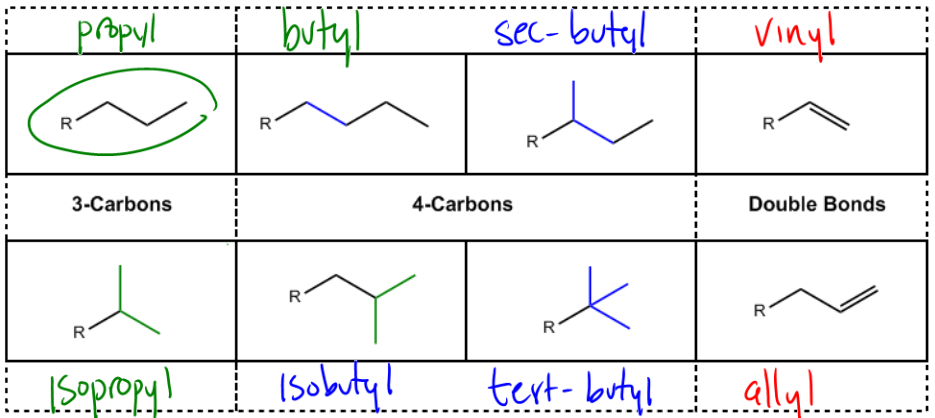
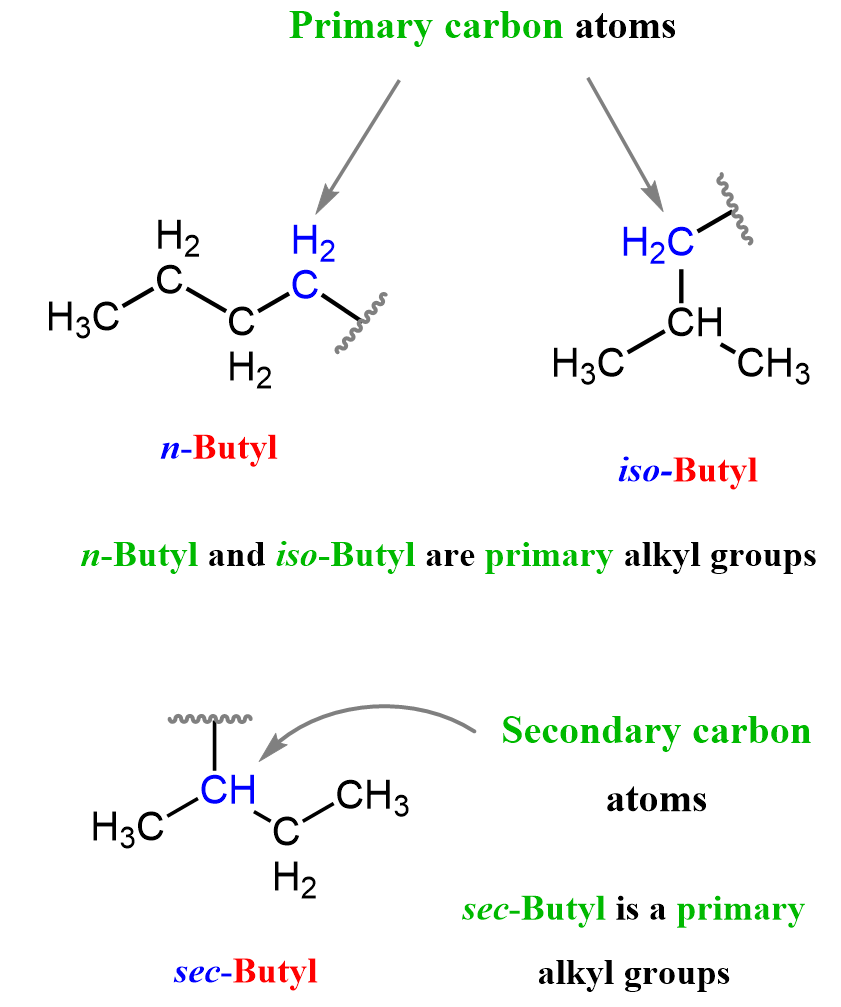
An alkyl group is what you get when you remove one hydrogen from an alkane. That is the textbook definition, and it is technically correct, but it does not prepare you for how messy these things behave once you are actually running reactions in a lab. I spent years watching people treat alkyl groups as if they were interchangeable placeholders when, in practice, the size and branching of an alkyl substituent can completely change reaction outcomes, selectivity, and sometimes whether a reaction works at all. When you strip a hydrogen from methane you get methyl, CH<3>. Remove one from ethane and you get ethyl, CH<2>CH<3>. Propane gives you n-propyl or isopropyl depending on which carbon loses the hydrogen. Butane gets interesting fast, giving you n-butyl, sec-butyl, isobutyl, and tert-butyl. Each of these has a different shape, different steric profile, and different electronic effect even though they are all technically just carbon chains attached to something else. The way alkyl groups are introduced into molecules matters enormously. Alkylation reactions, whether they are Friedel-Crafts alkylations, SN2 displacements, enolate alkylations, or transition-metal-catalyzed C-H functionalization, all rely on alkyl electrophiles or nucleophiles. The reagents you pick, the leaving groups, the solvent, the temperature, these all interact with the alkyl structure in ways that are not always obvious from a reaction scheme.
I once ran a simple benzylic alkylation on an electron-rich aromatic substrate using an alkyl bromide that should have been straightforward. The alkyl halide was 1-bromo-2,2-dimethylpropane, commonly called neopentyl bromide. On paper this is a primary halide, so SN2 should work fine. In practice, neopentyl systems are notoriously unreactive in SN2 conditions because the two methyl groups right next to the reaction center create a massive steric wall. I tried standard conditions with a base and a nucleophile, heated it, cooled it, tried different solvents, and got almost nothing but starting material and some elimination byproducts. It took me three days before I stopped trying to force the SN2 pathway and switched to a radical-based approach using a tin hydride reduction with AIBN initiation. That gave the product in decent yield in about four hours. The lesson was not that neopentyl halides cannot be alkylated, it was that I had been applying the wrong mechanism to a substrate that actively resists that mechanism.
How Alkyl Groups Affect Reactivity Beyond the Basics
Most introductory courses teach that alkyl groups are electron-donating through inductive effects and hyperconjugation. This is true but incomplete. The electron donation from an alkyl group stabilizes carbocations, which is why tertiary carbocations form more readily than primary ones. But the same electron-donating character also affects nucleophilicity, basicity, and the stability of radical intermediates in ways that are easy to miss if you are only thinking about one reaction type. Here is something people often get wrong. A bulkier alkyl group does not always slow down a reaction just because of sterics. In some cases, a bulky alkyl group like tert-butyl can actually accelerate certain steps by preventing unwanted side reactions or by locking a molecule into a conformation that favors the desired pathway. I have seen this clearly in ester hydrolysis reactions where a tert-butyl ester serves as a protecting group specifically because the bulk around the carbonyl makes nucleophilic attack difficult under mild conditions. The same bulk that would block an SN2 reaction becomes an advantage when you want controlled deprotection. Hyperconjugation deserves more attention than it gets. When an alkyl group is adjacent to a double bond or a positively charged center, the C-H sigma bonds can donate electron density into the empty or antibonding orbital. This is not just theory. It explains why Zaitsev products dominate in elimination reactions and why certain carbocations rearrange the way they do. The more alpha hydrogens an alkyl group has, the stronger its hyperconjugative stabilization. A methyl group has three, a methylene has two, and a methine has one. This difference matters when you are predicting regioselectivity in reactions like E2 eliminations or electrophilic additions to alkenes.
Get the Full Details
Common Mistakes When Working With Alkyl Substituents
One of the most frequent errors I see is assuming that all alkyl halides of the same class will behave identically. Primary alkyl halides are generally considered good SN2 substrates, but allylic and benzylic primary halides are in a different category entirely. They react much faster in SN2 conditions because the adjacent pi system stabilizes the transition state. At the same time, they are also more prone to SN1 reactions than typical primary halides would be. If you are designing a synthesis and you assume reactivity based purely on primary versus secondary versus tertiary classification, you will make mistakes. Another pitfall involves alkyl groups on aromatic rings in electrophilic aromatic substitution. Methyl and other small alkyl groups are ortho-para directors, which is standard textbook knowledge. But the ratio of ortho to para product is not always what beginners expect. For toluene nitration, the ortho and para products form in roughly equal amounts, but as the alkyl group gets bulkier, the ortho position becomes increasingly blocked. With tert-butylbenzene, para substitution dominates almost completely because the tert-butyl group is too large to allow significant ortho attack. This is a practical consideration that affects yield and purification time. Enolate alkylation is another area where alkyl group choice can make or break a reaction. When you deprotonate a ketone to form an enolate and then add an alkyl halide, the choice of alkyl halide determines whether you get clean monoalkylation or a mess of polyalkylation and elimination products. Methyl iodide and ethyl iodide tend to work well. Larger primary alkyl halides like propyl and butyl still work but slower. Secondary alkyl halides are risky because elimination competes strongly. Tertiary alkyl halides basically never work in enolate alkylation because E2 elimination is the only pathway available. I learned this the hard way when a grad student in my group tried to use isopropyl bromide with a lithium enolate and spent two days purifying a reaction that was mostly propene and unreacted starting material.
Practical Approaches For Alkylation Reactions
If you need to install an alkyl group on a molecule, the first decision is which carbon you are alkylating and what mechanism suits that position. For sp3 carbons in aliphatic systems, SN2 with a primary alkyl halide is the go-to approach. Use a polar aprotic solvent like DMF or acetonitrile, pick a strong nucleophile, and keep the temperature moderate to avoid elimination. For sp2 carbons in aromatic systems, Friedel-Crafts alkylation works but has significant limitations including carbocation rearrangement and polyalkylation. Friedel-Crafts acylation followed by reduction is often the better route if you need a specific alkyl chain without rearrangement. For alkylation at alpha carbons of carbonyl compounds, consider whether you need kinetic or thermodynamic enolate control. LDA at low temperature gives the kinetic enolate, which is the less substituted one. A weaker base at higher temperature gives the thermodynamic enolate, which is the more substituted one. The alkyl halide you add will react with whichever enolate you generate, so the choice of base and conditions directly controls where the alkyl group ends up on your molecule. Transition metal-catalyzed cross-coupling reactions have expanded what is possible with alkyl groups significantly. Modern methods using nickel or palladium catalysts can couple alkyl halides or alkyl boron reagents with aryl or vinyl partners. These reactions handle secondary and even primary alkyl groups that would fail in traditional SN2 conditions. The key is choosing the right ligand and catalyst system. Buchwald-type phosphine ligands work well for alkyl palladium coupling, and nickel catalysts with bipyridine or N-heterocyclic carbene ligands are effective for alkyl-alkyl cross couplings. These methods have made alkylation far more reliable than they were ten or fifteen years ago.
Limitations And When Alkylation Strategies Fail
Not every alkylation you attempt will work, and knowing when to stop trying the same conditions is part of the skill. Alkylation of hindered nucleophiles is one area where standard approaches consistently fail. If your nucleophile is surrounded by bulky groups, even a methyl halide might react extremely slowly or not at all. In these cases, switching to a more reactive electrophile like a triflate or using a phase transfer catalyst can help, but sometimes the only solution is to redesign the synthetic route to introduce the alkyl group at an earlier stage when steric congestion is lower. Polyalkylation is another persistent problem, especially in Friedel-Crafts reactions and amine alkylation. Once you add an alkyl group to an aromatic ring or an amine, the product is often more reactive than the starting material, so the reaction continues past the point you want. This is why excess starting material is typically used in Friedel-Crafts alkylation, and why reductive amination is generally preferred over direct alkylation for amines when you want a specific degree of substitution. Carbocation rearrangement remains a serious issue in reactions that proceed through carbocation intermediates. If your alkyl group can migrate to form a more stable carbocation, it will, and you will get a product that does not match your intended structure. This happens frequently in Friedel-Crafts alkylation with propyl or butyl halides, where the initially formed primary carbocation rearranges to a secondary one before the electrophilic attack occurs. The workaround is usually to use acylation instead of alkylation, or to employ a directing group strategy that controls regiochemistry.

Recognizing Alkyl Groups In Spectroscopic Data
When you are analyzing a compound, identifying alkyl substituents comes down to interpreting NMR and IR data with an understanding of what different alkyl environments look like. In proton NMR, methyl groups typically appear as singlets, doublets, or broad singlets depending on their neighbors, usually in the 0.7 to 1.5 ppm range for simple alkyl chains. Methylenes and methines appear slightly downfield, around 1.2 to 1.8 ppm, and any alkyl group adjacent to an electronegative atom or pi system shifts further downfield. Carbon NMR is more straightforward for distinguishing alkyl types. Primary carbons show up around 10 to 30 ppm, secondary carbons around 20 to 45 ppm, tertiary carbons around 30 to 50 ppm, and quaternary carbons around 30 to 40 ppm. The exact values depend on what the alkyl group is attached to, but the general trend holds. A tert-butyl group, for example, gives a distinctive pattern in both proton and carbon NMR because all nine methyl protons are equivalent and the quaternary carbon is clearly visible in the carbon spectrum. IR spectroscopy can confirm the presence of alkyl groups through C-H stretching vibrations. Sp3 C-H stretches appear just below 3000 cm-1, typically around 2850 to 2960 cm-1, while sp2 C-H stretches appear just above 3000 cm-1. The bending vibrations of methyl and methylene groups show up in the fingerprint region around 1375 and 1450 cm-1 respectively. A strong doublet near 1375 cm-1 is characteristic of gem-dimethyl groups, which is useful for identifying isopropyl or tert-butyl substituents quickly.
Understanding alkyl groups goes beyond memorizing names and structures. The practical reality is that every alkyl substituent carries steric, electronic, and conformational consequences that affect how a molecule behaves in every reaction it participates in. The difference between a reaction that works cleanly and one that produces a dozen byproducts is often the difference between a straight-chain alkyl group and a branched one, or between a primary and a secondary carbon at the reaction site. Paying attention to these details saves time, reagents, and a lot of unnecessary purification work.