Silver’s Oxidation State: What You Actually Need to Know
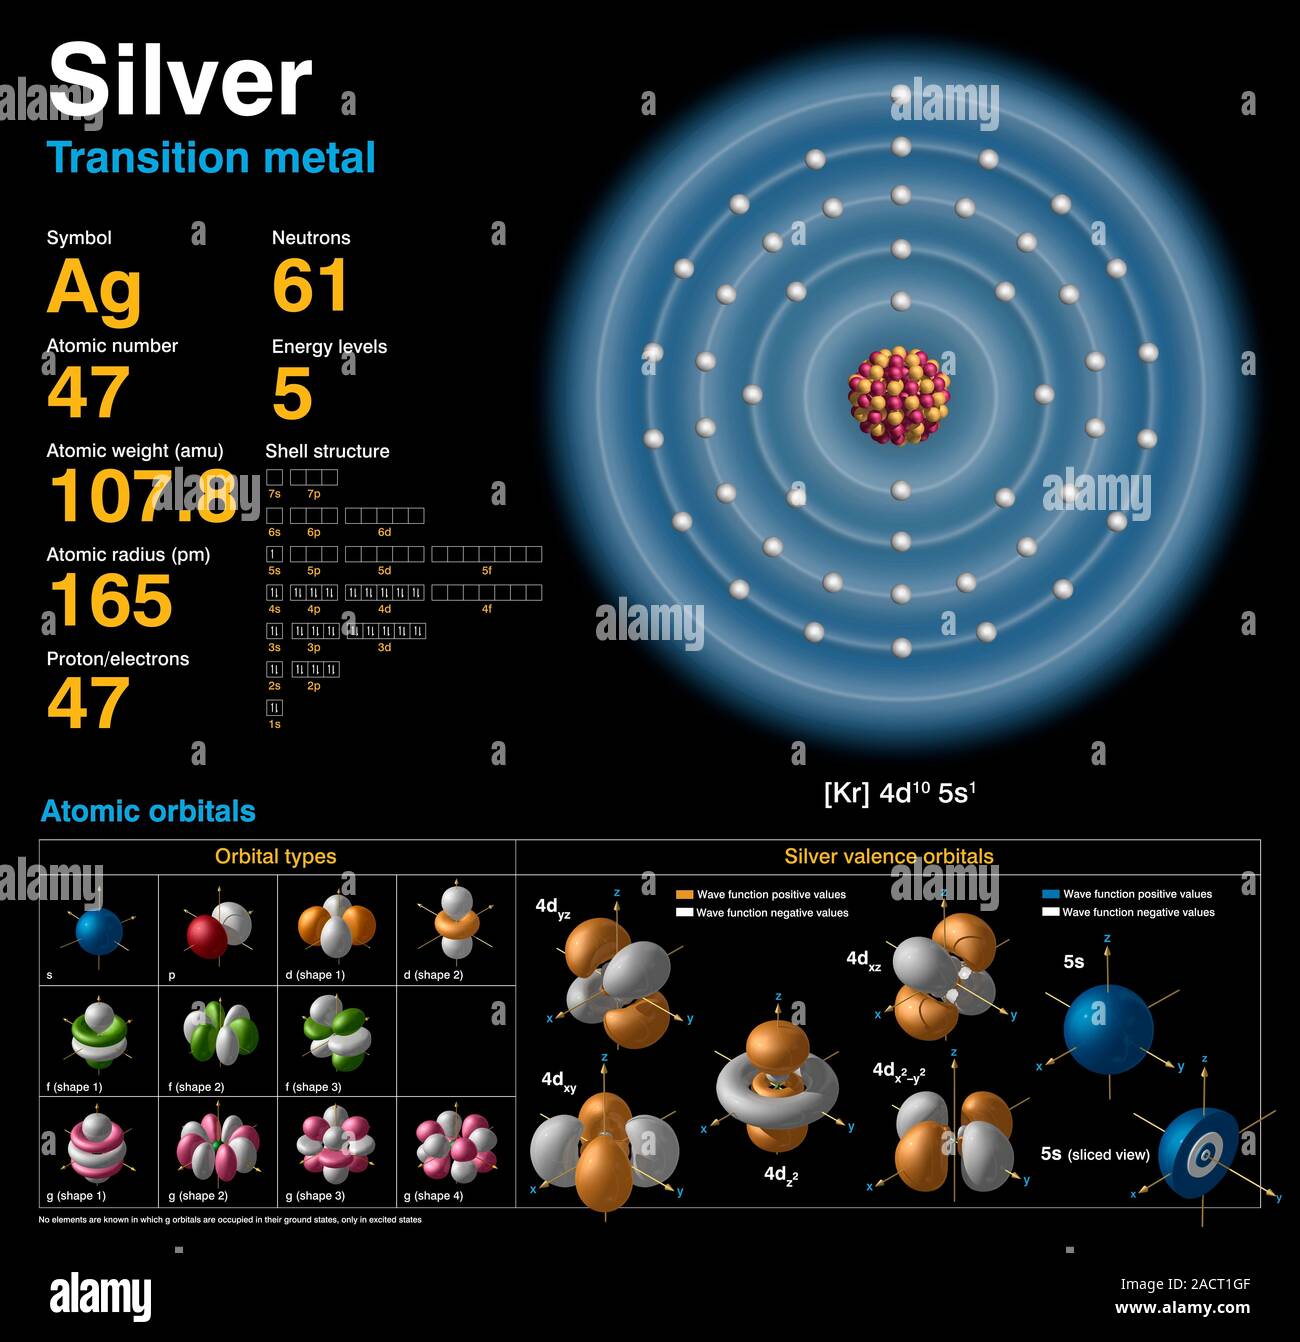
Silver most commonly forms a +1 charge, but the chemistry around it is messier than introductory textbooks suggest. If you’ve ever tried to precipitate silver halides or run a copper-silver electrochemical cell, you’ve probably noticed that things don’t always behave the way the periodic table promises. Let’s look at what’s actually happening in solution, what charges silver can take, and why your lab notebook might be lying to you. In the vast majority of everyday reactions, silver exists as Ag+. That’s its stable oxidation state in aqueous solution, and it’s the one you’ll see in silver nitrate, silver chloride, and the classic Tollens’ reagent. The electron configuration makes sense for this: silver’s ground state is [Kr] 4d10 5s1, so losing that single 5s electron gives you a filled d-subshell, which is relatively happy. This isn’t a theoretical curiosity. I’ve seen students fail titrations because they wrote Ag2+ in a redox equation, which doesn’t exist under normal aqueous conditions. Stick with +1 for 99% of your undergraduate work. But here’s where it gets annoying. Silver does have documented +2 and even +3 states, usually stabilized by strong oxidizing ligands like fluoride or peroxo groups. AgF2 contains Ag2+, and K3AgF6 contains Ag3+. These aren’t common lab chemicals. They’re powdery solids that decompose aggressively in moisture and require anhydrous conditions to handle. If you order a bottle of silver fluoride from a supplier, double-check the catalog. Some sell AgF (which is just Ag+ again), and some sell AgF2. Mixing them up will ruin your experiment fast.
The most counter-intuitive thing I encountered involved a routine silver recovery process from photographic waste. We were using sodium thiosulfate to dissolve unexposed AgBr, then adding zinc dust to precipitate metallic silver. The solution turned unexpectedly clear, and the yield dropped to almost nothing. Turns out the pH had drifted too low, and the silver was forming soluble Ag(S2O3)23- complexes instead of precipitating. Adjusting the pH back above 8 with a small amount of sodium hydro carbonate fixed it immediately. I’d spent three hours troubleshooting before I realized the complex ion equilibrium was the real problem, not the zinc reduction itself. This is the kind of thing you learn from burning through reagents, not from reading a summary table.
Why Silver’s +1 State Is So Reliable
The filled 4d10 subshell after losing the 5s electron gives Ag+ a pseudo-noble-gas configuration. It’s not as stable as a true noble gas, but it’s enough that the +1 state dominates in water. The standard reduction potential for Ag+ + e- Ag(s) is +0.80 V, which is high. That means Ag+ is easily reduced to metallic silver, which is why it’s used in so many analytical and electroplating applications. But that same high potential also means Ag+ won’t spontaneously disproportionate or oxidize further in most aqueous environments. You need external oxidizers to push it past +1. A practical implication: if you’re preparing a silver mirror reaction, the pH matters enormously. Tollens’ reagent requires basic conditions because the diamminesilver(I) complex, [Ag(NH3)2]+, only forms and stays stable in ammonia solution. If you acidify it, the ammonia protonates, the complex falls apart, and you get a precipitate of Ag2O or even metallic silver depending on the reducing agent present. I once neutralized a waste beaker too quickly with dilute HCl and got a sludge that took a week to clean out of the glassware. Slow addition, room temperature, and good stirring are not optional here.
Get the Full Details
Pitfalls That Cost Time and Money
The biggest mistake people make is assuming silver compounds follow the same solubility rules as other transition metals. Silver sulfate is moderately soluble, silver acetate is sparingly soluble, and silver carbonate decomposes before it dissolves. None of these behave like, say, zinc or iron salts. If you’re doing qualitative analysis and you see a white precipitate with HCl, don’t immediately conclude it’s AgCl without testing for mercury(I), which also precipitates as a white chloride. The solubility product of AgCl is 1.8 × 10⁻¹⁰, but Hg2Cl2 is 1.3 × 10⁻¹⁸, so they can co-precipitate and confuse an inexperienced analyst. Another issue that comes up in industry is silver passivation during electroplating. You’d think higher current density gives faster deposition, but silver deposits as a rough, dendritic mess past about 0.5 A/dm2 in most cyanide-free baths. The bath composition dictates everything. Citrate-based baths give smoother deposits at slightly higher currents but are sensitive to organic contamination. I once calibrated a plating line using old parameters from a different bath type and ended up with a brittle coating that flaked off after thermal cycling. Two days of rework. The fix was lowering the current density by half and adding a wetting agent to suppress hydrogen evolution at the cathode. There’s also the photochemical decomposition problem. Silver halides darken on exposure to light, which is useful for photography but terrible for storage. If you’re working with AgBr or AgI in solution and your yields are inconsistent, check whether your glassware is clear or amber. UV light drives Ag+ reduction to Ag0, and you lose active material to colloidal silver that won’t precipitate cleanly. Amber bottles and low-light conditions aren’t aesthetic choices. They’re necessary.
When +1 Isn’t Enough
If your application requires oxidative silver chemistry, you’re working with niche reagents and non-standard conditions. The +2 state appears in AgO, but even that compound is structurally complex. X-ray studies show it’s better described as AgI AgIII O2, containing both +1 and +3 silver ions in a mixed-valence lattice. Calling it “silver(II) oxide” is convenient shorthand, but it’s chemically inaccurate. This matters if you’re using AgO as an oxidant in organic synthesis, because the reactivity comes from the AgIII center, not a simple d9 Ag2+ ion. For most practical purposes, whether you’re balancing equations, running precipitation reactions, or setting up an electrochemical cell, treating silver as Ag+ is correct. The exceptions are specialty materials research, certain catalytic processes, and high-oxidation-state chemistry that requires fluorinating agents or electrochemical anodization in acidic media. Don’t let the existence of exotic oxidation states confuse the basics. The +1 charge is the answer to almost every question you’ll encounter in a standard chemistry curriculum or laboratory setting. If you need a reference table for silver compounds, solubility products, or standard potentials, the NIST Chemistry WebBook and the CRC Handbook of Chemistry and Physics are the most reliable sources I’ve found. Peer-reviewed journals get published data wrong occasionally, but those two don’t. They’re dry, they’re expensive, and they’re worth it.