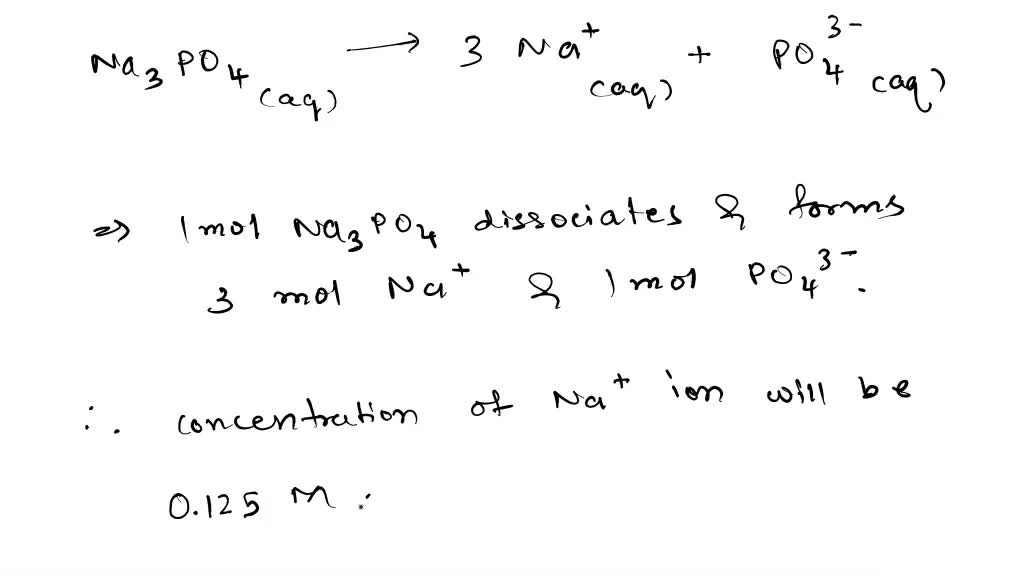
Ion Speciation in Sodium Phosphate Solutions
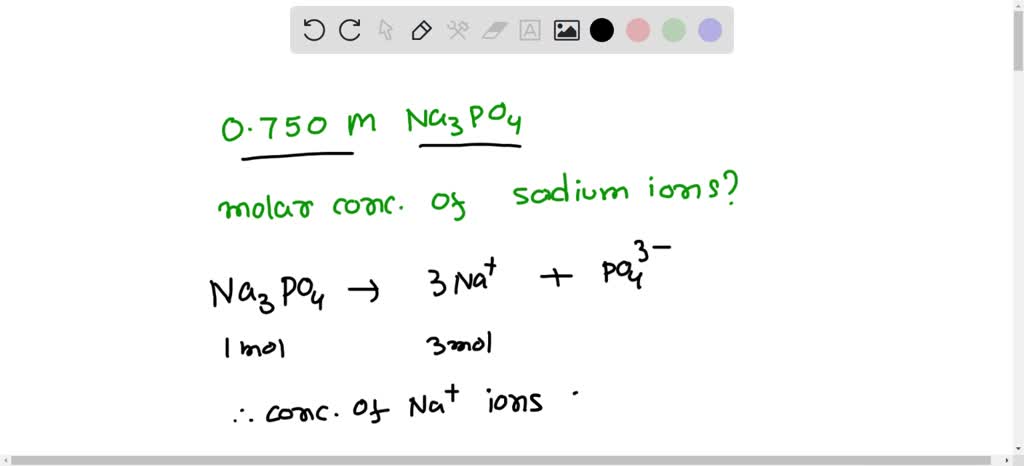
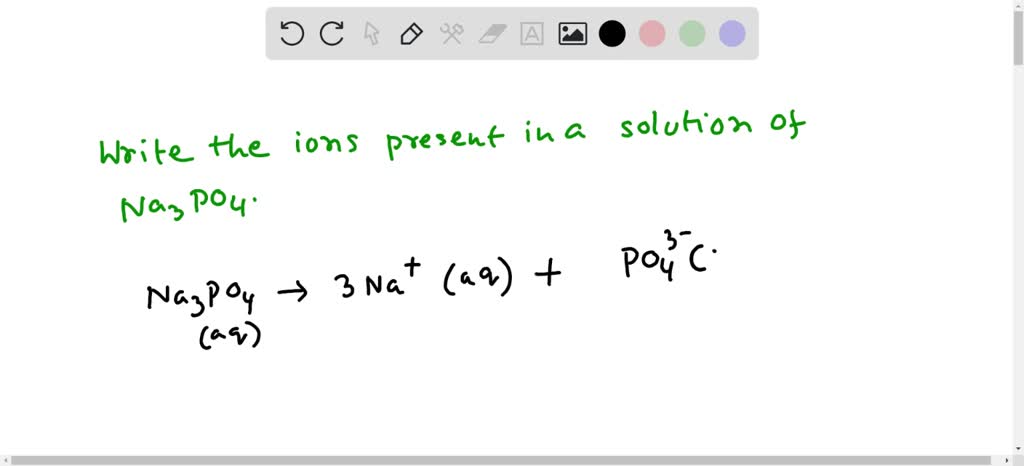
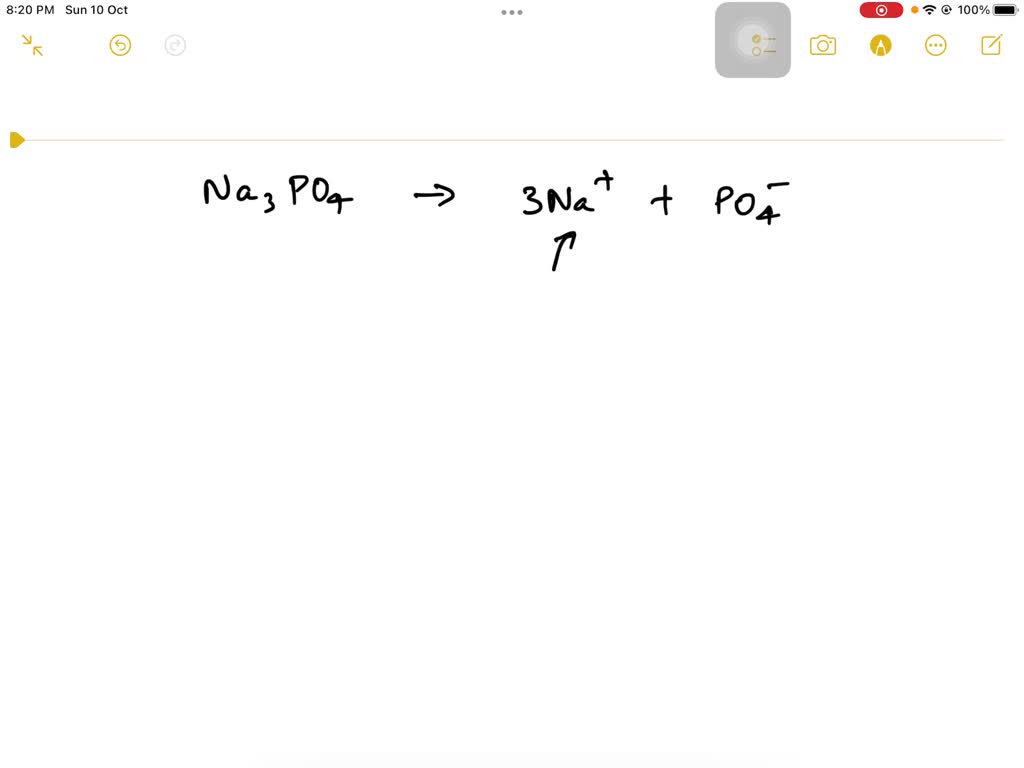
Sodium phosphate is one of those salts where the straightforward answer hides a bunch of equilibria you need to account for if you are doing anything that requires precision. Let me walk through how the dissociation actually plays out in water. When you dump Na3PO4 into water, the first thing that happens is complete dissociation of the ionic lattice. That part is simple and non-negotiable. You get three sodium cations and one phosphate anion for every formula unit that dissolves. The sodium ions just hang out. They are spectator ions in almost every practical sense. They do not hydrolyze. They do not form complexes at normal concentrations. You can treat them as fully free Na+ for any calculation that involves pH or precipitation.
Where things get interesting is the phosphate ion. PO4^3- is a strong base in water. It immediately starts pulling protons from the solvent. This is where the speciation drama begins.
Stepwise Protonation Equilibria
The phosphate system undergoes three successive protonation steps, each with its own equilibrium constant. I remember running into trouble with this back when I was doing buffer work for an enzyme assay. I prepared what I thought was a clean 0.1 M sodium phosphate buffer at pH 7.2, and the actual speciation was nowhere near what the textbook Henderson-Hasselbalch equation predicted. The issue was that at moderate concentrations, the simple approximation breaks down because you cannot ignore the second and third dissociation steps. Here are the equilibria: PO4^3- + H2O HPO4^2- + OH- with Kb1 = Kw/Ka3
Get the Full Details
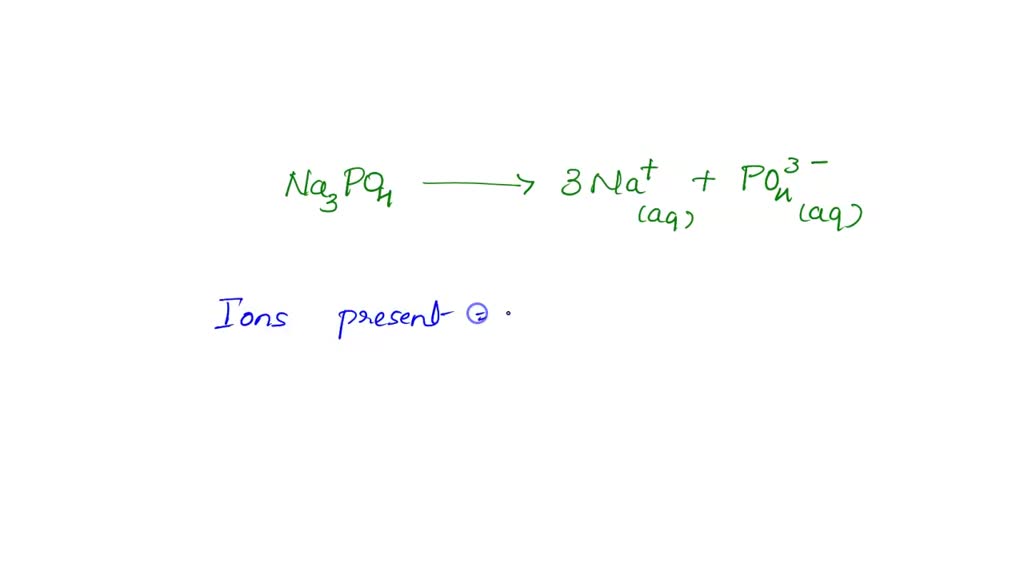
HPO4^2- + H2O H2PO4- + OH- with Kb2 = Kw/Ka2 H2PO4- + H2O H3PO4 + OH- with Kb3 = Kw/Ka1 The pKa values for phosphoric acid are approximately 2.15, 7.20, and 12.35 at 25 degrees Celsius. That means Ka3 is about 4.5 × 10^-13, making PO4^3- a quite strong conjugate base.
When I finally solved the full system of equations numerically instead of using the simplified approach, I discovered that my calculated pH was off by nearly 0.4 units. The workaround was straightforward: I set up the charge balance and mass balance equations and solved them iteratively. It took about five minutes in a spreadsheet compared to the ten minutes I was wasting on wrong calculations.
Major Ionic Species at Different pH Ranges
At very high pH, above 12, the dominant phosphate species is indeed PO4^3-. This is the region where your simple dissociation equation actually holds some water. The concentration of HPO4^2- is negligible here because there are essentially no free protons to protonate the phosphate. As you drop the pH toward 12.35, which is the pKa3 value, you enter the transition zone where PO4^3- and HPO4^2- exist in comparable amounts. This is a notoriously difficult region to buffer because the pKa is so far up. Most people who try to work in this pH range end up using strong base like NaOH to adjust, which just adds more sodium ions to the mix. Around pH 7.20, which is pKa2, the system shifts again. HPO4^2- and H2PO4- become the dominant species. This is the classic phosphate buffer region, and it is widely used in biochemistry. The buffer capacity here is reasonable, though you need to account for the fact that your total phosphate concentration affects the actual speciation.

Below pH 7, H2PO4- starts dominating. At pH 2.15, you transition into the H3PO4 region. Below pH 2, phosphoric acid itself is the main species, though in practice you are dealing with strongly acidic conditions where other reactions might interfere.
Countering the Simplistic View
Here is something beginners miss: the naive assumption that Na3PO4 simply gives you PO4^3- and Na+ ions is only valid at extremely high pH. In any practical aqueous solution, you must account for the hydrolysis of the phosphate ion. The actual distribution of species depends heavily on concentration, temperature, and the presence of other ions. Another counter-intuitive point is that ionic strength matters more than people expect. At high salt concentrations, activity coefficients deviate significantly from unity. I once prepared a 1 M sodium phosphate solution and found that the measured pH was 0.3 units lower than the calculated value. The fix was to use activity coefficients from the Davies equation rather than assuming ideal behavior. Temperature is another factor that gets overlooked. The pKa values shift with temperature. At 37 degrees Celsius, which is physiological temperature, pKa2 drops to about 6.8. This means phosphate buffers that work well at room temperature perform quite differently at body temperature. If you are doing biological work, you need to recalibrate your pH meter at the working temperature.
Common Pitfalls in Calculations
The most frequent error I see is treating the first hydrolysis step in isolation and ignoring the subsequent protonation equilibria. At 0.1 M Na3PO4, the pH is approximately 12.7, not the 13.5 you would calculate if you only considered the first Kb. The second hydrolysis step contributes enough OH- to make a meaningful difference. Another trap is assuming that the sodium ions do not participate in anything. In concentrated solutions, ion pairing between Na+ and HPO4^2- can form. The association constant is small but not negligible. For high-precision work, you need to consider these interactions. There is also the issue of carbon dioxide absorption. Open phosphate solutions gradually absorb CO2 from the air, which forms carbonic acid and shifts the pH. I learned this the hard way when a stored 0.5 M Na3PO4 solution showed a pH drift of 0.2 units over two weeks. The solution was to store under inert atmosphere or use freshly prepared solutions for sensitive work.
Practical Implications
If you are using sodium phosphate for precipitation reactions, the speciation determines which phosphate salt precipitates. At high pH, you might get Na3PO4 precipitation if the concentration is high enough. At neutral pH, you are more likely to see Na2HPO4 or NaH2PO4 depending on the exact conditions. For analytical chemistry applications, the ionic composition affects conductivity measurements, electrode responses, and chromatographic behavior. I once had trouble with ion exchange chromatography because I did not account for the fact that my phosphate sample contained significant amounts of H2PO4- rather than pure PO4^3-. The retention times were completely wrong until I adjusted the sample preparation. In industrial settings, sodium phosphate solutions are used for water treatment, food processing, and detergent formulation. The speciation determines the effectiveness for chelation, pH control, and precipitant action. Understanding the actual ionic composition helps optimize dosage and avoid unexpected side reactions.
Special Cases and Edge Conditions
At very low concentrations, below 10^-3 M, the simple approximations become more valid because the hydrolysis contribution to pH is smaller relative to the autoionization of water. However, at these dilute levels, contamination and atmospheric CO2 absorption become proportionally more significant. Mixed salt systems add complexity. If you have both Na3PO4 and NaCl in solution, the ionic strength increases, which affects activity coefficients and shifts the apparent pKa values. I encountered this when analyzing river water samples that contained elevated phosphate from agricultural runoff mixed with high ionic strength from road salt. The presence of polyvalent cations like Ca2+ or Mg2+ changes everything. These ions form insoluble phosphate precipitates, removing phosphate from solution and shifting the equilibria. Hard water phosphate problems are a classic example where speciation calculations alone are insufficient without considering precipitation.
For most routine laboratory work, I recommend preparing sodium phosphate solutions with care: use freshly boiled and cooled water to minimize CO2, standardize the pH with a calibrated meter at the working temperature, and calculate the actual speciation using a proper equilibrium solver rather than rough approximations. The extra five minutes of preparation saves hours of troubleshooting later.