What We Actually Got Wrong About Acids for Decades
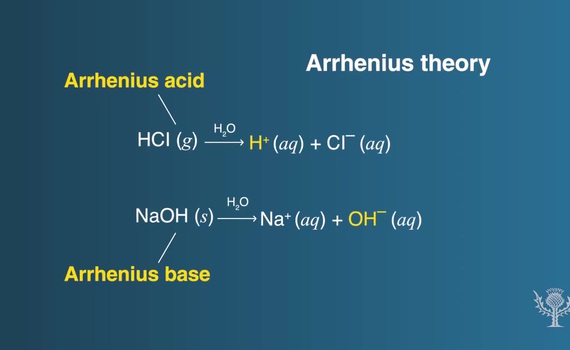
Svante Arrhenius published his doctoral thesis on this in 1884 when he was twenty-two years old. His committee barely passed him, giving it the lowest possible grade. They thought the idea was absurd. Dissociation of molecules in solution simply wasn't how people understood chemistry at the time. The core idea was deceptively simple. An Arrhenius acid is any substance that increases the concentration of hydrogen ions, H+, when dissolved in water. An Arrhenius base increases the concentration of hydroxide ions, OH-, under the same conditions. That's it. That's the entire definition. Nothing poetic about it. Where people run into trouble is when they try to apply this definition beyond water as a solvent or to substances that don't contain obvious hydrogen or hydroxide groups. I spent three days troubleshooting a reaction once because someone had classified ammonium chloride as an Arrhenius acid and then wondered why the pH readings were completely inconsistent with a strong acid. The NH4+ ion does donate a proton in water, sure, but Arrhenius's original framework was built around molecular dissociation, not Brønsted-Lowry proton transfer. The distinction matters when you're actually running experiments and trying to predict behavior.
Unveiling The Mysteries Of Arrhenius Acids The Pioneering Concept That Revolutionized Chemistry
The concept changed everything because before Arrhenius, acids were defined by what they did: they tasted sour, turned litmus red, reacted with metals. Those are observable properties, not explanations. Arrhenius gave us a mechanism. He said acids dissociate in water to produce H+ ions, and the strength of the acid corresponds directly to the extent of that dissociation. Strong acids dissociate completely. Weak acids establish an equilibrium. This was radical because it tied observable chemical behavior to something invisible: ions floating freely in solution. It gave chemists a way to quantify acidity with the pH scale, which we still use today. It made sense of neutralization as a simple combination of H+ and OH- to form water. It explained why some acids conducted electricity better than others. The practical formula you need to work with is the acid dissociation constant, Ka. For a generic acid HA dissociating in water:
HA H+ + A- Ka = [H+][A-] / [HA] A high Ka means strong acid. A low Ka means weak acid. The pKa is just negative log of Ka, which compresses the range into usable numbers. I use pKa values constantly when planning extractions or choosing buffering systems. A difference of one pKa unit means a tenfold difference in dissociation. That's the kind of detail that separates a working chemist from someone flipping through a textbook.
Get the Full Details

Here's where beginners routinely mess up. They assume that because acetic acid is called an acid, it behaves like hydrochloric acid in every situation. It doesn't. In a 0.1 M solution, HCl gives you a pH of about 1. Acetic acid at the same concentration gives you a pH around 2.9. That's nearly a hundredfold difference in actual H+ concentration. The names are the same. The behavior is wildly different. You need to check the Ka or pKa value before you design anything around a weak acid, period. I ran into a specific problem last year when a colleague was trying to use phenol as an acidic catalyst in an organic synthesis. Phenol has a pKa around 10, which makes it an extremely weak acid by Arrhenius standards. He expected it to protonate a carbonyl efficiently. It didn't. The reaction barely proceeded. We ended up switching to a stronger acid catalyst and got results in half the time. The lesson: Arrhenius acidity is a spectrum, and being labeled an acid doesn't mean you're useful as one in every context. Always check the numbers. Another thing the Arrhenius model handles poorly is amphoteric substances. Water itself, aluminum hydroxide, amino acids. These can act as either acid or base depending on what they're mixed with. Arrhenius couldn't account for that. It wasn't his fault, really. He was working with a very specific framework, and it served its purpose for aqueous solutions. But if you're working outside water or dealing with species that don't fit neatly into the H+ producer or OH- producer boxes, you need the Brønsted-Lowry definition or the Lewis definition instead.
The Lewis definition is worth mentioning briefly because it's the one most professionals actually use. It defines an acid as an electron pair acceptor and a base as an electron pair donor. No water required. No H+ or OH- required. This covers things like BF3 acting as an acid, which the Arrhenius model can't explain at all. If you're doing anything beyond basic aqueous chemistry, you'll eventually need this broader framework. The limitation of Arrhenius acids is that they only apply to aqueous solutions. Try dissolving HCl in benzene and you won't get any H+ ions because there's no water to facilitate the dissociation. The acid is still there, but the Arrhenius definition becomes meaningless. This isn't a flaw in chemistry, it's a boundary condition. Every model has them. The trick is knowing when you've crossed one. For most practical lab work, you don't need to think about Arrhenius versus Brønsted-Lowry versus Lewis unless you're designing a novel reaction or teaching an introductory course. The pH scale works. The Ka tables work. The Henderson-Hasselbalch equation works for buffers. Just remember that these are models, not laws of nature, and they have domains where they break down. When they break, you move to a better model. That's how this field has advanced for the last hundred and forty years since Arrhenius got roasted by his own thesis committee.