Why Medical Device Classification Drives Everything Else
You can't submit a 510(k), get De Novo clearance, or file a PMA without first knowing which class your product falls into. It sounds simple. It isn't. I've seen teams waste three months chasing the wrong regulatory pathway because they misclassified a minor software module at the start. The classification determines your submission type, your timeline, your cost, and whether you even need FDA involvement at all. Let me walk through how this actually works, the places where people regularly get stuck, and one edge case I ran into that isn't covered in any of the basic guides.
Unraveling The Mystery Of Us Medical Device Classifications A Decoding Guide
The FDA sorts devices into three classes based on risk and regulatory control. Class I is low risk. Class II is moderate risk. Class III is high risk, usually life-sustaining or implantable. Most of the market sits in Class II. You're more likely to encounter a 510(k) pathway than anything else. But classifying a device isn't just picking a number. You have to determine what the device is, what it does, how it's intended to be used, and whether a predicate device exists. These decisions stack on top of each other. Get one wrong and the whole pathway shifts.
The Classification Process in Practice
Start with the device description. Write it out in plain language before you open any FDA database. I know that sounds obvious, but most classification errors happen because the device description is vague or mixes multiple intended uses into one paragraph. Next, find the product code. This is the unique identifier the FDA assigns to every device. It lives inside the Classification Panel database at fda.gov. The product code links to your specific classification regulation, which tells you whether the device is exempt, subject to special controls, or requires premarket approval. Then check the regulation section. Each product code maps to a CFR citation, usually in Title 21. This is where you find the actual requirements. Some devices are exempt from premarket notification. Most Class II devices are not exempt but have special controls like performance standards, labeling requirements, or post-market surveillance. Class III devices almost always need a PMA.
Get the Full Details
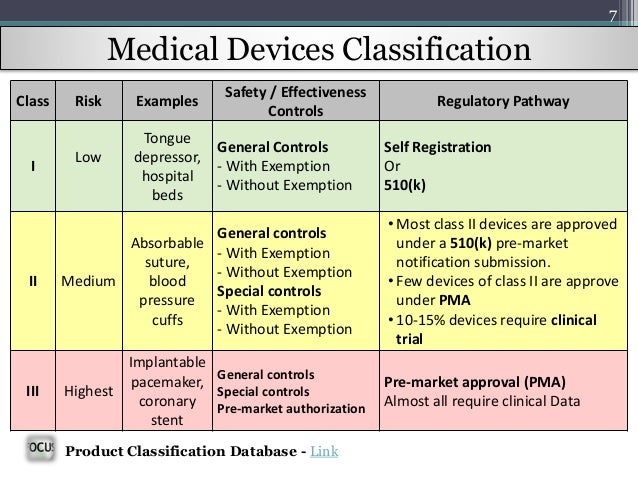
After that, determine your regulatory pathway. For Class II, that typically means a 510(k). You need to show your device is substantially equivalent to a predicate device that's already on the market. For novel devices without a predicate, you might go the De Novo route. Class III typically requires a PMA, which is the most expensive and time-consuming pathway available.
Edge Cases That Break the Standard Flow
Here's where it gets messy. Combined products. Software. Intended use statements that are too broad or too narrow. These scenarios don't fit neatly into the standard process, and the FDA often pushes back during review. Last year I worked with a team building a wound care dressing that included a small pH sensor embedded in the material. The dressing itself was Class I and exempt. The sensor, standing alone, would have been Class II and required a 510(k). But together, they formed a combination product. The FDA classified it as a drug-device combination, which moved it into Class III territory and triggered a PMA requirement. Three months of extra work and a completely different regulatory strategy. If we'd caught the combination product angle earlier, we could have designed around it or split the submission differently. The workaround we used was to restructure the product. We separated the sensor into a standalone accessory with its own labeling and intended use, and kept the dressing as a basic wound care product. That got us back to a simpler pathway. It wasn't ideal from a product design standpoint, but it was regulatory-wise the only move that made sense at the time.
Common Pitfalls to Avoid
The biggest mistake I see is relying on another company's classification without verifying it. Just because a competitor got away with Class I exemption doesn't mean your similar device is classified the same way. Intended use, labeling, and even minor design differences can change the classification entirely. Always pull the regulation yourself and read the full text. Another pitfall is assuming all software falls under the same category. The FDA has a specific guidance document for software as a medical device, and not all software is treated equally. Some software modules are Class I, some are Class II, and some fall outside device regulation altogether if they meet certain criteria. Check the guidance and the GMDN database if you're working with software. A third common error is misreading the exemption status. A device might be listed as exempt from premarket notification, but that doesn't mean it's exempt from everything. You still need to follow general controls, registration, listing, and labeling requirements. Exemption from 510(k) is not the same as being unregulated.

Tools and Resources That Actually Help
The FDA's online database is the primary tool. It covers product codes, classification regulations, and guidance documents. It's not the most user-friendly interface, but it's accurate and free. The device search tool at accessdata.fda.gov lets you look up existing cleared devices and their classification information. The Global Medical Device Nomenclature (GMDN) database is another useful resource, though it requires a subscription for full access. It assigns codes that can help you identify the right product code and classification faster than searching through CFR sections manually. I also keep a bookmarked copy of the FDA's "How to Determine Whether a Device is a Drug or a Combination Product" guidance. It's dry reading, but it saves time when you're dealing with borderline cases.
When Classification Guidance Falls Short
The system works well for standard devices. It breaks down when you're doing something genuinely novel. In those cases, you might need to request a classification determination through the FDA's Division of Industry and Consumer Education. There's a formal process for that, and it usually takes 60 to 90 days. It's slower than just guessing and finding out later, but cheaper than reworking your entire submission after the FDA sends you a deficiency letter. Sometimes the right move is a pre-submission meeting. These are optional but can save weeks of back-and-forth. You present your device description, intended use, and proposed classification, and the FDA tells you whether you're on the right track. A pre-sub takes about 30 days to schedule and a few days to prepare. The feedback you get is valuable, especially for complex devices. There's also the option of requesting a De Novo classification if your device is low-to-moderate risk but has no predicate. This pathway was created specifically for novel devices that don't fit into the traditional 510(k) framework. It's not common, but it exists, and it's worth knowing about if you're building something the FDA hasn't seen before.
What the Classification Determines Beyond the Submission Type
Class I devices generally need facility registration and device listing. That's it for most of them. Some have additional requirements depending on the specific regulation, but the bar is low. You'll still need to follow quality system regulations, though that changed with the QSR to QMSR alignment, and you should be tracking that transition. Class II devices carry more requirements. You need a 510(k) in most cases, you need to follow QSR/QMSR, you need to register and list, and you may need to comply with specific guidance documents or standards that the FDA references in your classification regulation. The special controls section of your CFR citation will list these. Read it carefully. Class III devices have the most requirements. A PMA is almost always necessary, along with QSR/QMSR compliance, registration and listing, and often post-approval studies. The PMA process typically takes 12 to 18 months and costs anywhere from $300,000 to over a million dollars depending on the complexity of the clinical data required.

Post-market requirements apply across all classes. Adverse event reporting, corrections and removals, and medical device reporting are mandatory. The stringency increases with the class, but even Class I devices aren't completely free from post-market obligations. Don't skip this part of your compliance plan.
A Note on International Devices
If your device is classified and marketed outside the US, that doesn't guarantee anything about your US classification. The FDA uses its own system. CE marking in Europe follows a different framework entirely. MDR classifications don't map 1:1 to FDA classes. I've seen companies assume their CE Class IIa device was automatically FDA Class II and skip the proper classification work. It came back to bite them during the 510(k) review. If you're translating a foreign classification into an FDA pathway, treat it as a starting point, not a conclusion. Do the full analysis from scratch for the US market.
Bottom Line
Classification is the foundation. Get it wrong and everything downstream gets harder, more expensive, and slower. Get it right and the rest of the regulatory process becomes a series of manageable steps instead of a mystery. Take the time to understand your device, read the actual regulation, verify against existing products, and don't assume anything without checking. The FDA database is your best friend here. It's not elegant, but it's comprehensive and authoritative. Use it. Supplement it with guidance documents. And when you're unsure, use the formal channels the FDA provides. A 60-day classification determination request is better than a 6-month 510(k) deficiency letter.