Understanding the Concerted Bond Formation
The Diels-Alder reaction is a [4+2] cycloaddition between a conjugated diene and a dienophile that proceeds through a single cyclic transition state. You form two new sigma bonds and one new pi bond in one step. No intermediates. No carbocations. The six-membered ring that results is a cyclohexene, and stereochemistry is preserved from the starting materials because everything happens at once. I used to think this was straightforward organic chemistry, but the reality is messier than most textbooks suggest. Getting good yields consistently requires understanding orbital symmetry, electronic matching, and the subtle ways substitution patterns shift outcomes. This isn't a reaction you can just set up and walk away from without surprises.
Unraveling The Mysteries Of The Dielsalder Reaction A Concise Analysis Of A Concerted Process
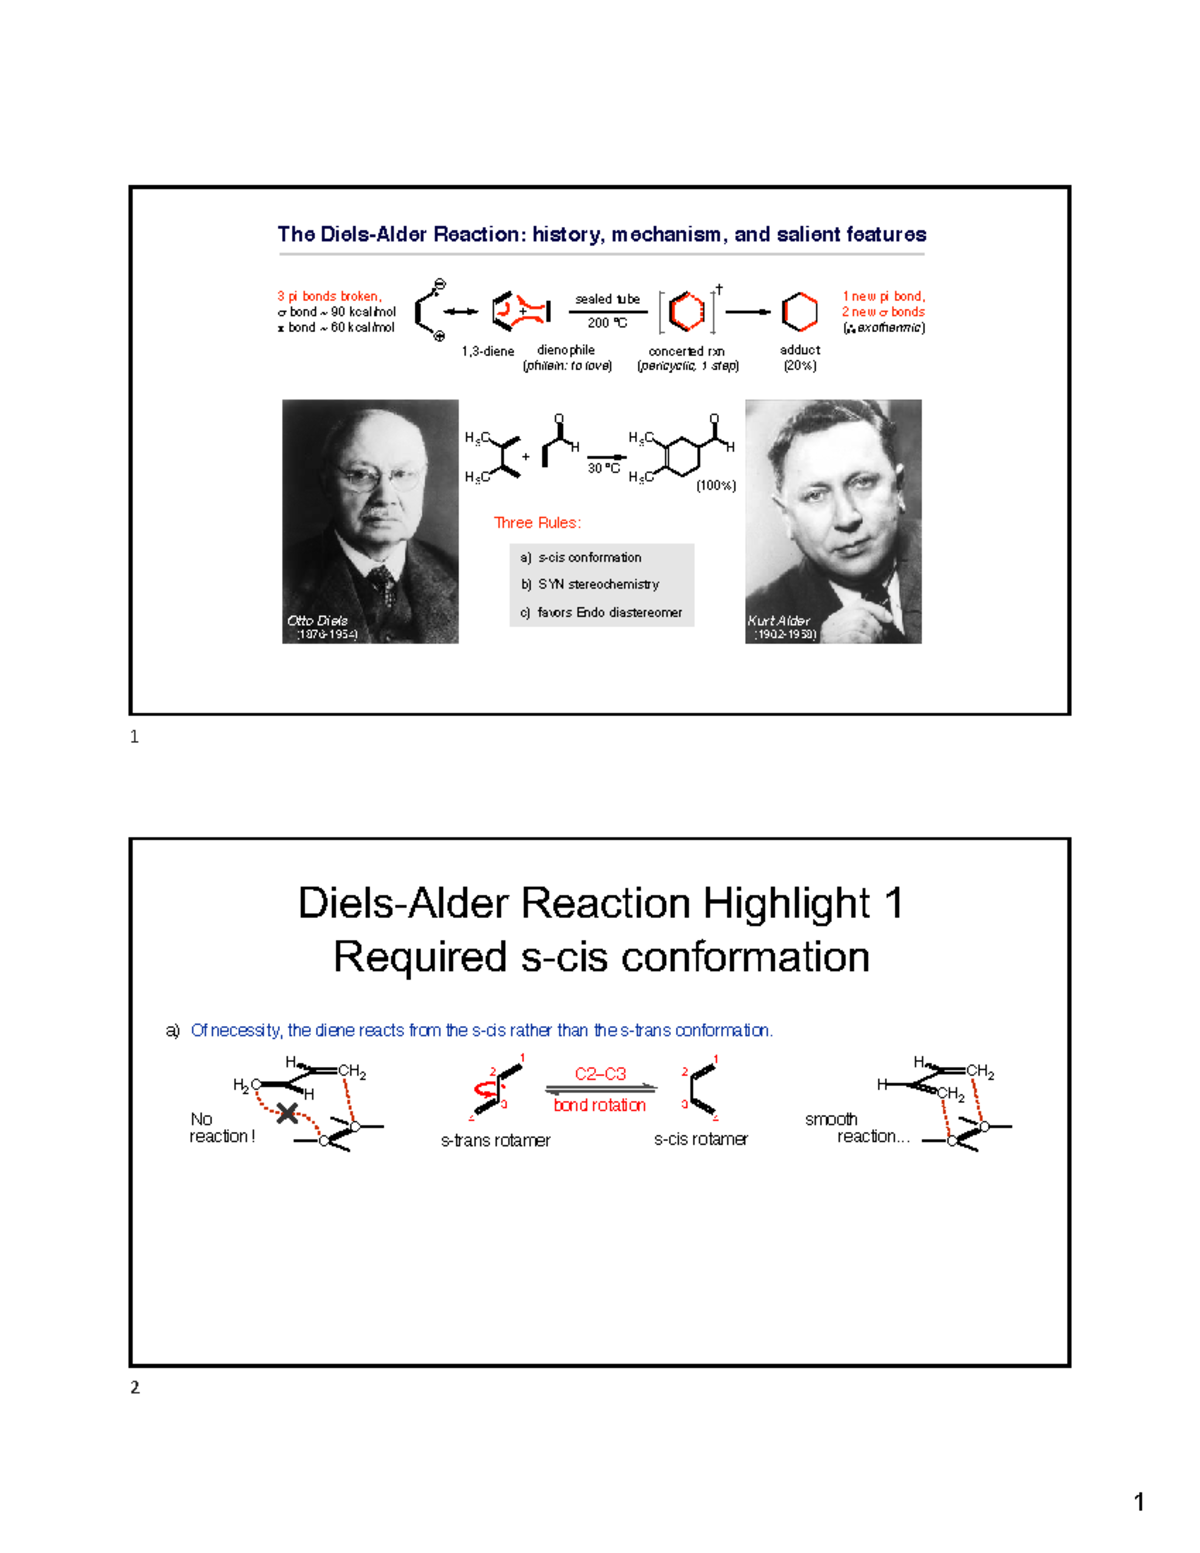
Let me start with how it actually works in practice rather than the textbook definition. You take your diene in an s-cis conformation and bring it close to an electron-poor alkene. The HOMO of the diene interacts with the LUMO of the dienophile. When those orbitals align properly, bonds form simultaneously across the six atoms involved. The transition state is roughly planar or slightly boat-like depending on substituents, and the activation barrier drops significantly when the energy gap between HOMO and LUMO is small. Electronic effects are the primary lever you control. Electron-donating groups on the diene raise its HOMO energy. Electron-withdrawing groups on the dienophile lower its LUMO energy. Both changes narrow the gap and accelerate the reaction dramatically. A simple acrylate dienophile paired with a Danishefsky-type diene can go to completion in hours at room temperature, while unactivated counterparts might require heat and still give poor conversion. Regioselectivity follows from orbital coefficient analysis. In an unsymmetrical diene-unsymmetrical dienophile pairing, the largest HOMO coefficient on the diene bonds to the largest LUMO coefficient on the dienophile. For an electron-rich diene with an EDG at C1 and an electron-poor dienophile with an EWG, you get the 1,2-product as major over the 1,4-product. The ortho/para analogy to aromatic substitution holds reasonably well here, but it is a rough guide, not a law. I have seen cases where computational prediction and actual product distribution disagreed by a wide margin because solvent effects and secondary orbital interactions mattered more than the simple frontier model suggested.
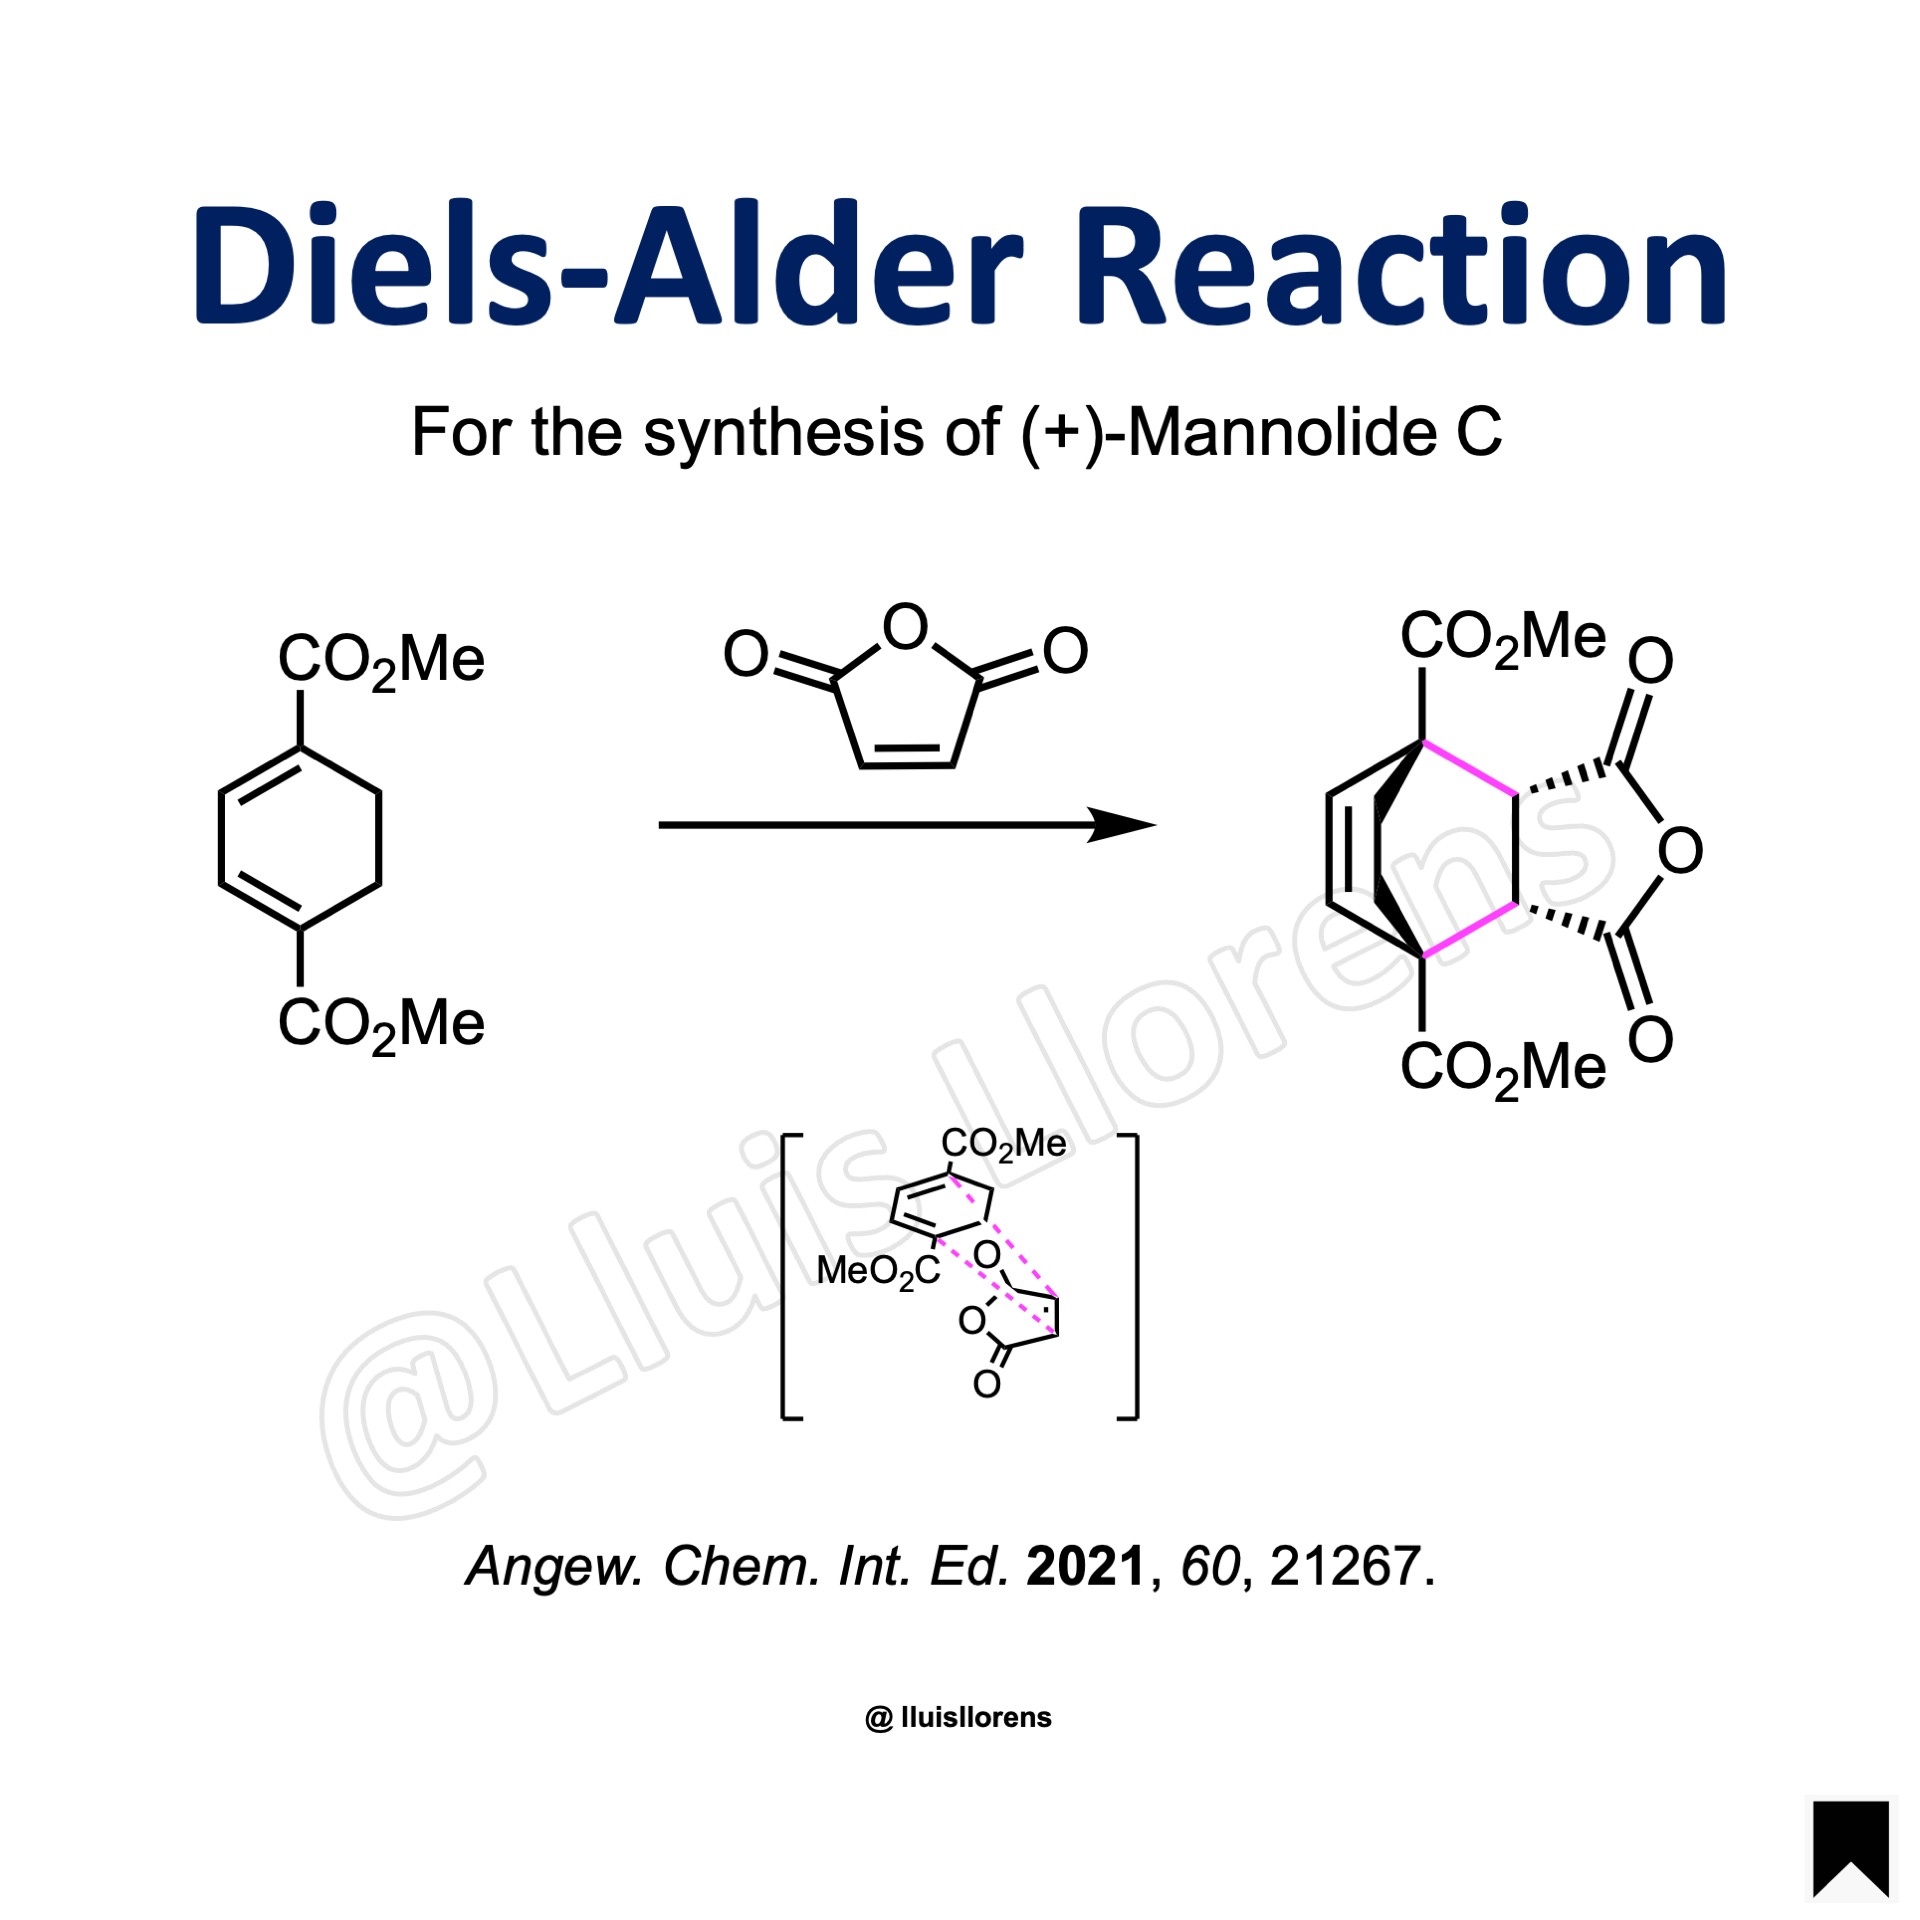
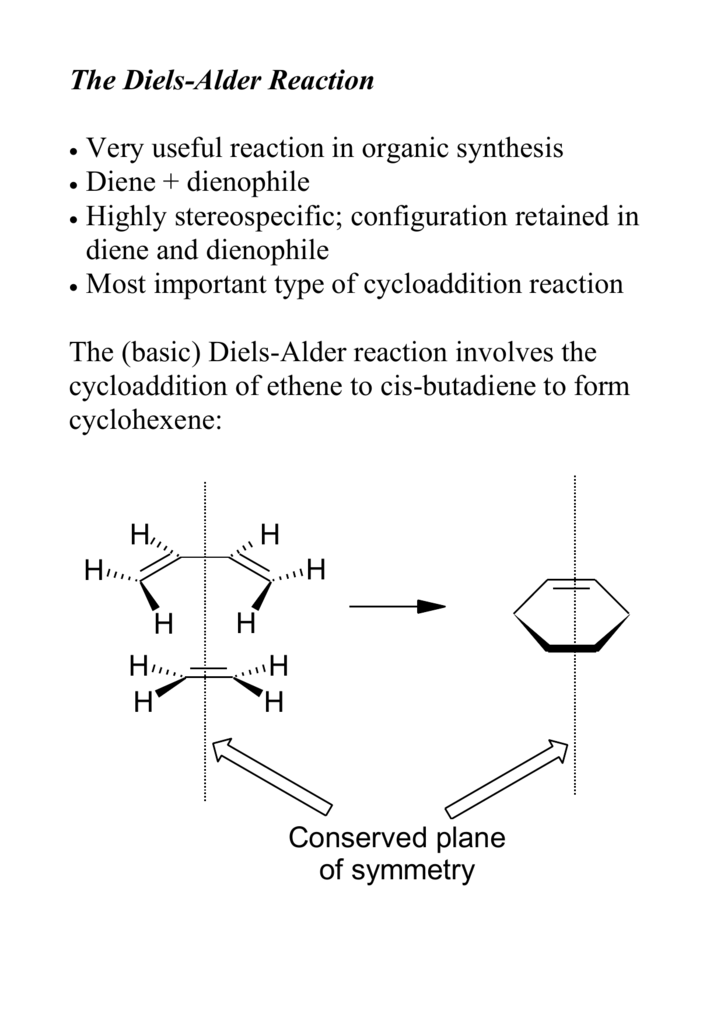
Endo selectivity is governed by secondary orbital interactions in the transition state. The endo product places the electron-withdrawing substituent of the dienophile underneath the pi system of the diene, allowing stabilizing overlap. This is typically the kinetic product and dominates at lower temperatures. The exo product is thermodynamically more stable because it minimizes steric strain, so under forcing conditions or prolonged reaction times, you may see endo-to-exo equilibration. I learned this the hard way during a synthesis campaign where heating a diels-alder adduct at reflux in toluene for 18 hours flipped the major product from endo to exo and cost me an entire batch I had been optimizing for three weeks. Here is a specific edge case that almost got me fired. I was running a hetero-Diels-Alder reaction with an aldehyde as the dienophile and a silyl enol ether as the diene equivalent, aiming for a dihydropyran. The standard procedure called for TiCl4 as a Lewis acid catalyst at low temperature. Everything looked fine on paper, but the reaction gave a complex mixture. After three days of troubleshooting, I realized the issue was trace moisture in the solvent. TiCl4 is aggressively hygroscopic, and even 100 ppm water in dichloromethane was enough to hydrolyze the catalyst and generate HCl, which protonated the carbonyl oxygen instead of coordinating to it cleanly. The workaround was strictly anhydrous conditions with distilled solvents over molecular sieves and pre-dried glassware under argon. Once I did that, the reaction went cleanly in two hours at -78 degrees Celsius with 87 percent yield. Moisture is the silent killer in Lewis acid-catalyzed cycloadditions, and most protocols gloss over how sensitive these systems actually are. STEREOCHEMISTRY deserves more attention than it gets. The reaction is stereospecific with respect to both partners. A cis-dienophile gives a cis-substituted product. A trans-dienophile gives trans substitution. The diene stereochemistry maps directly onto the ring. If you start with (E,E)-1,3-diene, the two substituents end up trans to each other on the new ring. This predictability is one of the greatest strengths of the method for complex molecule synthesis.
Get the Full Details
Intramolecular Diels-Alder reactions are another tool that worth understanding deeply. When the diene and dienophile are connected by a tether, the effective molarity skyrockets and the reaction becomes much faster and more selective than the intermolecular equivalent. Woodward's rule predicts the ring fusion stereochemistry based on the geometry of the tether, but I find it more practical to just run a few modeling calculations before committing to a route. The theoretical predictions are usually right, but the few times they are wrong, they are wrong in expensive ways. The major bottlenecks and failure modes are real and often underestimated. Not all dienes can adopt the required s-cis conformation. Cyclopentadiene locks into s-cis and reacts incredibly fast, while 1,3-butadiene exists predominantly in s-trans and requires higher temperatures and longer reaction times. Sterically hindered dienes like 2,3-di-tert-butyl-1,3-butadiene barely react at all because the bulky groups prevent the necessary orbital overlap. Electron-neutral or electron-rich dienophiles without strong withdrawing groups are essentially unreactive under standard conditions unless you push the temperature or use high pressure. Pressure is a legitimate tactic I recommend for stubborn cases, because the negative volume of activation means high pressure accelerates cycloadditions. Going from 1 atm to 10 kbar can increase rates by orders of magnitude. Retro-Diels-Alder is the reverse reaction, and it matters practically. Some adducts are thermally unstable and will break apart at elevated temperatures. If you need to purify a sensitive cyclohexene product by column chromatography, running silica gel at ambient temperature might be fine, but heating during concentration under reduced pressure can decompose your material. I have lost product to this more than once by concentrating a crude reaction mixture too aggressively.
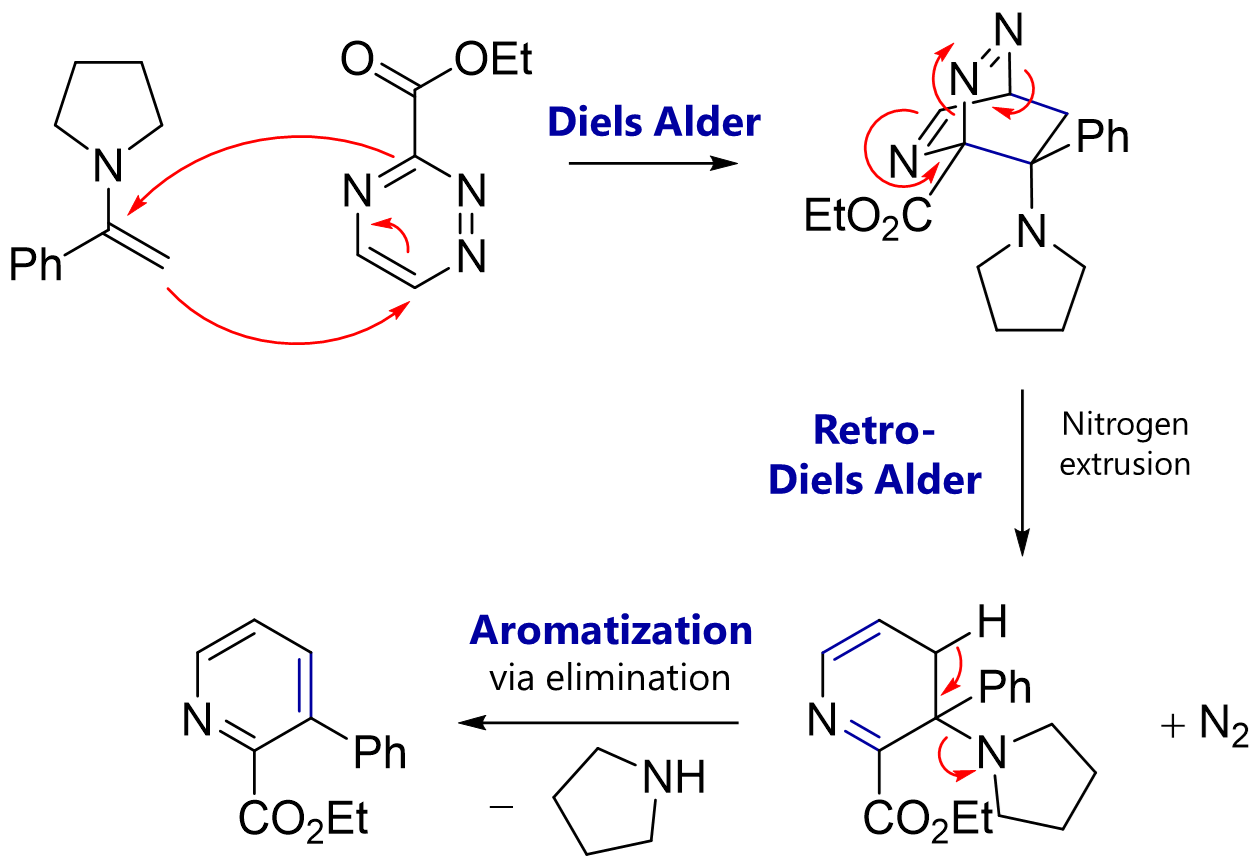
The inverse electron demand variant flips the orbital roles. An electron-poor diene pairs with an electron-rich dienophile. This is the basis for tetrazine ligation chemistry, which has become huge in bioconjugation. The reaction is exceptionally fast, sometimes diffusion-controlled, and proceeds under aqueous conditions at physiological temperature. It is probably the most useful industrial application of Diels-Alder chemistry right now, used in antibody-drug conjugates and polymer modification. Solvent effects are generally modest but not negligible. Polar solvents can accelerate reactions when there is significant charge development in the transition state, especially in Lewis acid-catalyzed variants. Water sometimes gives unusual rate enhancements and selectivity shifts, an effect attributed to hydrogen bonding networks and hydrophobic compression, though the mechanism is debated. I rarely optimize solvent for Diels-Alder reactions unless I am hitting a specific selectivity problem, because the choice usually does not make or break the reaction. Computational modeling has made predicting outcomes more reliable. DFT calculations at the M06-2X/6-31G* level can predict regioselectivity and endo-exo ratios within a few kilocalories of experimental values for most standard cases. If you are designing a novel diene-dienophile pair, running a quick computational screen before going to the bench saves reagents and time. The calculations will not replace actual experimentation, but they eliminate the clearly wrong approaches early.
Workup is straightforward in most cases. Quench with aqueous sodium bicarbonate or satiated ammonium chloride, extract with ethyl acetate or dichloromethane, dry over magnesium sulfate, and concentrate. Recrystallization or flash chromatography on silica gel handles purification for the vast majority of products. The only tricky cases involve thermally sensitive adducts or products that co-elute with starting material, which is rare but annoying when it happens. Scale-up introduces its own considerations. Exotherms are generally mild compared to other cycloadditions, but on multi-kilogram scale, heat dissipation matters. I have run these reactions from milligram to 50-gram scale without major issues, but above that, you need to evaluate thermal profile properly. Calorimetry data is worthwhile if you are committing to a process route. The reaction itself does not introduce unusual safety hazards, but some dienophiles like maleic anhydride are respiratory irritants, and Lewis acid catalysts require careful handling procedures. The reaction has genuine limitations that prevent it from being a universal solution. It does not work well for aromatic substrates because the aromatic stability is too high to disrupt under normal conditions. Heteroaromatic dienes are generally unreactive unless activated. Vinyl cyclopropanes can participate but undergo ring-opening first, which complicates the outcome. And for forming highly substituted or strained cyclohexenes, the thermodynamic and kinetic barriers often make alternative disconnections more practical.

If the Diels-Alder route is failing, alternatives include stepwise conjugate addition followed by ring closure, transition-metal-catalyzed cyclopropanation or cyclopropane ring expansion, or using different pericyclic pathways like the 1,3-dipolar cycloaddition when the target ring system allows it. No single method covers every synthetic need, and recognizing when to move away from a Diels-Alder approach is part of knowing the reaction honestly.