The Synaptonemal Complex and Why It Keeps Breaking Your Spreadsheets
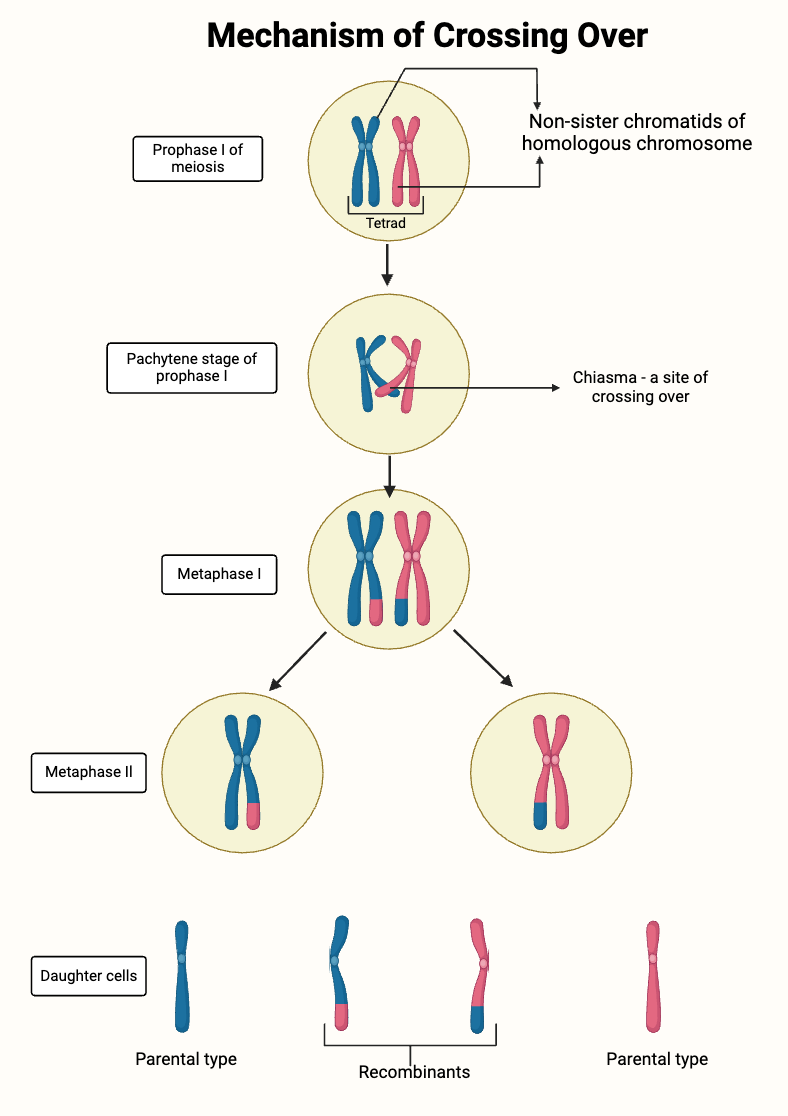
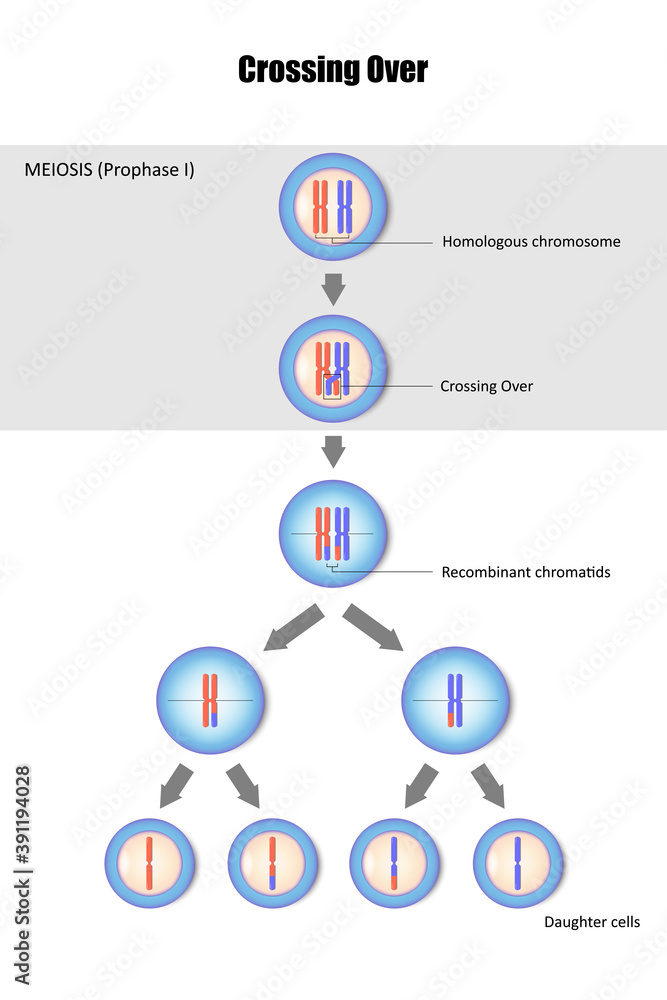
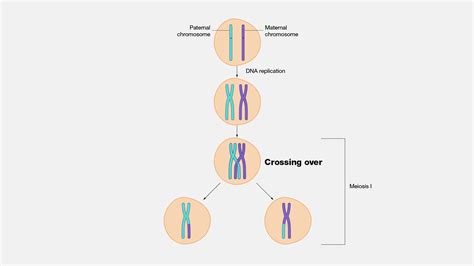
I spent three weeks last year trying to map recombination frequencies in Arabidopsis using custom PCR primers, and every single assay came back showing zero crossover products in a region that textbook literature said had one of the highest rates in the entire genome. I had to go back and look at the chromosome squash under a proper fluorescence microscope instead of relying on genetic markers alone, and what I found was that the crossover was happening, just not in the region my primers were designed to amplify because there was a small inversion that shifted where the chiasmata actually formed. Genetic mapping data alone doesn't always tell you where the physical exchange is happening, and if you only look at offspring phenotypes without karyotype or cytological confirmation, you're essentially guessing at half the picture. Crossing over is the reciprocal exchange of genetic material between homologous chromosomes during prophase I of meiosis. It happens specifically during the pachytene substage, after the synaptonemal complex has fully assembled between the paired homologues. The molecular machinery involves a cascade of proteins starting with SPO11 creating programmed double-strand breaks, followed by resection by nucleases like MRE11 and RAD51, strand invasion into the homologous chromosome, and eventually the formation and resolution of Holliday junctions into either crossover or non-crossover products. About one in three repair events becomes a crossover; the rest are resolved as gene conversion without exchange of flanking markers. There is a common misconception that crossing over occurs randomly along the chromosome. It doesn't. In humans, crossovers are strongly suppressed near centromeres and telomeres and are enriched in promoter-proximal regions and recombination hotspots driven by PRDM9 binding sites. PRDM9 is a zinc finger protein that methylates histone H3 at lysine 4, creating a chromatin signature that recruits the recombinational machinery. Different alleles of PRDM9 bind different DNA motifs, which is why crossover hotspots vary substantially between individuals and between species. In dogs, for instance, PRDM9 has gone almost completely pseudogenized, and they use alternative hotspot determinants that humans don't have.
The biological purpose is straightforward but often understated. Crossovers generate new allele combinations on each chromosome arm, which is the primary source of genetic diversity in sexually reproducing organisms. They also serve a mechanical function. Chiasmata, the visible manifestations of crossovers, are what hold homologous chromosomes together after the synaptonemal complex disassembles during diplotene. Without at least one chiasma per chromosome arm, the homologues can't achieve proper bivalent orientation on the metaphase I spindle, and nondisjunction becomes much more likely. That's why achiasmate chromosomes are a leading cause of aneuploidy, including trisomy 21 in human oocytes. Here's something most introductory courses don't cover adequately: crossover interference. A crossover in one region of a chromosome actively suppresses another crossover from forming nearby. The spacing between adjacent crossovers is more regular than would be expected by chance, and this is quantified as the coefficient of coincidence, which is typically between 0.2 and 0.5 in most eukaryotes. The molecular mechanism behind interference isn't fully settled, but the leading model involves the zippering behavior of the synaptonemal complex and tension-dependent signaling from the completed crossover site propagating along the chromosome axis. This matters practically because it means you can't model crossover distribution as a simple Poisson process when you're doing genetic mapping or constructing linkage maps. When I was working on that Arabidopsis project, I also ran into the issue of sex-specific recombination rates. In many organisms, including humans and Drosophila, males and females have very different crossover landscapes. Human females average about 41 crossovers per meiosis while human males average about 27. The female map is roughly 1.6 times longer than the male map. If you're building a genetic map from population data and you pool sexes together, your map distances will be inaccurate. You need to sex-ify the map or use sex-averaged distances with the appropriate confidence intervals, which widen considerably when the two sexes disagree on hotspot usage.
Another practical consideration is the difference between obligate and facilitative crossovers. Every chromosome pair needs at least one obligate crossover to ensure proper segregation. Beyond that minimum, additional facilitative crossovers can form and contribute to diversity without being strictly necessary for segregation. In fission yeast, for example, most cells get exactly one crossover per chromosome pair, which is the bare minimum. In mammals, you regularly see two or more per arm, especially on the larger chromosomes. This has implications for linkage disequilibrium patterns in population genetics because regions with high crossover rates will break down linkage faster over generations.
Get the Full Details

What Goes Wrong When You Try to Measure It
Recombination fraction underestimates true genetic distance because of multiple crossovers. If two crossovers happen between two markers in the same meiotic event, they cancel each other out genetically and the gametes look parental. This is why mapping functions like Haldane's or Kosambi's exist. Haldane's assumes no interference and uses the formula d = -0.5 * ln(1 - 2), where is the recombination fraction. Kosambi's incorporates interference and is generally more accurate for most eukaryotic genomes. Neither is perfect, and both break down at large distances where approaches 0.5 and you can no longer distinguish linked from unlinked loci. Chromosomal rearrangements complicate everything. Inversions suppress apparent recombination in heterozygotes because crossovers within the inverted region produce dicentric bridges and acentric fragments that are mostly nonviable. What you observe is not a true absence of crossing over but a selective loss of recombinant products. This is a classic pitfall when interpreting recombination data from wild populations or breeding experiments, and it's why cytological examination or whole-genome sequencing is really necessary if you suspect structural variation is present. There's also the issue of gene conversion tracts that occur without crossover. These can be 500 to 2000 base pairs long in mammals and create apparent violations of Mendelian ratios at individual loci without any exchange of flanking markers. In yeast, you can detect these directly by tetrad analysis. In outcrossing organisms, you infer them from distorted segregation ratios or from sequence variation in pedigrees. If you're doing any kind of fine-scale recombination mapping and you don't account for gene conversion, your hotspot calls will be noisy and your estimated crossover positions will be systematically offset.
The practical takeaway is that crossing over is neither random nor uniform, it's tightly regulated at multiple levels, and the regulatory machinery overlaps with DNA repair pathways that operate throughout the cell cycle. That overlap is why mutations in genes like BRCA1, BRCA2, and PALB2, which are best known for their role in homologous recombination repair of double-strand breaks in somatic cells, also cause meiotic defects and infertility when disrupted in the germline. The same molecular machinery handles both contexts, and errors in either one have predictable consequences.