Working With H2O2-Derived Hydroxyl Radicals in the Lab
Hydrogen peroxide is cheap, stable on the shelf, and practically useless until you figure out how to split it into something reactive. The hydroxyl radical (•OH) is the product you actually want. It is non-selective, extremely short-lived, and impossible to store. You generate it in situ and deal with the consequences immediately. Most people who ask about "H2O2 conjugate base hydroxyl radical insights" are trying to optimize a Fenton-type system or a photo-Fenton process for degradation, synthesis, or analytical detection. Here is what I have learned from running these reactions repeatedly and watching them fail in ways that are not obvious from the literature. The conjugate base of hydrogen peroxide is the hydroperoxide anion, HO2. Its pKa is approximately 11.6 at 25°C. This means at neutral pH, the dominant species is molecular H2O2. As you push pH above 11, deprotonation becomes significant, and you now have HO2 in solution. This is not just a textbook equilibrium — it changes the entire reaction pathway. HO2 is a much stronger nucleophile and reductant than H2O2. It reacts with transition metals at different rates, undergoes single-electron oxidation more readily, and can generate hydroxyl radicals through pathways that do not require the classic Fenton mechanism. I spent about three weeks chasing an irreproducibility problem before realizing my "standard" Fenton conditions were drifting into hydroperoxide-anion territory because I was using a carbonate buffer that was slowly pushing pH upward over the course of the reaction. Once I switched to a phosphate buffer and monitored pH in real time, the data became consistent again. The key practical insight nobody emphasizes enough is that the hydroxyl radical yield from H2O2 is not a simple function of concentration. It is a function of pH, metal speciation, and the presence of radical scavengers that you might not even realize you added. Here is how I approach these systems now.
Generating the Radicals — The Actual Procedure
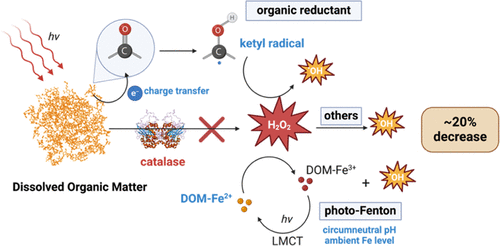
For a standard Fenton system, you need ferrous iron and hydrogen peroxide in an acidic medium. The typical range is pH 2.5 to 3.5. Outside this window, the iron precipitates as hydroxide or the H2O2 starts decomposing through a non-radical pathway. I use FeSO4·7H2O as the ferrous source because it is stable enough in acidic solution and affordable. The H2O2 should be fresh — old bottles that have sat open or been exposed to light lose concentration through catalytic decomposition. I check the actual concentration by titration before each batch of experiments rather than trusting the label. The mixing order matters more than most protocols admit. If you add H2O2 to a solution containing both Fe2+ and your substrate, the initial radical burst attacks everything in solution including your target compound, but also the solvent and any impurities. I pre-mix the Fe2+ with the buffer at the target pH, then add H2O2 as the final step while stirring. This gives me a reproducible initiation point. The reaction proceeds for a defined time — anywhere from 30 seconds to 30 minutes depending on what I am trying to achieve — and is typically quenched by raising the pH above 8 to precipitate the iron and stop radical production. When I work in the alkaline regime where HO2 dominates, the chemistry shifts. At pH above 11, the hydroperoxide anion can reduce Fe3+ back to Fe2+, which creates a catalytic cycle that is different from the classical Fenton stoichiometry. The net result is that you can generate hydroxyl radicals with less iron and less H2O2 waste, but you also get significant amounts of superoxide and singlet oxygen as byproducts. These secondary species complicate interpretation if you are trying to prove that •OH is the sole active agent. I run DMPO spin-trap EPR spectroscopy whenever I need to confirm the radical identity, and I include scavenger controls — methanol for hydroxyl radicals, catalase for H2O2, and SOD for superoxide.
Common Failure Modes
The most frequent problem I see is iron chelation. Citrate, EDTA, phosphate at high concentration, and even chloride at elevated levels can coordinate Fe2+ and Fe3+ and dramatically alter the redox potential. A citrate-Fe system operates efficiently at near-neutral pH, which is useful for environmental applications, but the radical flux is different and the mechanism involves ligand-to-metal charge transfer rather than simple Fenton chemistry. If you are comparing results across papers and the pH ranges do not match, the difference is almost certainly speciation, not methodology. Another issue is H2O2 self-decomposition. At pH above 4, especially in the presence of trace transition metal contaminants, H2O2 decomposes to water and oxygen without producing hydroxyl radicals. This is the disproportionation pathway and it is catalyzed by the same metals you are intentionally adding. The workaround is to use a slight excess of H2O2 — typically 2 to 5 times the stoichiometric amount relative to the target substrate — and to keep the reaction time as short as practical. I usually find that 5 to 10 minutes is sufficient for most degradation applications, and extending beyond 20 minutes gives diminishing returns because the H2O2 is being consumed by non-productive pathways. The third problem is pH drift during the reaction. The Fenton reaction produces protons: Fe2+ + H2O2 + H+ Fe3+ + •OH + H2O. As the reaction proceeds, pH drops. If you are not buffering adequately, the system can shift from optimal acidic conditions into a regime where iron precipitates and radical generation slows or stops. I use an acetate buffer at 0.1 M for reactions in the pH 3 to 5 range, and I verify the pH before and after the reaction to confirm it has not drifted more than 0.2 units.
Get the Full Details

Quantifying What You Actually Produced
If you need to measure hydroxyl radical output rather than just assuming it, the terephthalic acid fluorescence method is the most practical for routine lab work. Terephthalic acid reacts with •OH to form 2-hydroxyterephthalic acid, which fluoresces at 425 nm when excited at 315 nm. The response is reasonably linear in the micromolar range. I run a calibration curve with known •OH generation rates using a standard Fenton system, then apply that to my experimental conditions. The alternative is EPR with a spin trap like DMPO, but that requires equipment most synthesis labs do not have and the DMPO-OH adduct has a half-life of only a few minutes, so timing is critical.
When H2O2-Based •OH Generation Is the Wrong Tool
Hydroxyl radicals from H2O2 are indiscriminate. If you are trying to selectively functionalize a specific bond in a complex organic molecule, this approach will destroy your selectivity. UV photolysis of H2O2 at 254 nm produces •OH more cleanly and allows spatial control, but it requires a quartz vessel and a mercury lamp. Electrochemical generation of •OH on a boron-doped diamond anode is another option that gives better control over the radical flux, though the electrode cost is significant. For environmental remediation of recalcitrant organics, Fenton and photo-Fenton systems are well established and cost-effective. For precision organic synthesis, I recommend generating •OH through photolysis or using a more selective oxidant like a oxone-based system instead.