What Actually Happens When You Talk About Sp Hybridization
Most people learn sp hybridization as a set of rules: one s orbital mixes with one p orbital, you get two sp hybrids, linear geometry, 180 degrees. That part is fine. The problem is that this framing makes the whole concept feel arbitrary, like chemistry decided to invent a convenient shortcut. It didn't. There's a real physical reason it happens, and understanding that reason changes how you approach molecular orbital problems entirely. Here's how I got here. I was tutoring an undergrad who could draw acetylene on command but couldn't explain why the carbon-hydrogen bonds were different from the carbon-carbon bonds. Not just different in length, but different in everything — ionization energy, reactivity, orbital composition. She had memorized the diagram and moved on. That's the gap we're dealing with.
Unlocking The Secrets Of Molecules A Visual Guide To Understanding Sp Hybridization
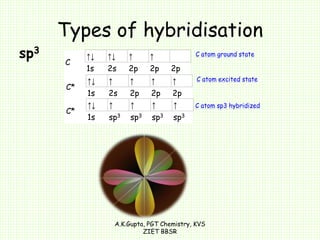
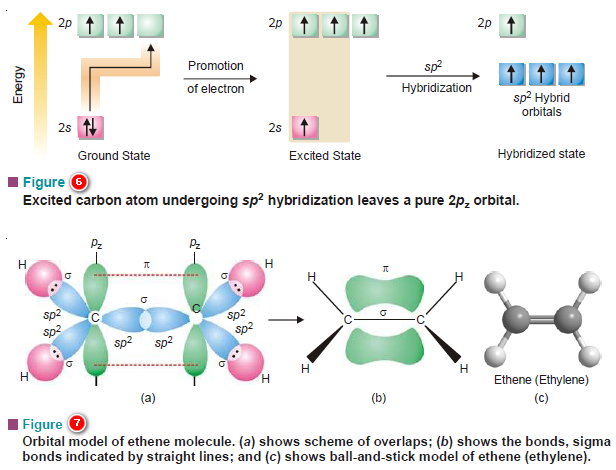
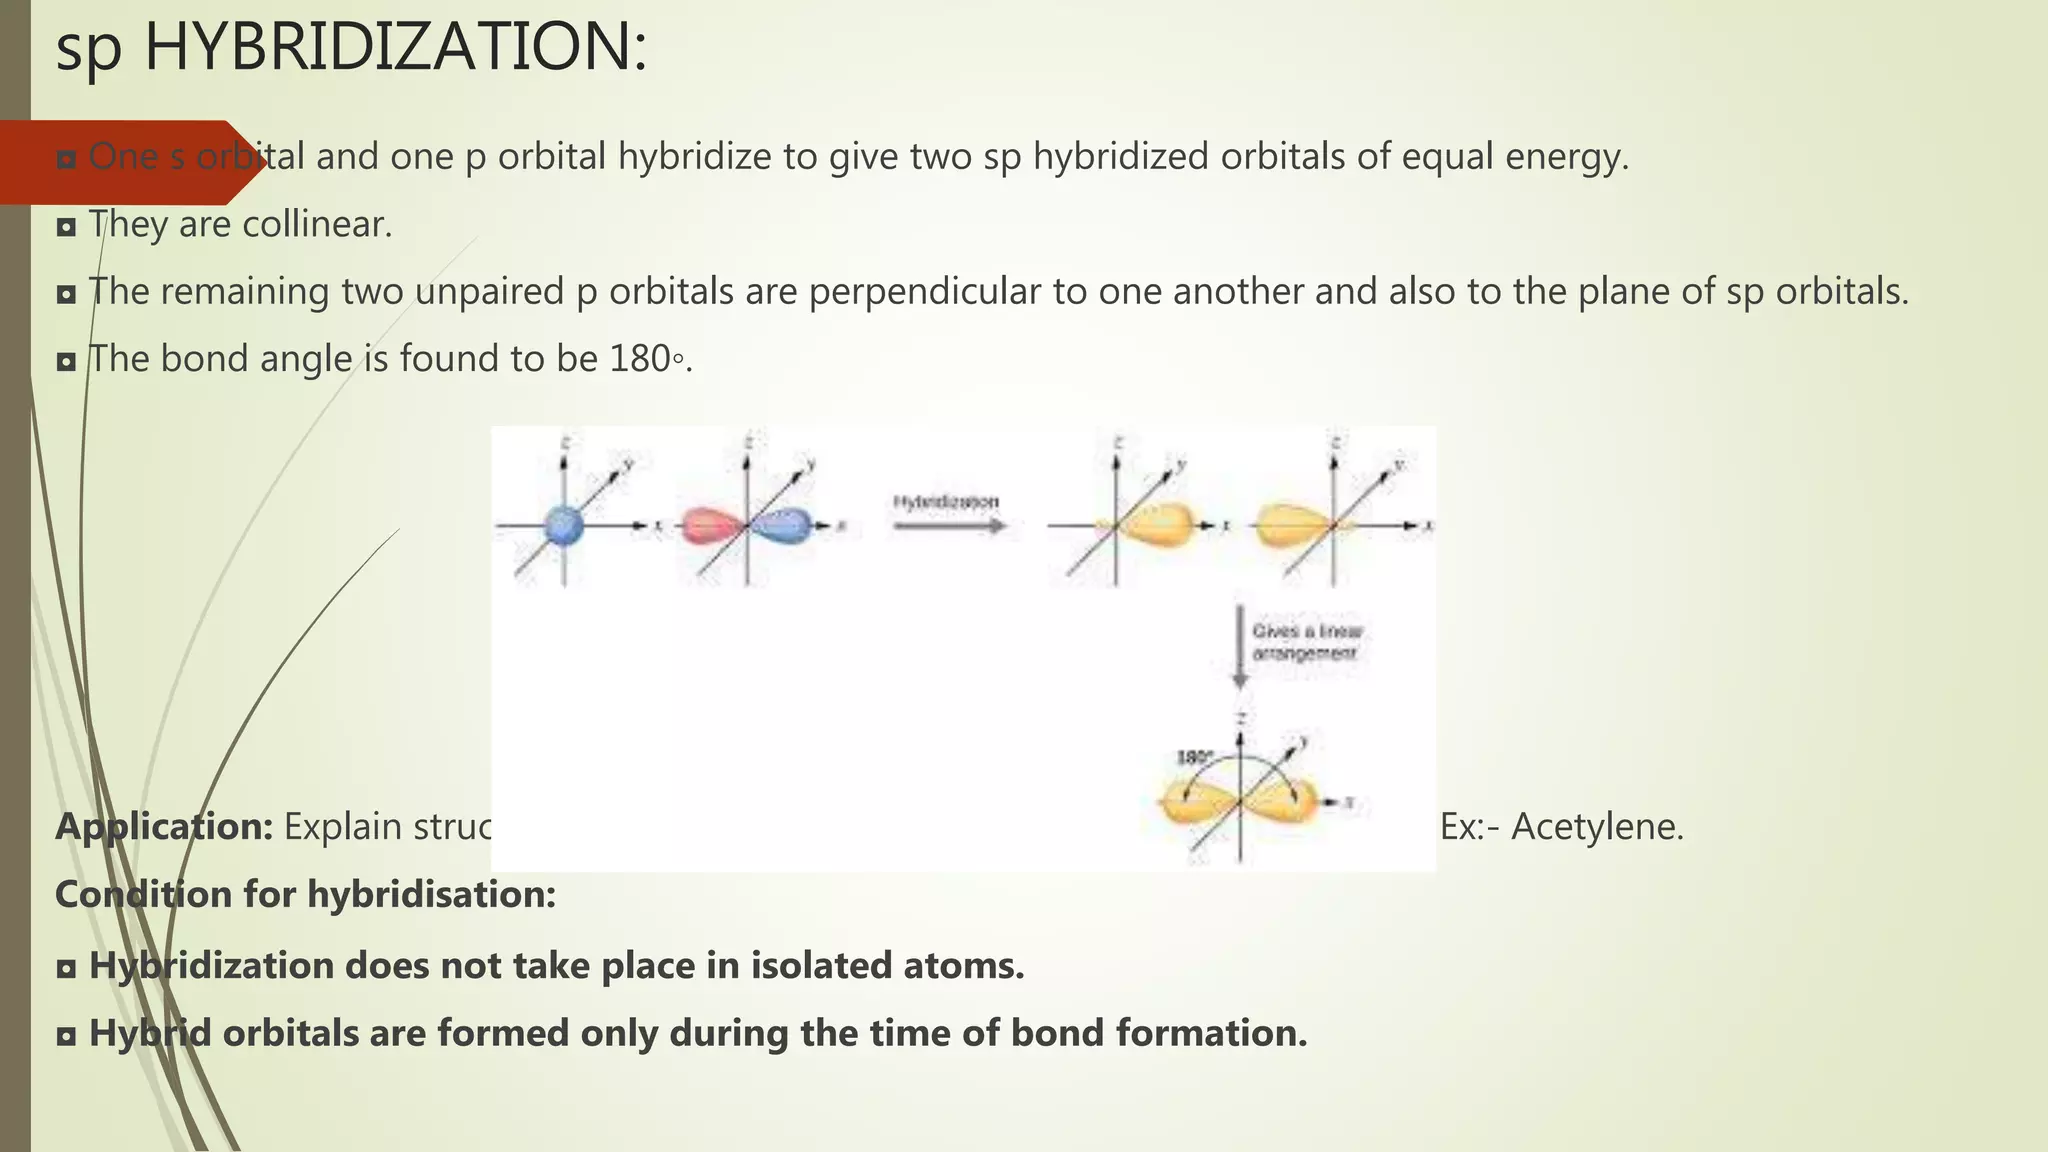
The visual part matters more than the math at this stage. Draw the carbon atom with its four valence electrons. Two in the 2s orbital, two unpaired in 2p. Now imagine what happens when you bring another carbon close enough for bonding. The system doesn't want to waste the 2s electrons on a single sigma bond if it can avoid it. So it reorganizes. The 2s and one 2p orbital merge into two new orbitals of equal energy — the sp hybrids. They point in opposite directions because that's the configuration that minimizes electron-electron repulsion. The remaining two p orbitals stay untouched and perpendicular to the hybrid axis. That's the model. Now let's talk about what breaks when you try to apply it blindly. I spent three hours once debugging a computational chemistry output where a molecule I was studying kept converging to the wrong geometry. The issue turned out to be that the default hybridization assignment in the software was forcing sp2 on an atom that should have been sp. The program assumed trigonal planar because it saw three attached atoms, but one of those "atoms" was a lone pair occupying a pure p orbital, not a hybrid. The geometry wasn't planar. It was linear. The fix was explicit hybridization specification in the input file. I learned to never trust automatic assignments when you're dealing with nitrogen in nitriles or terminal alkynes — those cases trip up the defaults every time.
Another thing nobody emphasizes enough: sp hybrid orbitals are not just any two mixed orbitals. They have a specific character. Each sp hybrid is 50% s and 50% p. This matters because s character dictates bond properties. Higher s character means the electrons are held closer to the nucleus. An sp C-H bond (50% s) is shorter and stronger than an sp2 C-H bond (33% s), which is shorter and stronger than an sp3 C-H bond (25% s). The pKa of acetylene is about 25. The pKa of ethylene is about 44. The pKa of ethane is about 50. That difference isn't subtle. It's enormous, and it comes directly from the s character of the orbital holding the electrons. So when someone tells you hybridization is just a model and not "real," they're technically right but practically misleading. The model predicts real, measurable differences in bond length, bond energy, and acidity. The numbers work. The model works because it captures something true about electron distribution. Here's a practical workflow I use when I'm trying to determine hybridization for an unfamiliar molecule. Count the number of sigma bonds and lone pairs around the atom in question. Add them together. Two regions means sp. Three means sp2. Four means sp3. That's the standard approach. But here's where it gets tricky: the lone pair doesn't always go into a hybrid orbital. In ammonia, the lone pair sits in an sp3 hybrid. In pyridine, the lone pair sits in an sp2 orbital in the plane of the ring, while the pi electrons are in the perpendicular p orbitals. In a nitrile group, the nitrogen lone pair is in an sp orbital. The geometry tells you where it should be, but you have to look at the actual bonding situation, not just count regions.
Get the Full Details
I once had a colleague who insisted that all lone pairs occupy hybrid orbitals by definition. This led to some genuinely confused explanations about why the lone pair in CO wasn't behaving the way his model predicted. The reality is that lone pair placement depends on the energetic balance between sigma and pi interactions, and sometimes the lowest energy configuration puts a lone pair in a pure p orbital. Not always. Sometimes. You have to check. For a visual guide, start with the simplest cases and work up. BeCl2: linear, sp on beryllium, no lone pairs, straightforward. CO2: linear, sp on carbon, two double bonds where each double bond consists of one sigma (from sp) and one pi (from unhybridized p). HCN: sp on both carbon and nitrogen, triple bond between them. Each example reinforces the pattern while revealing a slightly different face of the same underlying principle. The limitation I keep running into is that sp hybridization breaks down in certain transition metal complexes and in molecules with significant electron correlation effects. In those cases, the simple orbital mixing picture gives you qualitatively wrong answers. DFT calculations or multiconfigurational methods are needed. If you're working with main group organics, hybridization theory is still perfectly adequate. If you're dealing with something exotic, it's not going to save you.
Bottom line: draw the orbitals, track the s character, check your assumptions about lone pairs, and verify with experimental data whenever you can. The visual guide works because it makes the invisible geometry visible, not because it simplifies the chemistry away.