Epoxidation in Practice
The standard workhorse here is the Michaelis-Arbuzov variant or just plain mCPBA on a suitably activated double bond. If you're working with an allyl phenyl ether or a styrenic system, getting clean epoxide formation without over-oxidation or ring opening is the real challenge, not the mechanism itself. I spend most of my time figuring out why the reaction isn't proceeding the way the textbook says it should. For aromatic substrates specifically, the double bond you're epoxidizing is usually exocyclic or part of a side chain rather than the ring itself, since direct epoxidation of the aromatic pi system gives you an unstable arene oxide that rapidly rearranges or polymerizes before you can isolate anything useful.
Unlocking The Secrets Of Epoxidation How This Chemical Reaction Revolutionizes Aromatic Ring Synthesis
The practical approach I use involves treating the alkene-substituted aromatic compound with meta-chloroperbenzoic acid in dichloromethane at zero degrees Celsius. The key is keeping the temperature low and the reaction mixture dilute, typically around 0.1 molar concentration. At higher concentrations the exotherm becomes difficult to control and you start seeing ring-opened byproducts and chlorinated material from solvent participation. When the substrate is an electron-rich aromatic system like a methoxyphenyl variant, the epoxidation proceeds faster than for unsubstituted styrene. I once ran a reaction with a 3,4-dimethoxystyrene derivative and got nearly quantitative conversion in twenty minutes at room temperature instead of the two hours the procedure specified. The opposite is also true: electron-poor substrates like a nitrostyrene analogue needed extended reaction times and sometimes a catalytic amount of titanium isopropoxide to push the conversion above sixty percent. The workup is straightforward but matters more than people admit. Quench with aqueous sodium sulfite to destroy excess peracid, wash the organic layer with sodium bicarbonate to remove the resulting m-chlorobenzoic acid, dry over magnesium sulfate, and concentrate under reduced pressure below thirty degrees Celsius. The epoxides tend to degrade if you apply heat during solvent removal, so I keep the water bath temperature low and use a vacuum pump rather than a rotary evaporator for the final solvent stripping.
Isolation usually involves flash chromatography on silica gel using hexanes and ethyl acetate. The epoxide elutes earlier than the starting material and the carboxylic acid byproduct comes off last. I typically get between seventy and eighty-five percent yield depending on substrate electronics and steric bulk near the double bond.
Get the Full Details

Why This Matters for Aromatic Synthesis
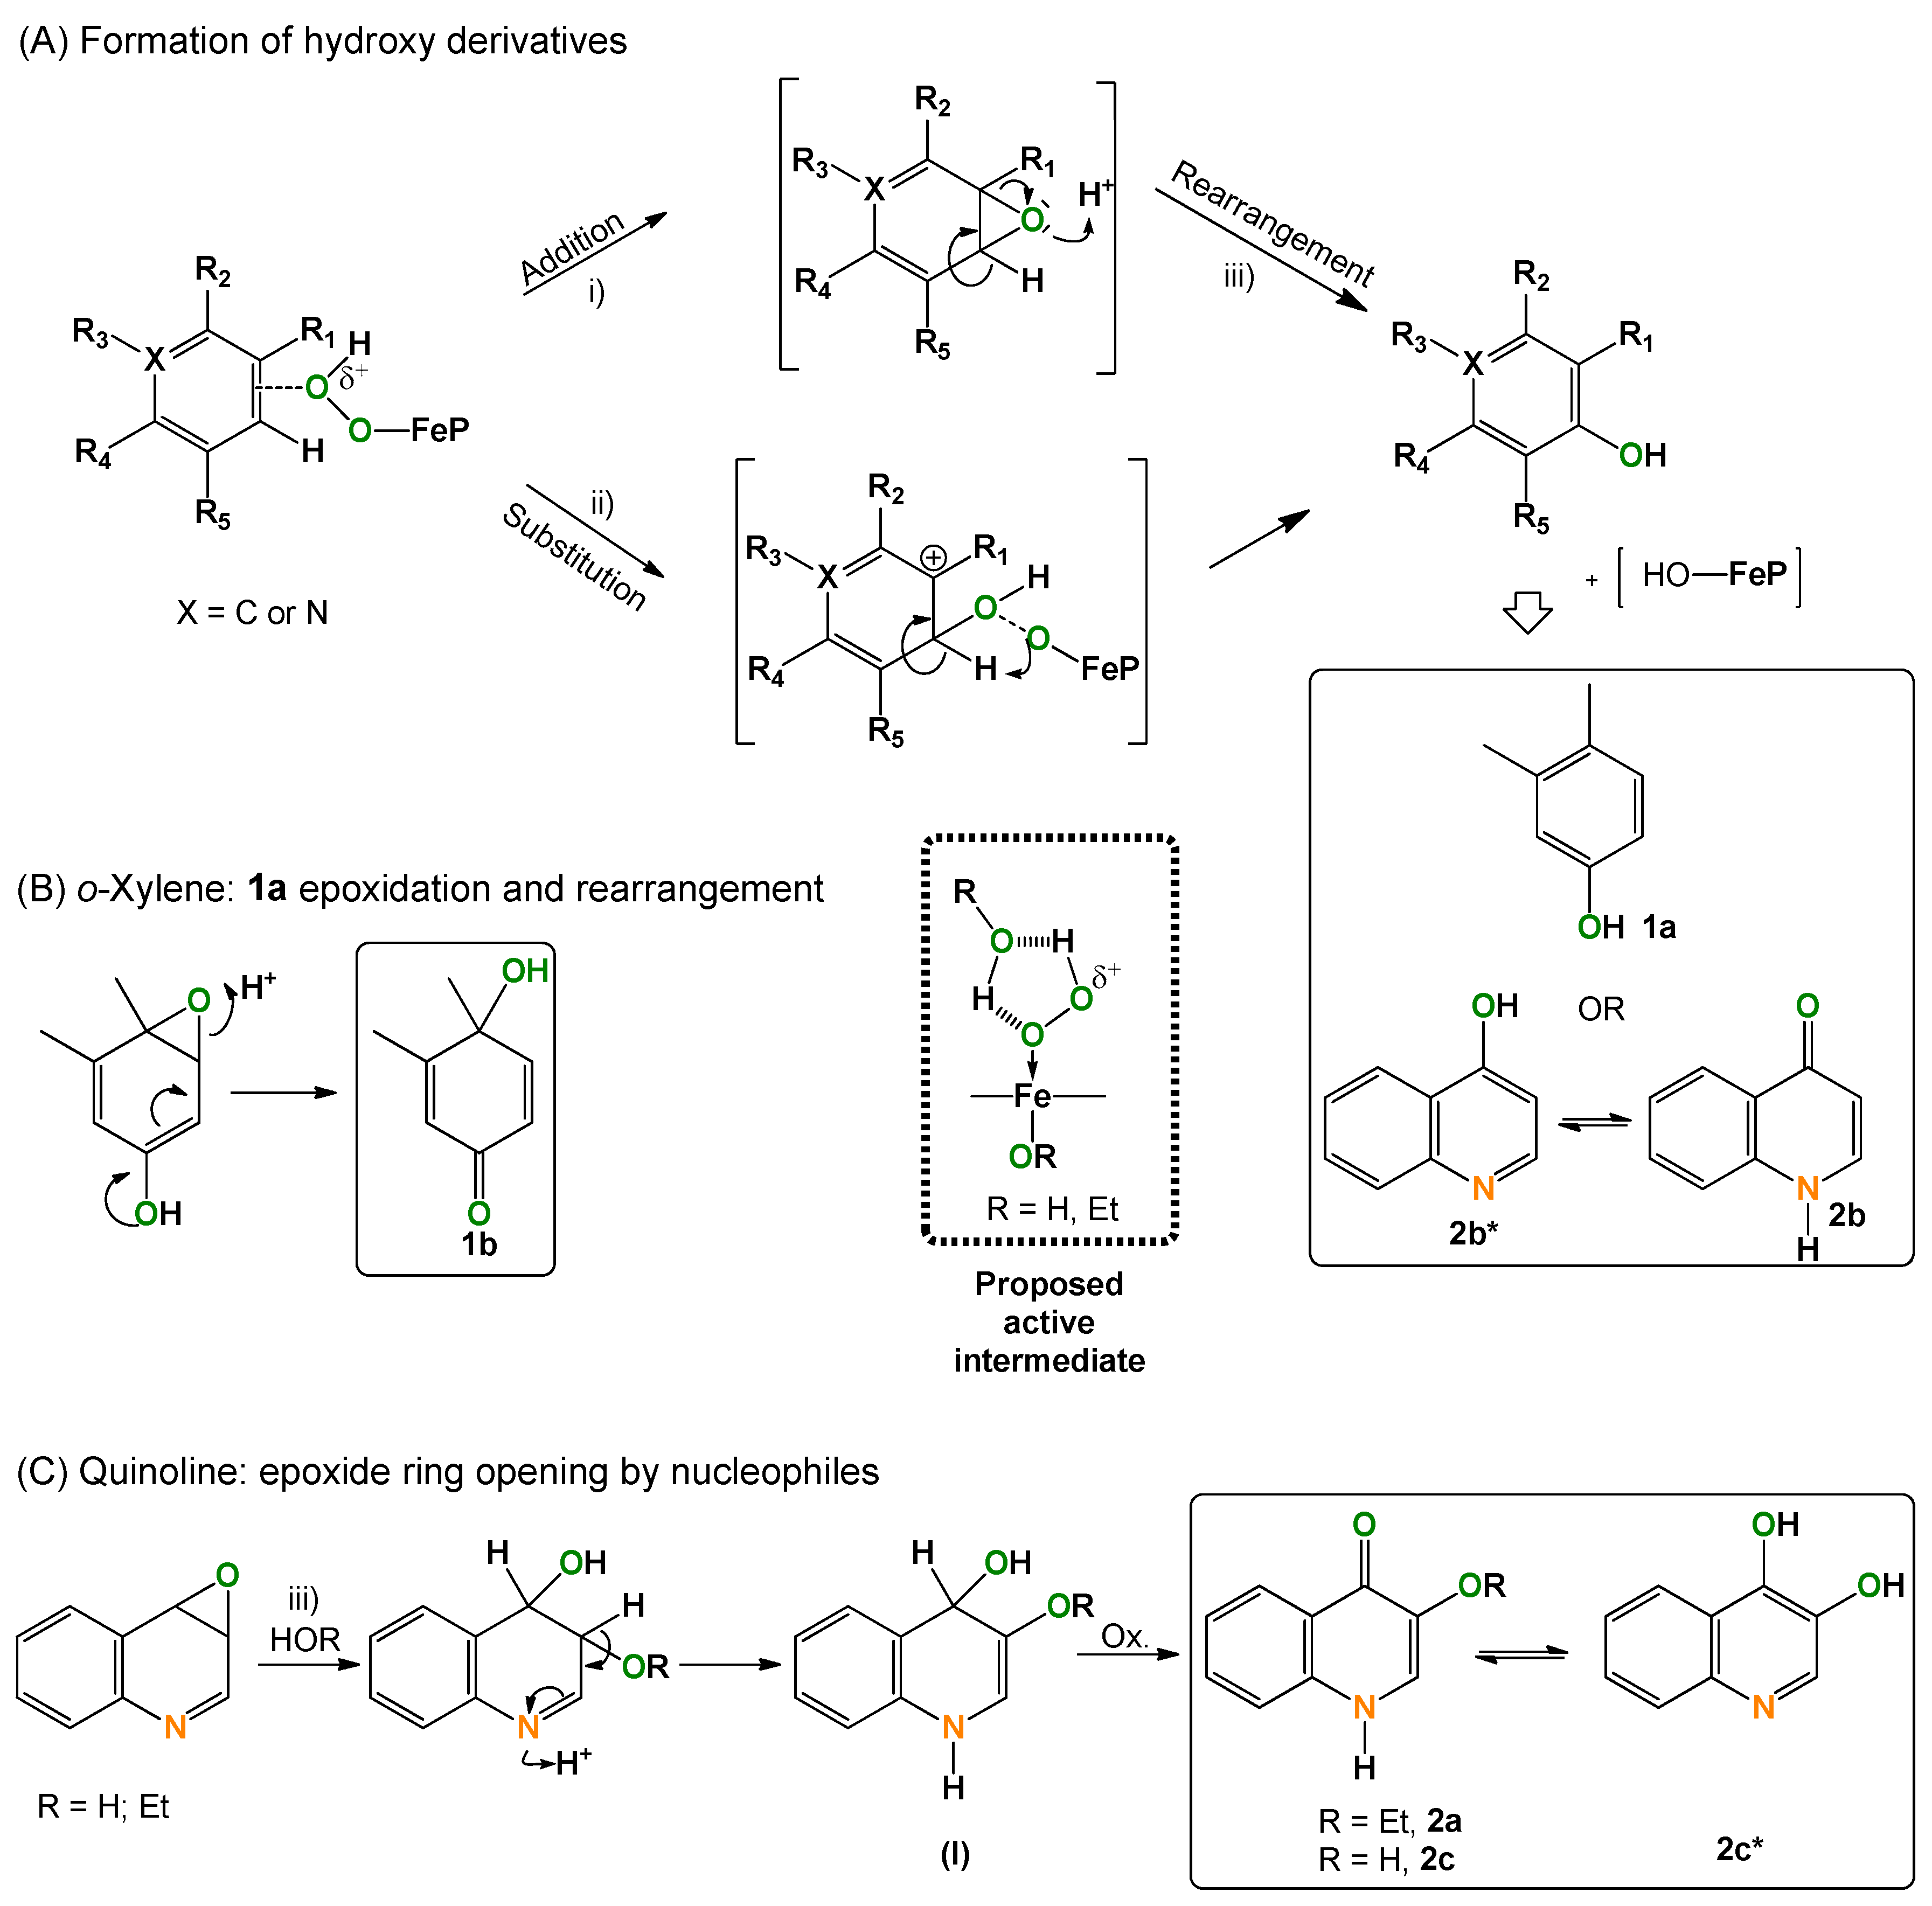
Once you have the epoxide, it becomes a versatile intermediate. Acid-catalyzed ring opening gives you regioselective hydroxylation patterns that are difficult to achieve through direct aromatic substitution. Base-catalyzed opening with nucleophiles like azide or thiolate provides different regiochemical outcomes because the nucleophile attacks the less hindered carbon. This is the step that most procedural guides skip over without explaining why you would choose one conditions set over the other. The aren oxide pathway deserves a mention even though it is less commonly used in my work. Treatment of benzene or substituted benzenes with peroxy acids under specific conditions generates the three-membered oxirane fused to the ring. Benzene oxide tautomerizes to oxepin or rearranges to phenol through the [2,3]-Wittig rearrangement. In practice this means that if you're trying to make phenol from benzene via epoxidation, you are really just doing a Baeyer-Villiger-style oxidation with a poor atom economy compared to the cumene process. What makes epoxidation valuable for aromatic ring synthesis is the ability to generate vicinal diols, amino alcohols, and other functionalized motifs that serve as precursors for ring-contracted or ring-expanded products. The Meinwald rearrangement of aryl epoxides gives you aldehydes and ketones with the aromatic ring intact. From there you can do ring closure reactions or functional group manipulations that build more complex architectures.
Pitfalls I Have Encountered
The biggest issue with epoxidation of aromatic substrates is substrate sensitivity. Phenols and anilines often interfere with mCPBA oxidations because the peracid reacts with the free hydroxyl or amino group. Protecting these groups before epoxidation is necessary, but deprotection afterward can sometimes open the epoxide prematurely. I use silyl ethers for phenols and acetamides for anilines because both survive the epoxidation conditions and can be removed under mild conditions that leave the epoxide intact. Another problem is epoxide polymerization. Certain substrates with adjacent electron-donating groups catalyze cationic ring-opening polymerization during concentration or chromatography. I discovered this when a batch of product turned into a sticky tar on the chromatography column. The fix was to run the purification cold using a pre-chilled column and avoid silica that had been dried at high temperatures, which can introduce trace acidity. Stereochemistry is also a consideration when the alkene is trisubstituted or part of a constrained system. mCPBA delivers the oxygen from the less hindered face, but steric bulk from the aromatic substituent can flip the selectivity in unexpected ways. I verify the stereochemistry of each new substrate using NOE NMR experiments rather than assuming the model will predict correctly.
Alternative Reagents
When mCPBA gives poor results or generates too much m-chlorobenzoic acid waste, I switch to the Jacobsen-Katsuki epoxidation for electron-poor alkenes or the Sharpless epoxidation for allylic alcohols. Both require more expensive catalysts and longer reaction times but give cleaner conversions on problematic substrates. For large scale work, I use hydrogen peroxide with a vanadium catalyst in acetonitrile because the only byproduct is water and the cost per gram of reagent is negligible compared to peracid methods. The tradeoff with aqueous peroxide systems is that water can promote epoxide ring opening during the reaction itself. I use molecular sieves and run the reaction under anhydrous conditions to minimize this, and I monitor the reaction by TLC more frequently than with mCPBA since the kinetics are less predictable.

Practical Notes
Store mCPBA at minus twenty degrees Celsius and check the potency before each use by iodometric titration. Commercial grades vary widely in actual peracid content, and a sample that shows only seventy percent purity will give you reproducibility problems that are hard to diagnose. I prepare a fresh standard solution of sodium thiosulfate and titrate a small aliquot of the peracid against it before starting any reaction that depends on stoichiometric oxidant. For the epoxidation step itself, add the mCPBA solution dropwise to the stirred substrate solution rather than the reverse. Adding substrate to oxidant creates a local excess of peracid that increases the chance of over-oxidation and side reactions. The addition rate should be slow enough that the reaction temperature does not rise more than five degrees above the set point. Characterize the product epoxide by proton NMR, looking for the characteristic signals of the oxirane protons between two and three ppm. The coupling pattern between these protons can tell you about the stereochemistry if the epoxide is disubstituted. Mass spectrometry confirms the molecular weight and the loss of water during ionization is a common artifact that should not be mistaken for a separate product peak.