Why You're Probably Doing Alkylation Wrong
Enolate cross-coupling is not a single reaction. It is a family of transformations where an enolate or enolate equivalent replaces its oxygen-bound counterion with a carbon-based nucleophile through a transition metal catalyzed pathway. The core idea replaces the classical direct alkylation that has been failing you for years, especially when your electrophile is unreactive or your substrate is sensitive. Instead of using a strong base and hoping for the best, you pre-form the enolate, trap it as a metal species, and let the catalyst do the selective carbon-carbon bond formation. The approach has been around since the early 2000s and gained serious traction around 2010 when palladium and nickel catalyst systems became reliable enough for synthetic use. Before that, enolate cross-coupling was mostly a curiosity in the literature with marginal yields and narrow scope.
Unlocking The Secrets Of Enolate Cross Couplings A Comprehensive Guide To The Most Useful Methods
Let me go through the methods that actually work in practice, in the order I reach for them. This is the workhorse. You generate the zinc enolate by treating a ketone or ester with an organozinc reagent or by transmetallation from a magnesium enolate. The most common route uses a silyl ketene acetal precursor or direct deprotonation with diisopropylamine followed by zinc chloride addition. Then you add the electrophile, typically an aryl or vinyl triflate, bromide, or chloride, along with a palladium catalyst. I use this when I need high stereochemical fidelity and the substrate tolerates the mildly basic conditions. The yield is usually in the 70 to 90 percent range for simple substrates. The real advantage over classical enolate alkylation is that you avoid the competing O-alkylation pathway entirely. The zinc enolate is stable enough to isolate and store at low temperature for several hours if your schedule requires it.
The catalyst system matters enormously here. Pd(dba)2 with a bulky phosphine ligand like XPhos or SPhos gives better results than triphenylphosphine for most electrophiles. I found this out the hard way when a reaction with Pd(PPh3)4 gave me 12 percent yield while switching to Pd-118 gave 84 percent on the same substrate. Don't skip the ligand screening step. It will save you weeks of troubleshooting. A practical detail people miss: the zinc enolate should be generated at low temperature, ideally between -78 and -40 degrees Celsius. If you add the electrophile too early or at too high a temperature, you get significant homocoupling of the electrophile and protodezincation side products. I keep the reaction mixture cold until the palladium catalyst is introduced, then allow it to warm to room temperature gradually over about thirty minutes.
Get the Full Details

Suzuki-Miyaura Coupling With Boron Enolates
Boron enolates are commercially available in some cases and easy to generate from silyl enol ethers using catecholboron or pinacolborane with a fluoride source. The Suzuki coupling uses palladium or sometimes nickel catalysis with a base such as potassium carbonate, cesium fluoride, or barium hydroxide. The boron route is attractive because the enolate equivalents are more stable than zinc versions and can be stored as solid boronate esters. The trade-off is that boron enolates are less nucleophilic, so the coupling requires more activated electrophiles. Aryl triflates and iodides work well. Aryl chlorides usually do not unless you use a specialized catalyst system like Pd-PEPPSI or a nickel system with bipyridine ligands. I reached for the Suzuki method last year when my substrate contained a free hydroxyl group that would have been incompatible with the basic conditions of the Negishi approach. The cesium fluoride mediated variant gave clean conversion in tetrahydrofuran at 60 degrees Celsius. The yield was around 71 percent after chromatography. The hydroxyl group stayed put. That would not have worked with the Negishi protocol.
One pitfall with boron enolates is protodeboration. If your workup is too slow or the aqueous phase is too acidic, the boron group falls off and you are left with the starting ketone or ester. I quench with saturated sodium bicarbonate solution instead of dilute acid. It takes one extra minute and prevents the side reaction entirely.
Copper-Catalyzed Conjugate Addition and Alkylation
Copper catalysis is relevant when you are working with enolates derived from -ketoesters or malonates and need to perform a conjugate addition or direct alkylation. The Gilman-type reagents or organocopper enolates couple with allylic or benzylic electrophiles under mild conditions. This method is not as broadly applicable as the palladium routes. It excels with soft electrophiles. Hard electrophiles like primary alkyl bromides tend to give poor yields or require stoichiometric copper, which is wasteful and difficult to remove from the product. The copper route is also the method of choice when you need to preserve an adjacent stereocenter that would epimerize under palladium conditions. I had a case where a secondary alkyl bromide substrate underwent -hydride elimination almost exclusively with palladium. Switching to a copper iodide catalyst with a thioether ligand at 0 degrees Celsius gave the desired product in 63 percent yield with no detectable elimination byproduct. The same substrate with Pd(dppf)Cl2 gave a mixture of elimination and homocoupled material. Copper was the only viable option.
Hiyama Coupling With Silyl Enol Ethers
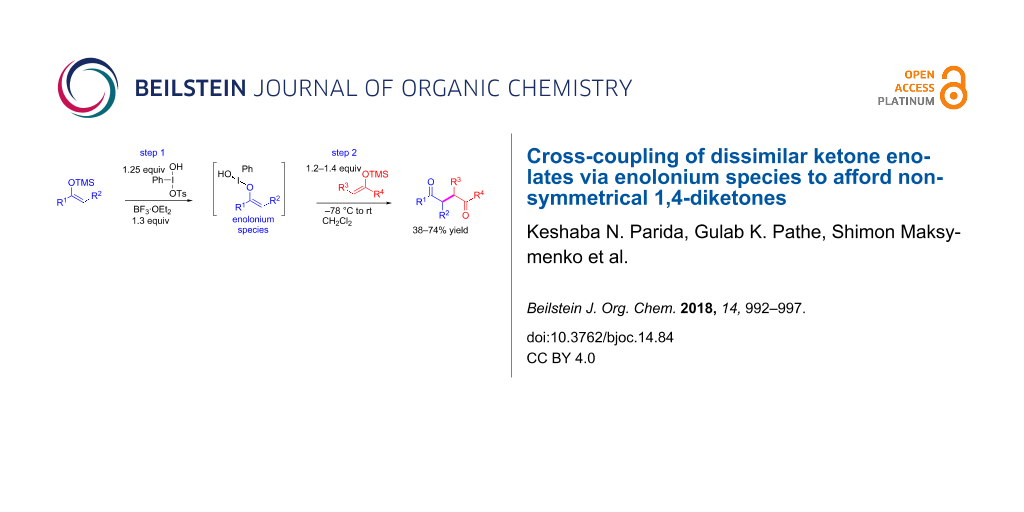
The Hiyama coupling uses silyl enol ethers activated by fluoride or a Lewis acid. The original protocol required stoichiometric fluorosilicate activation, but modern variants use organotrialkylsilane enol ethers with palladium catalysis and minimal activator. The reaction is cleaner than the boron version in terms of byproducts but slower and generally lower yielding. I rarely use this method anymore. The setup is more finicky than the Suzuki alternative and the scope is narrower. The one scenario where Hiyama still makes sense is when your substrate contains functional groups that are incompatible with fluoride activation in Suzuki coupling. Silicon-based activation with TBAF is gentler than cesium fluoride in some cases, though not always.
PRACTICAL PROCEDURE: Negishi-Type Enolate Coupling
Here is the exact procedure I follow for a typical ketone-derived enolate cross-coupling. Adjust the scale and conditions based on your substrate. Dissolve the ketone or ester (1.0 mmol) in anhydrous tetrahydrofuran (5 mL) under nitrogen at -40 degrees Celsius. Add diisopropylamine (1.1 mmol) dropwise. Stir for ten minutes. Add zinc chloride solution (1.2 M in tetrahydrofuran, 1.1 mmol) and stir for another twenty minutes at the same temperature. This generates the zinc enolate in situ. Add the electrophile, typically an aryl triflate (1.2 mmol), followed by the palladium catalyst (2 mol percent Pd-118) and additional ligand if needed (4 mol percent). Warm the reaction to room temperature over forty-five minutes. Monitor by thin layer chromatography. The reaction is usually complete within one to two hours.
Quench with saturated ammonium chloride solution. Extract with ethyl acetate. Wash the organic layer with brine, dry over magnesium sulfate, filter, and concentrate. Purify by column chromatography on silica gel using hexanes and ethyl acetate as the eluent. This sequence typically takes about two hours from start to workup. The yield depends heavily on the electrophile and the substrate electronics. Electron-rich aryl triflates couple faster. Electron-poor ones can be sluggish and may require extended reaction times or higher catalyst loading.

Common Failure Modes and How to Fix Them
The most common problem is complete lack of reactivity. This usually means the electrophile is not activated enough. Switch from a chloride to a triflate or use a more reactive palladium catalyst system. If the electrophile is already a triflate and nothing happens, the enolate generation step is likely incomplete. Check the temperature and the order of addition. The base must precede the zinc source. Another frequent issue is protodehalogenation of the electrophile. The aryl or vinyl halide converts back to the arene or alkene without coupling. This indicates the zinc enolate is decomposing before it can transmetallate. Lower the temperature during enolate formation and ensure the zinc salt is freshly prepared. Old zinc chloride solutions absorb moisture and lose reactivity. -Hydride elimination is the third major problem, especially with secondary alkyl electrophiles. The palladium intermediate undergoes elimination instead of reductive elimination, giving an alkene byproduct and recovering the starting material. Switching to a bulkier ligand or a nickel catalyst can suppress this pathway. In some cases, changing the solvent to toluene or 1,4-dioxane instead of tetrahydrofuran improves selectivity.
I encountered a stubborn case where the desired cross-coupling product formed in only 28 percent yield alongside 41 percent -hydride elimination product. The substrate was a secondary alkyl triflate with an adjacent stereocenter. After trying three different ligand systems, I switched to a nickel bromide catalyst with 4,4-di-tert-butyl-2,2'-bipyridine. The yield jumped to 79 percent with less than 5 percent elimination. The change in metal and ligand altered the oxidative addition and reductive elimination kinetics enough to favor coupling over elimination.
When These Methods Fail Completely
Enolate cross-coupling is not a universal solution. It fails with certain substrate classes. Sterically hindered enolates from disubstituted ketones at the -position resist enolate formation and transmetallation. Highly electron-deficient electrophiles can coordinate to the catalyst and poison it. Substrates with multiple reactive functional groups, especially free carboxylic acids and unprotected amines, complicate the reaction significantly and often require protecting group strategies that add steps and reduce overall yield. If your substrate has a free carboxylic acid near the reaction center, the carboxylate will coordinate to the metal catalyst and inhibit the coupling cycle. I typically protect it as a methyl or benzyl ester before attempting the cross-coupling. The extra two steps are usually worth it compared to failing the reaction entirely.

Alternative Approaches When Cross-Coupling Is Not Viable
If enolate cross-coupling does not work for your substrate, consider the direct alkylation route with phase transfer catalysis. This is older and less selective but works for simple substrates. Another option is the Mannich-type reaction using activated imines as electrophiles. For particularly challenging cases, the Mukaiyama aldol reaction with a silyl enol ether and a Lewis acid can achieve similar structural outcomes through a different mechanistic pathway. The direct aldol condensation using stoichiometric base remains the simplest approach when selectivity is not a concern. It is fast, inexpensive, and requires no transition metal catalysts. The downside is poor control over regiochemistry and stereochemistry with unsymmetrical ketones. My general rule is to attempt enolate cross-coupling first for substrates that require precise stereochemical control or contain sensitive functional groups. For simple alkylations of symmetrical enolates, classical methods are sufficient and faster. The cross-coupling approach is not inherently superior in all cases. It is superior in specific cases where the alternative methods fail due to selectivity or functional group compatibility issues.
The landscape has improved significantly since these methods were first reported. Modern catalyst systems handle substrates that would have been impossible ten years ago. But the core principles remain the same: control the enolate generation, choose the right metal and ligand for your electrophile, and watch for the predictable failure modes before they consume your time.

