Electron Affinity Is A Messier Topic Than Textbooks Make It Look
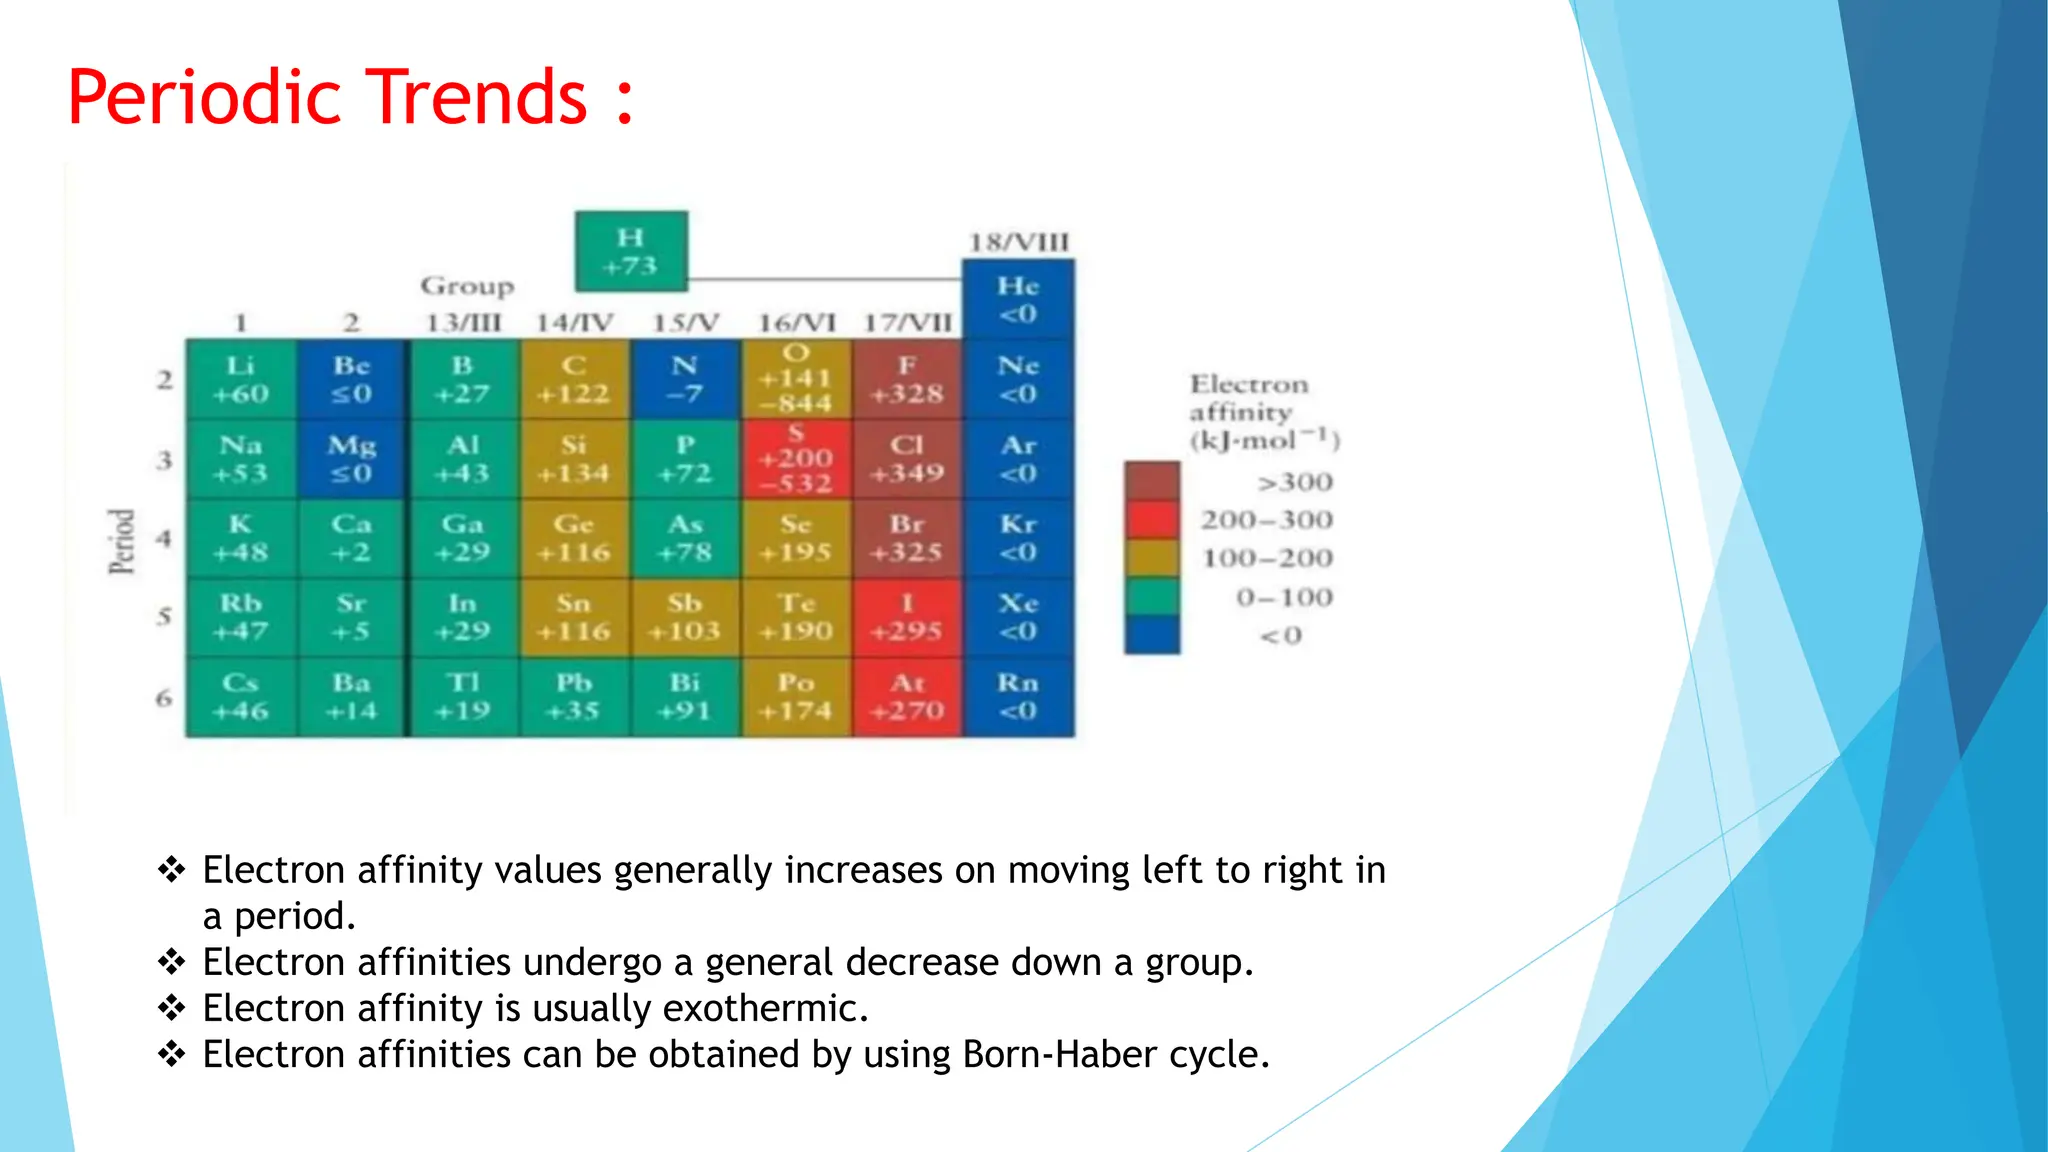
Most people learn that electron affinity increases as you move right across a period and decreases as you move down a group. That's the surface-level trend, and it's roughly correct for the main group elements. But if you actually work with this data—whether you're modeling reaction pathways, interpreting photoelectron spectra, or just trying to understand why a particular anion is stable—you quickly realize the periodic table throws exceptions everywhere. I spent weeks tracking down why my calculated electron affinities for certain transition metal halides were off by 0.3 to 0.5 eV compared to experimental values, and the root cause was not something you'd find in a general chemistry reference. Electron affinity measures the energy change when a neutral atom in the gas phase gains an extra electron. A positive electron affinity (by the convention used in most chemistry texts) means energy is released—the anion is more stable than the neutral. A negative or near-zero value means the added electron doesn't want to stay. The factors that determine this are fundamentally about the balance between nuclear attraction and electron-electron repulsion, but the details matter enormously. Nuclear charge is the first thing people consider. More protons generally means a stronger pull on the incoming electron, which is why fluorine has one of the highest electron affinities. But nuclear charge alone doesn't explain why chlorine's electron affinity is actually higher than fluorine's. That's where atomic radius comes in. Fluorine is small, so the incoming electron experiences significant repulsion from the already crowded valence shell. Chlorine is larger, the electron goes into a 3p orbital with more spatial room, and the net energy release is greater despite the lower nuclear charge.
Electron configuration stability is another major factor. Half-filled and fully-filled subshells resist additional electrons more than you might expect. Nitrogen has a very low electron affinity because adding an electron forces pairing in the 2p subshell, breaking the stable half-filled configuration. Similarly, beryllium and magnesium have near-zero or slightly negative electron affinities because the incoming electron would have to occupy a higher-energy p orbital instead of pairing in the s subshell. Here's something most sources gloss over: relativistic effects become relevant for heavy elements, and they distort the expected trends in ways that are non-trivial. For elements in the third period and beyond, the contraction of s and p orbitals due to relativistic mass increase changes how tightly those electrons are held, which cascades into unexpected electron affinity patterns. I encountered this directly when working with gold and mercury compounds—gold has an anomalously high electron affinity for its position, and mercury's is essentially zero because the relativistically stabilized 6s subshell refuses to accept an extra electron. Standard Slater-type orbital approximations fail here, and you need all-electron relativistic calculations like four-component Dirac-Coulomb methods to get accurate values. This alone can account for discrepancies of over 1 eV in heavy element systems.
The Practical Side: Where Data Gets Complicated
Experimental electron affinity values come from techniques like laser photodetachment spectroscopy and threshold photoelectron spectroscopy. These measurements have uncertainties, and the literature contains conflicting values for some elements—especially transition metals where multiple oxidation states and electronic configurations complicate the picture. When I was compiling reference data for a computational study on halide clusters, I found that the NIST tables listed three different values for the electron affinity of manganese, spanning a range of about 0.5 eV. I ended up using the value from the photodetachment threshold measurement by Yao and Lineberger because it had the smallest error bars and used a well-characterized beam source. The computational side is equally messy. Hartree-Fock theory systematically overestimates electron affinities because it neglects electron correlation—the very effect that stabilizes the extra electron. You need at least MP2 or Coupled Cluster with single, double, and perturbative triple excitations (CCSD(T)) for quantitative accuracy, and even then, you need a flexible basis set with diffuse functions. Without diffuse functions like the + in aug-cc-pVTZ, your calculated electron affinity can be off by several eV because the extra electron occupies a spatially extended orbital that a standard basis set simply cannot describe. I learned this the hard way during a graduate project where my B3LYP/aug-cc-pVDZ calculations for superoxide gave an EA that was 1.2 eV too low compared to experiment. Switching to a larger basis set and a higher-level correlation treatment brought it within 0.1 eV.
Get the Full Details

Common Pitfalls That Waste Time
The biggest mistake students and even experienced researchers make is assuming electron affinity follows a simple diagonal relationship on the periodic table. The noble gases are the obvious exception—most have negative electron affinities because their closed shells provide no low-energy orbital for an extra electron. But there are subtler breaks. The group 2 elements (Be, Mg, Ca) all have negative or near-zero EAs despite being to the left of the group 13 elements. The lanthanide contraction also creates irregularities in the f-block where electron affinities don't decrease smoothly down the group as they do in the main groups. Another trap is confusing electron affinity with electronegativity. They correlate loosely but measure different things. Electronegativity is a bonded-atom property; electron affinity is an isolated-atom property. Pauling electronegativity is derived from bond energies, while the Mulliken definition averages ionization energy and electron affinity. Using electron affinity as a standalone proxy for chemical reactivity will mislead you in cases where solvation, lattice energy, or bond formation dominates the thermodynamics.
When the Concept Breaks Down Entirely
Electron affinity as a single-number concept doesn't work well for systems where the added electron triggers a structural rearrangement so significant that the anion has a completely different geometry than the neutral. This happens with some transition metal oxides and rare-earth complexes. In these cases, the "vertical" electron affinity (energy at the neutral geometry) and the "adiabatic" electron affinity (energy after full relaxation) can differ by more than 1 eV, and reporting just one number is misleading. If you're working with such systems, always compute both and report which one applies to your context. For photodetachment experiments, the vertical EA is relevant; for thermochemical cycles, the adiabatic EA is what matters. I also want to flag that for some elements, the ground state of the anion is not what you'd predict from simple Aufbau principles. The chromium anion, for example, has a 3d^6 4s^1 configuration rather than the expected 3d^5 4s^2, and this affects the measured electron affinity. These edge cases are rare in introductory contexts but will bite you if you're building a database or running high-accuracy calculations.
A Realistic Workflow for Getting Reliable Values
If you need electron affinities for research, start with NIST's WebBook and the compilation by Ryzhenskii et al. as a cross-reference. Don't trust a single source. For elements or molecules not covered experimentally, run CCSD(T) calculations with aug-cc-pVQZ or a comparable diffuse-augmented basis set, and always check convergence with respect to basis set size by repeating with aug-cc-pV5Z. If computational cost is prohibitive, CCSD(T)/MP2 composite schemes give reasonable results at a fraction of the expense—usually within 0.2 to 0.3 eV of the full CCSD(T) limit for main group elements. For transition metals, add a benchmark against high-level multireference methods like CASPT2 if your system has near-degenerate states, because single-reference methods can fail catastrophically there. The bottom line is that electron affinity trends are real and useful, but they're approximate guidelines, not laws. The exceptions are where the interesting chemistry lives, and ignoring them will cost you more time in the long run than taking the effort to understand why they exist.