The Chemistry Behind the Cycle
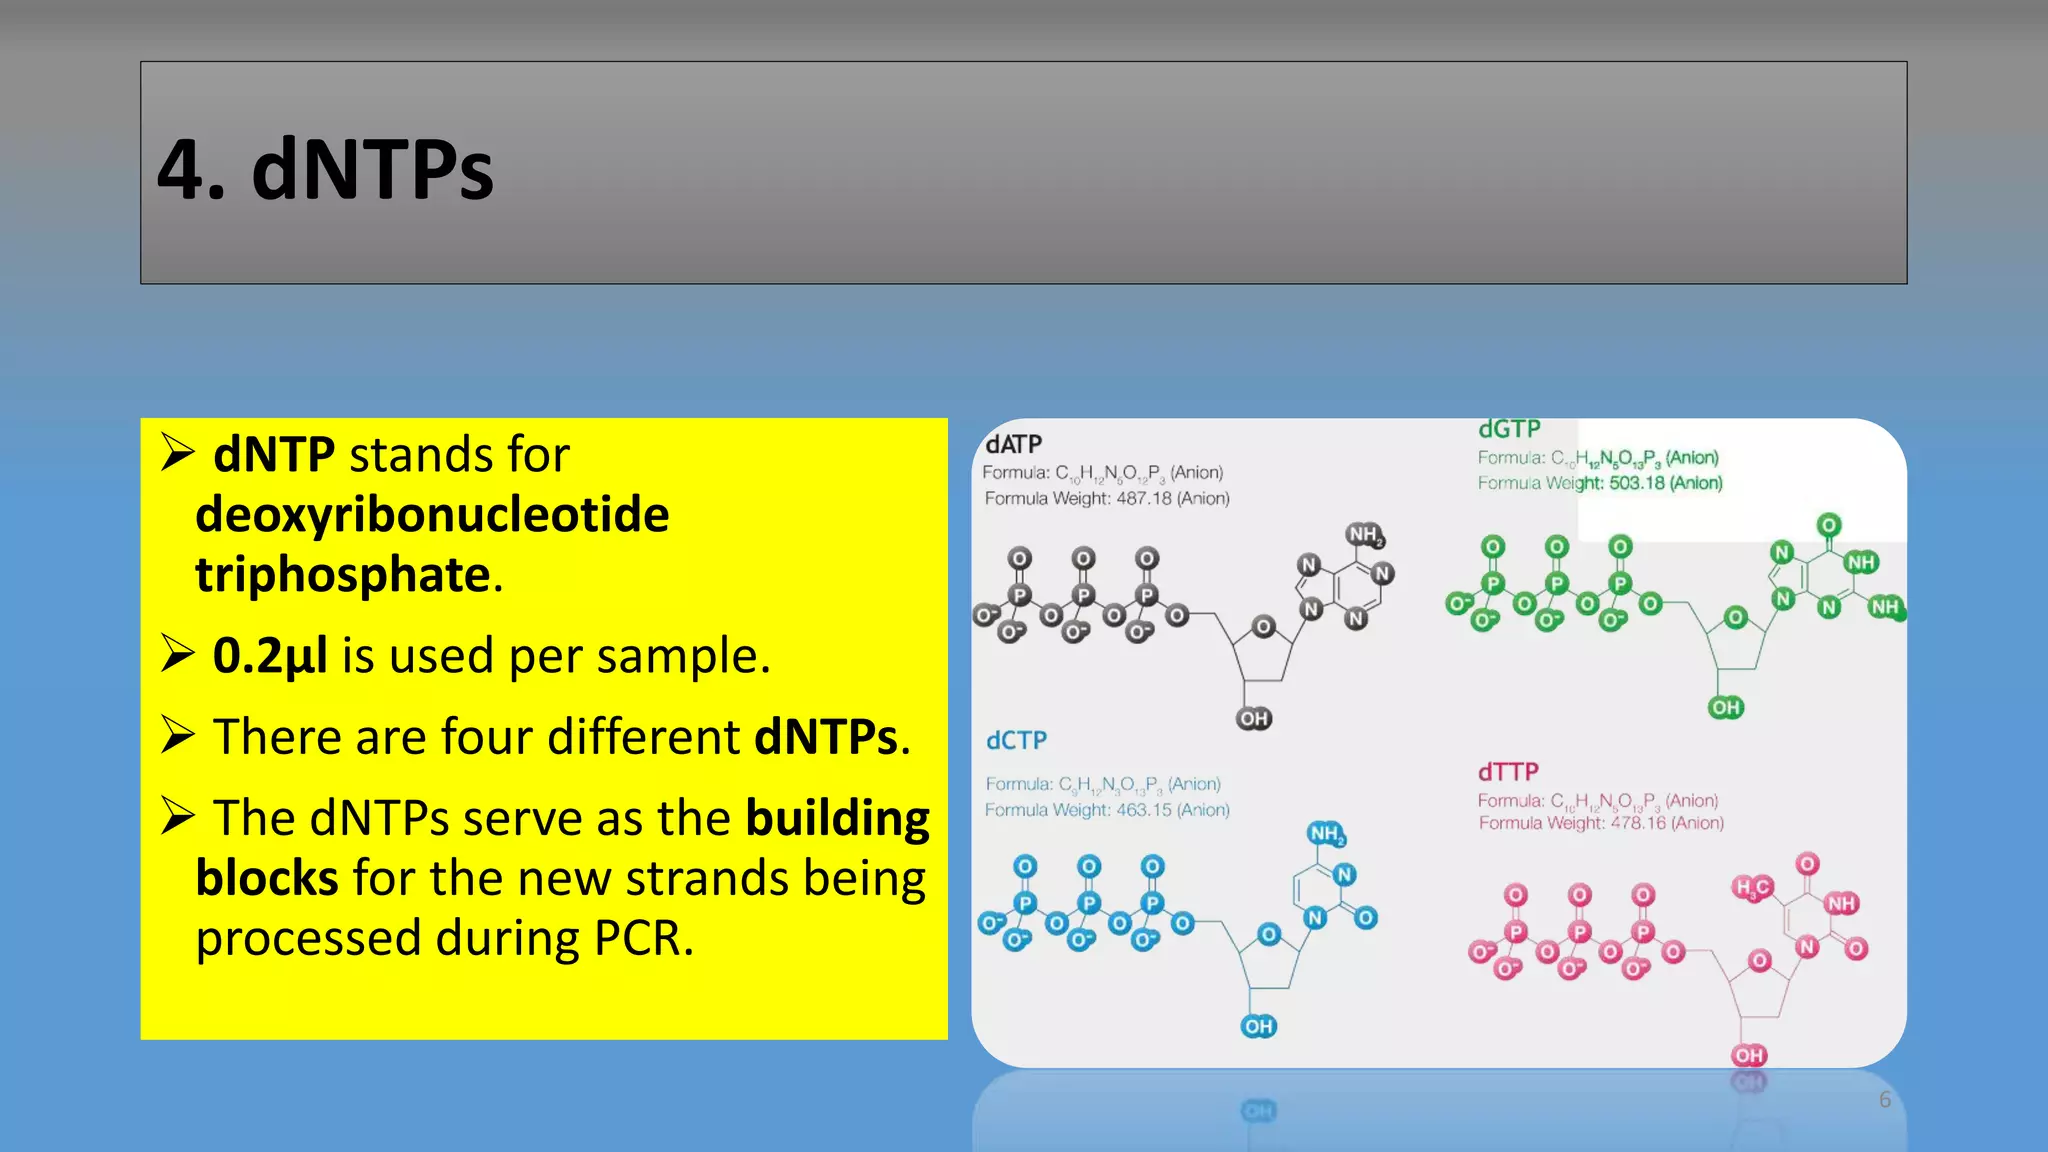
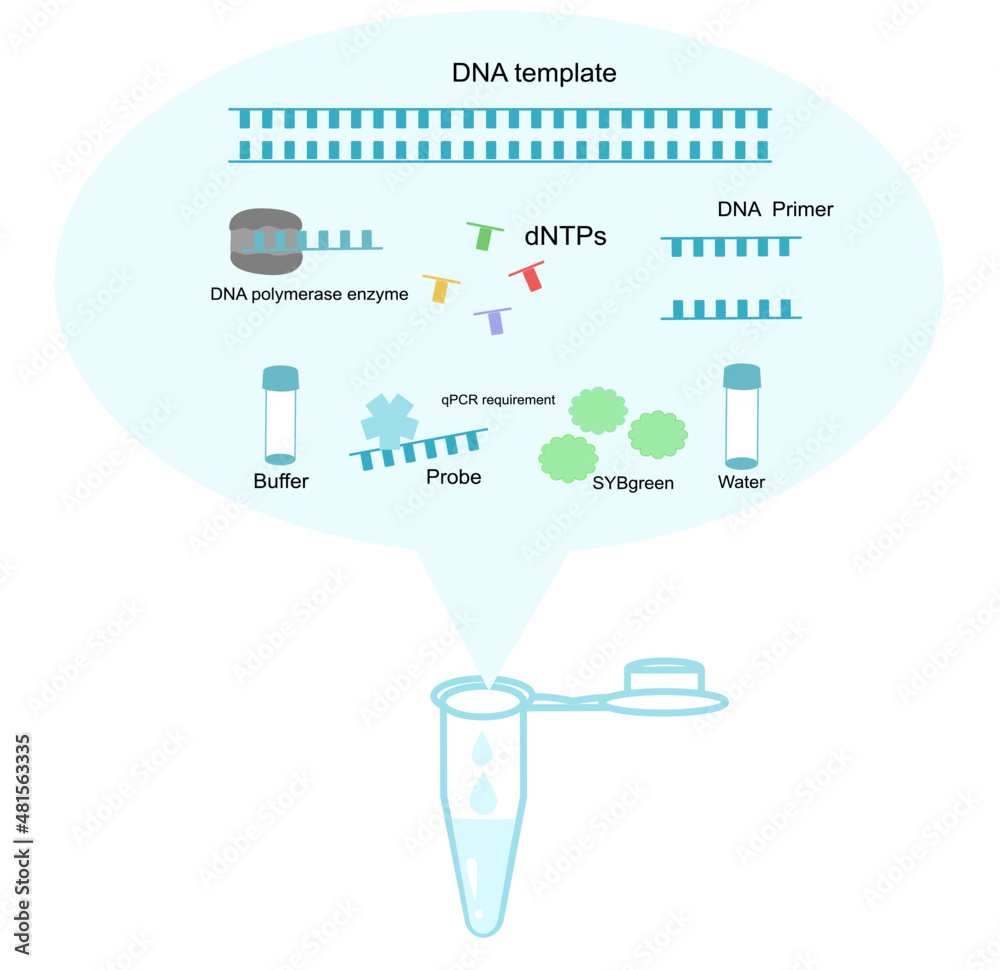
Thermal cyclers do one thing repeatedly. They heat and cool a tube full of water, salts, primers, template, and four nucleotide triphosphates. Everything else is theater. If you have understood dNTPs, you have understood the rate-limiting step of polymerase chain reaction. The acronym stands for deoxyribonucleoside triphosphates. That means dATP, dTTP, dGTP, and dCTP. Each carries three phosphate groups. During extension, the polymerase cleaves off the alpha and beta phosphates as pyrophosphate. The remaining alpha phosphate forms a phosphodiester bond with the growing strand. This releases energy and drives the reaction forward. Without the triphosphate, you have monophosphate nucleotides, which cannot be incorporated. The difference matters enormously in practice.
Why Concentration Is Non-negotiable
Most commercial PCR kits supply 10 mM of each dNTP in a mixed solution. You typically add 0.2 mM final concentration of each dNTP per reaction. That means a 50 microliter reaction receives 1 microliter of the 10 mM mix. Simple bookkeeping. The problem is that this number shifts when you change anything else. High-fidelity enzymes like Q5 or Phusion tolerate lower dNTP concentrations than Taq. Taq needs 0.2 mM for robust activity. These modified enzymes perform better at 0.05 to 0.1 mM. The reason relates to fidelity. Lower dNTP concentrations reduce misincorporation rates because the polymerase has more time to reject mismatched nucleotides before phosphodiester bond formation locks the error in place. I learned this the hard way when switching protocols without adjusting the mix. My first run with Phusion at standard 0.2 mM dNTPs showed a threefold increase in point mutations across 800 base pairs. Dropping to 0.05 mM cut the error rate by half again. That change came from consulting the manufacturer datasheet, not from any textbook recommendation. Another consequence you will encounter involves reaction volume scaling. When you move from 50 microliters to 100 microliters, you must double the dNTP volume, not just add more template or enzyme. The molar concentration must remain constant or the kinetics change. A common mistake is keeping the same dNTP volume while doubling everything else. This halves the effective dNTP concentration and kills yield. I fixed this by always calculating millimolar final concentration rather than trusting volume ratios. It takes thirty seconds and prevents a full troubleshooting cycle.
Balance Matters More Than Total Amount
Equal concentrations of all four dNTPs is the gold standard. 0.2 mM each means 0.8 mM total. Unequal ratios cause problems. Excess dATP and dTTP relative to dGTP and dCTP favors AT-rich regions during amplification but stalls at GC tracts. The polymerase encounters fewer matching partners and slows down. Conversely, excess dGTP and dCTP creates misincorporation bias because the enzyme incorporates G and C more readily even at mismatched positions. The real issue is that dNTPs compete with each other at the active site. Polymerases have a selectivity filter that measures geometric compatibility between the incoming nucleotide and the template strand. When one species dominates, the competition shifts. Misincorporation rates increase for the abundant nucleotide. This becomes critical during long amplicons. A 3 kilobase product with unequal dNTPs shows progressive truncation. The polymerase falls off more frequently when it encounters homopolymer runs of the limiting nucleotide. I encountered this with a primer set designed for a 2.5 kilobase GC-rich target. The amplification failed completely until I rechecked the dNTP mix. The supplier had shipped a batch where dGTP was at 7.5 mM instead of 10 mM due to a calibration error. Running the reaction with corrected equimolar dNTPs restored yield. The fix took two hours. Preventing the problem takes five minutes of verifying the certificate of analysis on incoming reagent batches.
Get the Full Details

Unlocking The Secrets Of Dna The Crucial Role Of Dntps In Pcr
The secret is not complicated. dNTPs are the substrate. Substrate concentration determines reaction velocity according to Michaelis-Menten kinetics. The Km value for most polymerases ranges from 10 to 50 micromolar. At 0.2 mM, you are operating well above saturation. The reaction proceeds at near Vmax. Going much higher provides no benefit and introduces new problems. dNTPs degrade over time. Hydrolysis cleaves the terminal phosphates. This happens faster at room temperature and in acidic conditions. The recommended storage is minus twenty degrees Celsius in aliquots. Thawing and refreezing accelerates degradation. Each freeze-thaw cycle reduces concentration by approximately 5 percent. After ten cycles, your stock may be at 60 percent of labeled concentration. This matters when you are running sensitive applications like quantitative PCR or cloning. I stopped relying on stored dNTP stocks after my sequencing results showed increasing heterogeneity across replicate runs. The problem was not the polymerase or the primers. The dNTP mix had degraded. Running a fresh aliquot restored clean chromatograms. The workaround was to purchase smaller volume stocks and use them within six months. The cost increase was negligible compared to wasted reagents and time.
Anion exchange chromatography can purify degraded dNTPs. This removes hydrolysis products like dNMP and pyrophosphate. The process takes approximately forty-five minutes using a small spin column. The recovered yield is usually 80 to 90 percent of original concentration. This is practical when you have a large investment in a bulk stock and cannot afford to replace it immediately.
Interference from Chelators
Magnesium ions are essential cofactors. The polymerase requires Mg2+ at the active site to coordinate the incoming dNTP and the primer terminus. EDTA and other chelators bind magnesium and inhibit the enzyme. If your buffer contains EDTA, you must compensate by increasing magnesium concentration. Typical PCR buffers supply 1.5 mM MgCl2. With 0.2 mM dNTPs, approximately 0.4 mM magnesium becomes sequestered by the triphosphate groups. This reduces free magnesium to about 1.1 mM, which may be below the optimal range for some polymerases. The solution is to add 0.5 to 1.0 mM extra MgCl2 when running standard dNTP concentrations. This ensures sufficient free magnesium for catalysis. I discovered this during a troubleshooting session where my GC-rich templates refused to amplify despite optimal primer design and cycling conditions. Increasing MgCl2 from 1.5 mM to 2.5 mM restored amplification. The insight came from calculating magnesium binding by dNTPs rather than guessing.
Common Pitfalls
Using dUTP instead of dTTP is a frequent contamination source. Some protocols deliberately substitute dUTP to prevent carryover contamination through uracil DNA glycosylase treatment. If you accidentally use a dUTP-containing mix in a standard Taq reaction, the polymerase incorporates uracil into the product. This does not affect amplification. It becomes a problem during cloning or sequencing because uracil triggers repair enzymes and causes fragmentation. I made this error once by confusing two similar looking vials. The fix was to rerun the PCR with a fresh dTTP-containing mix. The lesson was to label every aliquot clearly and store dUTP and dTTP stocks in separate compartments of the freezer. Another pitfall involves assuming all dNTP mixes are equal. Different manufacturers use different buffers. Some include stabilizers like trehalose or glycerol. Others contain trace amounts of contaminants that inhibit certain polymerases. When switching brands, I always run a control amplification before committing to expensive downstream applications. This takes one hour and prevents days of troubleshooting.
When dNTPs Are Not the Solution
No-templete control amplification indicates contamination, not dNTP issues. Adding more dNTPs will not fix this. Similarly, non-specific bands result from primer dimer formation or insufficient annealing temperature. No amount of additional substrate corrects these problems. The troubleshooting path should address the actual cause rather than adjusting dNTP concentration. For metagenomic or low-input samples, dNTP concentration may need reduction rather than increase. Competitive inhibition from diverse template sequences requires lower substrate concentrations to maintain specificity. I reduced dNTPs to 0.025 mM each when amplifying environmental DNA with degenerate primers. This improved specificity at the cost of yield. The tradeoff was acceptable because subsequent cloning and sequencing provided sufficient coverage. Sanger sequencing reactions use a completely different nucleotide mix. Dideoxynucleotides terminate chain elongation. Using standard dNTPs in a sequencing reaction produces full-length products instead of terminated fragments. The result is unreadable chromatograms. This is not a dNTP problem. It is a protocol mismatch. Always verify which nucleotide mix a protocol requires before setting up the reaction.
The fundamental principle remains constant regardless of application. dNTPs are the building blocks. Their concentration, balance, and integrity determine reaction efficiency and fidelity. Understanding this chemistry prevents most common PCR failures. The rest is optimization within the constraints of enzyme kinetics and template complexity.