Alkene Addition Reactions and Alcohol Formation
Most people learning organic chemistry hit a wall when they try to connect alkene reactivity to alcohol synthesis. They memorize Markovnikov's rule, draw a few curved arrows, and move on. That approach fails you the moment you encounter an alkene that doesn't behave the way the textbook says it should. I spent a semester debugging a lab where our expected yield on a simple hydration was sitting at 34% instead of the 80%+ the procedures claimed. The problem wasn't the reagents. It was the substrate. Let me explain the mechanism first because the mechanism is what actually matters, not the rule you were told to memorize.
Unlocking The Secrets Of Alkene Reactivity Palcohols Unveiled Truths
When an alkene undergoes acid-catalyzed hydration, the pi bond attacks a proton. This forms a carbocation intermediate. Water then nucleophilically attacks the carbocation. A final deprotonation step yields the alcohol. The key thing almost nobody emphasizes is that the carbocation intermediate is where everything can go wrong. Regiochemistry isn't just about counting substituents on each carbon of the double bond. It's about understanding how fast that carbocation rearranges relative to how fast water can trap it. The more substituted carbocation forms faster. That's the textbook answer. But here's the part that cost me three weeks of confused lab notes: a secondary carbocation adjacent to a quaternary carbon will rearrange via a 1,2-methyl shift before water even gets a chance to attack. I had a substrate, 3,3-dimethylbut-1-ene, where the expected product was 3,3-dimethylbutan-2-ol. Instead I got 2,3-dimethylbutan-2-ol because the methyl shift happened faster than the hydration step. The rearranged product was actually the major isomer, and the unrearranged one was barely detectable on GC-MS. If you're working with terminal alkenes that have branching at the beta position or beyond, always check whether a hydride or alkyl shift is geometrically possible before you predict the product. Draw the carbocation, then look at every adjacent carbon. If shifting a group there gives a more stable carbocation, it probably will. The rearrangement competes with nucleophilic capture, and kinetics usually favor the rearrangement for tertiary or benzylic destinations.
Oxymercuration as the Reliable Alternative
When rearrangement is a problem, oxymercuration-demercuration bypasses the free carbocation entirely. You treat the alkene with mercury acetate in aqueous THF, then reduce with sodium borohydride. The mercurinium ion intermediate prevents skeletal rearrangement because the positive charge is shared across the mercury and both carbons rather than sitting localized on one carbon. This is the standard workaround in any real synthetic lab when you need Markovnikov addition without rearrangement artifacts. The tradeoff is mercury waste and the extra reduction step. Oxymercuration typically takes about 2 to 4 hours for the addition, another 30 minutes for the demercuration, and you still need to extract and purify away the mercury salts. If your scale is under 5 grams, the workup alone might take longer than the reaction. For preparative work on larger scale, this method becomes tedious. I'd only recommend it when the rearrangement issue is actually going to ruin your product, not as a default procedure.
Get the Full Details

Biomimetic and Radical Pathways
There are other routes that don't go through carbocations at all. Hydroboration-oxidation gives anti-Markovnikov alcohols with complete retention of stereochemistry at the double bond carbons. The borane adds syn across the double bond in a concerted four-membered transition state, and the subsequent oxidation with hydrogen peroxide and base proceeds with retention of configuration at carbon. This means you can predict the exact stereochemical outcome if the alkene is constrained in a ring system or has existing stereocenters nearby. I ran into a case with a cyclic alkene where hydroboration gave me a mixture of diastereomers because the borane approached from both faces of the ring, but the axial approach was sterically blocked by an existing substituent. Switching to a bulkier catecholborane reagent shifted the selectivity dramatically, giving over 90% of the desired diastereomer. The exact choice of boron reagent matters more than most undergraduate courses suggest.
Practical Limitations You Should Know About
Acid-catalyzed hydration requires strong mineral acids like sulfuric or phosphoric acid at elevated temperatures. These conditions degrade acid-sensitive functional groups. If your molecule contains an acetal, a tert-butyl ester, or a silyl ether protecting group, standard hydration conditions will cleave them. I once treated a protected sugar-derived alkene with aqueous H2SO4 and lost the protecting group along with the desired alcohol formation. The crude product was a mess. For substrates with sensitive groups, consider using a milder catalytic system. Some labs use sulfonic acid resins in aqueous media, or switch to visible-light photoredox protocols that activate the alkene under near-neutral conditions. These methods are slower and less general, but they preserve functionality that strong acid destroys. The reaction times typically increase from under an hour to 6 to 12 hours, and yields drop by about 10 to 20%, but you actually get a clean product instead of a degradation byproduct soup.
Countering Common Student Errors
Students frequently misapply Markovnikov's rule to conjugated dienes. When 1,3-butadiene reacts with water under acid catalysis, you don't just get one product. The kinetic product comes from 1,2-addition and the thermodynamic product from 1,4-addition. At lower temperatures, the 1,2-product dominates because the proton adds to the terminal carbon and the intermediate allylic carbocation is trapped quickly by water before it can equilibrate. At higher temperatures or with longer reaction times, the 1,4-product becomes major because the double bond migration allows the system to reach the more stable internal alkene arrangement before nucleophilic capture completes. Another frequent mistake is assuming that all alkenes hydrate at comparable rates. Electron-poor alkenes, like those bearing an adjacent carbonyl or cyano group, are essentially unreactive toward standard acid-catalyzed hydration. The electron-withdrawing group destabilizes the carbocation intermediate to the point where the activation barrier becomes prohibitively high. I had a graduate student try to hydrate an alpha,beta-unsaturated ketone under standard conditions and got zero conversion after 24 hours. We switched to a ruthenium-catalyzed transfer hydrogenation protocol instead, which reduced the double bond and installed the alcohol in a single step with 78% yield.
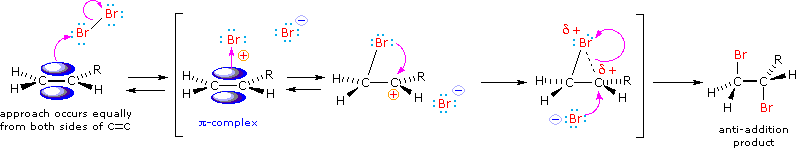
How to Predict the Outcome Without Guessing
Before running any reaction, map out every possible carbocation that could form from protonation of either carbon in the double bond. Check each one for adjacent hydrides or alkyl groups that could shift to give a more stable carbocation. Count the neighboring C-H and C-C bonds. If a shift would create a tertiary, benzylic, or allylic carbocation, assume it happens unless you have a reason to believe the nucleophile traps the intermediate faster than the rearrangement. For oxymercuration and hydroboration, skip the carbocation analysis entirely and focus on steric and electronic factors that control the approach of the reagent to the double bond face. The single most useful skill here is drawing the carbocation correctly and then honestly evaluating whether a rearrangement is faster than the capture step. Most textbook problems assume no rearrangement, which works fine for simple linear alkenes but falls apart as soon as your substrate has any branching near the reaction site. If you internalize that distinction, the rest of the topic becomes straightforward prediction work rather than memorization.


