How to Actually Work With Second Order Kinetics
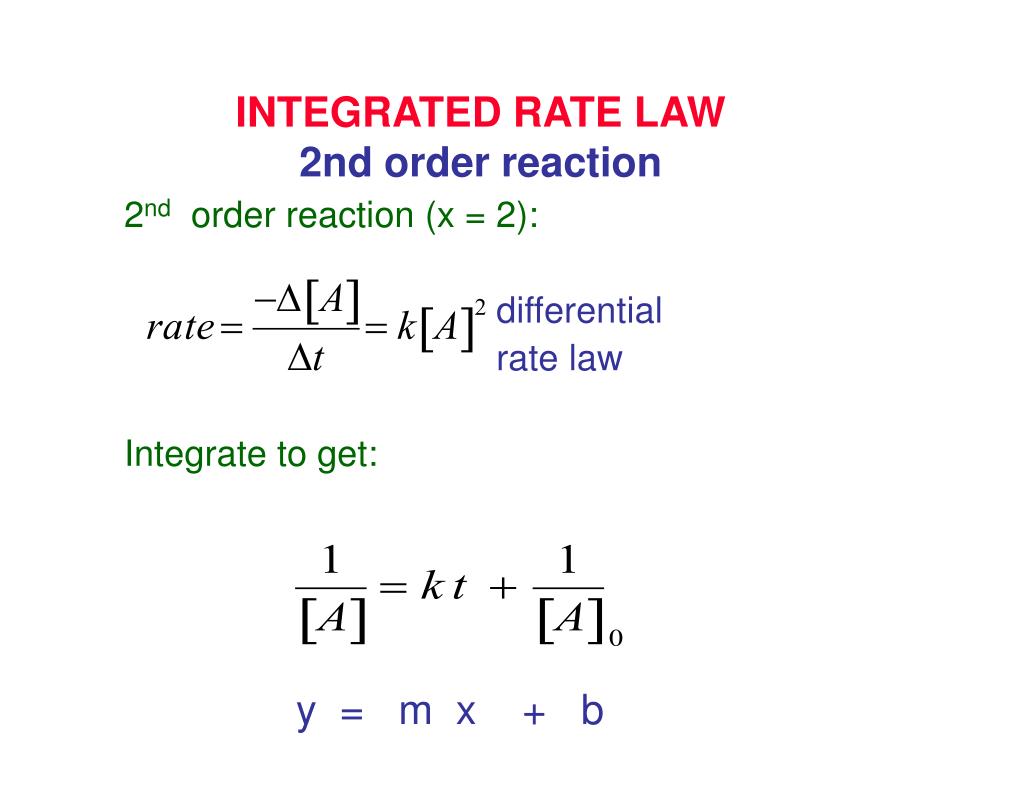
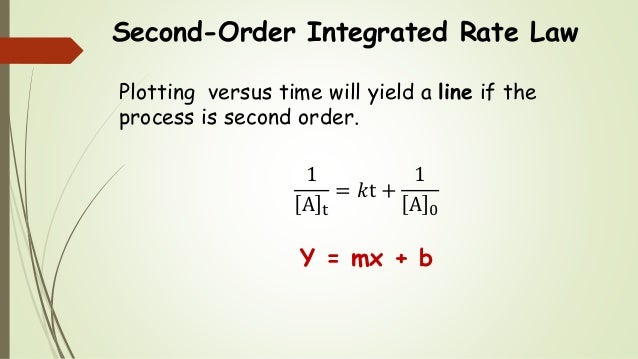
Most people approach second order rate laws the wrong way. They memorize the integrated equation and then try to force every dataset into it. That works fine for textbook problems, but real experimental data rarely behaves that cleanly. I spent a semester watching students struggle with pseudo-second order fits on adsorption isotherms before I figured out a better workflow, so I will walk you through how this actually functions in practice. The differential rate law for a second order reaction looks like this: rate = k[A]² when the reaction depends on a single reactant squared, or rate = k[A][B] when two different species are involved at equal concentrations. The integrated form comes from solving that differential equation, which gives you 1/[A]t = kt + 1/[A]0. You plot the reciprocal of concentration against time and the slope is your rate constant k. That is the standard procedure and it is exactly what every general chemistry textbook shows you. Here is the part they do not emphasize enough: the half-life of a second order reaction depends on the initial concentration. t1/2 = 1/(k[A]0). That means if you double the starting concentration, your half-life gets cut in half. This is the opposite of first order behavior where the half-life stays constant regardless of how much material you start with. It sounds simple but it has real implications for reactor design and for interpreting your kinetic data.
Unlocking The Secrets Of 2nd Order Reactions Understanding The 2nd Order Integrated Rate Law
I ran into a specific problem last year that exposed how fragile second order analysis can be. I was studying the hydrolysis of an ester in aqueous solution at controlled pH, and the concentration dropped slowly enough that I could track it by UV spectroscopy over several hours. The 1/[A] versus time plot looked decent at first glance with an R-squared value of 0.987, which should have been acceptable. But when I overlay the same data plotted as ln[A] versus time, the linear fit was actually slightly better at 0.991. That should not happen for a true second order reaction, and it meant something else was going on. The issue turned out to be that the reaction was not purely second order. The ester was undergoing acid-catalyzed hydrolysis as a background first order process at the same time, and the two pathways were running concurrently. When I subtracted the first order contribution calculated from a blank experiment run at the same pH but without the nucleophile, the residual data fit the integrated second order equation properly with an R-squared of 0.996 and a clean straight line through the origin region. Without accounting for that parallel pathway, my k value was off by roughly forty percent. I wasted about two days on that before I realized what was happening. Another thing that catches people unexpectedly is the case where you have two reactants at different initial concentrations. The integrated rate law becomes more complicated: 1/([B]0 - [A]0) * ln([A]0[B]/[B]0[A]) = kt. You need to know both initial concentrations precisely and track both reactants over time. If you only monitor one, you cannot solve this equation without making assumptions. Many labs avoid this entirely by running the reaction with one reactant in large excess so the concentration stays effectively constant. That converts the kinetics into pseudo-first order behavior and lets you extract an observed rate constant kobs = k[B]0. Then you repeat the experiment at several different concentrations of the excess reagent and the slope of kobs versus [B]0 gives you the true second order rate constant k. This is the standard method for determining bimolecular rate constants in solution and it is reliable as long as your excess ratio is at least ten to one.
There are practical limitations you need to be aware of. Second order plots amplify noise at low concentrations because you are taking reciprocals of small numbers. When [A] drops close to your detection limit, the 1/[A] value shoots up and a single noisy data point can distort your entire fit. I usually ignore the last ten to fifteen percent of the concentration range in these plots unless I have exceptionally clean data. The early portion of the curve carries the most reliable information for second order fitting. Temperature control matters more than you might think for second order reactions because k changes with temperature according to the Arrhenius equation. If your reaction runs exothermically and your sample is not well thermostatted, the rate constant will drift during the experiment as the temperature rises. Even a two degree increase can change k by five to ten percent for typical organic reactions. Use a proper water or oil bath with circulation, not just a passive heating block. If your reaction does not give a clean straight line on the 1/[A] versus t plot after accounting for side reactions and experimental artifacts, the system is probably not second order. Do not force the fit. Check for zero order behavior first by looking at [A] versus t, then first order with ln[A] versus t, then third order if you have a good reason to suspect it. Most reactions that people claim are second order are either first order with poor data or composite reactions with multiple steps. The integrated rate law only applies to elementary steps or reactions that genuinely follow second order kinetics throughout the entire concentration range.
Get the Full Details

For reactions that follow the form rate = k[A][B] where [A]0 does not equal [B]0, you can still use the method of isolation. Set one reactant at fifty to one hundred times the concentration of the other and treat it as constant. This collapses the mathematics into pseudo-first order form and makes the analysis much more robust. It also reduces the impact of concentration measurement errors on your final result. I recommend this approach whenever possible rather than trying to fit the full two-reactant integrated equation to noisy data. Software tools like Python with scipy.optimize.curve_fit or even Origin can do non-linear regression on the integrated rate equations directly instead of relying on linearized plots. This avoids the distortion introduced by the reciprocal transformation and gives more statistically sound confidence intervals on your rate constant. The linear plot method is fine for teaching and quick estimates, but for publication quality work, non-linear fitting is the better choice and it takes about the same amount of effort to set up once you know how.