Running the wet bench before you trust the instrument
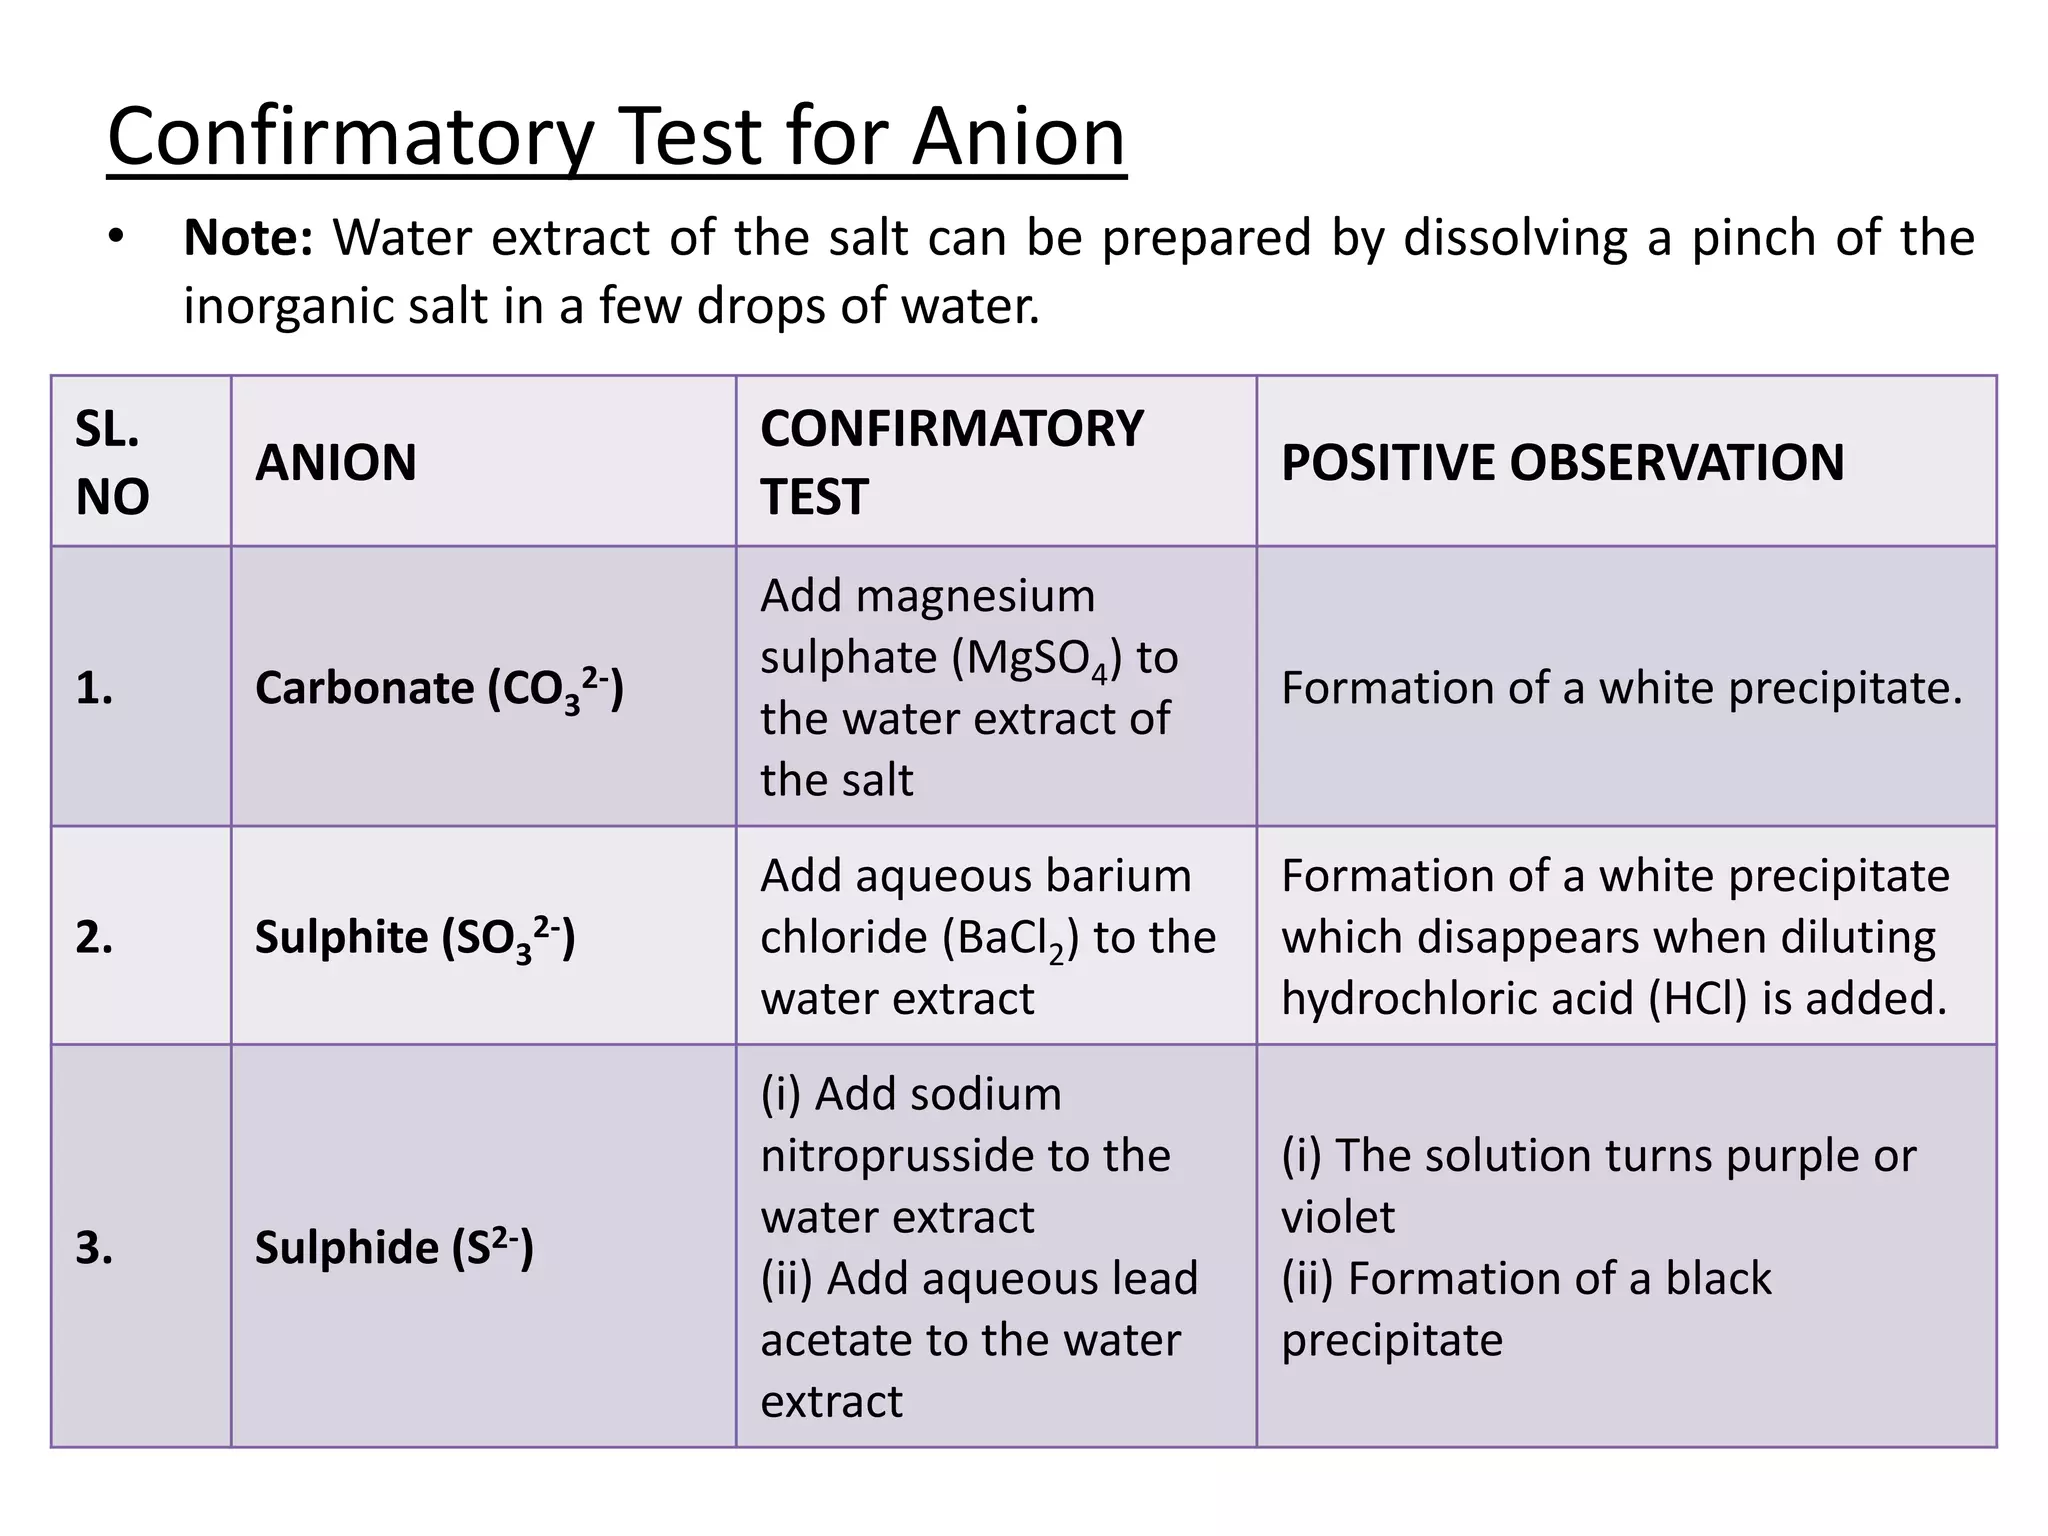
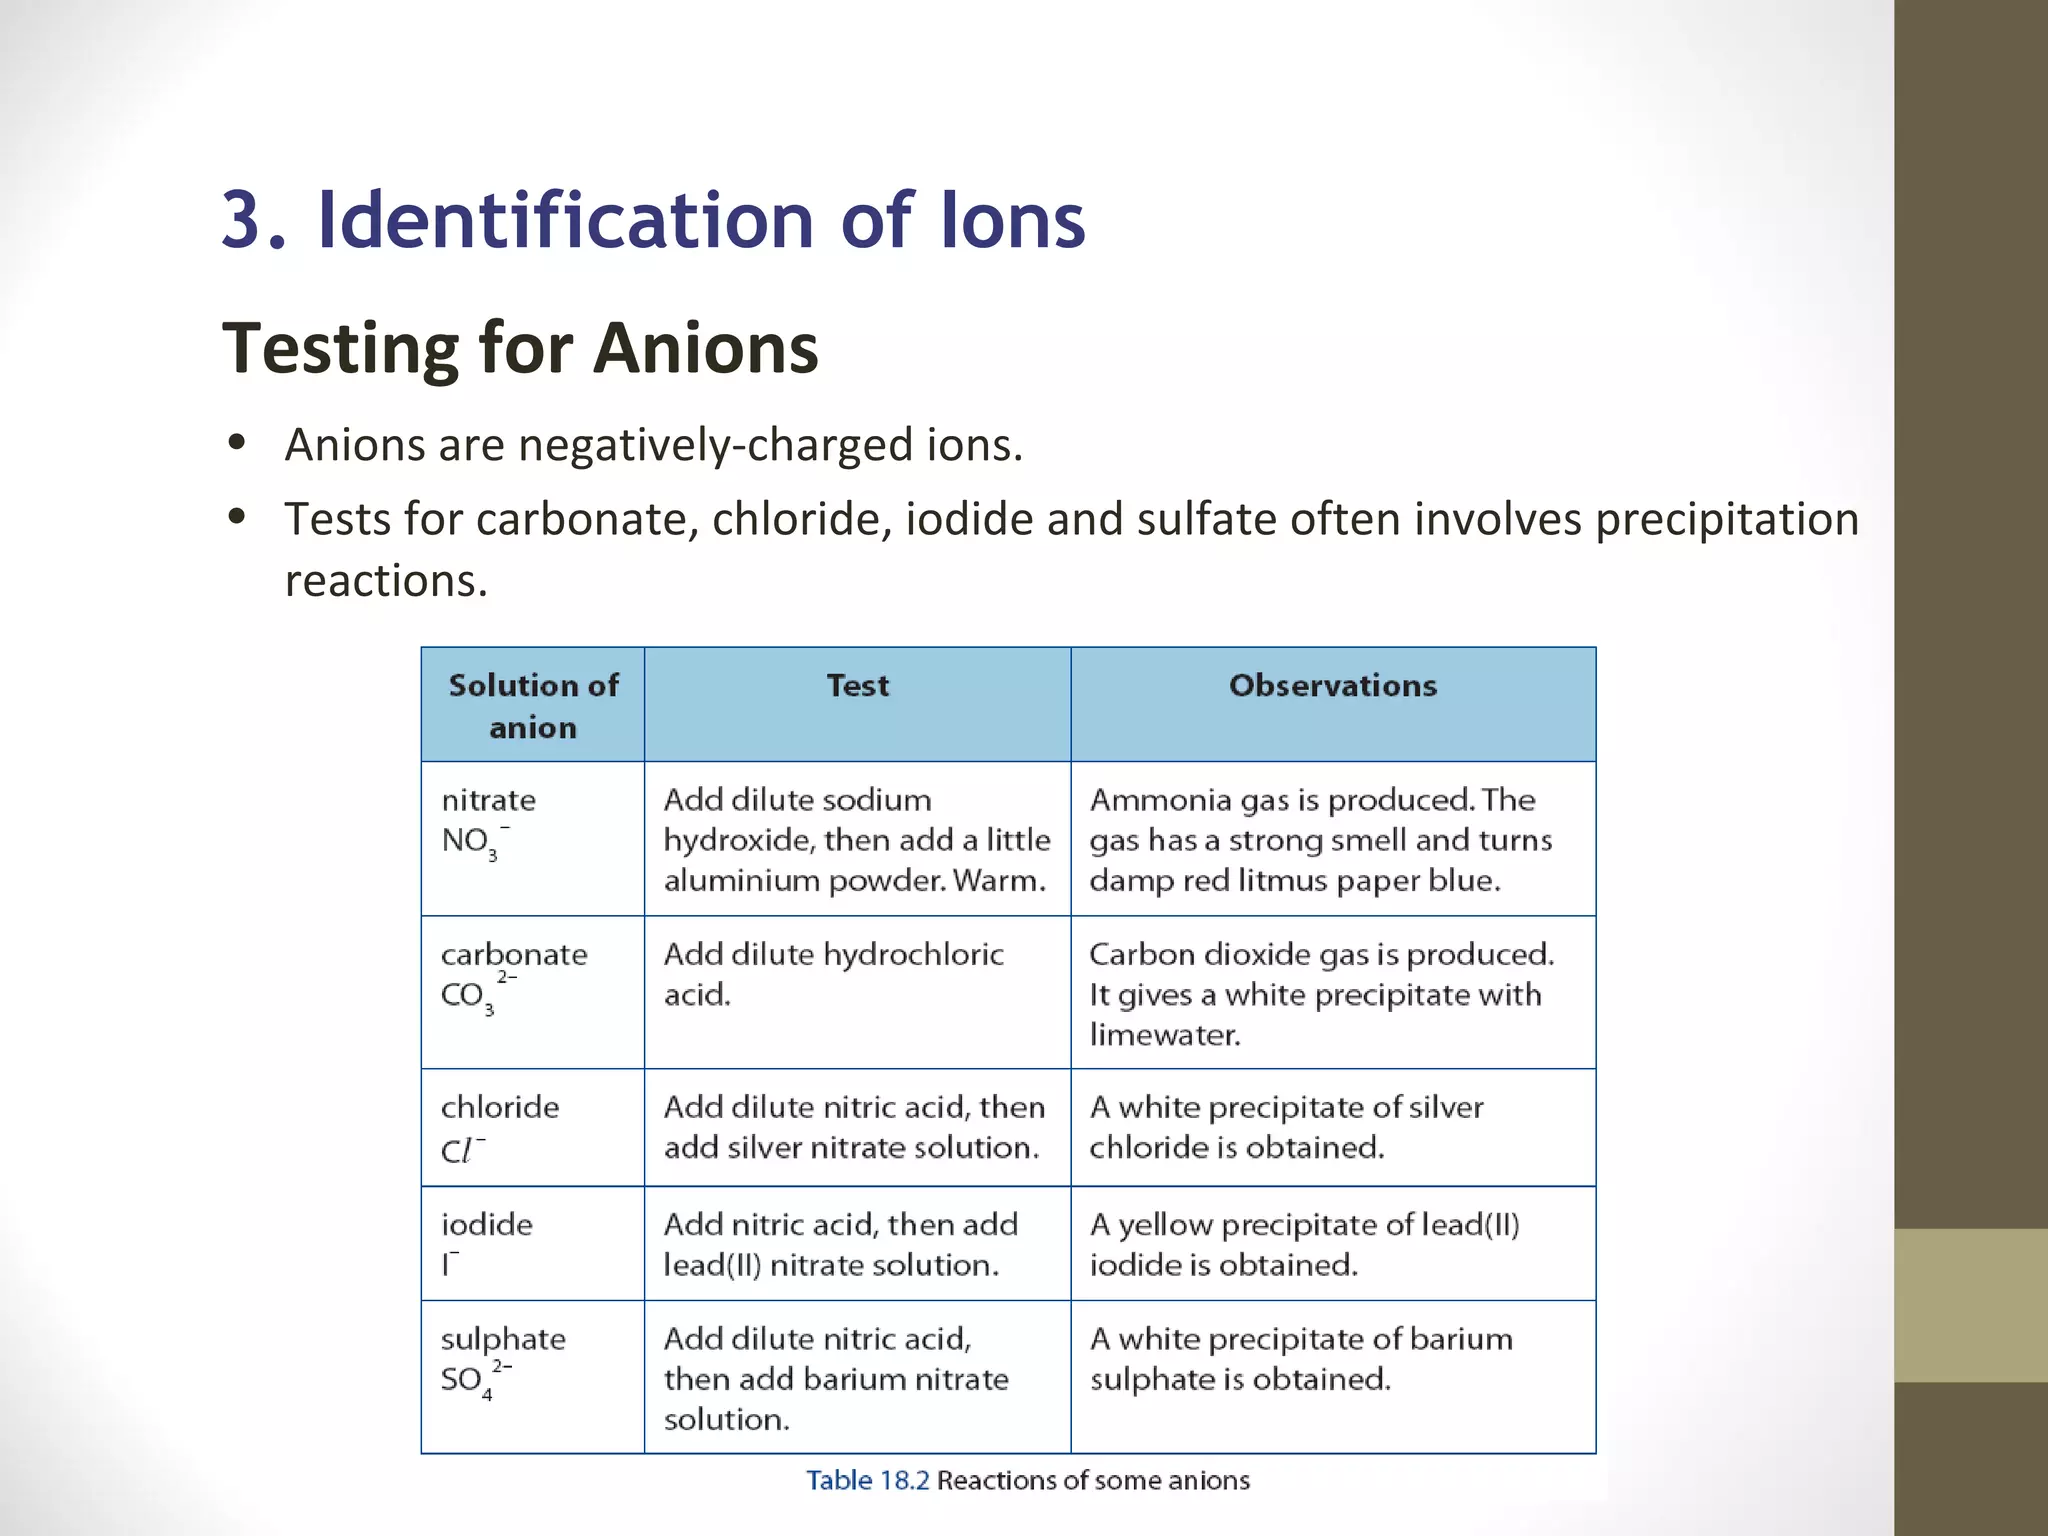
The first step in any anion identification scheme is always a preliminary screen. You run the ignition test on the solid, add dilute sulfuric acid to an aliquot, and watch what happens. Effervescence tells you something volatile is leaving the solution. The smell of the gas, its effect on damp litmus paper, and whether it blackens filter paper treated with lead acetate are the data points you actually need right now. I spent years skipping this part because we had an ion chromatograph sitting in the corner. It cost us three days of trouble-shooting a stubborn baseline before someone remembered that the unknown sample was a decomposing organic sludge that was fouling the column. The preliminary tests would have told us it was full of sulfite and thiosulfate in about twenty minutes. After the preliminary screen comes the confirmatory stage, and this is where most people make mistakes. You do not run every confirmatory test on the same solution. Each confirmation requires specific pH conditions, and the reagents used for one anion often interfere with another. I keep a separate spot plate for each anion class. When I am running a water sample from an industrial discharge permit, I allocate two clean 50 mL portions of the filtrate. One goes to the halide group, the other to the sulfate and nitrate group. I never mix them. The chemistry itself is straightforward if you respect solubility rules. Adding silver nitrate to a nitric acidified solution precipitates halides as silver halides. The color of the precipitate is your first clue. Silver chloride is white and soluble in dilute ammonia. Silver bromide is cream colored and only sparingly soluble. Silver iodide is yellow and does not dissolve in ammonia. The trick is the ammonia concentration. If you use concentrated ammonia, you will dissolve the bromide precipitate and falsely assign your sample as pure chloride. I standardize my ammonia by titration once per month because the stock bottle loses strength over time and the variation ruins the selectivity.
Sulfate identification relies on barium chloride in acidic medium. The white barium sulfate precipitate is unmistakable, but so is its stubbornness. It co-precipitates other species and traps them inside the crystal lattice. I learned this the hard way when a pharmaceutical client reported failing sulfate tests on a batch that I knew was clean. The sample contained trace amounts of chromate, and the barium chromate was hiding inside the sulfate matrix. The precipitate looked pure. It took me two days to figure out that a washing step with warm dilute hydrochloric acid stripped the chromate from the surface while leaving the sulfate intact. Without that wash, the confirmatory test was giving a false negative for interference and masking the real composition. Carbonate and bicarbonate detection is usually handled by acid evolution. Adding hydrochloric acid and passing the gas through barium hydroxide gives you a white precipitate of barium carbonate. The problem is that sulfite produces sulfur dioxide under the same conditions, and sulfur dioxide also precipitates barium hydroxide. The two reactions look identical to a novice. I oxidize the evolved gas by passing it through a trap containing hydrogen peroxide before it reaches the barium hydroxide. The peroxide converts any sulfur dioxide to sulfuric acid, which does not precipitate barium. Only the carbon dioxide produces the white turbidity. This single modification eliminated a whole class of false positives in our lab. Nitrate identification gets messy quickly. The brown ring test is the classic method, using iron(II) sulfate and concentrated sulfuric acid. But nitrite interferes directly because it also reduces the iron under those conditions. If your sample contains nitrite, you destroy it first by adding sulfamic acid and waiting for the nitrogen evolution to stop. Then you proceed with the brown ring test on the treated aliquot. Skipping the nitrite destruction step is the most common error I see in undergraduate reports and industrial QC checklists. The result is a brown ring that tells you nothing about whether nitrate is actually present.
Nitrite itself is detected using Griess reagent, which produces a pink azo dye. The sensitivity is high enough that ambient contamination from laboratory reagents can give you a false read. I prepare fresh reagent weekly and run a blank with every batch of samples. The blank absorbs about 0.02 absorbance units at 540 nanometers. Any sample reading below 0.05 above the blank is effectively background noise and should be reported as non-detect rather than as a trace quantity. Reporting those false lows creates compliance issues downstream. Phosphate detection uses ammonium molybdate in acidic solution to form ammonium phosphomolybdate, which appears as a pale yellow precipitate. The reaction is sensitive to pH and temperature. If the solution is too alkaline, you get molybdate precipitating on its own. If it is too cold, the reaction takes hours instead of minutes. I maintain the acidified molybdate reagent at 40 degrees Celsius in a water bath during the test. The color development is complete within five minutes and the precipitate is stable for at least thirty minutes before it starts redissolving. This timing window matters when you are processing forty samples in a shift. Halide separation by fractional precipitation is another area where practice diverges from theory. The solubility product values for silver chloride, bromide, and iodide suggest you can separate them cleanly by controlling the silver ion concentration. In reality, mixed halide samples form solid solutions rather than pure separate precipitates. The precipitate you collect at low silver concentration contains chloride, bromide, and a fraction of the iodide all mixed together. If you need individual concentrations rather than just presence or absence, you need ion chromatography or potentiometric titration with silver nitrate. Wet chemistry alone will not resolve a mixture of all three halides to acceptable accuracy. I use the precipitation method as a screening tool and send ambiguous samples to the IC lab for quantification.
Get the Full Details

In industrial settings the matrix is almost always worse than in textbook problems. Cooling tower water contains silicate, which forms a gelatinous precipitate with molybdate reagent and masks phosphate detection. Boiler feedwater contains sulfide, which precipitates silver and barium reagents indiscriminately. I remove sulfide by bubbling nitrogen through the acidified sample for ten minutes before running any confirmatory tests. The nitrogen strips the hydrogen sulfide without oxidizing the other anions. Silicate interference is handled by adding fluorosilicate or by adjusting the molybdate reagent composition to suppress the silicomolybdate formation. These are not elegant solutions but they are practical. Regulatory environments in particular industries demand documented detection limits for each anion method. The EPA methods for anion analysis specify minimum reporting levels that are often lower than what a standard spot plate procedure can achieve. If you are working under a consent decree or an industrial wastewater permit, the lab SOP must account for that. A qualitative identification of chloride at 5 milligrams per liter using silver nitrate on a spot plate is not sufficient documentation. You need either a calibrated titration or instrumental confirmation to meet the reporting requirement. Understanding which level of verification your application demands before you start the analysis saves time and prevents rework. The biggest limitation of classical anion identification is that it is inherently qualitative. You can say a certain anion is present or absent with reasonable confidence. You can estimate order of magnitude. You cannot produce a precise concentration without moving to a quantitative technique. This is not a flaw in the method, it is a boundary condition. Treating it as a complete analytical solution leads to incorrect conclusions, especially when comparing samples across different time points. I pair the classical scheme with periodic IC runs on representative samples to build a correlation between the two methods for each matrix type we encounter.
Another practical limitation is the skill ceiling. A technician who has only run these tests twice will miss subtle color differences between silver bromide and silver chloride. The distinction is real but it requires pattern recognition that comes from repetition. I budget roughly six months of supervised runs before allowing a junior analyst to sign off on identification results independently. The shortcut of training someone faster produces errors that cost more in corrective actions than the extra training time ever would. For the records, there is no single downloadable protocol that covers every scenario because the method depends entirely on your sample matrix and the anions you are targeting. The standard reference is the APHA Standard Methods for the Examination of Water and Wastewater, which has dedicated chapters for each anion. Those are available through institutional subscriptions. Commercial suppliers like Sigma and Merck publish analytical guides that include step-by-step procedures for their reagent systems. My personal working reference is a compiled binder of internal SOPs built from those sources, updated whenever a method deviation occurs in practice. The binder lives on the bench, not online.