Understanding London Dispersion Forces Without The Hype
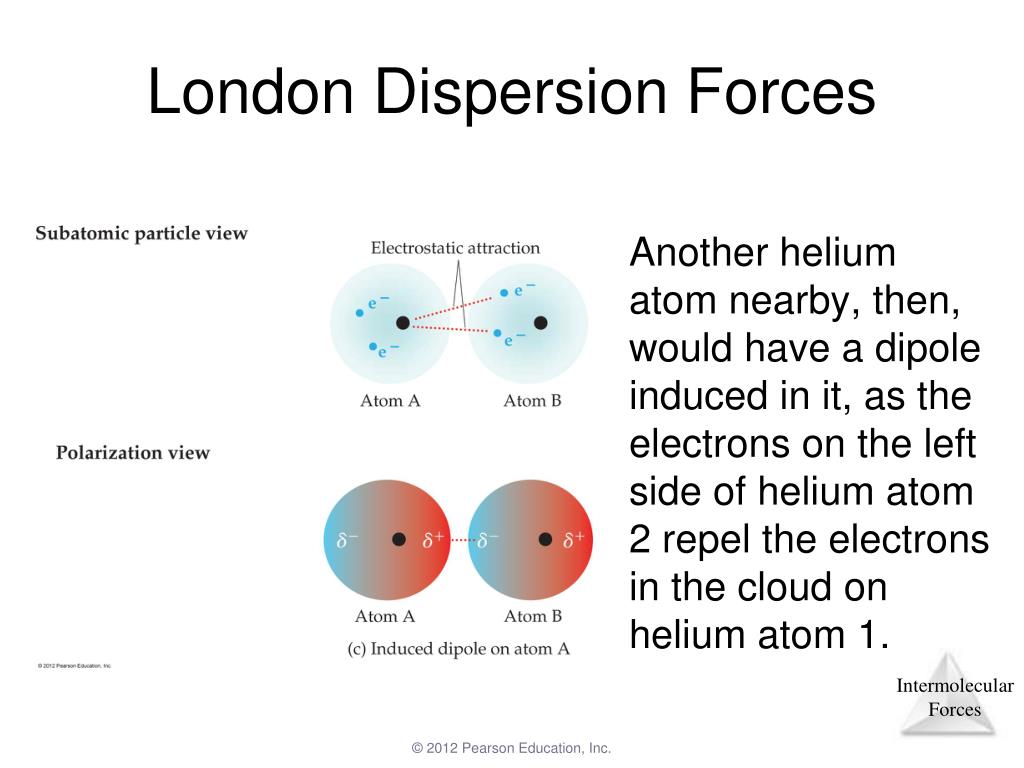
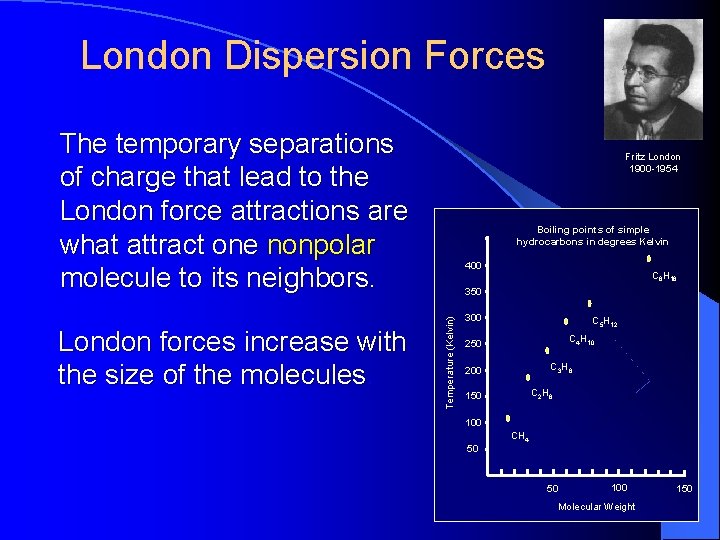
London dispersion forces are the weakest of the van der Waals interactions, but they dominate whenever you have nonpolar molecules or large electron clouds. I used to watch students completely dismiss them in intro chemistry because the textbook called them "temporary" and "weak," which makes them sound harmless. That assumption gets people in trouble fast. Here is how they actually work, stripped of the drama. Electrons are moving constantly. At any given instant, there is a random fluctuation in electron density that creates a temporary dipole. That temporary dipole induces a matching dipole in a neighboring molecule. The two dipoles attract each other briefly until the electrons shift again. This happens trillions of times per second across every nonpolar substance on earth. The strength of this interaction scales with polarizability. Larger atoms and molecules with more diffuse electron clouds distort more easily, which means stronger dispersion forces. This is why iodine is a solid at room temperature while fluorine is a gas, even though both are diatomic and nonpolar. The difference comes down entirely to how many electrons each molecule carries and how far out those valence electrons sit from the nucleus.
Surface area matters too, and people overlook that regularly. Consider two isomers: n-pentane and neopentane. Both have the same molecular formula, C5H12. N-pentane boils at 36 degrees Celsius. Neopentane boils at 9.5 degrees Celsius. They share the same number of electrons, yet their boiling points differ by nearly 27 degrees. The linear chain of n-pentane allows more surface contact between molecules, which multiplies the number of simultaneous dispersion interactions. Spherical neopentane minimizes that contact area. The electron count is identical. Shape does the heavy lifting. I ran into a practical problem a few years back while working on a separation process involving a mixture of long-chain hydrocarbons and a fluorinated solvent. The standard solubility predictions based on polarity parameters were completely wrong. The fluorinated solvent should not have dissolved those hydrocarbons in any meaningful amount according to textbook rules, but they were miscible at elevated temperatures. The calculations failed because they did not account for the dispersion contribution properly. The long alkyl chains had enough polarizable electrons that the dispersion term outweighed the unfavorable polar interactions. I ended up using Hansen solubility parameters instead of simple polarity models, which broke the total cohesive energy density into three components: dispersion, polar, and hydrogen bonding. That decomposition let me actually predict solubility behavior without spending weeks on trial-and-error experiments.
Why This Matters In Practice
If you are modeling intermolecular interactions in any software, default force fields often underestimate dispersion unless you specifically parameterize for it. The standard Lennard-Jones potential captures the attraction-repulsion balance, but the C6 coefficient determines how strong the attraction actually is. Using generic C6 values for everything will give you garbage results if your system involves heavy atoms or extended pi systems. I have seen simulations produce incorrect boiling points, wrong liquid densities, and even incorrect phase behavior because someone copied parameters from a small-molecule force field and applied them to a polymer without adjustment. A common pitfall is assuming dispersion forces are negligible at room temperature. They are not. In fact, they are the primary cohesive force holding together most organic liquids and solids. Water has hydrogen bonding, yes, but liquid methane relies entirely on dispersion. Liquid bromine relies entirely on dispersion. The entire category of alkanes, from methane to paraffin wax, exists as liquids or solids because of dispersion. Without it, every nonpolar compound would be a gas at standard conditions. Another thing beginners miss: dispersion forces are additive. A single C-H bond contributes a small amount, but a molecule with thirty C-H bonds on a surface that can closely approach another molecule accumulates enough total attraction to matter enormously. This is why graphene sheets stick together so tightly despite having no permanent dipoles. It is also why carbon nanotubes bundle into fibers rather than staying isolated unless you functionalize the surface.
Get the Full Details
When Dispersion Fails You
There are situations where dispersion alone cannot explain observed behavior. Highly polar molecules with strong dipole-dipole interactions or hydrogen bonding networks will show properties that dispersion models alone cannot reproduce. Water is the obvious example. Its boiling point is far higher than what dispersion calculations based on its size would predict. In those cases, you need to combine dispersion terms with electrostatic terms and sometimes specific directional interaction terms. There is no single universal formula that handles all cases equally well. If you are doing computational chemistry work and need to account for dispersion accurately, modern methods like DFT-D3 or DFT-D4 corrections are the standard approach. They add empirical dispersion terms to density functional theory calculations and usually improve accuracy significantly. Without them, many common functionals actually underestimate binding energies for weakly interacting systems. I once spent three days debugging a reaction barrier calculation that looked reasonable until I realized the base functional was missing dispersion entirely. Adding the D3 correction shifted the transition state energy by about 4 kilocalories per mole, which completely changed the predicted rate. That is a real difference, not a rounding error. The takeaway is straightforward. London dispersion forces are not mysterious. They are quantum mechanical in origin, rooted in electron correlation, and they follow predictable scaling laws. But treating them as a minor detail is a mistake. They shape boiling points, solubility, crystal packing, protein folding, and material properties across the board. Pay attention to polarizability, consider molecular shape, and use the right parameterization when you are simulating anything that depends on weak intermolecular attraction.