Understanding Sigma And Pi Bonds In Organic Chemistry
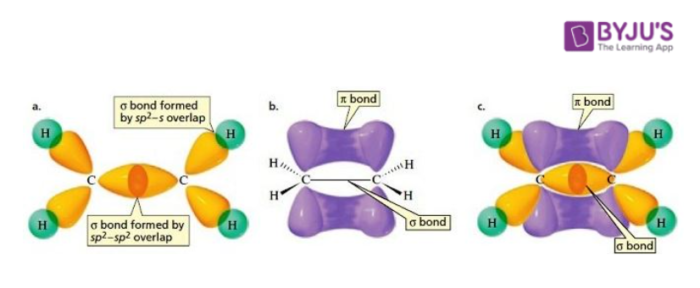
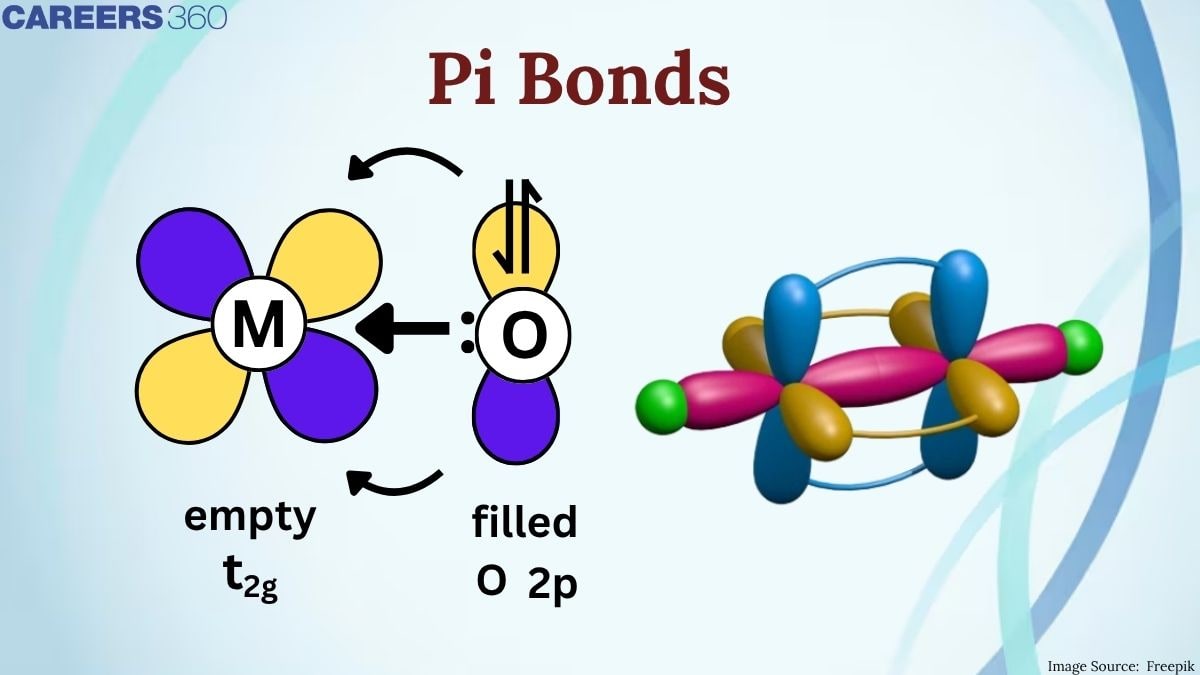
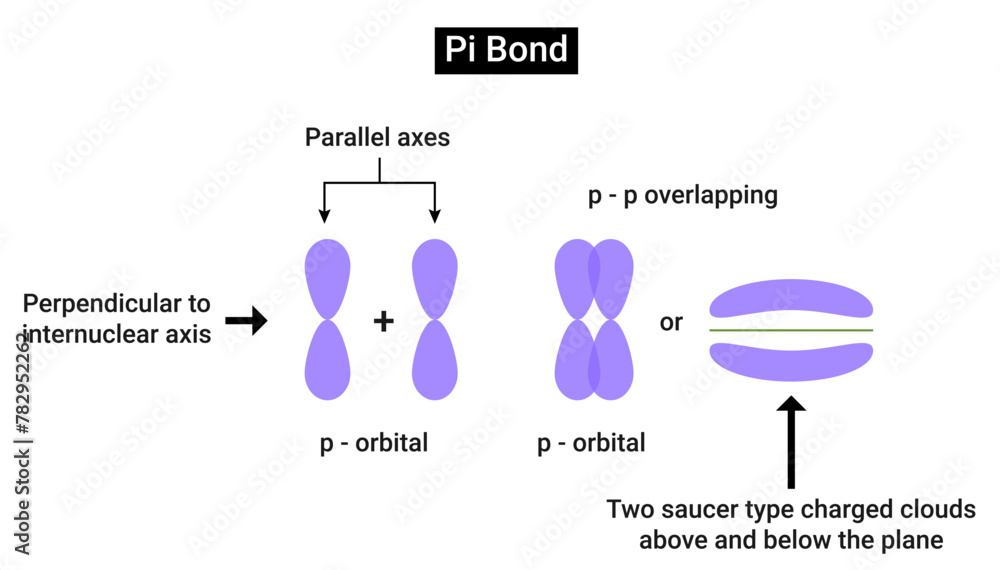
Most organic chemistry courses introduce sigma bonds first because they're simpler to visualize. Everything starts with head-on orbital overlap between two atoms. Once you understand that foundation, pi bonds are just what happens when p orbitals sit parallel to each other and share electron density above and below the bonding axis. That's it. It's not mystical. It's geometry. A pi bond forms when two adjacent p orbitals overlap laterally, meaning side to side rather than end to end. This creates electron density concentrated in two lobes, one above and one below the plane of the bonded atoms. A double bond is never just one bond. It's always one sigma bond plus one pi bond. A triple bond stacks a second pi bond at a right angle to the first, so you get two perpendicular pi systems alongside the sigma framework. The common mistake students make is thinking pi bonds are just weaker versions of sigma bonds. They're fundamentally different in how they behave. Pi bonds restrict rotation. That's why cis and trans isomers exist in alkenes. A sigma bond alone would let the substituents spin freely. The pi bond locks things in place because rotating the atoms would break the parallel alignment of the p orbitals and lose that overlap entirely. Breaking a pi bond requires energy—typically around 60 to 65 kcal/mol for a carbon-carbon double bond—while breaking a sigma bond usually needs closer to 85 to 100 kcal/mol. The pi bond is weaker, yes, but calling it weak is misleading. It's structurally significant in ways sigma bonds aren't.
I remember spending three days stuck on a problem involving the hydrogenation of a substituted cyclohexene where the reaction didn't proceed the way the textbook predicted. The molecule had a methyl group blocking one face of the double bond. I kept drawing the mechanism with the hydrogen approaching from the wrong side. The workaround was switching to a molecular model kit and physically building the molecule. Once I could see the steric hindrance from the methyl group directing the catalyst surface, everything clicked. You can't really understand pi bond reactivity from 2D drawings alone. The three-dimensional topology matters more than the electron count. Another thing beginners consistently miss is the relationship between pi bonds and hybridization. A carbon involved in a pi bond is sp2 hybridized, which means it has one unhybridized p orbital remaining perpendicular to the trigonal plane. That leftover p orbital is the one forming the pi bond. If a carbon is part of a triple bond, it's sp hybridized with two unhybridized p orbitals available, creating two pi bonds. The hybridization state determines bond angles, which determines molecular shape, which determines reactivity. These aren't separate topics. They're the same system viewed from different angles. Conjugation is where pi bonds become genuinely interesting. When pi bonds alternate with single bonds across a chain of carbons, the p orbitals overlap across three or more atoms simultaneously. This delocalization lowers the overall energy of the molecule. But conjugation only works when the p orbitals can align. If a single bond in the chain rotates and the p orbitals twist out of alignment, the conjugation breaks. This is why s-cis and s-trans conformations matter in dienes, and why 1,3-butadiene isn't just two isolated double bonds connected by a single bond. The central bond has partial double bond character because of the delocalized electrons running through it.
There are real limitations to consider when relying on pi bond theory. Molecular orbital theory gives a more accurate picture than simple valence bond descriptions, but it's computationally expensive. For large molecules, semi-empirical methods or density functional theory approximations are often necessary, and those come with their own accuracy trade-offs. Pi bond character also gets complicated in heteroatomic systems where electronegativity differences distort the orbital overlap. A C=O pi bond behaves very differently from a C=C pi bond because oxygen pulls electron density toward itself, polarizing the entire system. Simple pictures break down here. If you're working with pi bonds in a practical setting, whether that's predicting reaction outcomes or interpreting spectroscopic data, focus on the geometry first. Draw the molecule with realistic bond angles. Identify which atoms have p orbitals and whether those orbitals can actually overlap. Then think about what would happen if that overlap changed. The reactivity follows from the structure, not the other way around.
Get the Full Details