How We Actually Use Rf Values in the Lab
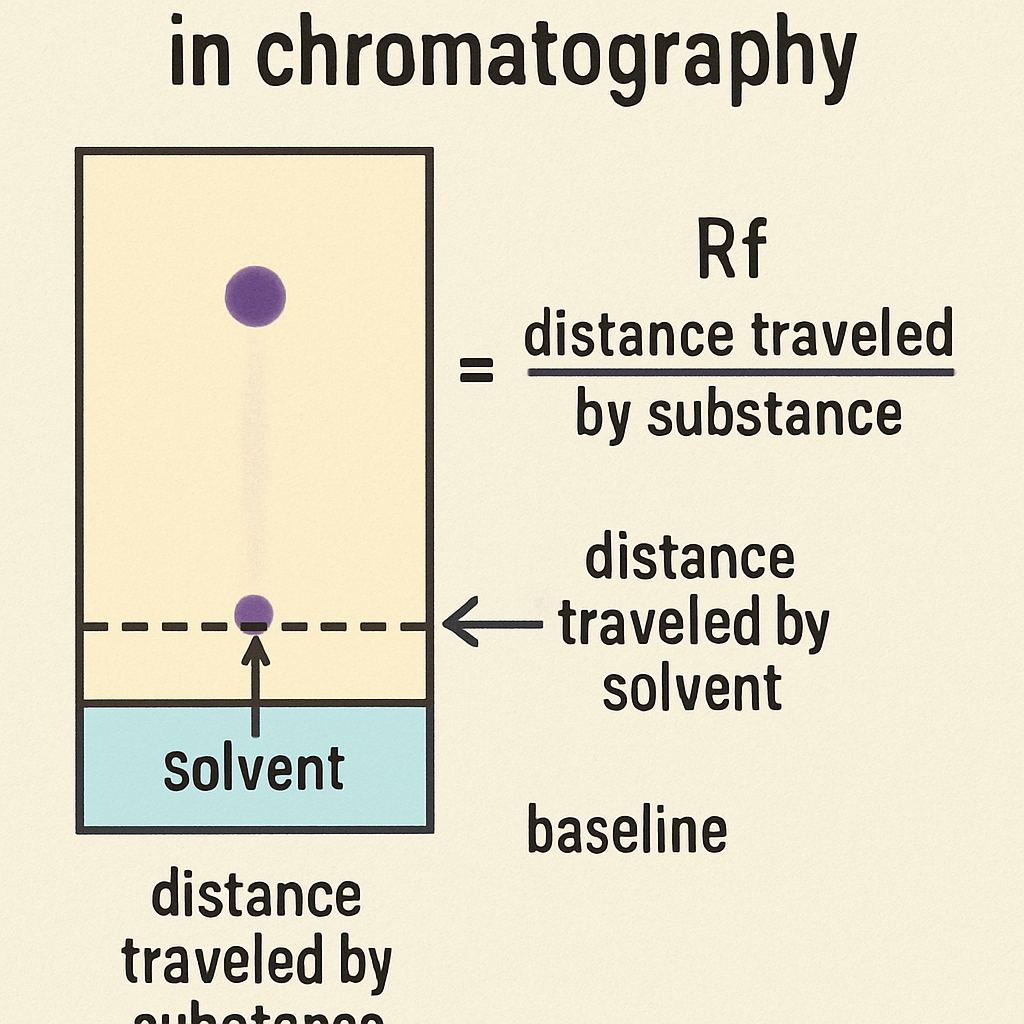
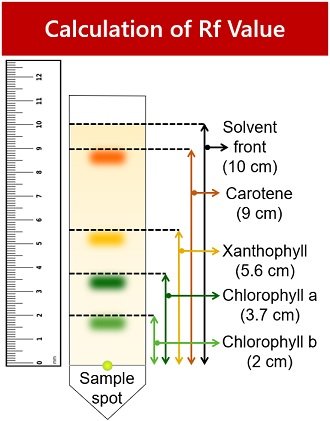
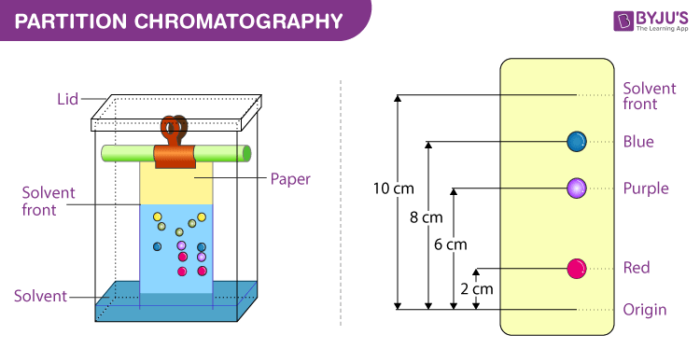
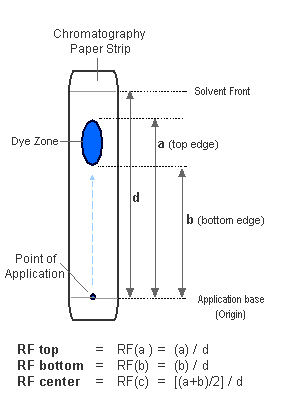
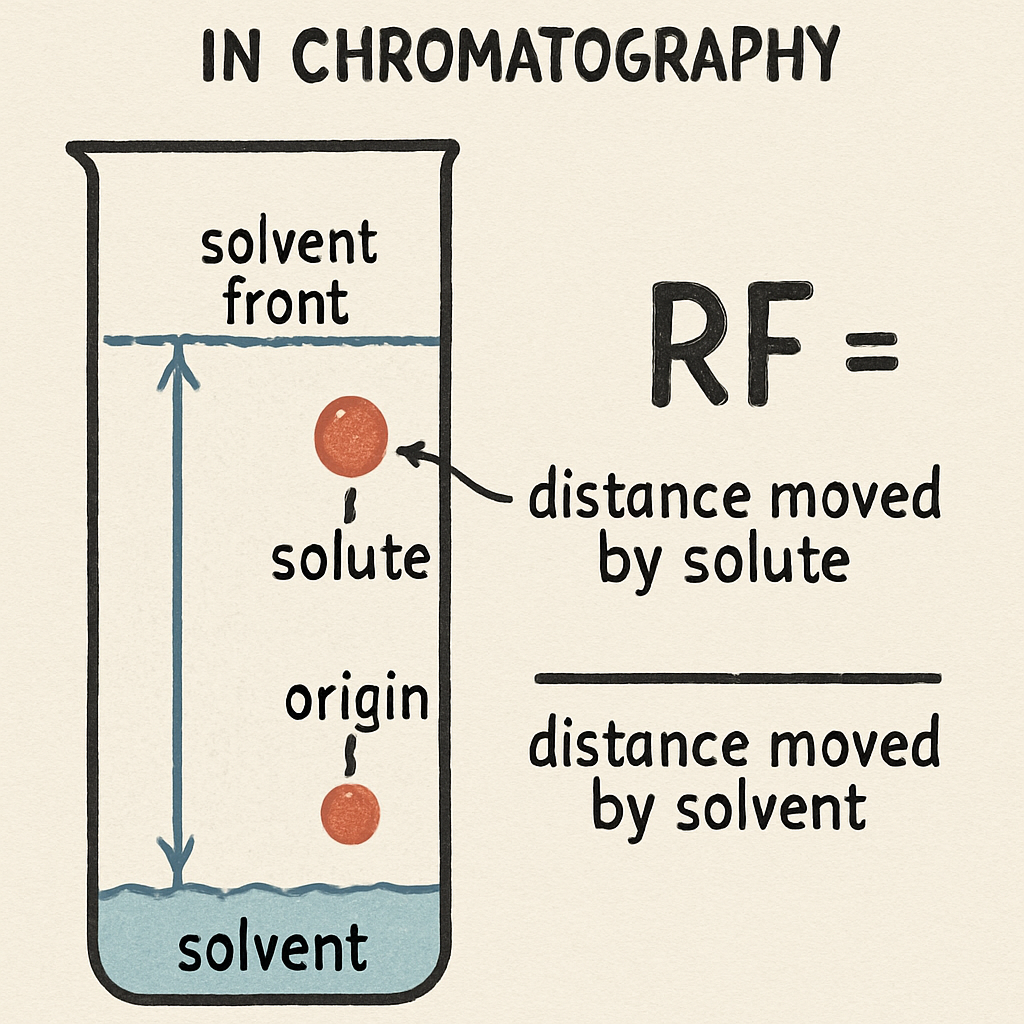
Rf values are one of those things everyone learns in sophomore organic chemistry and then kind of forgets until they need them again. The formula is straightforward enough — distance traveled by the compound divided by distance traveled by the solvent front. What people don't always grasp is how much real-world variation creeps into that number before you even spot the plate. I spent last week trying to reproduce a TLC result from a paper that had been running for three years. The reported Rf was 0.42 in 30% ethyl acetate/hexanes. My plate read 0.51. Same reagents, same supplier, same brand of silica gel plates. The difference wasn't a mistake on my end. It was humidity. The silica had absorbed enough moisture from the lab air to change its adsorption characteristics. I dried the plates at 110°C for twenty minutes before use and got 0.44. Close enough to call it matched.
Unlocking Separation Precision The Critical Role Of Rf Value In Chromatography
Before we get into the practical side, let me say something that might surprise people who only run TLC occasionally. Rf is not a constant. Not for any compound. It shifts with solvent composition, stationary phase batch, temperature, chamber saturation, and how long you let the plate develop. Treating an Rf value like a molecular weight — something fixed and reproducible across labs — is one of the most common rookie mistakes I see. It's a relative measurement, period. The utility comes from consistency within your own conditions, not from expecting someone else's number to map directly onto yours. That said, there are real workflows where getting the Rf right matters a great deal. Flash column chromatography is the main one. If your target compound runs at an Rf of 0.15 on TLC in a given solvent system, you're probably going to need something close to that ratio when you scale up to a column. The rule of thumb most people learn is aiming for an Rf between 0.2 and 0.35 for good separation on flash. Below 0.2 and you're spending too much time eluting. Above 0.35 and everything co-elutes and you're wasting solvent doing nothing useful. The second place where Rf becomes genuinely important is method development for HPLC. Retention time correlates loosely with Rf, and knowing your compound's behavior on silica gives you a starting point for choosing a reverse-phase gradient. A compound that flies through a normal-phase TLC plate at Rf 0.7 is probably going to be stubborn on C18. Not always, but often enough that the TLC becomes a useful screening tool before you commit to running HPLC method trials.
Here's where it gets tricky and where most people slide into bad data. Visualization. If you're using UV and your compound doesn't absorb well at 254 nm, you might see nothing or a ghost of a spot and record an Rf that's completely wrong. I once spent an afternoon chasing a compound that I thought had an Rf of 0.6, only to realize the spot I was measuring was an impurity. The actual product wasn't showing up under UV at all. Switched to KMnO4 stain and found the real spot at Rf 0.28. That changes your column strategy significantly. Another issue that people overlook is spot size. If you load a big concentrated spot, it tails. The measured distance from the origin to the center of a tailing spot will be higher than the true Rf. Load smaller amounts. Use a fine capillary. The spot should be no wider than 2 millimeters when you start the run. This takes practice but it matters more than most people think when you're trying to get reproducible numbers. Chamber saturation is another one that makes or breaks reproducibility. An unsaturated TLC chamber gives you a lower Rf because the solvent evaporates from the plate as it travels up, concentrating the mobile phase and changing the effective polarity. Run a development in a closed jar lined with filter paper soaked in the same solvent system and watch your Rf values climb by 0.05 to 0.15 compared to an open beaker. That's not noise. That's a systematic error that compounds every time you skip the saturation step.
Get the Full Details
Temperature is less talked about but it does matter. A plate developed at 25°C will give a different Rf than one developed at 35°C. The effect is usually small for non-volatile solvent systems but it adds up when you're working with volatile components like dichloromethane or ether in your mobile phase. I keep my TLC drawer away from the fume hood exhaust and the radiator. It's a minor detail that most protocols ignore and most people never think about until their results drift week to week. There are honest limitations to Rf that nobody likes to admit. It doesn't work well for ionic compounds on standard silica — they streak or stay at the origin no matter what you do. You need reverse-phase TLC plates or you need to add a small percentage of acid or base to the mobile phase to suppress ionization. Even then, the numbers are noisy. For those compounds, Rf is basically useless and you should move straight to HPLC or capillary electrophoresis instead of wasting time trying to force a normal-phase result. Stereoisomers are another case where Rf falls apart. Enantiomers have identical Rf values on plain silica. Diastereomers sometimes separate, sometimes don't, and there's no reliable way to predict which. If you're working with a new chiral compound and need to check diastereomeric purity, TLC is the wrong tool. Use chiral HPLC or NMR. Don't let an Rf of zero separation convince you that your reaction is clean.
For anyone actually using this stuff day to day, here's a practical workflow I've settled on over the years. Run a grid. Don't test one solvent ratio and call it done. Set up four plates with the same compound and develop them in 10%, 20%, 30%, and 40% ethyl acetate in hexanes. Within ten minutes you've got a sense of where your compound sits and whether there's a useful separation window. Then pick the ratio that puts your target between 0.2 and 0.3 and run a second plate with a neighbor ratio for fine-tuning. This usually takes fifteen minutes and saves you three hours of column chromatography later. Always run a co-spot. Place a dot of your reaction mixture next to a dot of the starting material and a dot of an isolated product if you have one. Develop the plate. If the product spot aligns perfectly with the starting material spot, you haven't made anything. If they're separate, you're looking at actual conversion. This single step catches half the problems I see in people's data. Record everything. Solvent ratio, plate type, saturation method, development time, visualization method, ambient temperature if you're feeling precise. Your future self will thank you when you come back to this reaction six months later and need to reproduce it. I have notebooks full of Rf values with no context that are completely useless for that purpose. Don't be that person.
The bottom line is that Rf is a rough but practical tool when you understand what it can and can't tell you. It's not precise by nature. It's not a physical constant. But used carefully — with controlled conditions, proper visualization, and a grid-based approach to method development — it saves more time in the lab than almost anything else. Just don't treat it like gospel and don't blame the method when your technique is the variable that's drifting.