CRISPR Is Not Magic, It Is Molecular Machinery That Breaks When You Don't Respect Its Limits
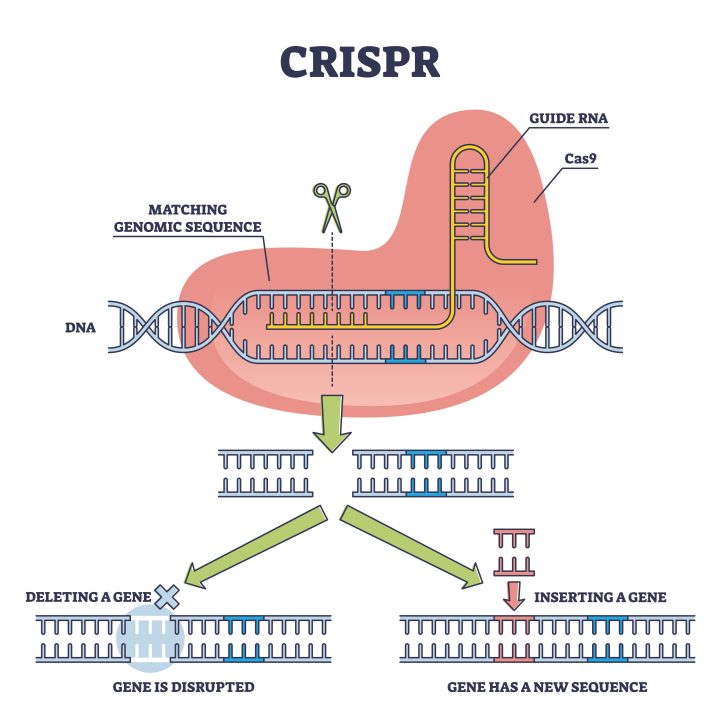
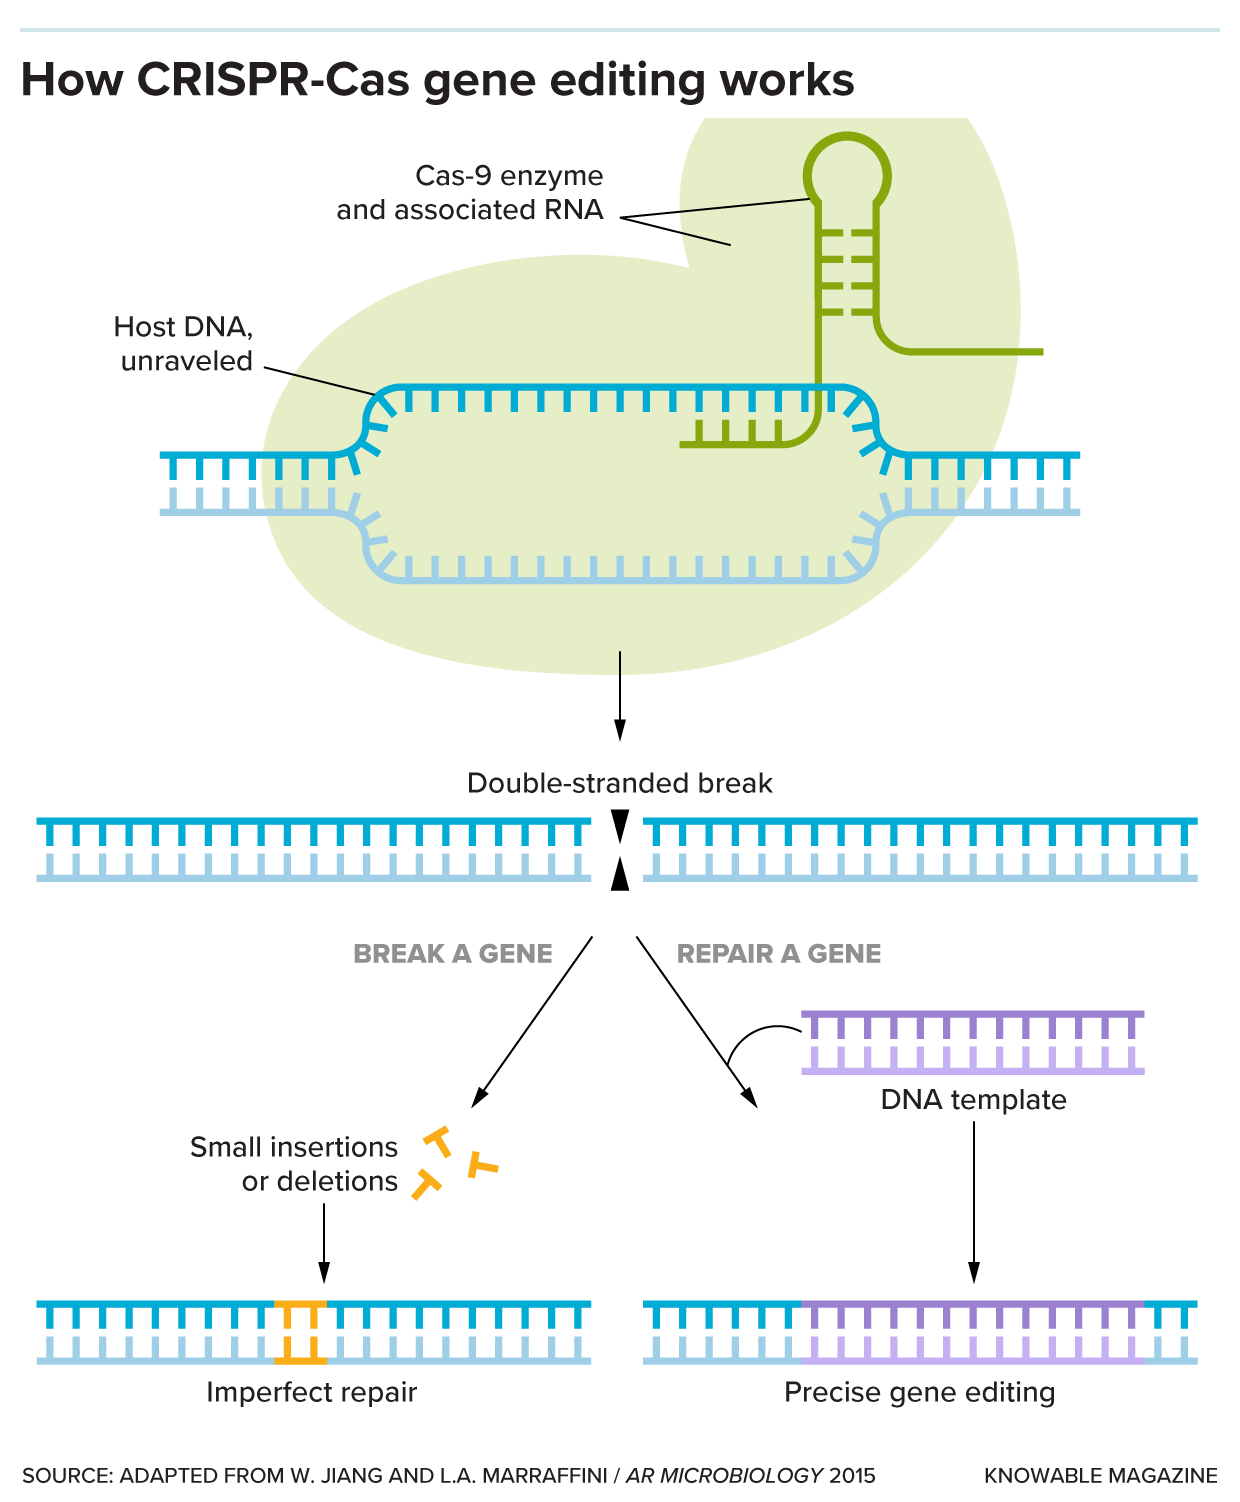
Everyone wants to know how gene regulation works now that CRISPR has replaced RNA interference as the dominant tool. The technology arrived and suddenly every molecular biology lab looked like it had won the lottery. A lot of people still treat CRISPR like a universal edit button. It isn't. It is a bacterial immune system fragment that has been adapted into a programmable DNA targeting platform, and understanding the difference matters every single time you design an experiment. The core mechanism depends on a guide RNA complementary to your target sequence and a Cas nuclease that physically cuts the DNA. dCas9 is the catalytically dead version that binds but does not cleave. That is the version you use when your goal is regulation rather than mutation. Fuse a repressor domain like KRAB to dCas9 and you get stable transcriptional silencing. Fuse an activator like VP64-p65-Rta or the newer SunTag system and you get upregulation. The difference between knocking out a gene and tuning its expression is a single amino acid change in the Cas protein and a slightly different delivery strategy. That is the whole secret people are chasing, and most of them skip straight to ordering oligos without reading the protein manual.
Unlock The Secrets Of Gene Regulation The Rise Of Crispr Gene Editing
I ran into a specific problem last year that illustrates everything most beginners miss. I was doing a time-course repression experiment in primary human T cells using dCas9-KRAB paired with a custom sgRNA against a ~1200 base pair region upstream of a promoter. Standard design tools put the target binding site at position minus 400 from the transcription start site, which looked perfect on paper. After 72 hours the knockdown was sitting at about 15 percent instead of the 80 to 90 percent I had seen repeatedly in cell lines. The sgRNA was binding correctly. The KRAB domain was functional. The problem was chromatin accessibility. That region sat inside a heterochromatin zone marked by H3K9me3 and dense nucleosome packing. dCas9-KRAB could bind, but the repressive complex could not spread efficiently from that position because the local architecture resisted it. I solved it by shifting the target site approximately 200 base pairs closer to the TSS into a region that mapped to an open chromatin peak from an ATAC-seq dataset I pulled from ENCODE, and I switched to a bipartite sgRNA strategy with two guides spaced about 50 base pairs apart. Repression jumped to around 85 percent within 96 hours. You cannot design CRISPR regulatory experiments purely from sequence. You have to layer in epigenomic context or you waste weeks on guide RNAs that look ideal on a computer and fail in the tube. Guide RNA design deserves the same blunt treatment. Rule one: keep the target sequence between 18 and 22 nucleotides with a 5-prime NGG PAM. Rule two: avoid run-lengths of four or more identical nucleotides, especially poly-T tracts, because RNA polymerase III promoters used in standard vectors will terminate prematurely. Rule three: specificity is not the same as efficiency. A guide with zero predicted off-targets in the human genome is useless if it also has weak on-target binding due to secondary structure or unfavorable dinucleotide composition. Tools like CRISPOR, CHOPCHOP, and Benchling do a reasonable job of ranking guides, but their models assume standard cell line conditions. They will misrank heavily in primary cells, embryos, and hard-to-transfect tissues.
The delivery question usually decides the outcome before anyone writes a protocol. Lipid nanoparticles have matured rapidly since 2020 and now dominate for in vivo work, especially with liver and immune cell targets. Electroporation remains the default for ex vivo applications like CAR-T engineering because it gives you transient expression, which matters when you want to minimize off-target cuts. Plasmid transfection is essentially obsolete for most regulatory work outside of stable cell line generation, and even then viral vectors outcompete it because plasmid integration is stochastic and your selection pressure will favor clones with low dCas9 expression rather than high target engagement. If you need long-term regulation without repeated dosing, lentiviral delivery of a pooled dCas9 plus sgRNA system is the only reliable route, but you pay for it with insertional mutagenesis risk and clonal variability that requires single-cell sorting and validation. Off-target effects in CRISPR gene regulation are different from off-target effects in gene editing. With knockout experiments you can sequence the locus and see indels. With dCas9-KRAB or activation systems you are dealing with epigenetic remodeling, so off-target gene expression changes manifest as transcriptome-wide noise rather than mutations. RNA-seq is mandatory if you care about this, not optional. I have seen people skip RNA-seq and rely solely on qPCR at the intended target, then publish clean results while their pathway analysis shows dozens of unexpected gene expression shifts caused by guide RNA spread. A common workaround is to use truncated sgRNAs with 17 or 18 nucleotides instead of the standard 20. They reduce off-target binding because mismatch tolerance drops sharply with shorter guides, though you trade a bit of on-target efficiency. Another option is fused dCas9 with high-fidelity mutant variants like eSpCas9 or SpCas9-HF1, which have been engineered to reduce non-specific DNA contacts. Those variants cost roughly 20 to 30 percent of your maximum repression or activation strength, so they are not a universal fix. Here is the part most promotional material avoids entirely: CRISPR gene regulation has hard ceilings. It works best as a switch, not a dimmer. dCas9-KRAB repression typically lands between 60 and 90 percent depending on the target and the delivery context. You will rarely achieve complete knockdown because the promoter retains some baseline activity from residual unbound molecules and spontaneous chromatin opening. Activation with single-domain fusions usually tops out around a tenfold increase, which sounds impressive until you compare it to endogenous promoters that can drive hundredfold to thousandfold induction under physiological conditions. If your biological question requires precise titration across orders of magnitude, CRISPRa and CRISPRi are awkward tools. You would be better served by inducible systems like tet-on or chemical dimerizers, or by RNAi for moderate downregulation where complete elimination is not necessary.
Get the Full Details

Another limitation that people encounter repeatedly is persistence. A single round of dCas9 delivery in dividing cells produces repression or activation that decays over roughly five to ten population doublings as the protein turns over and the sgRNA is diluted. Non-dividing cells can maintain regulation for weeks with the right vector, but that advantage evaporates the moment you move to an in vivo setting where clearance rates and immune responses shorten the effective window. If you need chronic regulation, you have to accept stable integration and the associated risks, or return to pharmacological interventions that are reversible by design. For anyone actually running these experiments, the practical workflow starts with defining the biological readout before touching a guide design tool. Sequence the gene, pull the nearest open chromatin data from a relevant tissue, pick three to five guides flanking the regulatory region, order them as synthetic sgRNAs or clone them into a U6-driven vector, transfect or transduce at a low multiplicity to keep expression manageable, and validate at both the RNA and protein level before drawing conclusions. Skipping any step because it feels redundant is how projects go sideways. The technology is powerful, but it rewards careful planning more than it rewards speed.