Why Hydrogen Bonding Keeps Your Computational Chemistry Projects Alive
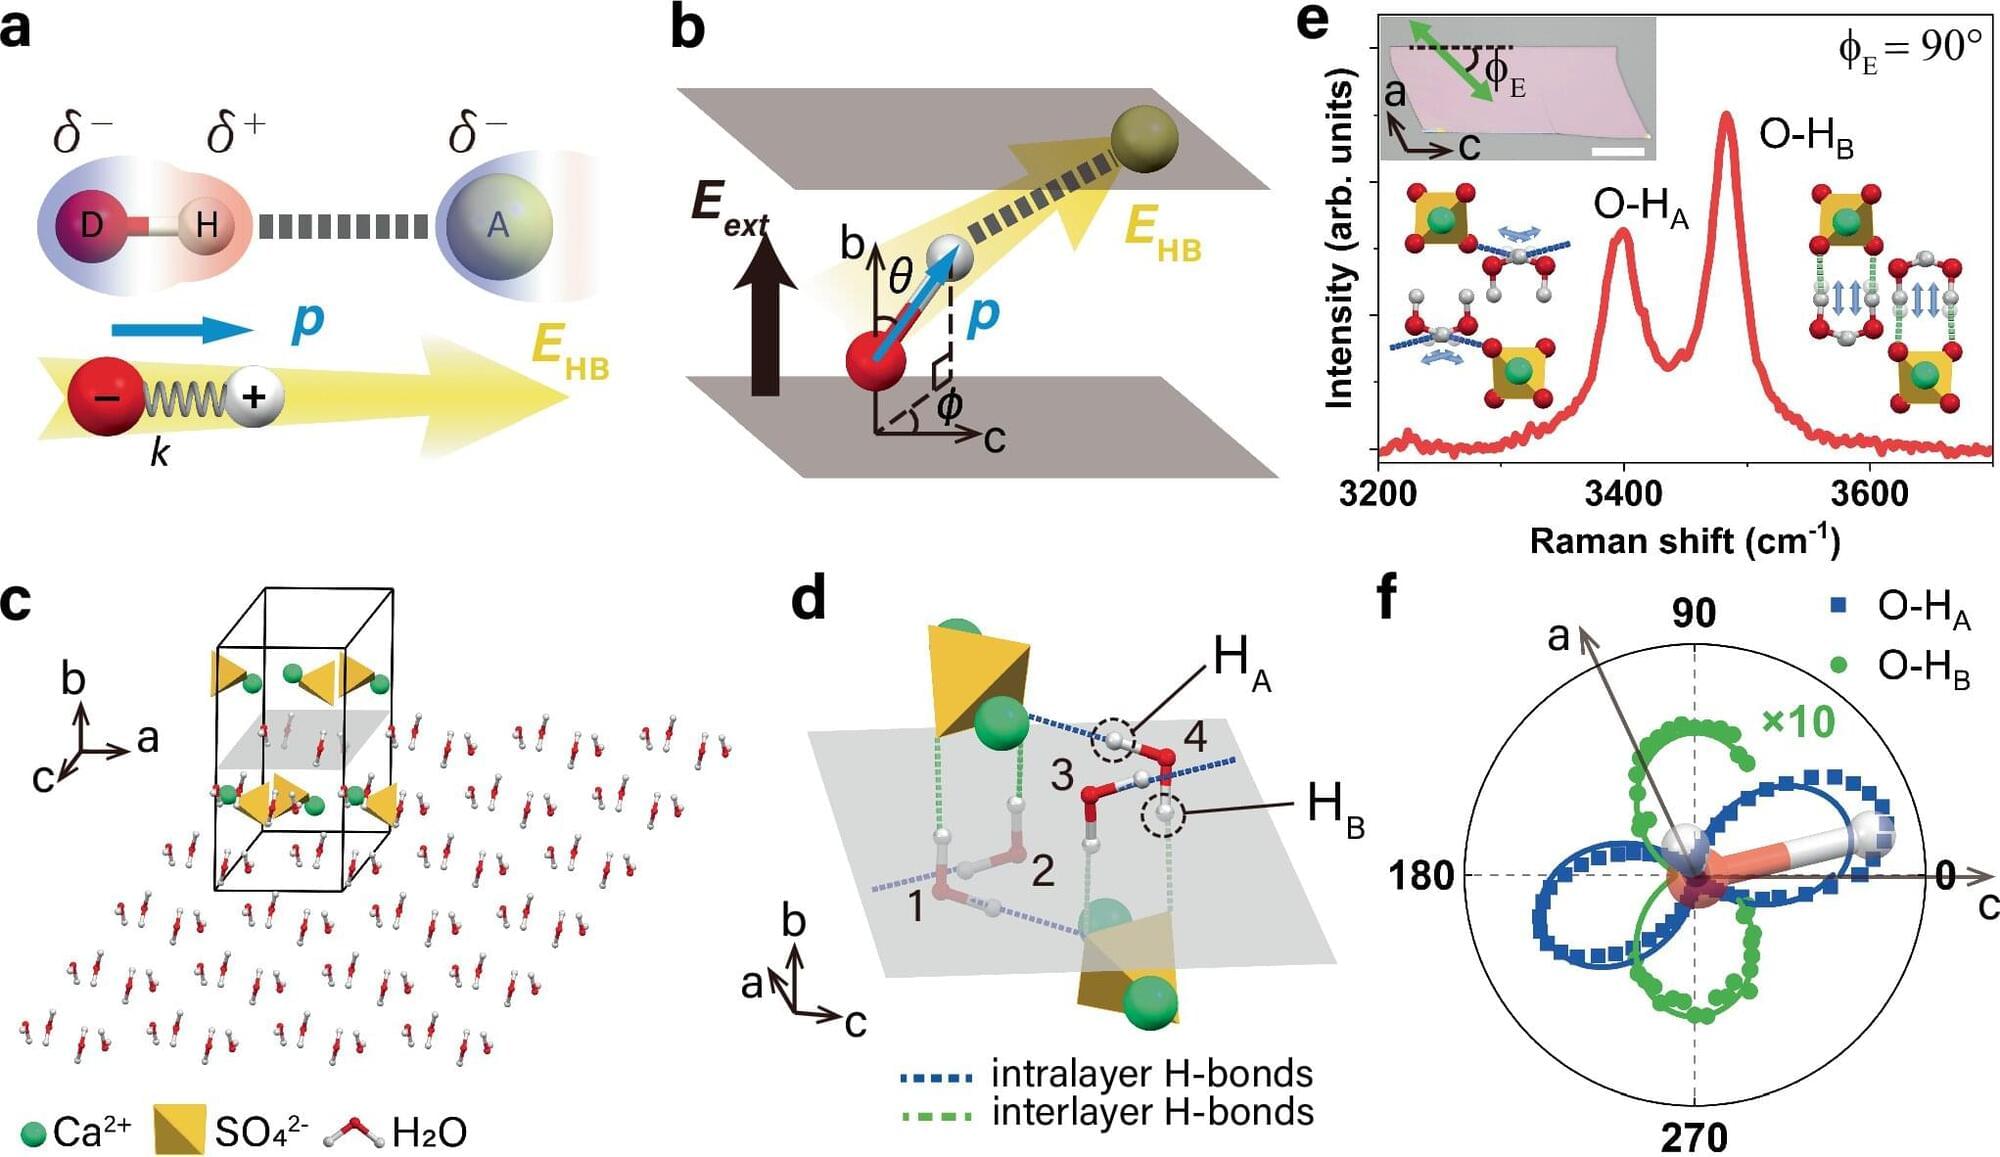
You run a simulation, you optimize a geometry, and then you look at the structure and realize it fell apart somewhere around the active site. The culprit is almost always how you handle hydrogen bonds. People treat them like a checkbox in their methodology section. They are not. They are the reason your protein-ligand binding energy is off by 15 kcal/mol or why your water cluster geometry looks like something a toddler assembled. A hydrogen bond sits in a weird energy window. It is stronger than van der Waals dispersion but weaker than a covalent bond, typically landing between 1 and 10 kcal/mol depending on geometry and environment. That range matters because it means hydrogen bonds are easily disrupted by thermal motion, dielectric screening, or an inadequate level of theory. When you underestimate that window, your results drift. The common textbook description tells you hydrogen bonds are electrostatic attractions between a hydrogen attached to an electronegative atom and another electronegative lone pair. That is true but incomplete. There is a significant charge-transfer component, especially in strong hydrogen bonds like the fluoride-hydrogen fluoride complex. You can have a hydrogen bond where the donor and acceptor orbitals actually overlap enough that you are describing it more accurately as a partial covalent interaction. That happens in short, strong hydrogen bonds where the donor-acceptor distance drops below 2.5 angstroms for O-H...O systems. Standard force fields do not capture any of this.
I learned this the hard way when I was modeling a protease inhibitor binding to a serine active site. The ligand formed what looked like a standard water-bridged hydrogen bond network in the X-ray structure. My first molecular dynamics run with a standard AMBER force field showed the bridge collapsing within 2 nanoseconds. The protein collapsed around it in a way that no one would consider realistic. The problem was not the force field being broken. It was that the bridge involved a low-barrier hydrogen bond where the proton sits nearly equidistant between two oxygen atoms. The harmonic bond parameters in the force field cannot model a delocalized proton in that regime. The energy well is too flat. The workaround was to isolate that specific interaction and run a quantum mechanical calculation on a truncated model. I took the relevant residues, capped the peptide bonds with acetyl and N-methyl groups, and ran a DFT optimization at the B3LYP-D3BJ/def2-TZVP level. That gave me the proper potential energy surface for the proton position. I then used those distances and angles to manually adjust the restraint potentials in the MD setup rather than trying to force the classical force field to describe something it physically cannot. The simulation stabilized immediately after that change. The binding energy shifted by about 4 kcal/mol compared to the uncorrected setup, which is the difference between a compound that looks promising and one that looks like noise. Here is what most people miss when they start working with hydrogen bonds computationally. Dispersion corrections matter more than you think. A lot of people throw in a generic continuum solvation model and call it good. But the hydrogen bond network in proteins and nucleic acids is heavily dependent on the local dielectric environment. If you model a salt bridge in water using a simple implicit solvent model with a high dielectric constant, you will artificially weaken or eliminate the interaction. The salt bridge might register as zero binding contribution when it should contribute roughly 3 to 5 kcal/mol. I switched to explicit solvent molecules around the key interaction sites and recalibrated. The binding profile changed enough that I had to re-rank three compounds that were originally predicted to be lead candidates.
Another thing nobody warns you about is the temperature dependence. Hydrogen bonds weaken as temperature increases, and the effect is non-linear. The bond energy does not simply scale with kT. For a typical O-H...O hydrogen bond at room temperature, thermal fluctuations can break and reform the bond on a picosecond timescale. That is fast enough that if your simulation time is shorter than 50 nanoseconds, you are not sampling the interaction properly. You are seeing a snapshot, not a distribution. I ran a series of test simulations at 298K, 310K, and 325K on the same system and saw the average hydrogen bond lifetime drop from about 12 picoseconds to roughly 4 picoseconds. That is not a small shift. It changes how you interpret RMSD plots and binding stability metrics. If you are doing this kind of work regularly, your toolkit should include at least one DFT package with good dispersion support, a molecular dynamics engine that handles explicit solvent well, and something like Multiwfn or AIMAll for analyzing the electron density topology around hydrogen bonds. The quantum theory of atoms in molecules approach gives you a Laplacian of the electron density at the bond critical point. That is a more reliable indicator of whether a hydrogen bond is genuinely present than looking at distances alone. A positive Laplacian with a moderate electron density value at the critical point usually confirms a closed-shell interaction typical of hydrogen bonding. If the density is too low, you might be looking at weak van der Waals contact masquerading as a hydrogen bond. There are limits to what you can do here. Not every system can be treated with QM/MM. The computational cost scales badly, and the boundary treatment between the quantum region and the molecular mechanics region introduces artifacts that are hard to quantify. If your hydrogen bond network spans a large area, like in a hydrated ion channel or a protein-protein interface, you are often stuck with classical approaches and you have to accept that your accuracy has a floor. In those cases, using polarizable force fields like AMOEBA or Drude oscillator models can help, but they are significantly more expensive and not always better depending on the system.
Get the Full Details

The practical takeaway is straightforward. Check your hydrogen bonds explicitly before you trust any binding energy or structural result. Verify that your method can actually describe the interaction regime you are studying. Don't assume a standard force field covers everything. The effort to validate that single step saves you from building an entire project on a flawed foundation.