
So you need to figure out when a molecule is actually flat.
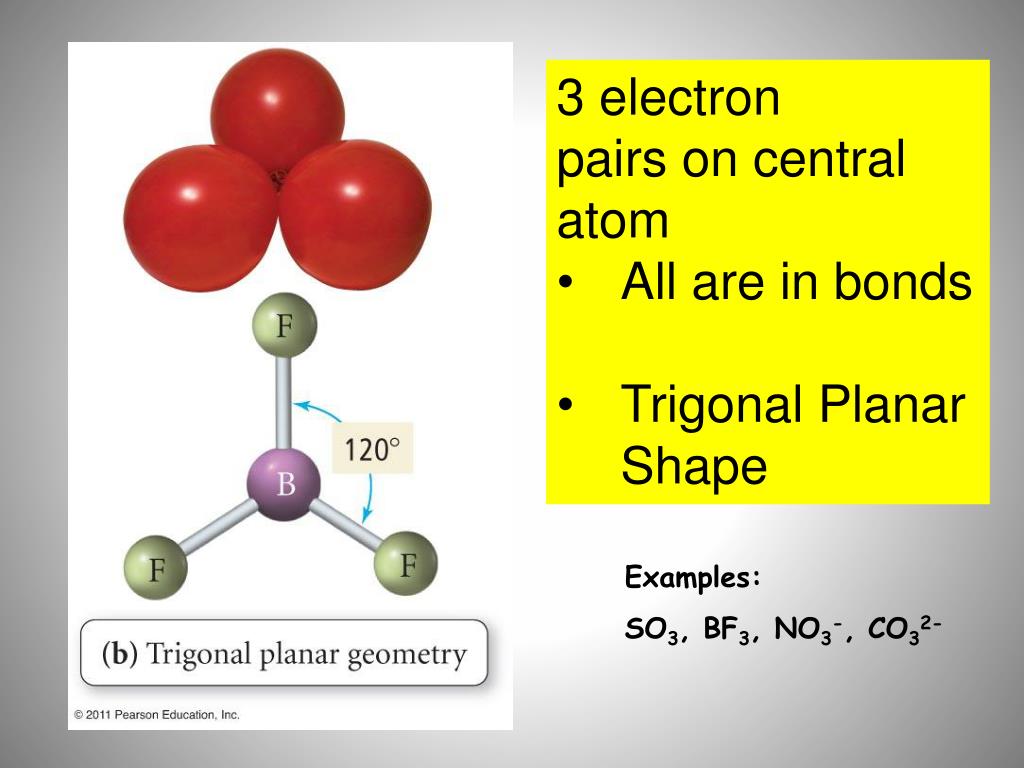
I spend a lot of time going back to VSEPR theory and finding out it gives you the wrong answer more often than you'd think. The standard textbook approach works fine for textbook problems, but real molecules don't always cooperate. Trigonal planar geometry is one of those cases where the rule is simple to state and nearly impossible to apply correctly without knowing where it breaks. The basic shape involves a central atom bonded to three surrounding atoms with no lone pairs taking up space. All four atoms sit in the same plane. Bond angles are ideally 120 degrees. The hybridization is sp2, which means one s orbital and two p orbitals mix to form three equivalent hybrid orbitals pointing outward at those 120-degree intervals. A fourth p orbital stays perpendicular to the plane and is available for pi bonding.
Trigonal Planar Geometry The Perfect Balance Of Atoms In Core Molecules
BF3 is the classic example. Boron has three valence electrons, each bonds to fluorine, and there is no lone pair left. The molecule is flat. SO3 has sulfur double-bonded to three oxygens in a resonance hybrid. The effective geometry is still trigonal planar. The carbonate ion (CO3^2-) is another straightforward case with three resonance structures distributing the double bond character evenly. Here is where things get messy in practice. I was running DFT calculations on a substituted boron compound last year — a boron center with two aromatic amines and one alkyl group. The molecule should have been trigonal planar, and the initial geometry came out nearly flat. But when I relaxed it further, the boron started tilting out of plane by about 8 degrees. Not a huge amount, but enough to mess with my orbital analysis. The workaround was forcing a symmetry constraint during the optimization step. Without that, the optimizer would find a slightly pyramidal local minimum that looked fine on paper but didn't match experimental UV-Vis data. This is worth paying attention to because many molecules that look trigonal planar at first glance actually have a shallow energy surface near the planar geometry. Small perturbations — solvent effects, crystal packing forces, substituent bulk — can push the central atom out of plane. If you're working with computational chemistry, always check the imaginary frequencies after optimization. If you see one, your structure didn't actually converge to a true planar minimum. Run the job again with tighter convergence criteria or add a dispersion correction if you aren't already using one. Most standard functionals underbind in these systems by about 2 to 4 kcal/mol, which sounds small until it ruins your barrier height predictions.
The sp2 hybridization model also has a limit that beginners consistently miss. It works well for molecules where all three ligands are identical or nearly identical. But as soon as you introduce ligands with very different electronegativities or steric demands, the bond angles shift. CH2O has a C=O double bond and two hydrogens, and the H-C-H angle is about 116.5 degrees while the O-C-H angles are around 121.7 degrees. The double bond takes up more space. The simple 120-degree model is wrong by several degrees, and those several degrees matter when you're trying to predict reactivity patterns or interpret NMR coupling constants. Another counter-intuitive point: not every molecule with three substituents is trigonal planar. Nitrogen trifluoride (NF3) has three bonds and one lone pair. The lone pair occupies the fourth sp3 orbital and pushes the bonds down into a pyramidal shape with angles around 102 degrees. The common mistake is looking at the number of bonded atoms and assuming geometry. You always need to count the electron domains, not just the bonds. That distinction collapses quickly when you start looking at transition metal complexes or hypervalent main group compounds where the rules change entirely. For people who need a quick reference, most introductory chemistry platforms offer geometry prediction tools, but they tend to assume idealized conditions. I usually just run through the VSEPR steps manually: count valence electrons, draw the Lewis structure, count bonding and lone pair domains, assign the base geometry, then adjust for multiple bonds and lone pairs. It takes about 30 seconds once you've done it enough times and is more reliable than most online calculators for anything outside the standard cases.
Get the Full Details
The main downsides to relying on this model are straightforward. It breaks down for molecules with significant d-orbital participation, which becomes relevant past the second row of the periodic table. It doesn't handle radical species well without modification. And it gives you no information about the electronic structure beyond geometry — you still need molecular orbital theory or a computational method to understand bonding properly. If you need to model larger systems where planar geometries are in flux, consider combining VSEPR predictions with a quick semiempirical calculation. It adds maybe 10 to 15 minutes to your workflow but saves you from building a model that looks right on paper and fails immediately in simulation. For actual computational work, Gaussian or ORCA with a modest basis set like 6-31G* is enough for initial geometry determination. Tighten the convergence and switch to a larger basis set only if you need accurate energies. Don't waste CPU time running a triple-zeta calculation on a geometry that hasn't been verified at a lower level first. That habit alone saved me roughly a week of compute time on a project last semester.


