
What You Actually Need To Know About Molecular Polarity
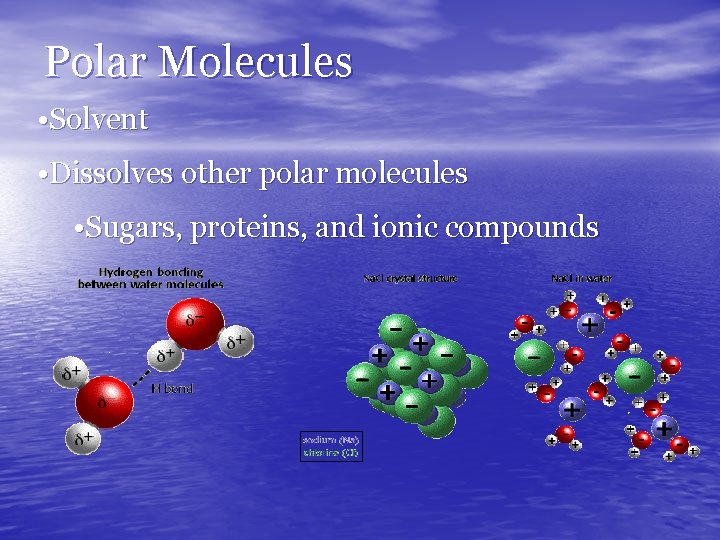
Polarity comes down to electronegativity differences between atoms in a bond. When one atom pulls electrons harder than its neighbor, you get a partial positive and partial negative charge sitting across the molecule. That's it. But the consequences are where things get messy, because molecular geometry also matters. A CO2 molecule has polar C=O bonds, but they're arranged symmetrically, so the dipoles cancel and the molecule itself is non-polar. Water has two O-H bonds at roughly 104.5 degrees, which means the dipoles reinforce each other instead of canceling. Most people stop at the definition and never actually deal with the real problems this causes in a lab setting. Here's how it hits you when you're working with actual chemicals rather than textbook diagrams.
The Secret World Of Polar Molecules Unlocking The Mysteries Of Chemistry
Let me walk through a practical workflow first, then fill in what's happening underneath. Say you need to separate a polar product from a reaction mixture containing unreacted starting material and some non-polar byproducts. Flash chromatography on silica is your standard move. Load your crude mixture onto a silica column and elute with an increasing polarity gradient. Start with something like 10% ethyl acetate in hexanes and ramp up to 50-80% over the course of the run. Collect fractions, analyze by TLC, and pool the ones containing your product. The silica gel is polar itself, which is why this works. Polar compounds stick to it harder. Non-polar compounds zip right through. The trick is knowing when your compound is going to be stubborn. If you're running a compound that has multiple hydroxyl groups or an amine, it's going to tail badly on silica, sometimes irreversibly. I spent three solid days chasing a polyhydroxylated natural product through a silica column last year, wasting about two grams of material before I realized the compound was literally adsorbing to the glassware. I switched to a reversed-phase C18 column with an acetonitrile-water gradient and got clean separation in forty minutes. Silica wasn't the problem. The compound just didn't want to leave it. Here's the technical explanation for why that happened, coming back to it. Silica has surface silanol groups (Si-OH) that form hydrogen bonds with polar functional groups on your molecule. Multiple hydroxyls on your compound mean multiple hydrogen bonding sites, which means strong adsorption and slow elution. Reversed-phase chromatography uses a non-polar stationary phase (C18 chains bonded to silica), so your polar compound spends more time in the mobile phase and moves faster. The elution order essentially inverts.
Choosing The Right Solvent System
Solvent selection is where most people make mistakes. The dielectric constant is the number you want to look at, but it's only part of the picture. Dimethyl sulfoxide has a dielectric constant of 47, which makes it excellent for dissolving ionic and highly polar compounds. But it's nearly impossible to remove completely from your product. I've seen people try to rotary evaporate DMSO off at 40 millibar and 60 degrees Celsius for six hours, only to still have residue. It boils at 189 degrees Celsius at atmospheric pressure, and even under vacuum it leaves traces that show up in NMR and mess with downstream reactions. Acetonitrile, on the other hand, has a dielectric constant around 37 and evaporates cleanly. It's a better general-purpose polar solvent for workup and concentration. Methanol is another option, but it's protic, which means it can participate in hydrogen bonding with your compound and sometimes creates emulsions during aqueous workup. I learned that the hard way with a peptide synthesis where the methanol-water mixture formed a stable emulsion that wouldn't break for hours. Switching to ethyl acetate and water resolved it in ten minutes. The rule of thumb is: use the least polar solvent that still dissolves your compound. This gives you better crystallization outcomes and easier purification later. If your compound won't dissolve in anything reasonable, you may have a structural problem rather than a solvent problem.
Get the Full Details

Understanding Dipole Moments And Their Measurement
A dipole moment is measured in Debyes. Water has one of 1.85 D. Chloroform is 1.04 D. Carbon tetrachloride is zero because of symmetry. These numbers aren't just academic — they tell you how a molecule will behave in an electric field, how it packs in the solid state, and how it interacts with other molecules in solution. When you're measuring dipole moments experimentally, you typically use the Debye equation, which relates the dielectric constant of a solution to the concentration and temperature. The procedure involves preparing several solutions of your compound in a non-polar solvent like benzene or cyclohexane, measuring the dielectric constant of each, and then extrapolating to infinite dilution. The extrapolation step is critical because intermolecular interactions at higher concentrations distort the measurement. I once skipped the extrapolation and got a dipole moment that was about 15% too high because my compound had some dipole-dipole association in solution. That error propagated into my calculation of the molecular structure and I had to redo the whole series. There's also the matter of bond dipoles versus molecular dipoles. A C-Cl bond has a dipole moment of about 1.5 D, but chloroform's molecular dipole is 1.04 D because the three C-Cl bonds don't point in the same direction. Tetrahedral geometry means the vector sum is less than any individual bond contribution. This is why understanding molecular geometry is non-negotiable if you want to predict polarity from structure alone.
Hydrogen Bonding And Its Practical Implications
Hydrogen bonds are directional intermolecular interactions that occur when hydrogen is bonded to nitrogen, oxygen, or fluorine. They're stronger than van der Waals forces but weaker than covalent bonds, typically in the range of 4 to 30 kJ/mol depending on the partners involved. This strength range is exactly why they dominate the behavior of water, alcohols, amines, and carboxylic acids. Hydrogen bonding affects boiling points dramatically. Compare dimethyl ether (boiling point -24°C) to ethanol (boiling point 78°C). Same molecular formula, C2H6O. The only difference is that ethanol has an O-H group capable of hydrogen bonding while dimethyl ether doesn't. That's a 102-degree difference caused entirely by one functional group arrangement. For chromatography, hydrogen bonding is the primary mechanism by which compounds retain on silica. If your compound has both a hydrogen bond donor and acceptor, it's going to interact strongly with the silanol groups. This is why amino acids are difficult to separate on silica — they have both amine and carboxylic acid groups. I usually add a small amount of triethylamine or acetic acid to the mobile phase to block silanol interactions and sharpen the peaks. Without that additive, your amino acid peak will be broad and asymmetrical, making fraction collection imprecise.
Spectroscopic Identification Of Polar Groups
Infrared spectroscopy is the fastest way to identify polar functional groups. An O-H stretch appears as a broad band around 3200-3600 cm¹. A C=O stretch shows up around 1650-1750 cm¹ depending on the exact group. N-H stretches appear around 3300-3500 cm¹ but are usually sharper than O-H bands. If you're working in CDCl3 for NMR and your compound has an exchangeable proton, adding a drop of D2O to the NMR tube will make that signal disappear through deuterium exchange. This is a quick confirmatory test for hydroxyl, amine, and carboxylic acid protons. Mass spectrometry can also give you clues. Polar compounds tend to ionize well with electrospray ionization (ESI) in positive or negative mode, depending on whether they have basic or acidic groups. Electronspray is soft ionization, so you usually see the molecular ion [M+H] or [M-H] rather than extensive fragmentation. This is useful for determining molecular weight without the complications you get from electron impact ionization, which produces a lot of fragment ions that can make interpretation harder.
Common Pitfalls And Where People Go Wrong
The biggest mistake I see is assuming that polarity is a binary property. It isn't. Every molecule exists on a continuum, and the context determines how polar it behaves. A molecule with a long hydrocarbon chain and one hydroxyl group may behave more like a non-polar compound in chromatography than a polar one, simply because the hydrophobic portion dominates the interaction with the stationary phase. This is why C18 columns are used for reverse-phase separations of seemingly polar molecules like fatty acids. Another issue is temperature dependence. Polarity isn't constant across temperatures. Dielectric constants decrease as temperature increases, which means a solvent that works fine at room temperature may not dissolve your compound at elevated temperatures the way you expect. I ran a reaction at 80°C in DMF and found that my product precipitated out as it cooled, which actually simplified the workup. If I'd anticipated that, I could have skipped a purification step entirely. Temperature effects are often overlooked in method development. Water content is another factor that people don't account for enough. Most polar solvents are hygroscopic. Acetonitrile, DMF, and DMSO all absorb water from the air over time. If you're running reactions that are sensitive to water, this absorbed moisture can interfere. I once had a Grignard reaction fail because I used acetonitrile that had been sitting open for a few days. The water content was low enough that it didn't kill the reaction outright, but it was enough to reduce the yield significantly. Molecular sieves or freshly opened solvent bottles solve this.
When Polarity Predictions Fail
Computational chemistry tools can predict dipole moments and partial charges fairly accurately these days. Software like Gaussian or ORCA using DFT functionals will give you numbers that are usually within 5-10% of experimental values. But there are cases where computation breaks down. Highly conjugated systems with charge transfer character, molecules in excited states, and systems with strong electron correlation effects can produce unreliable results. I worked on a project involving a push-pull chromophore where the calculated dipole moment was off by about 30% compared to the experimental value in solution. The issue was that the ground-state geometry optimization didn't account for solvent effects adequately. Using a PCM (Polarizable Continuum Model) to simulate solvent corrected most of the error, but it required running multiple calculations at different dielectric constants to match the actual solvent. For most practical purposes in the lab, though, you don't need computational chemistry. You need to know whether your compound will dissolve in a given solvent, how it will behave on a column, and what purification strategy to use. That comes from experience with similar structures and a basic understanding of the underlying principles. TheSecret World Of Polar Molecules Unlocking The Mysteries Of Chemistry isn't really a secret at all — it's a set of patterns that become obvious once you've dealt with enough compounds to recognize them. If you're just starting out, focus on memorizing the common polar functional groups and their characteristic behaviors: alcohols and acids dissolve in water, amines are basic and can be extracted into aqueous acid, halides are generally non-polar unless they're polyhalogenated. Then build from there by running thin-layer chromatography on compounds with known polarity and comparing Rf values. This hands-on approach is faster and more reliable than trying to predict everything from first principles.


