Converting Alkenes to Alcohols
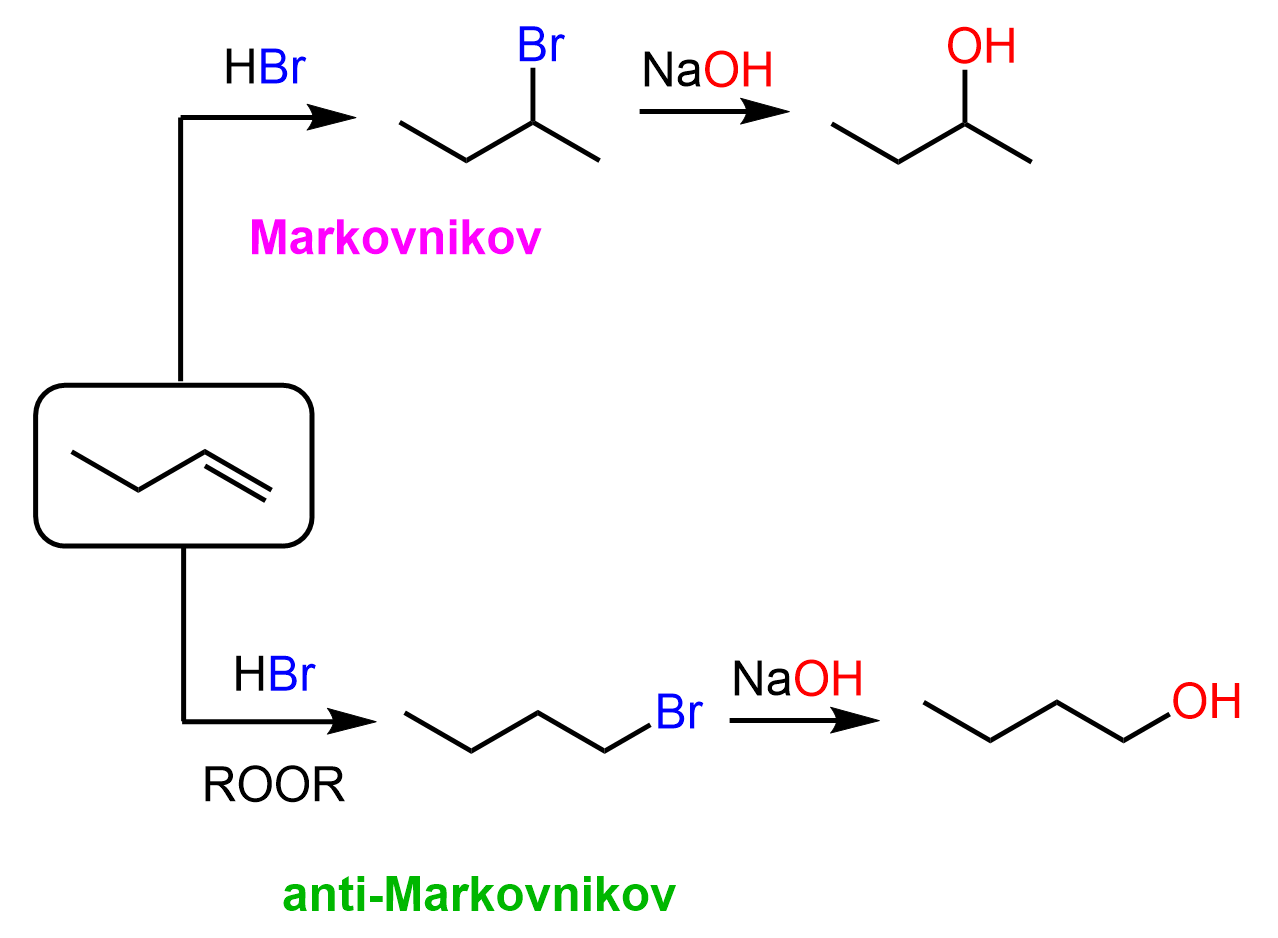
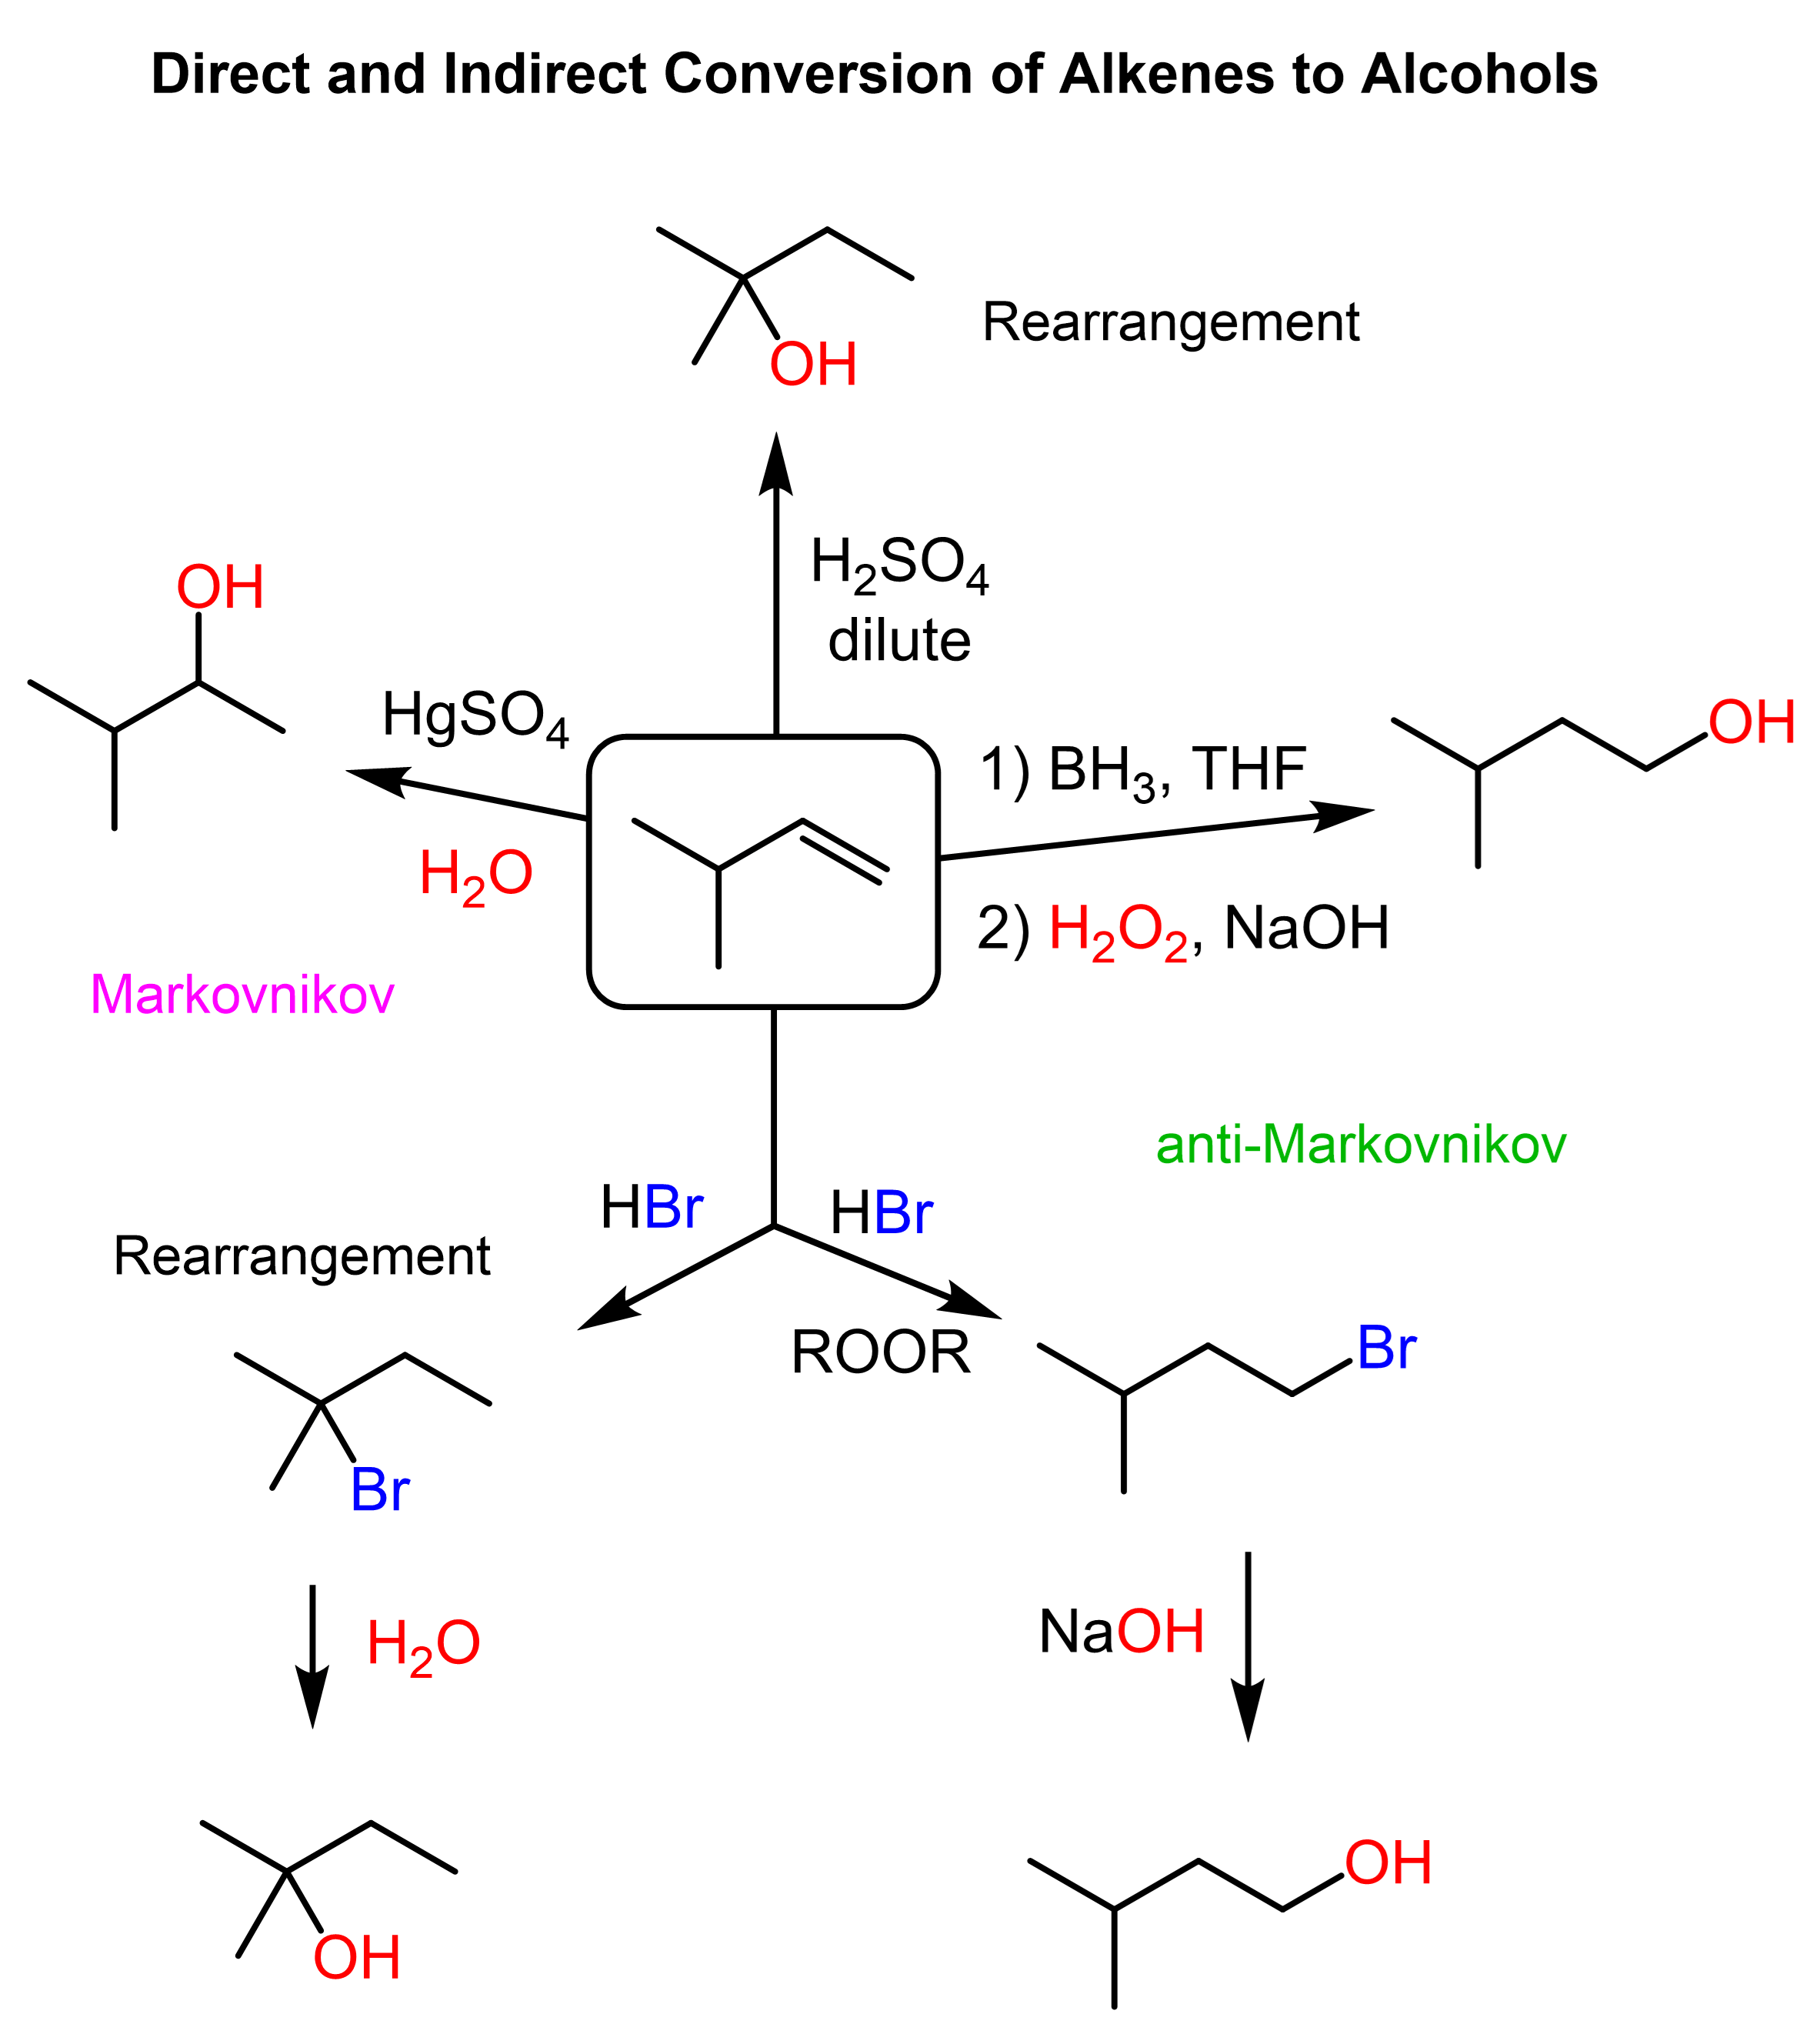
The most common question I see in practice is how to reliably turn a double bond into an alcohol group. It sounds simple because the textbook answer is simple. It is only simple if you know which pathway to reach for and you do not force a reagent into a substrate where it will not cooperate. The three main methods each give you a different regiochemical outcome. That outcome is usually the only thing that matters when you are trying to hit a specific product. If you strip away the textbook language, what you are really doing is adding water across a carbon-carbon double bond. The bond breaks, a hydrogen attaches to one carbon, a hydroxyl group attaches to the other, and you have your alcohol. The whole problem is just figuring out which carbon gets which piece of water, and then picking the right set of conditions so you do not spend six hours watching nothing happen in the flask. Acid-catalyzed hydration is the first method people learn. You treat the alkene with water in the presence of a strong acid like sulfuric acid or phosphoric acid. The mechanism goes through a carbocation intermediate, which means the hydroxyl group lands on the more substituted carbon. That is Markovnikov selectivity. The practical version is: mix the alkene with dilute sulfuric acid and water, warm it gently, and let it sit. For terminal alkenes like 1-hexene, you get 2-hexanol as the major product. If your substrate has the potential to rearrange, that rearrangement will happen. I learned this the hard way with a cyclohexenyl substrate where a hydride shift scrambled the expected product. The workaround was abandoning acid catalysis entirely and switching to oxymercuration.
Oxymercuration-demercuration gives you Markovnikov addition without carbocation rearrangement. You treat the alkene with mercuric acetate in water and tetrahydrofuran, then reduce the organomercury intermediate with sodium borohydride. The hydroxyl still ends up on the more substituted carbon, but the reaction proceeds through a mercurinium ion rather than a free carbocation. That is the whole advantage. I once ran this on a substrate with a neighboring tertiary center that kept wanting to rearrange under acidic conditions. The oxymercuration route gave clean conversion in about 45 minutes at room temperature. The yield was around 82 percent after workup. Mercury waste is a pain to handle and dispose of, but the selectivity is usually worth the extra step. Hydroboration-oxidation is the method you use when you need the opposite regiochemistry. You add borane or a substituted borane across the double bond first, then oxidize with hydrogen peroxide and base. The hydroxyl group lands on the less substituted carbon. That is anti-Markovnikov selectivity. The reaction is also stereospecific, giving syn addition of the boron and hydrogen. For terminal alkenes this means you get primary alcohols where acid-catalyzed hydration would give you a secondary product. I routinely use 9-borabicyclo[3.3.1]nonane, commonly called 9-BBN, when I need better selectivity than plain borane gives me. It is more expensive, but the regioselectivity is noticeably sharper on disubstituted alkenes. Here is a detail beginners tend to miss. The choice of solvent matters more than the literature suggests. In oxymercuration, using methanol instead of water as the cosolvent changes the product entirely. You get a methoxymercurial instead of a hydroxymercurial, and after demercuration you have an ether, not an alcohol. I have seen this come up repeatedly when people reuse solvent from a previous procedure without checking what was in it. Same problem with hydroboration. If your borane reagent has been sitting open to the air, the concentration is lower than the label says and your reaction time stretches out unpredictably.
Epoxidation followed by ring opening is the fourth option worth knowing about. You convert the alkene to an epoxide using a peracid like mCPBA, then open the ring with aqueous acid or base to get the alcohol. This gives you anti addition overall and works well when you need two functional groups on adjacent carbons. The limitation is that epoxides can polymerize under acidic conditions if the substrate is strained or if you heat the reaction too long. I keep the pH near neutral and run the opening at low temperature whenever possible. Workup is where most people lose yield, not the reaction itself. After hydroboration-oxidation, you typically have boron byproducts that form a gummy emulsion during extraction. Adding a saturated brine wash and letting the layers separate fully before collecting the organic layer makes a noticeable difference. With oxymercuration, you need to remove mercury thoroughly. A wash with dilute sodium sulfide solution precipitates residual mercury and pulls it into the aqueous phase. Skip this step and you will have mercury contamination in your organic layer that shows up later as a persistent impurity on the NMR. The harsh truth about these methods is that none of them handle every substrate equally well. Acid-catalyzed hydration fails on sterically hindered alkenes and rearranges sensitive skeletons. Oxymercuration works broadly but requires mercury handling and generates toxic waste. Hydroboration is clean but borane reagents are pyrophoric and the oxidation step can over-oxidize if you are not careful about temperature and stoichiometry. Epoxidation opens another whole set of problems with peracid handling and epoxide stability.
Get the Full Details

If your alkene is electron-poor or conjugated with a carbonyl group, standard hydration conditions slow down considerably. In those cases I switch to a Lewis acid assisted pathway or use a phase-transfer catalyst to push the reaction forward. It adds a step to the procedure but saves you from running the reaction for 24 hours and getting 30 percent yield. That trade-off is almost always worth it. The bottom line is that alkene to alcohol conversion is routine once you stop treating it as a single method. Pick the regiochemistry you need, choose the conditions that avoid rearrangement for your specific substrate, and pay attention to workup. The chemistry itself is straightforward. The details are where the time goes.