Working With Nucleotides When You Are Not Sure What You Are Looking For
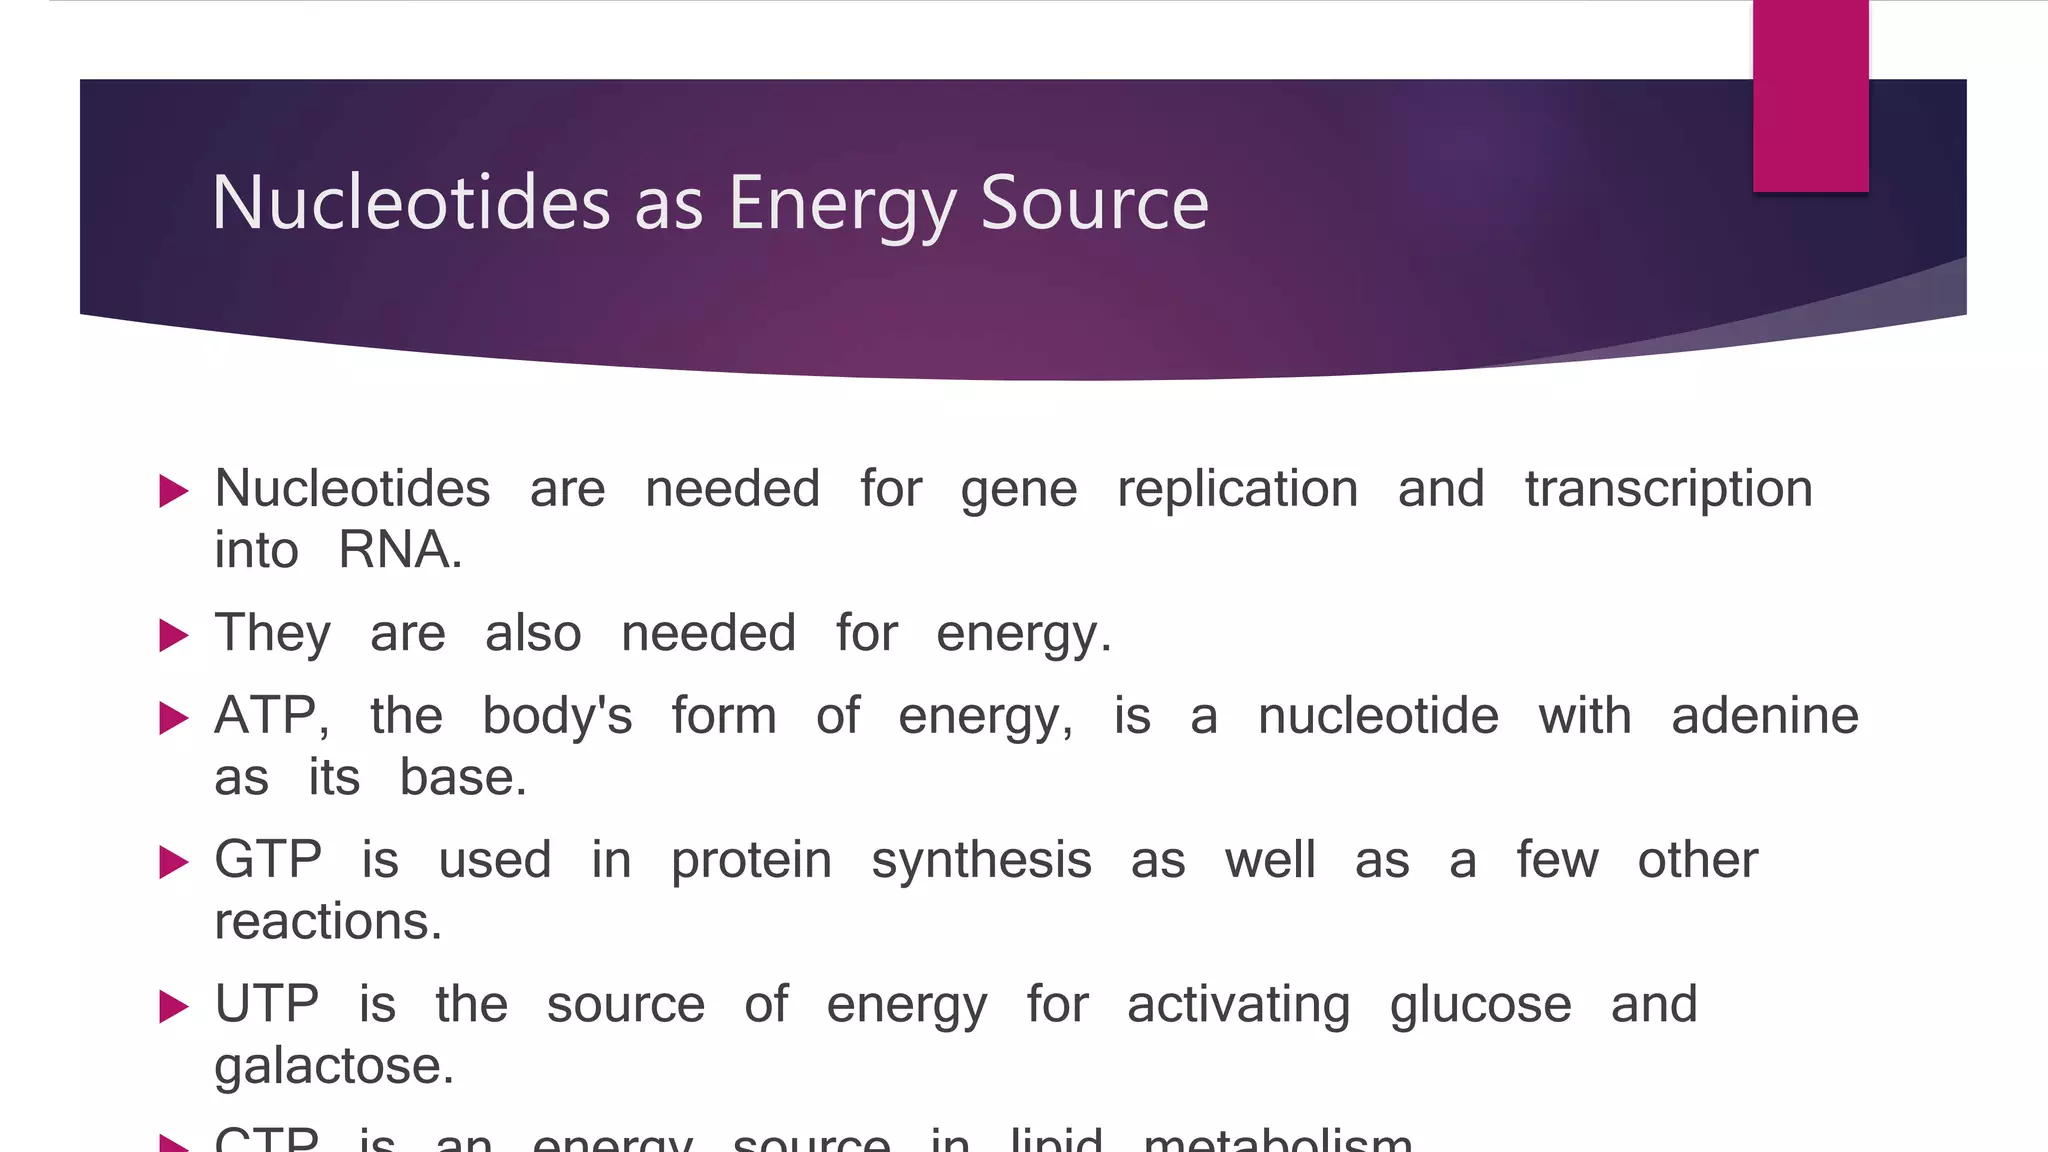
I spent three days troubleshooting a PCR reaction that refused to amplify. The primers were fine, the template was clean, the thermocycler was calibrated. The issue turned out to be the dNTP concentration in my master mix. I had used a stock that had been frozen and thawed roughly twenty times, and the ATP component had degraded just enough to throw off the balance. The reaction wasn't failing because of anything complicated. It was failing because nucleotides are fragile in ways that most protocols gloss over. That is the kind of thing you learn after you have ruined enough samples. Nucleotides are molecules made of three parts: a nitrogenous base, a five-carbon sugar, and one or more phosphate groups. The bases are adenine, guanine, cytosine, thymine, and uracil. When you string them together, you get DNA or RNA. When you add phosphates, you get energy carriers like ATP, GTP, CTP, and UTP. They do both things at once in every cell. That dual role is what makes them important outside of a textbook diagram. In the body, ATP is the primary energy currency, but it is not the only one. GTP drives protein synthesis at the ribosome. CTP is required for lipid metabolism, specifically in the Kennedy pathway for phospholipid production. UTP activates sugar residues during glycogen synthesis. If any of these fall out of balance, metabolic pathways slow down or stall. The body recycles them constantly. A single ATP molecule may be hydrolyzed and regenerated thousands of times per day in an active cell. You are not running on a battery. You are running on a spinning wheel.
I learned this the hard way when I was preparing cell lysates for a metabolomics run. The lab had a standard protocol that said to snap-freeze cells in liquid nitrogen and store them at minus eighty. Simple enough. But the protocol did not mention that ATP degrades within minutes if you let the sample warm even slightly during homogenization. My first batch showed ATP levels that looked plausible until I ran a control. The control was a fresh sample processed in under two minutes. The difference was a factor of four. I had essentially measured nothing but degradation products. The workaround was straightforward: I added an ice-cold quenching buffer containing perchloric acid directly to the culture dish before harvesting, then neutralized and centrifuged. This locked the metabolite profile in place almost instantly. It added twenty minutes to the workflow but saved the data. Beyond basic biochemistry, nucleotides are central to nearly every major molecular biology technique. PCR depends on dNTPs. RT-qPCR depends on them too, plus a reverse transcriptase that occasionally misincorporates if the mix is old. Sanger sequencing uses dideoxynucleotides, which are just modified nucleotides that terminate chain elongation. Next-generation sequencing libraries are built by ligating adapters to fragmented DNA, a process that requires ATP for the ligase enzyme. CRISPR-Cas9 cutting efficiency varies with nucleotide availability in the cell because the repair machinery that follows uses nucleotide substrates for homology-directed repair. If you are working in a nutrient-poor environment, HDR efficiency drops significantly compared to nucleotide-replete conditions. There is a misconception that more nucleotides in a reaction mix always means better results. It does not. Excess dNTPs can chelate magnesium ions, which are required as cofactors for polymerases. Most protocols call for 200 micromolar of each dNTP. Going higher often reduces fidelity and increases misincorporation rates. Going lower causes premature strand dissociation. The sweet spot is narrow, and different polymerases have different tolerances. Some high-fidelity enzymes prefer 100 micromolar. Standard Taq prefers around 200. Check the manufacturer sheet. The default recipe on the internet is not universal.
Another thing people overlook is the difference between nucleotide supplements and what your cells actually need. Over-the-counter nucleotide supplements claim to boost immunity and repair tissue. The evidence for systemic benefit in healthy adults is weak. Your body synthesizes purine and pyrimidine nucleotides de novo through well-characterized pathways, and it has salvage pathways that recycle bases from degraded RNA and DNA. Oral nucleotides get broken down by intestinal enzymes and hepatic metabolism before they reach circulation in any meaningful form. The claim that swallowing inosine powder boosts your immune function is not supported by clinical data. It is not completely baseless either. There is modest evidence that nucleotide supplementation helps in specific contexts: critically ill patients, people with malabsorption, or athletes undergoing extreme volume training where turnover exceeds normal synthesis capacity. For the average person, the body already handles it. From a research perspective, the useful applications are far more concrete than supplement marketing suggests. Nucleotide analogs are the backbone of antiviral drugs. Acyclovir, for example, is a guanosine analog that gets phosphorylated by viral thymidine kinase and then incorporated into viral DNA, causing termination. Remdesivir is an adenosine analog that interferes with RNA-dependent RNA polymerase in coronaviruses. These drugs work because viruses lack the proofreading mechanisms that eukaryotic cells have, making them far more susceptible to nucleotide misincorporation. The same principle applies to cancer chemotherapy. 5-fluorouracil is a pyrimidine analog that inhibits thymidylate synthase, blocking DNA synthesis in rapidly dividing cells. It is one of the oldest anticancer drugs still in use because it targets a fundamental dependency. If you are running experiments that involve nucleotides, the practical details matter more than the theory. Store dNTP stocks at minus twenty in small aliquots. Never freeze-thaw more than three times. Aliquot sizes should match your typical usage to avoid unnecessary handling. Check absorbance at 260 nanometers to verify concentration before a critical experiment. A nominal 100 millimolar stock can easily drift to 80 or 70 if it has been sitting open on the bench. Use Nuclease-free water, not regular water. RNases and DNases are everywhere, and they degrade your nucleotides and your templates without warning. Use low-binding tubes. Nucleotides stick to plastic, and the loss is measurable over time, especially at low concentrations.
Get the Full Details
One edge case that almost got me fired from a collaboration was the interaction between nucleotide concentration and magnesium in gel electrophoresis. I was running a TBE gel with a high RNA load and noticed smeared bands. The issue was not the gel percentage or the voltage. It was that the RNA was chelating the magnesium in the running buffer, and the complex was migrating unpredictably. Adding extra magnesium to the buffer corrected it. This is a niche problem, but it happens when you are working with concentrated RNA samples and you assume the buffer composition is fixed. It is not fixed if your sample changes the chemistry mid-run. The broader scientific breakthroughs driven by nucleotide research are easier to point to than the daily lab frustrations. The development of CRISPR itself depended on understanding how bacterial cells use short RNA guides, which are nucleotide polymers, to recognize and cut invading DNA. The entire field of synthetic biology is built on the ability to chemically synthesize nucleotide sequences of arbitrary length. You can now order a two-hundred-nucleotide single-stranded DNA oligo online and have it delivered in a few days. That would have been impossible thirty years ago. Gene editing, gene therapy, mRNA vaccines, single-cell sequencing, spatial transcriptomics. All of it rests on the chemistry of nucleotides. The limitation nobody talks about is scaling. Synthesizing long nucleotide sequences remains expensive and error-prone. Current technology reliably produces oligos up to about three hundred nucleotides. Beyond that, yield drops and errors accumulate. Full-length gene synthesis requires assembly of overlapping fragments, which introduces its own failure points. For therapeutic applications, this is a real bottleneck. We can design a gene, but building it with perfect accuracy at scale is still costly and time-consuming. Enzymatic DNA synthesis is promising, but it is not yet a drop-in replacement for chemical methods. If you are planning a project that depends on custom nucleotide reagents, budget for synthesis failures and plan alternative cloning strategies.
Another practical reality is that nucleotide chemistry does not play well with every environment. Polymerase chain reactions fail in the presence of hemoglobin, heparin, or high concentrations of phenol. Forensic samples, bloodstains, and formalin-fixed tissue often contain inhibitors that bind nucleotides or block polymerase activity. Pretreatment with purification columns or bovine serum albumin in the reaction mix usually resolves this, but it adds steps and variables. There is no single fix that works for every sample type. You have to test. The literature is full of protocols that assume clean DNA. Real samples are rarely clean. What I would tell someone starting out is to respect the molecule and stop treating it like an inert reagent. Nucleotides are reactive. They degrade. They chelate. They interact with buffers, metals, and enzymes in ways that are not always obvious from a protocol. I keep a notebook of failed reactions. Most of the failures trace back to something trivial: an old stock, a warm sample, a wrong pH, a missing cofactor. The ones that do not are the ones that lead to something worth writing about. Nucleotides power the body through continuous recycling and precise regulation. They drive science because they are the medium through which information is stored, copied, and manipulated. Understanding them is less about memorizing structures and more about learning how they behave when things go wrong.