Why Boron Always Gives You Trouble
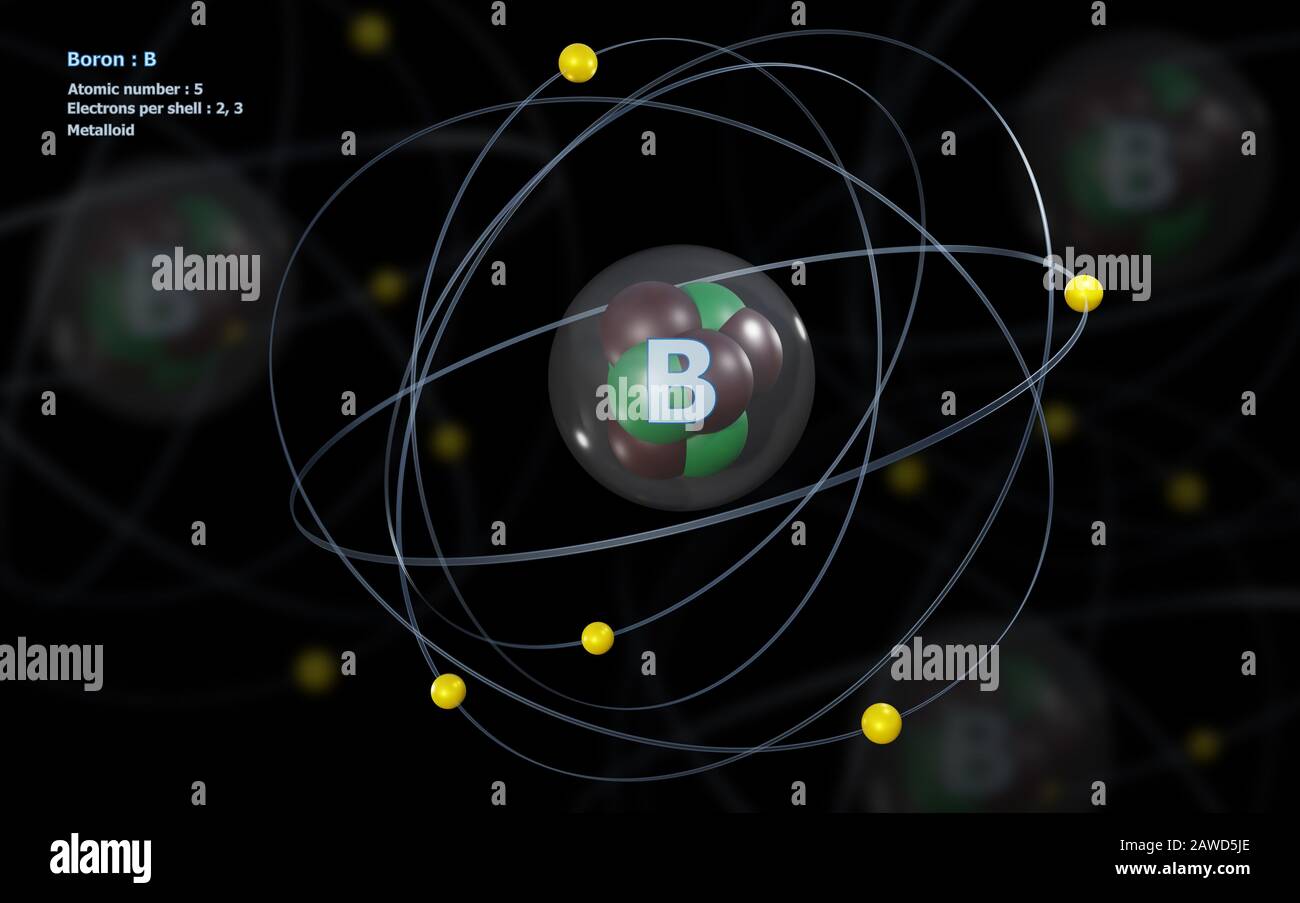
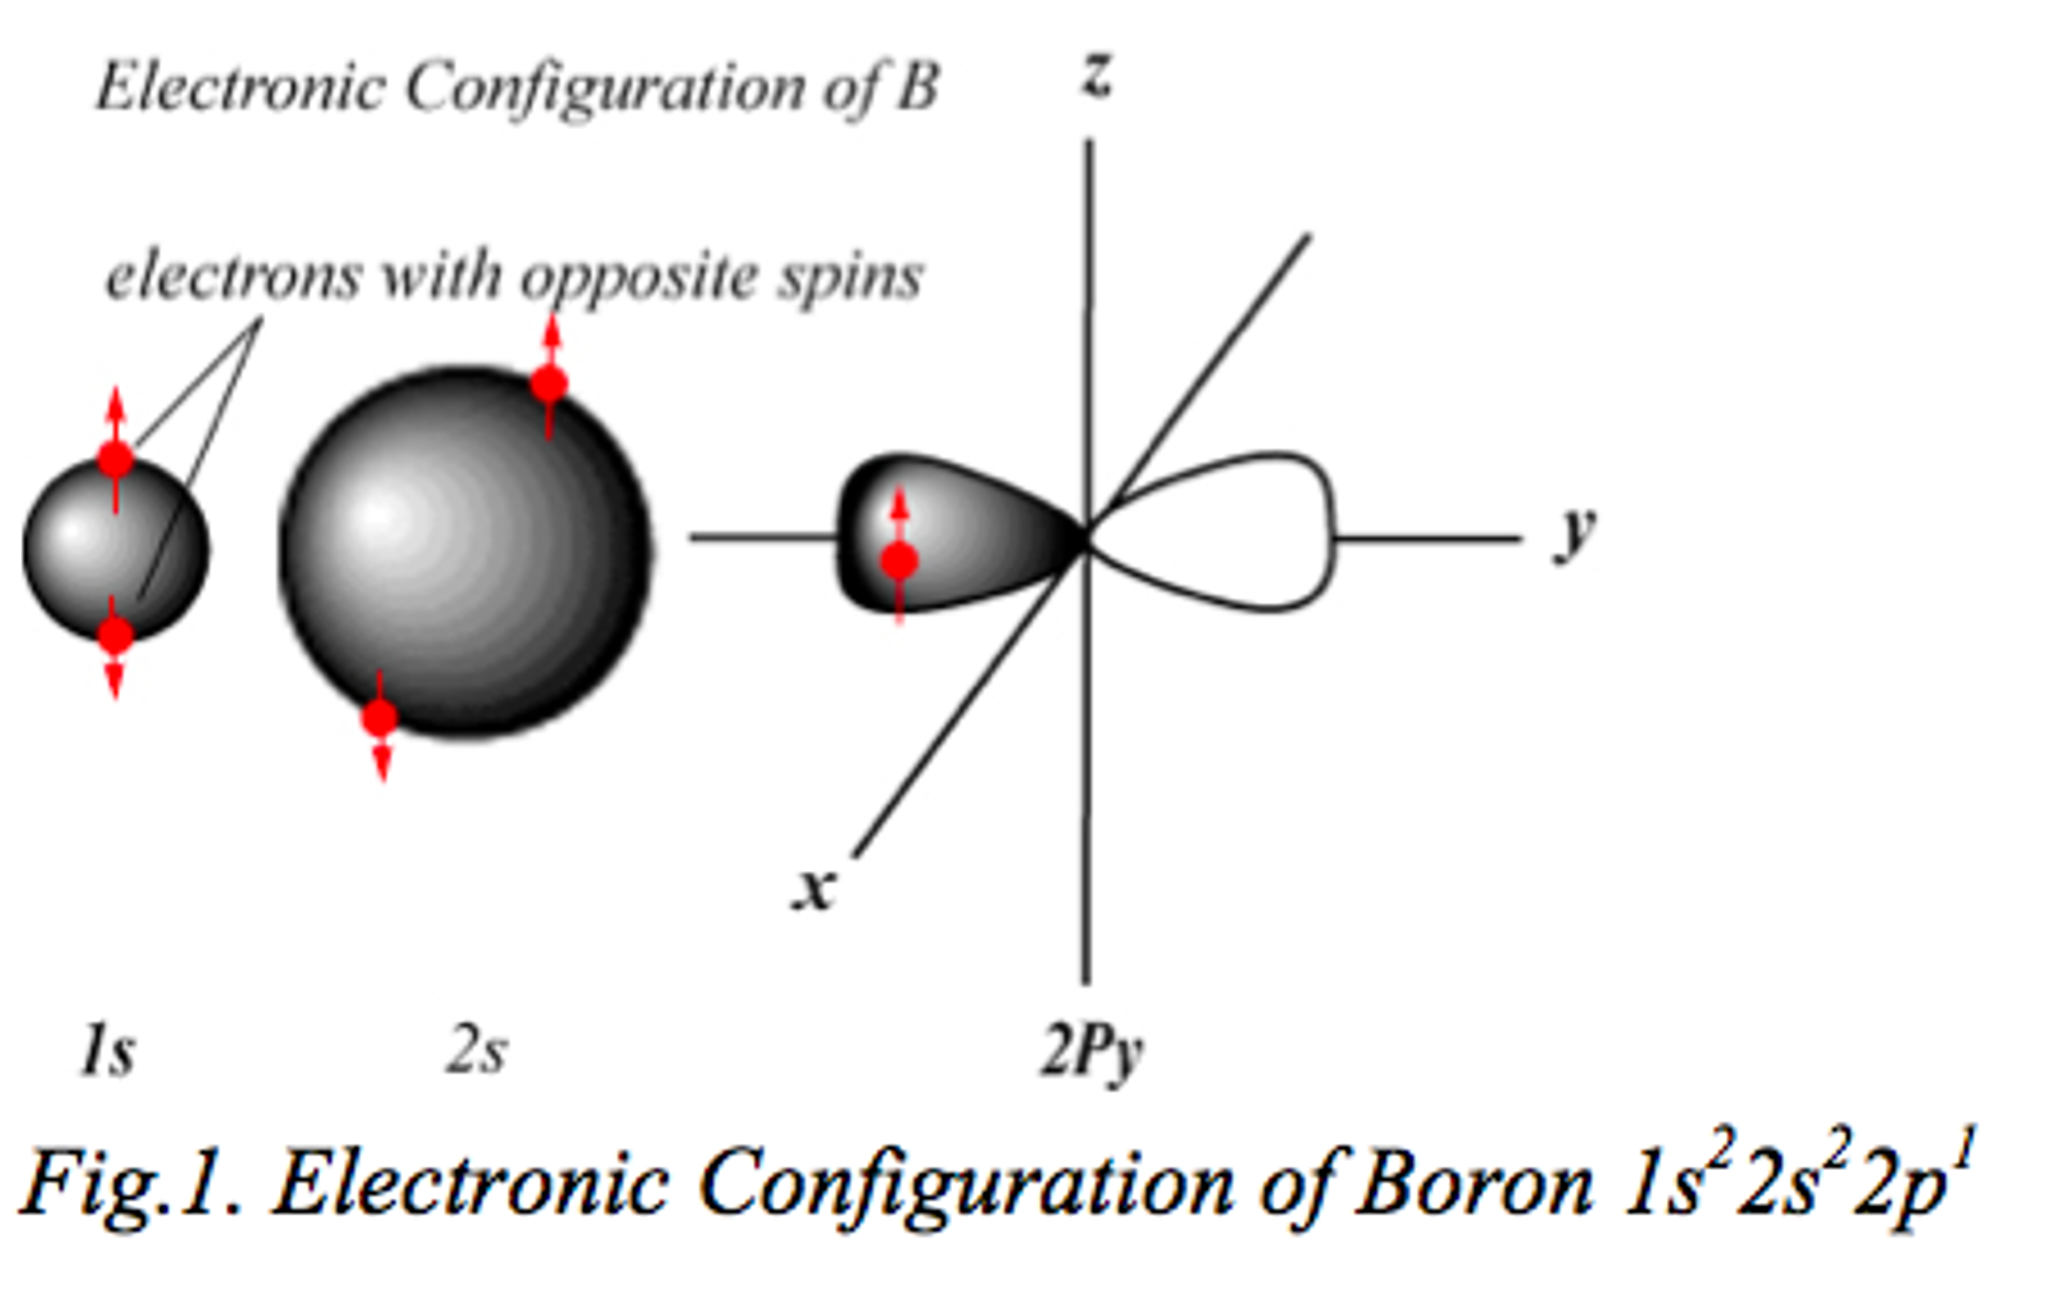
If you have ever run a DFT calculation on a boron-containing compound and watched it fail to converge, you are not imagining things. Boron is weird. It sits right at the edge of the periodic table between stable octet chemistry and everything that falls apart when you try to force it into standard bonding models. The quantum blueprint of Boron and how its electron structure dictates reality is something most textbooks gloss over because the standard treatment works well enough for general chemistry classes. But once you step outside those boundaries, you start seeing the cracks. Boron has an atomic number of 5. That means five protons and, in its neutral state, five electrons. The ground state configuration is 1s2 2s2 2p1. That 2p1 is the whole story. Everything about boron comes down to the fact that it has exactly one electron in its valence p-orbital and a filled s-orbital sitting right underneath it. It wants three bonds. It always wants three bonds. And it hates having a complete octet, which is why so many boron compounds are electron-deficient Lewis acids.
The Quantum Blueprint Of Boron How Its Electron Structure Dictates Reality
Let me walk through how I actually approach boron systems in practice. I do not treat it like carbon with a missing methyl group. That shortcut works for quick sketches but falls apart quickly when you need real numbers. The first thing I check is whether I am dealing with a trigonal planar sp2 center or something more complex like a borane cluster where the geometry changes completely. The electron configuration tells you what to expect, but the actual molecular environment decides what you get. When boron forms three sigma bonds in a neutral molecule like BF3 or BCl3, it is sp2 hybridized. The empty p-orbital sticks up perpendicular to the plane. This is the part that matters: that empty orbital makes boron extremely electrophilic. Fluorine back-bonds into it, which is why BF3 is less reactive than you might expect from the empty orbital argument alone. Chlorine does not back-bond as effectively. That single difference explains why BCl3 hydrolyzes violently in air while BF3 stays relatively calm until you add water intentionally. This is not theoretical. I have seen people underestimate BCl3 handling because they generalized from BF3 behavior. It does not work that way. The real complications start when you leave behind simple trihalides and move into boranes and organoboron compounds. Diborane B2H6 is the classic example and the one that breaks most students first. You cannot draw a normal Lewis structure for it. The bridging hydrogens create three-center two-electron bonds, which is a bonding model that does not exist for carbon compounds at all. I remember spending an afternoon debugging a Gaussian job on a substituted diborane where the initial geometry guess assumed standard two-center bonds. The optimizer ran for twenty minutes, found nothing meaningful, and then crashed. The fix was building the bridge structure from crystallographic data instead of guessing. About ten minutes total once I stopped trying to outthink the software.
For organoboron chemistry, the Suzuki-Miyaura coupling changed everything, but it also introduced a set of practical problems that are easy to miss. The boronic acid B(OH)2 group is the standard pinacol boronate ester is more stable and more common in cross-coupling work. Here is the counter-intuitive part that nobody warns you about: boronic acids tend to form cyclic boroxines B3O3 six-membered rings in the solid state and sometimes even in solution, depending on concentration and solvent. If you are running a reaction and the analytical data does not match the expected molecular weight, check for boroxine formation. I lost two weeks to this once because the NMR peaks looked reasonable but the elemental analysis was off by exactly the amount you would expect from a cyclic trimer. Converting to the pinacol ester fixed the problem permanently. Let me address something that comes up regularly in computational work. When you model boron compounds with standard functionals, you will often get poor geometries for species where the empty p-orbital plays a role. The problem is that most functionals were parameterized on systems where every atom has a complete octet. Boron breaks that assumption. If you are doing DFT on a boron catalyst or a boron-containing organic intermediate, test at least two different functionals and compare results. I usually run B3LYP and wB97X-D side by side. If they disagree on bond lengths by more than a few hundredths of an angstrom, you need to dig deeper. Sometimes you need to include dispersion corrections explicitly, sometimes you need a higher basis set. The range-separated hybrids tend to perform better for boron systems generally, but there is no universal rule. Another edge case that deserves mention: boron's isotope effects. Natural boron is roughly twenty percent B10 and eighty percent B11. In most synthetic work this is irrelevant. In neutron capture therapy research, it is the entire point. But there is a niche application that comes up in mechanistic studies where the small mass difference between isotopes actually matters. Kinetic isotope effects for boron are tiny compared to hydrogen deuterium systems, but they are measurable with high resolution mass spectrometry. I have used this to track whether a boron center is directly involved in the rate determining step of a transformation. It is not a routine experiment, but when you need it, it works.
Get the Full Details

If you are working with boron in a lab setting, here are the practical realities. Boron trifluoride etherate is a liquid and significantly easier to handle than the anhydrous gas, but it still decomposes many organic materials on contact. Keep it away from moisture and strong bases unless that is what you want. Boron triiodide is even more moisture sensitive and practically unusable without a rigorous dry box setup. For most organic synthesis applications, the trialkoxy boron species or the pinacol ester route is far more forgiving. The tradeoff is that you need an extra activation step to transmetallate in coupling reactions, usually with a base like potassium carbonate or sodium tert-butoxide. The electron deficiency of boron also shows up in ways that are not immediately obvious. Trivalent boron compounds can act as Lewis acid catalysts in a range of transformations, from Diels-Alder reactions to aldol condensations. The activity depends heavily on the substituents attached to the boron. Tris(pentafluorophenyl)borane B(C6F5)3 is one of the strongest Lewis acids that does not contain a transition metal, and it has found uses in frustrated Lewis pair chemistry where it partners with a bulky amine to activate small molecules like hydrogen and CO2. This is an area where the simple electron configuration picture gets complicated quickly because steric effects compete directly with electronic effects. A strong Lewis acid that is too sterically hindered will not bind substrates effectively. For anyone modeling or synthesizing boron materials, particularly for battery or semiconductor applications, the bonding flexibility of boron is both a feature and a liability. Boron doping in graphene creates p-type behavior because boron has one fewer valence electron than carbon. The mechanism is straightforward from the quantum perspective. But controlling the doping level precisely is harder than it sounds because boron tends to cluster rather than distribute uniformly during CVD growth. I have seen papers claim uniform substitutional doping where TEM-EDS mapping showed clear boron-rich regions. X-ray photoelectron spectroscopy can help distinguish substitutional boron from interstitial or clustered boron, and it is worth running even if the paper you are following did not include those data.
There is no clean summary here because boron simply does not behave in a clean way. It occupies a position in the periodic table where it borrows from both metals and nonmetals without fully committing to either. The single 2p electron is the starting point, and everything else follows from that. If you respect the electron deficiency and stop trying to force boron into carbon chemistry templates, most of the problems become manageable. If you do not, you will spend a lot of time chasing convergence failures and unexplained side products. Both approaches teach you something, but the first one teaches you faster.