Understanding Proton Transfer as the Core Mechanism
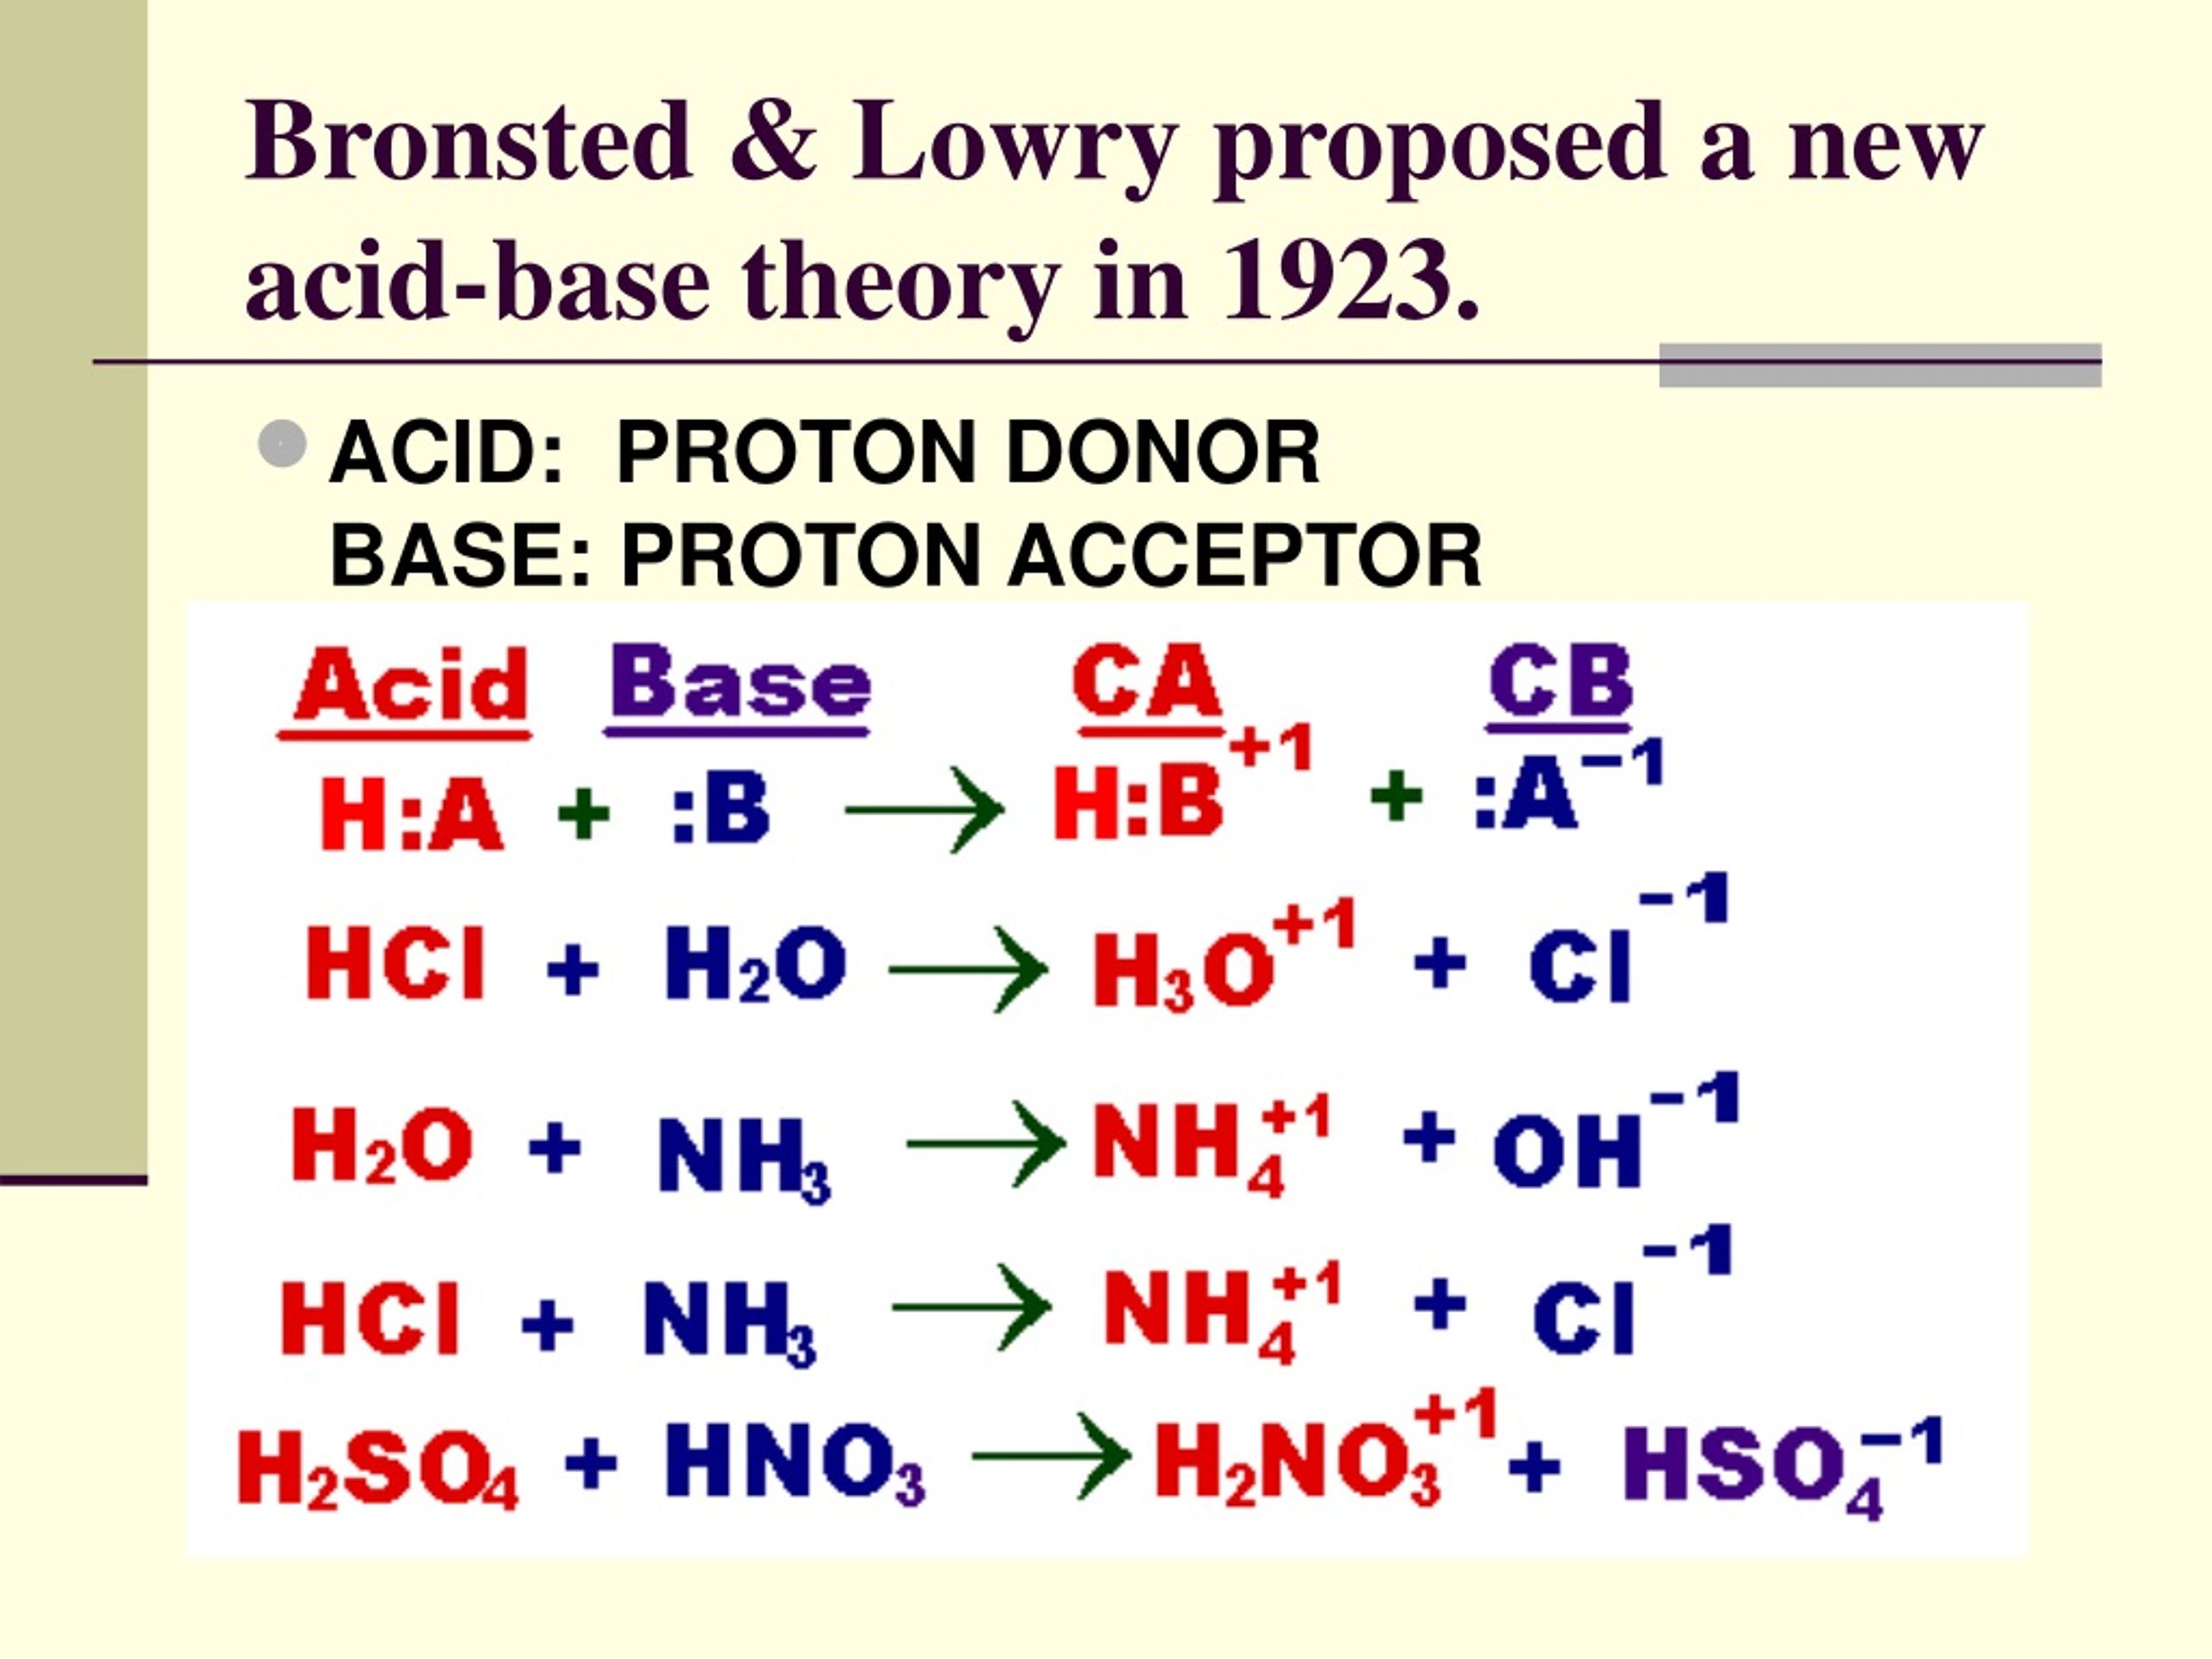
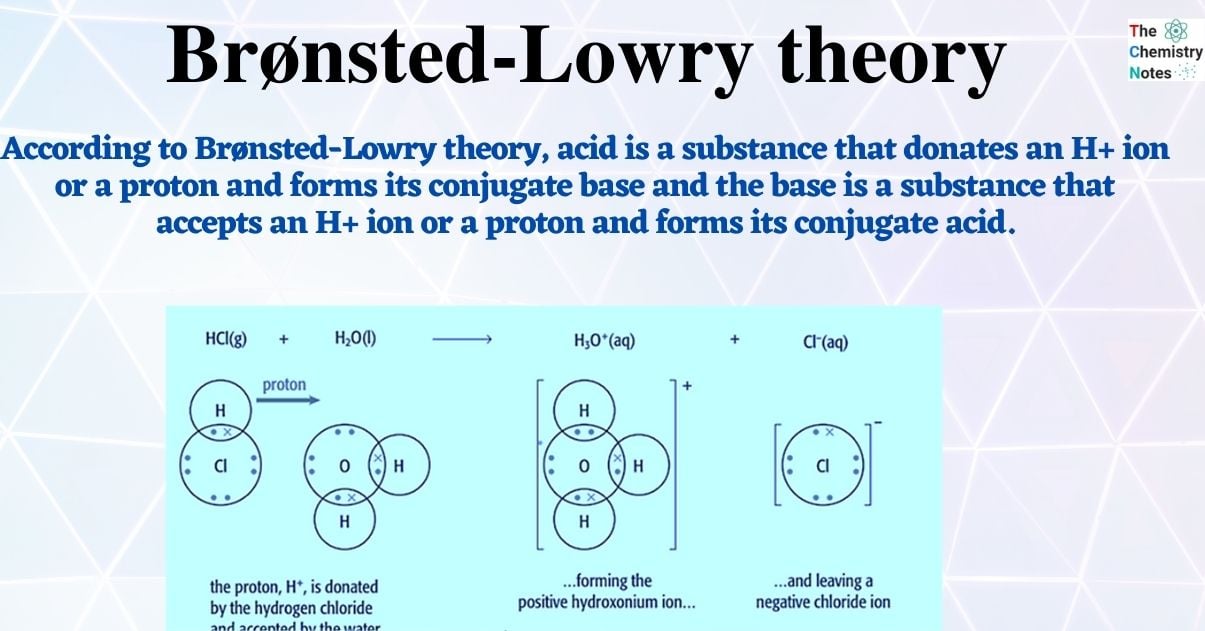
The Brønsted-Lowry model frames every acid-base reaction around one thing: a proton moving from one species to another. An acid donates a proton. A base accepts it. That's the entire definition. But the interesting part, and where most people trip up, is figuring out which direction the proton actually goes and how far it goes. I remember running into trouble once while trying to predict the outcome of a reaction between diisopropylamine and a carboxylic acid in dry DMSO. The pKa values on paper suggested complete deprotonation. What actually came out of the flask was a messy equilibrium mixture with significant amounts of starting material still present. The problem wasn't the theory. It was that pKa tables are built in water, and DMSO changes everything about how protons behave. I ended up using conductivity measurements and NMR integration to map the actual distribution instead of trusting the textbook numbers. If you're working in non-aqueous solvents, always verify with an experimental check rather than assuming the pKa values transfer directly. The practical way to approach these reactions is to look at the pKa of the acid on the left side and compare it to the pKa of the conjugate acid on the right side. The proton prefers to end up on the weaker acid, which means the side with the higher pKa value will be favored at equilibrium. A difference of about 3 pKa units typically gives you roughly 99% conversion in one direction under standard conditions. A difference of less than 1 unit means you're sitting at an equilibrium where both sides are significantly populated and your reaction yield is going to be mediocre unless you do something about it.
I've seen people treat pKa as if it were a fixed property of a molecule. It's not. It depends heavily on the solvent, temperature, and ionic strength of the solution. The pKa of acetic acid in water is 4.76. In DMSO it's 12.6. That's an eight-unit shift. If you're planning a synthesis and the solvent choice changes the pKa by eight units, your entire equilibrium picture flips upside down. This is why people occasionally get surprised when a reaction they expected to go to completion stalls out at around 50% conversion.
Working Through the Mechanism Step by Step
Take a simple reaction between hydrochloric acid and ammonia. HCl donates a proton to ammonia, forming chloride and ammonium. The pKa of HCl in water is roughly negative eight. The pKa of ammonium is about 9.25. That's a difference of about 17 pKa units. The equilibrium lies so far toward the products that for practical purposes the reaction goes to completion. You can write the equilibrium constant from that difference, which comes out to around 10 to the 17th power. That's not a reaction that has any meaningful reverse rate under normal conditions. Now consider something less decisive. Let's say you're mixing phenol, which has a pKa around 10, with hydroxide ion. The conjugate acid of hydroxide is water, with a pKa of 15.7. The difference is about 5.7 units, favoring the products. Hydroxide will deprotonate phenol, but not as overwhelmingly as it would deprotonate HCl. You still get mostly phenoxide and water, but a tiny fraction of phenol remains. In most lab work that doesn't matter. In a process where you need quantitative deprotonation, you'd push it with excess base or remove water to shift the equilibrium further. One thing beginners consistently miss is that the conjugate base of a weak acid is still a base, just a weaker one. The conjugate base of a strong acid is essentially inert in water. Chloride ion from HCl doesn't grab protons back from water because HCl is so much stronger. But acetate ion from acetic acid is a legitimate base that will accept protons from water to form hydroxide and acetic acid. That's why sodium acetate solutions are basic. The reverse reaction isn't negligible, it's just smaller than the forward reaction when you start with the strong acid.
Get the Full Details

Limitations and When the Model Falls Apart
The Brønsted-Lowry framework works well for proton transfer in protic solvents, but it has real blind spots. It doesn't handle reactions where no proton is involved at all, like the interaction between boron trifluoride and ammonia. That's a Lewis acid-base reaction, and trying to force it into the Brønsted-Lowry mold just creates confusion. Ammonia donates a lone pair to boron. No proton moves. Calling it an acid-base reaction under Brønsted-Lowry terms requires inventing a convoluted pathway that doesn't exist. The model also struggles with extremely concentrated solutions and highly acidic or basic media. In superacidic conditions, the concept of a discrete proton donor and acceptor blurs because the solvent itself participates actively in proton transfer networks. Water acts as both acid and base simultaneously, and you get autoprotolysis effects that make simple pKa comparisons unreliable. I've seen students try to use standard pKa tables to predict behavior in 98% sulfuric acid and get completely wrong answers because the reference state for pKa is dilute aqueous solution. Another common pitfall is assuming that a higher pKa always means a stronger base. That's true within a given class of compounds in the same solvent, but comparing across different classes can be misleading. For instance, tert-butoxide and cyanide have pKa values for their conjugate acids that might suggest one is significantly stronger, but in practice steric hindrance and solvation effects can invert that expectation. tert-Butoxide is a strong base but a poor nucleophile due to its bulk, while cyanide is a decent nucleophile despite being a moderate base. The pKa tells you about thermodynamics, not kinetics or steric access.
If you're dealing with reactions in non-polar solvents or with species that don't dissolve well in water, the Brønsted-Lowry pKa approach becomes unreliable. In those cases, you're better off using computational methods to estimate proton affinity or gas-phase basicity, or turning to experimental titration in the actual solvent system you're working with. There's no shortcut around that.
Practical Tips for Predicting Reaction Outcomes
When you need to predict whether a given base will deprotonate a given acid, start by identifying the conjugate acid of the base and looking up its pKa. Then look up the pKa of the acid you're trying to deprotonate. If the conjugate acid has a higher pKa, the base is strong enough. The larger the difference, the more complete the reaction. This is usually fast enough that you don't need to worry about kinetics for simple proton transfers. Most acid-base reactions in solution reach equilibrium in milliseconds. Keep in mind that solid bases behave differently from dissolved ones. Sodium hydride is a strong base, but if it's sitting as a solid and the proton source is in solution, the reaction happens only at the surface. Stirring rate, particle size, and surface area matter a lot more than the pKa values would suggest. I once ran a deprotonation with NaH that looked like it wasn't proceeding because the bubbles of hydrogen gas stopped evolving. Turns out the NaH had formed a passivated crust on the surface. Freshly prepared NaH under an inert atmosphere worked immediately. The theory said it should work either way, but the practical reality was different. For reactions where you need to drive an equilibrium toward products, removing one of the products often helps. If you're generating water as a byproduct and the reverse reaction is significant, using a drying agent or azeotropic removal can shift things in your favor. Molecular sieves, magnesium sulfate, or a Dean-Stark trap depending on the solvent system can make the difference between a 70% yield and a 90% yield on a deprotonation step. It's not glamorous, but it's the kind of detail that shows up in process chemistry where people actually need reliable outcomes rather than textbook predictions.