Understanding Lysine Side Chains in Protein Work
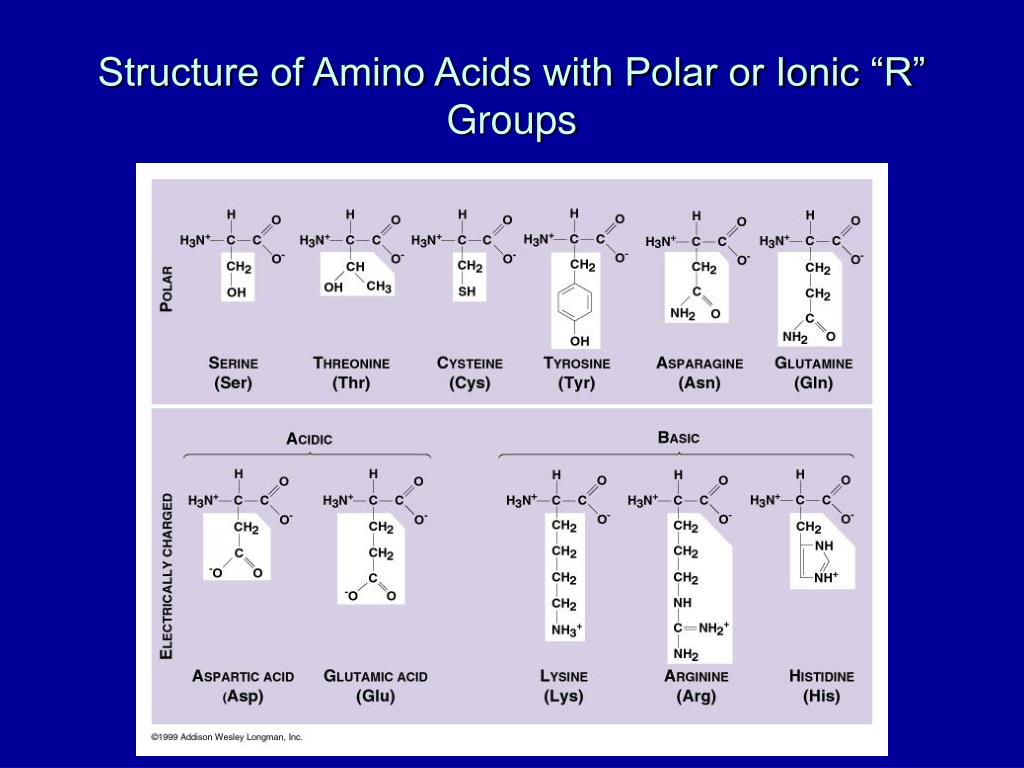
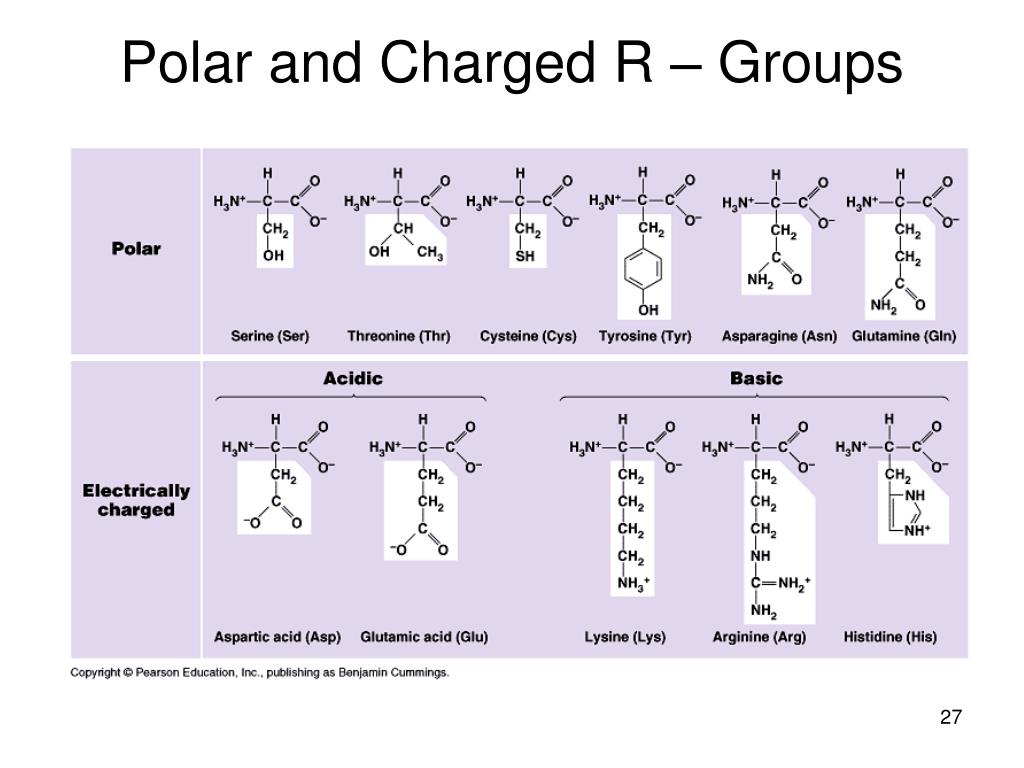
Lysine is one of the standard amino acids, and its side chain carries a primary amine group that gets protonated at physiological pH. That means it holds a positive charge most of the time inside a cell. This sounds simple, but the implications for how you design, model, or modify proteins are actually pretty deep. The pKa of the lysine epsilon-amino group is around 10.5, so even at pH 7.4 you are dealing with a fully charged residue. That charge drives salt bridges, influences local folding, and shows up in pretty much every interaction surface you will ever look at. I remember working on a project where we needed to introduce a post-translational modification simulation involving lysine acetylation. The standard molecular dynamics setup I used would have shown the lysine as permanently positively charged, which is wrong once that acetyl group is added. The side chain goes from charged to neutral, and that shift changes everything about how the protein behaves in solution. I had to manually reparameterize the force field terms for that residue, adjusting the partial charges and the torsion potentials. A lot of people skip this step and just run the simulation with the standard parameters, which gives you garbage results for any system where acetylation matters. Here is something most textbooks do not emphasize enough. The positive charge on lysine is not always available for interactions the way you would expect. In many protein structures, the side chain points inward and participates in a salt bridge with an aspartate or glutamate, and that interaction can be remarkably stable. The charge is effectively neutralized within the protein core, even though the pKa says it should be protonated. When you are analyzing a crystal structure and counting charged residues on a surface, you need to look at the actual geometry, not just assume every lysine is sticking out and available. I have seen people build binding models based on the assumption that every surface lysine contributes a +1 charge, when in reality half of them are buried in electrostatic partnerships that cancel each other out.
Another thing that comes up a lot in practice involves protein-protein docking and charge complementarity. Lysine rich patches on one protein surface will attract negatively charged regions on a partner, but the effect is highly dependent on ionic strength. If you are running docking simulations at a physiological salt concentration, the electrostatic steering is dampened significantly compared to low-salt conditions. I usually recommend checking what ionic strength your simulation or experiment actually represents before you make big claims about lysine-mediated binding. The charge is there, but it may not be doing as much work as you think at higher salt concentrations. The practical side of dealing with lysine also includes its role in cross-linking chemistry. If you are working with NHS-ester based conjugation or other carbodiimide chemistries, lysine is your primary target because of that free primary amine. The reaction works well at pH 7 to 9, which is convenient because the lysine is protonated and reactive at those conditions. But here is the catch: if a lysine is involved in a salt bridge or sits in a hydrophobic pocket, the pKa can shift downward significantly, sometimes into the range where it is partially deprotonated even at neutral pH. That makes it more reactive toward acylation, but it also means you cannot predict reactivity just from sequence. I once spent three weeks trying to get a consistent conjugation yield on a protein that had several buried lysines, and the only way forward was to map the structure first and avoid those residues by using site-specific labeling strategies instead of relying on random lysine modification. When you are modeling lysine in homology structures or AlphaFold predictions, keep in mind that the side chain conformation is not always well resolved, especially in lower-confidence regions. The chi angles for the epsilon amine can vary widely, and a poorly oriented lysine side chain in a predicted structure can mislead you about interaction potential. I always check the pLDDT scores and look at the rotamer quality before building any mutation or modification model that involves lysine positioning. A side chain pointing the wrong way in a prediction is not worth building experiments around without experimental validation.
There is also the matter of lysine methylation, which is a whole different ballgame from acetylation. Methylation does not remove the charge, which is a key distinction. Mono-, di-, and trimethylated lysine all remain positively charged because the nitrogen still has a proton attached. This means that in chromatin studies, people sometimes incorrectly assume that methylation neutralizes a charge the way acetylation does. It does not. The functional effect of methylation is entirely steric and hydrogen bonding related, not electrostatic. Mixing these up leads to incorrect mechanistic interpretations, and I see it come up in literature discussions regularly. If you are building a custom simulation system and need parameter files for modified lysine residues, the CHARMM and AMBER force fields both have documentation for acetyllysine and methylated variants. The parameters exist, but you need to be careful about which version you use because they have been updated across force field revisions. A parameter set from CHARMM36m will behave differently than one from earlier versions, and mixing parameter sets across residue types is a common source of artifacts. I always verify that every modified residue in my system comes from the same force field release before running production simulations. The takeaway here is not that lysine is complicated. It is straightforward. The complication comes from how the charge manifests differently depending on environment, modification state, and structural context. If you treat every lysine the same way in your models and experiments, you will miss the things that actually matter. Map the structure first. Check the environment around each residue. Know whether it is modified or capable of being modified under your conditions. Then decide whether the charge matters for what you are trying to do.
Get the Full Details