Why Your Organic Chemistry Notes Are Lying to You About Molecular Bonding
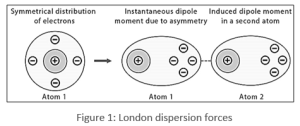
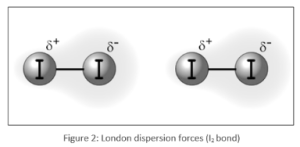
I spent three years trying to understand why nonpolar molecules stick together before I stopped treating it like a minor footnote. Most textbooks spend maybe two paragraphs on London dispersion forces, then rush into hydrogen bonding like it is the only interaction worth caring about. That approach leaves you completely unprepared for anything beyond simple diatomic molecules. The reality is that dispersion forces dominate the behavior of most organic compounds you will encounter in a lab setting, and they are the reason your distillation columns, chromatography separations, and crystallization attempts behave the way they do. The mechanism itself is straightforward once you stop treating it as some quantum mechanical mystery. Electrons are constantly moving around atomic nuclei, and at any given instant, their distribution is not perfectly symmetrical. This creates a temporary dipole, which then induces a complementary dipole in a neighboring molecule. The attraction between these fluctuating dipoles is what we call London dispersion. It works at every scale, from helium atoms barely holding onto each other at near absolute zero to massive hydrocarbon chains forming liquids with high boiling points. The strength scales with polarizability, which roughly correlates with the number of electrons and the surface area available for contact between molecules.
The Mysterious Forces Holding Molecules Together Uncovering The Secrets Of London Dispersion Forces
Here is where people who only read the textbook version get tripped up. They learn the formula E equals negative two-thirds times alpha squared times I divided by four pi epsilon naught times r to the sixth, and they think they understand dispersion. The formula tells you the energy decays with the sixth power of distance, which is correct but dangerously incomplete. It assumes isolated pairwise interactions, which breaks down almost immediately in condensed phases. In a real liquid or solid, you are dealing with many-body effects where the polarizability of one molecule is influenced by the presence of every other molecule nearby. Many computational chemistry packages handle this reasonably well with methods like DFT-D3 or DFT-D4, but if you are working without those corrections, your calculated binding energies for nonpolar systems can be off by factors of two or more. I ran into this problem explicitly when modeling the adsorption of long-chain alkanes on a graphene surface for a project a couple years back. Standard DFT with a common functional like PBE gave results that were qualitatively wrong, predicting adsorption energies that were essentially zero. The molecules weren't sticking to the surface at all in the simulation. Switching to PBE with D3 dispersion correction brought the calculated energies into line with experimental calorimetry data, shifting from near-zero to roughly negative forty kilojoules per mole for a C20 chain. That is not a small difference. It is the difference between predicting a molecule physisorbs weakly and predicting it desorbs entirely at room temperature. Polarizability is not just about electron count. The shape and flexibility of the electron cloud matters enormously. A linear alkane chain is significantly more polarizable than a compact spherical molecule with the same number of electrons because the electrons along the chain can shift more freely over a longer distance. This is why n-octane boils at one hundred twenty-five degrees Celsius while its isomer 2,2,3,3-tetramethylbutane boils at only one six degrees, despite being exact constitutional isomers with identical molecular weights. The surface area available for intermolecular contact changes the dispersion contribution dramatically. Branching reduces contact area, which weakens the cumulative dispersion interaction across the molecule.
This surface area principle explains a lot of practical separation problems that confuse people who rely solely on boiling point tables. In fractional distillation of isomeric mixtures, the difference in volatility between linear and branched isomers is almost entirely a dispersion effect. The linear isomer has more surface contact, stronger dispersion forces, and a higher boiling point. When you are trying to separate a mixture of C8 isomers, you are not fighting differences in dipole moments or hydrogen bonding. You are fighting differences in how efficiently the electron clouds of different molecular shapes can induce complementary dipoles in their neighbors. Another thing that trips people up is the assumption that dispersion forces are weak. Individual pairwise dispersion interactions between small molecules are indeed on the order of a few kilojoules per mole, comparable to thermal energy at room temperature. But these forces are additive, and they add up fast. The total dispersion contribution to the cohesive energy of liquid hexane is roughly forty kilojoules per mole. For larger molecules, the cumulative effect becomes the dominant term in determining boiling point, solubility, viscosity, and surface tension. Polyethylene, which has no permanent dipole and no hydrogen bonding capability, is a solid at room temperature precisely because the dispersion forces across the long polymer chains sum to something substantial. When you are working with solvent selection for extractions or recrystallizations, understanding dispersion helps you predict behavior that simple polarity scales cannot. A solvent like hexane is often dismissed as nonpolar and therefore uninteresting, but its dispersion interactions are the primary reason it selectively dissolves nonpolar impurities while leaving polar compounds behind. The selectivity comes from the mismatch between hexane's dispersion-dominated solvation and the stronger dipole-dipole or hydrogen bonding interactions within the polar solute. This mismatch makes the free energy of dissolution unfavorable for the polar compound, which is the entire basis of selective extraction.
Get the Full Details

There are cases where dispersion forces completely fail to explain observed behavior, and it is important to recognize those limits. When highly polar or charged species are involved, electrostatic interactions will overwhelm dispersion contributions. Salt bridges, ionic bonds, and strong hydrogen bonding networks operate on energy scales of fifty to two hundred kilojoules per mole, dwarfing the dispersion forces present in the same system. In protein folding, for example, the hydrophobic effect is often attributed to dispersion, but it is primarily an entropic phenomenon driven by water restructuring around nonpolar groups. The dispersion forces between the nonpolar side chains contribute, but they are secondary to the solvent entropy term. Treating hydrophobic interactions as purely a dispersion effect is a common oversimplification that leads to incorrect predictions in biochemistry. Computational chemists also hit a wall with dispersion when dealing with exotic systems. Standard semiempirical methods like PM6 or AM1 do not include dispersion corrections at all, and applying them to systems involving heavy elements or transition metals can give misleading results. The dispersion coefficient depends on the ionization potential and polarizability of the specific atoms involved, and for heavier elements, relativistic effects alter both parameters in ways that standard correction schemes do not capture. If you are modeling organometallic complexes or lanthanide compounds, you need specialized dispersion parameters or you should rely on high-level wavefunction methods like coupled cluster with perturbative triple excitations, which are computationally expensive but more reliable. The practical takeaway is that dispersion forces are not a footnote in molecular interactions. They are a fundamental component that dominates the physical properties of the majority of organic molecules, and ignoring them leads to qualitatively wrong predictions in both computational and experimental contexts. The key is recognizing when they matter most, which is in systems lacking strong dipoles or hydrogen bonding capability, and understanding how molecular shape and electron cloud flexibility modulate their strength. Once you internalize that, things like boiling point trends, solubility patterns, and adsorption behaviors become much more predictable.