Why Gas Laws Don't Add Up Without KMT
I spent three semesters watching students plug numbers into PV equals nRT and then get confused when their answers didn't match lab data. The problem was never the algebra. It was that nobody bothered to explain what the equation actually represents before asking them to calculate with it. The kinetic molecular theory is the framework that connects macroscopic observations to particle behavior. Without it, thermodynamics is just a collection of formulas. With it, you can predict what happens when conditions change.
The Kinetic Molecular Theory Unlocking The Secrets Of Matter And Energy
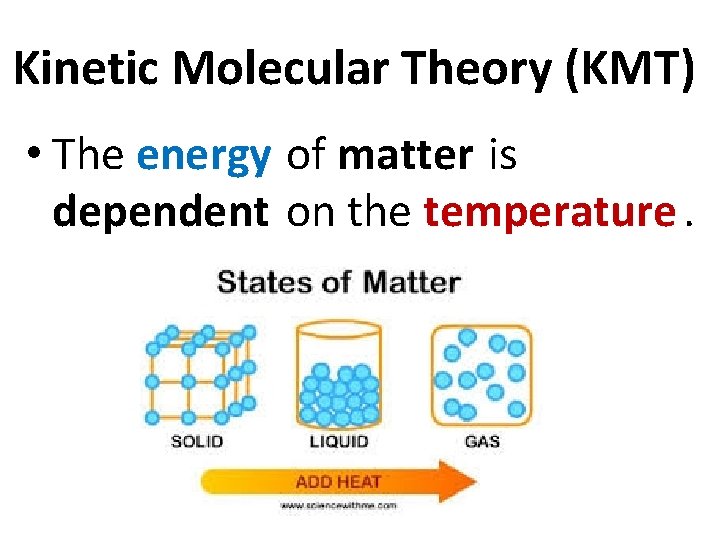
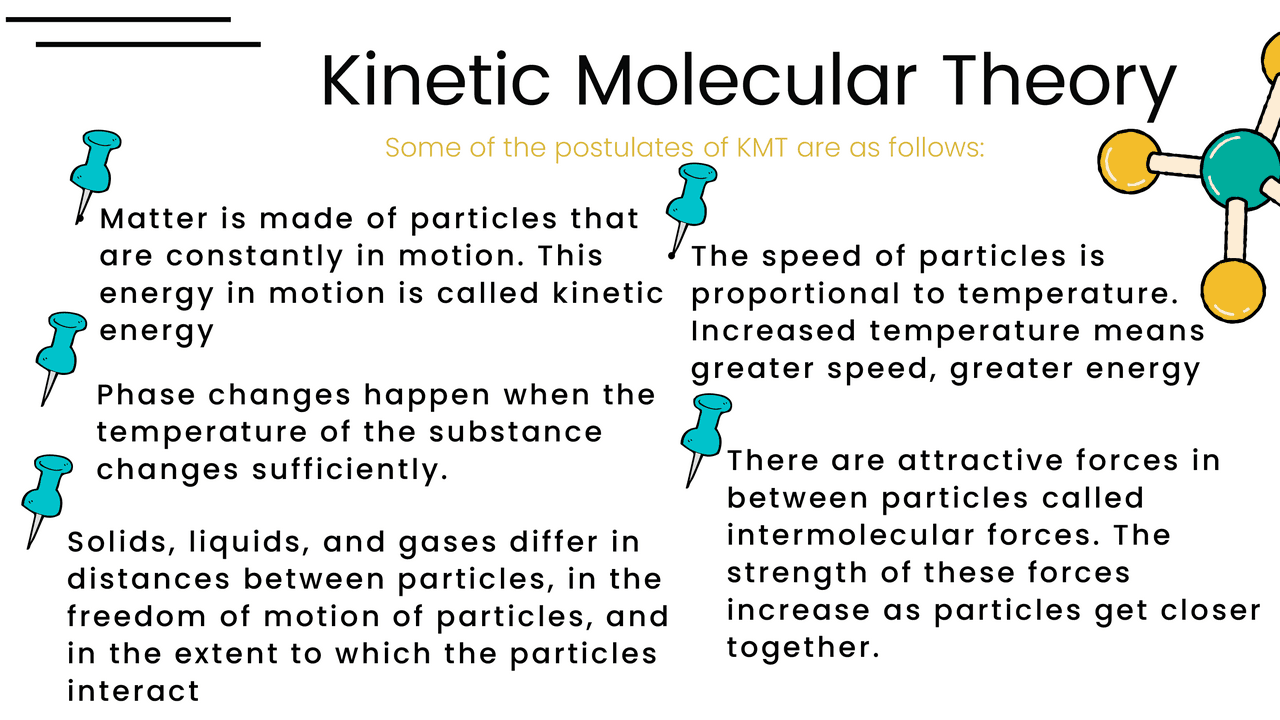
At its core, KMT makes five assumptions about ideal gas particles. They move in random straight lines. Collisions are perfectly elastic. The volume of individual particles is negligible compared to container volume. There are no intermolecular forces between particles. Average kinetic energy depends only on temperature. Here is the thing most textbooks skip: these assumptions hold reasonably well for gases at standard temperature and pressure. They break down aggressively under high pressure or low temperature. I learned this the hard way during a lab run where we were measuring the compressibility factor of nitrogen at 200 atmospheres. The ideal gas law was off by roughly eighteen percent. That is not a measurement error. That is the theory hitting its limits. When deviations appear, you switch to the van der Waals equation or the Redlich-Kwong equation of state. The correction terms account for particle volume and intermolecular attraction. It adds two constants to your calculation, but it drops the error from eighteen percent down to under two percent. Worth the extra five minutes of work.
Another practical detail: root mean square velocity. Students often confuse it with average velocity. The RMS speed formula is the square root of three RT divided by M, where M is molar mass in kilograms per mole. If you use grams per mole instead, your answer is off by a factor of ten. I catch this constantly in grad student reports. The unit conversion error is the single most common mistake I see in computational chemistry work involving gas phase kinetics. Understanding how energy distributes among particles matters more than memorizing the Maxwell-Boltzmann curve shape. The distribution skews right. Most particles cluster near the peak, but a significant tail extends toward higher energies. That tail is why reactions happen at temperatures well below the activation energy you would calculate from average kinetic energy alone. A substantial fraction of molecules always possess enough energy to overcome the barrier. This is not theory. This is what makes Arrhenius behavior work in practice. One nuance people miss: translational kinetic energy equals three-halves RT per mole regardless of molecular complexity. Rotational and vibrational modes contribute to internal energy but not to temperature in the simple sense. When you measure temperature with a thermocouple, you are sensing translational energy transfer through collisions. Rotational energy does not directly register. This distinction matters when you model polyatomic gases in simulation software. The heat capacity ratios change because those extra degrees of freedom store energy without raising temperature proportionally.
Get the Full Details
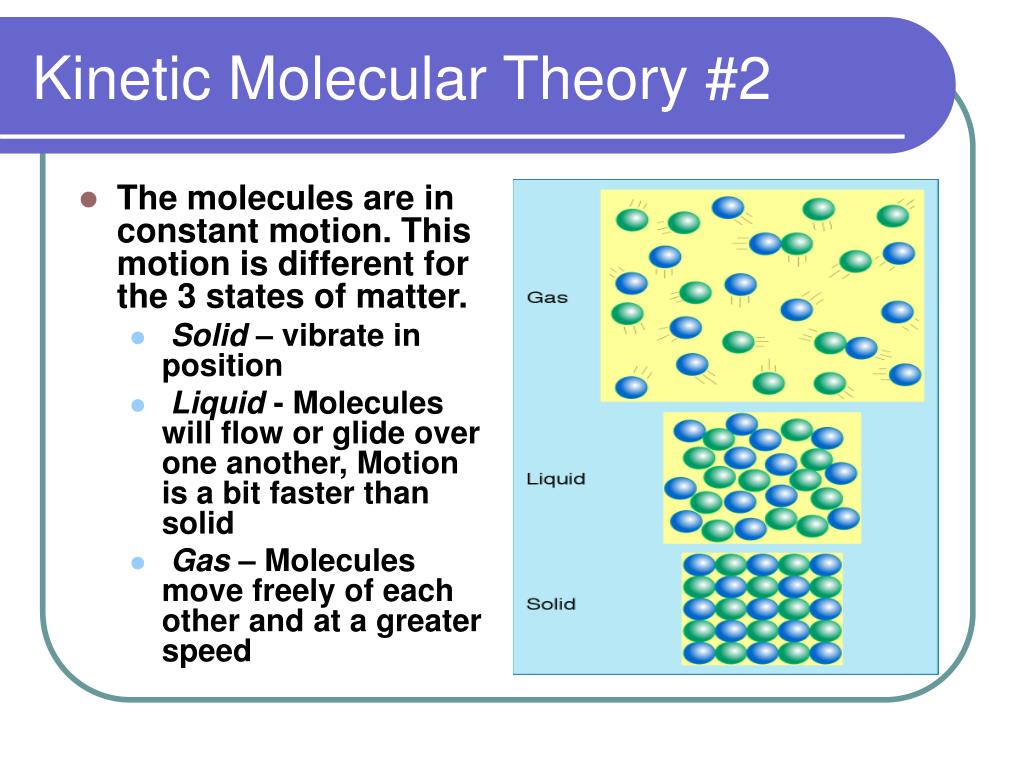
If you are working with real systems, remember that KMT assumes equilibrium. Flash processes, shock waves, and rapid expansions create non-equilibrium states where the theory does not apply cleanly. I worked on a project involving supersonic nozzle flow where local temperature was undefined for several microseconds after expansion. The gas had not redistributed energy among modes yet. Any calculation assuming equilibrium thermodynamics in that window produced garbage results. You have to use collision rate estimates to determine when equilibrium is reestablished, which typically takes on the order of ten to a hundred mean free paths downstream. The takeaway is straightforward. Learn the assumptions. Know when they fail. Use the right equation of state for the regime you are in. The theory is useful because it gives you intuition about particle behavior, not because it predicts real gas behavior with perfect accuracy across all conditions.