Understanding Molecular Kinetic Energy Beyond the Textbook
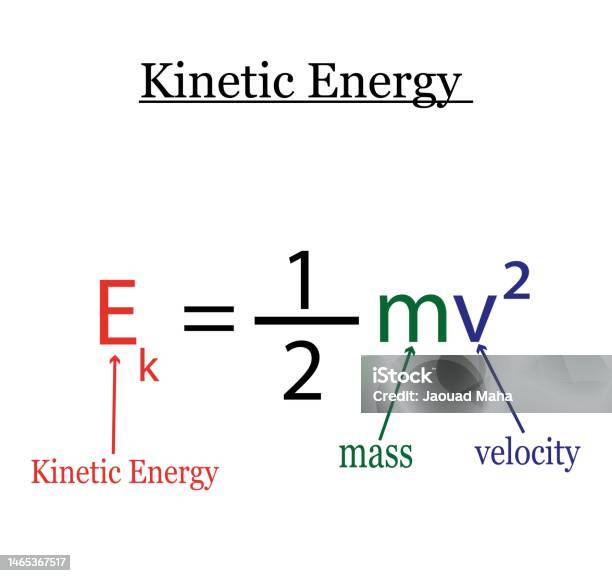
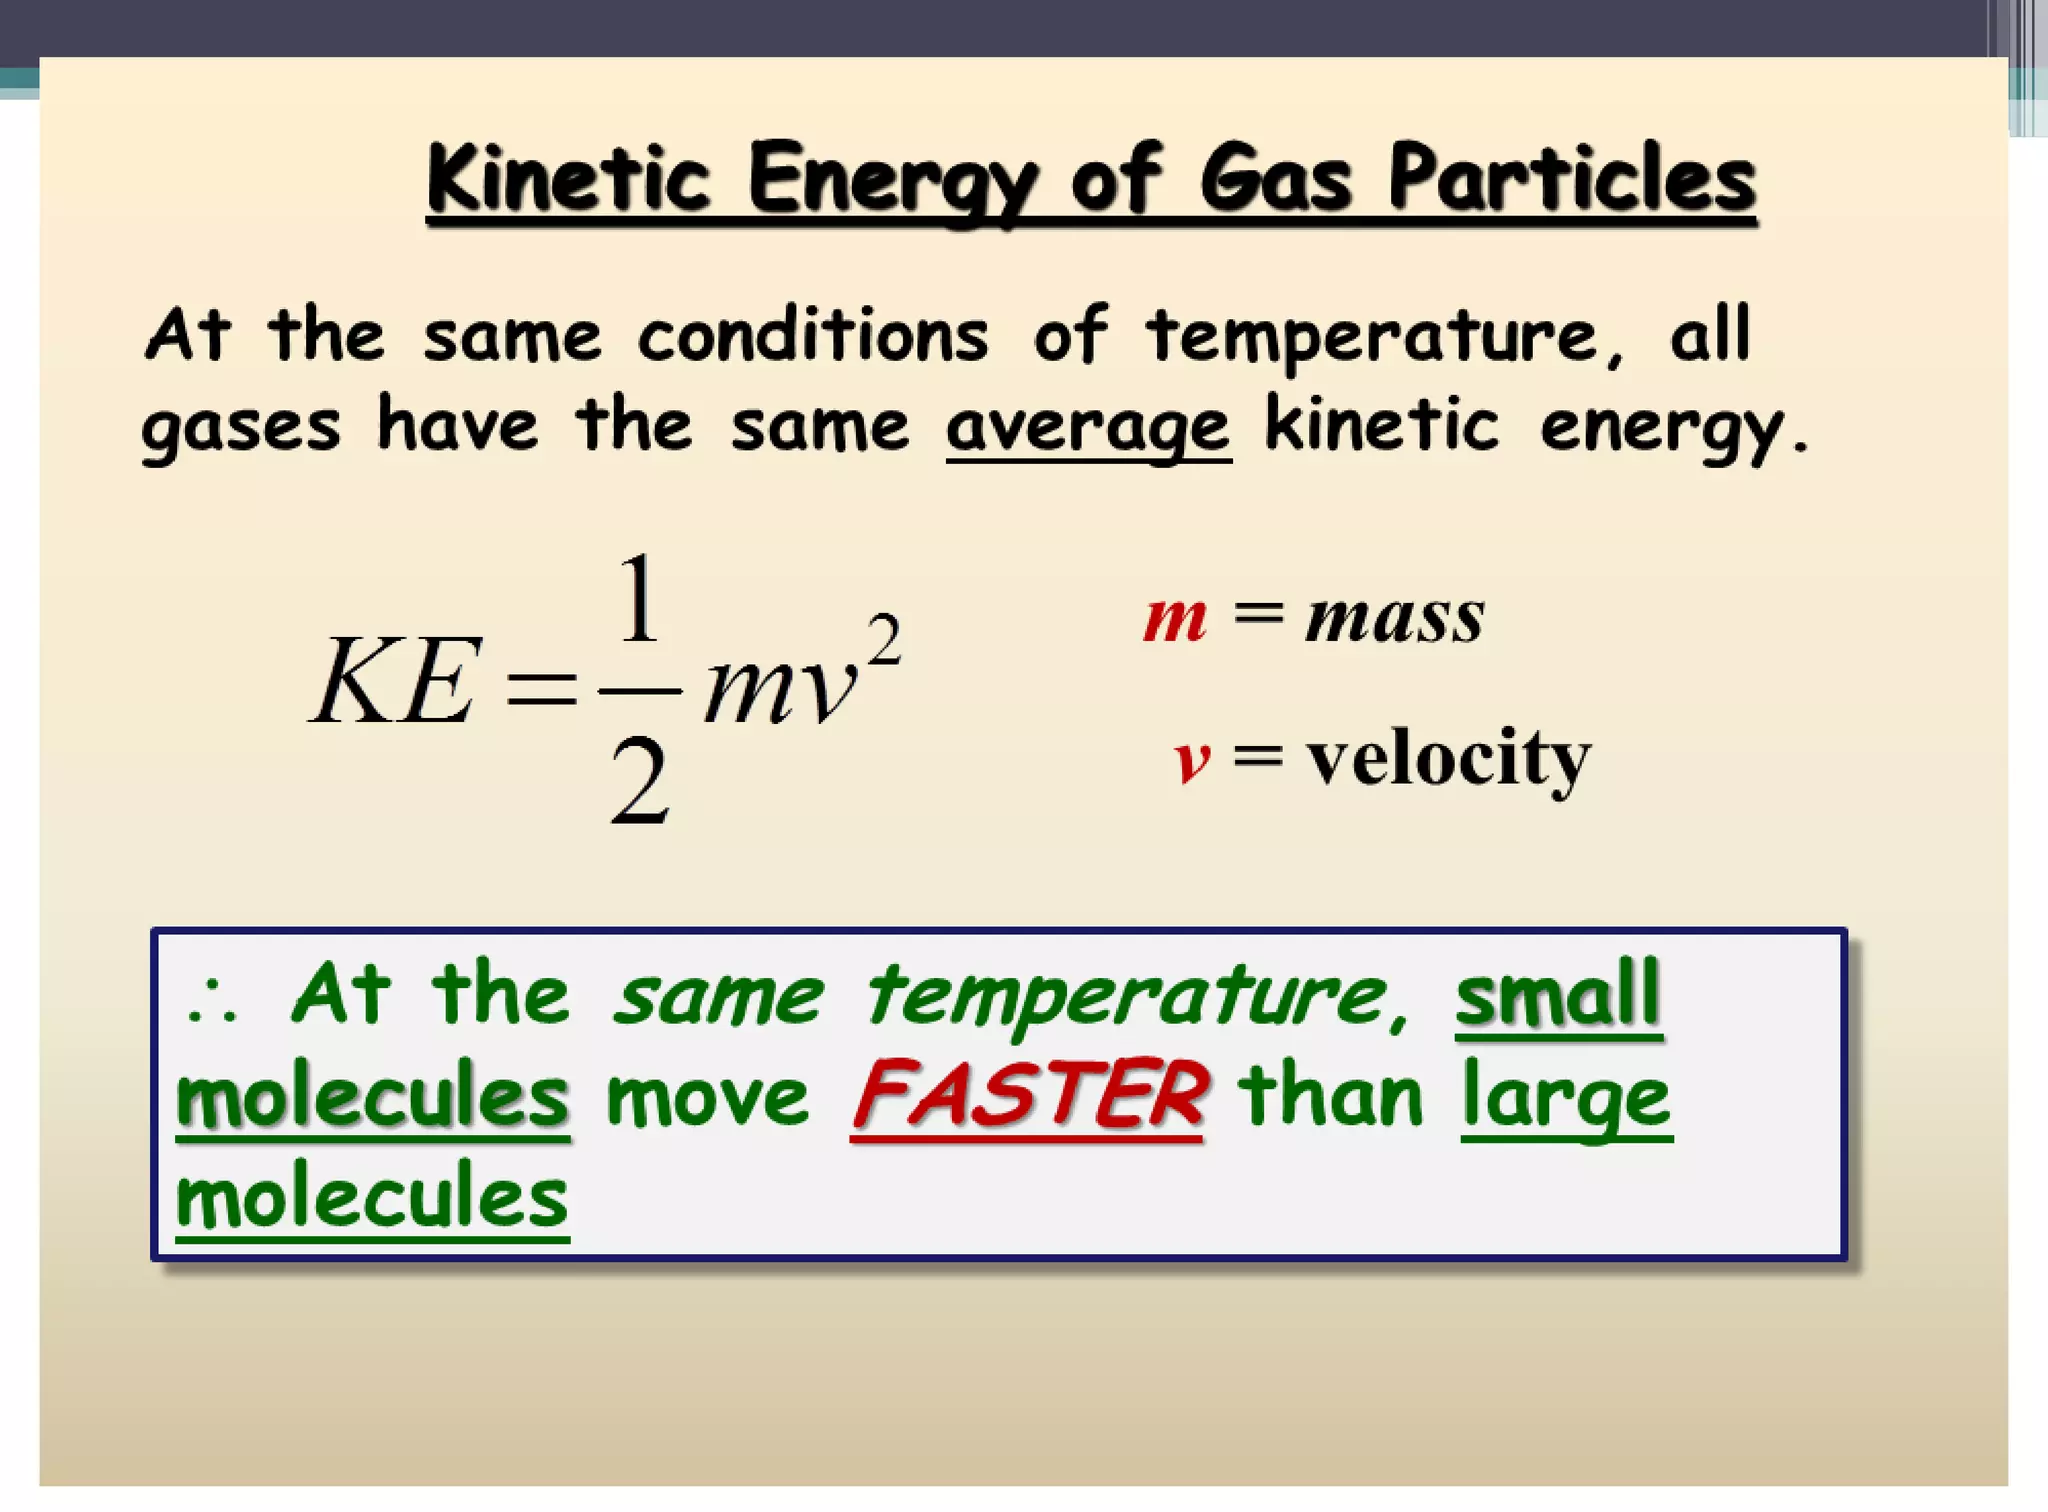
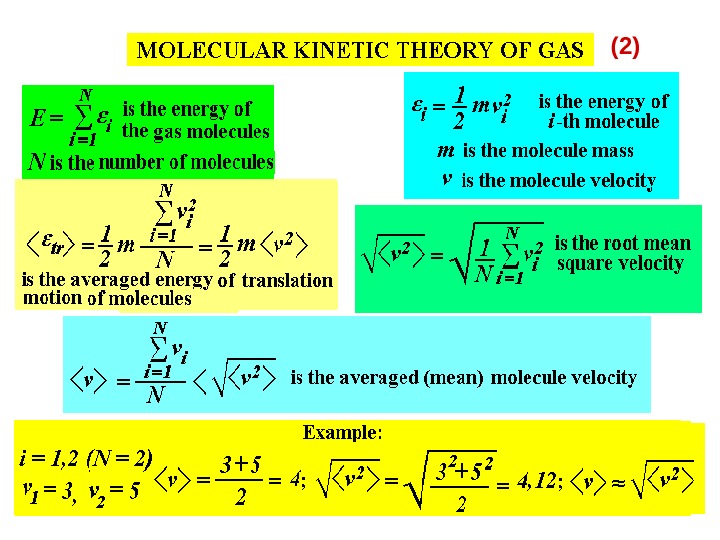
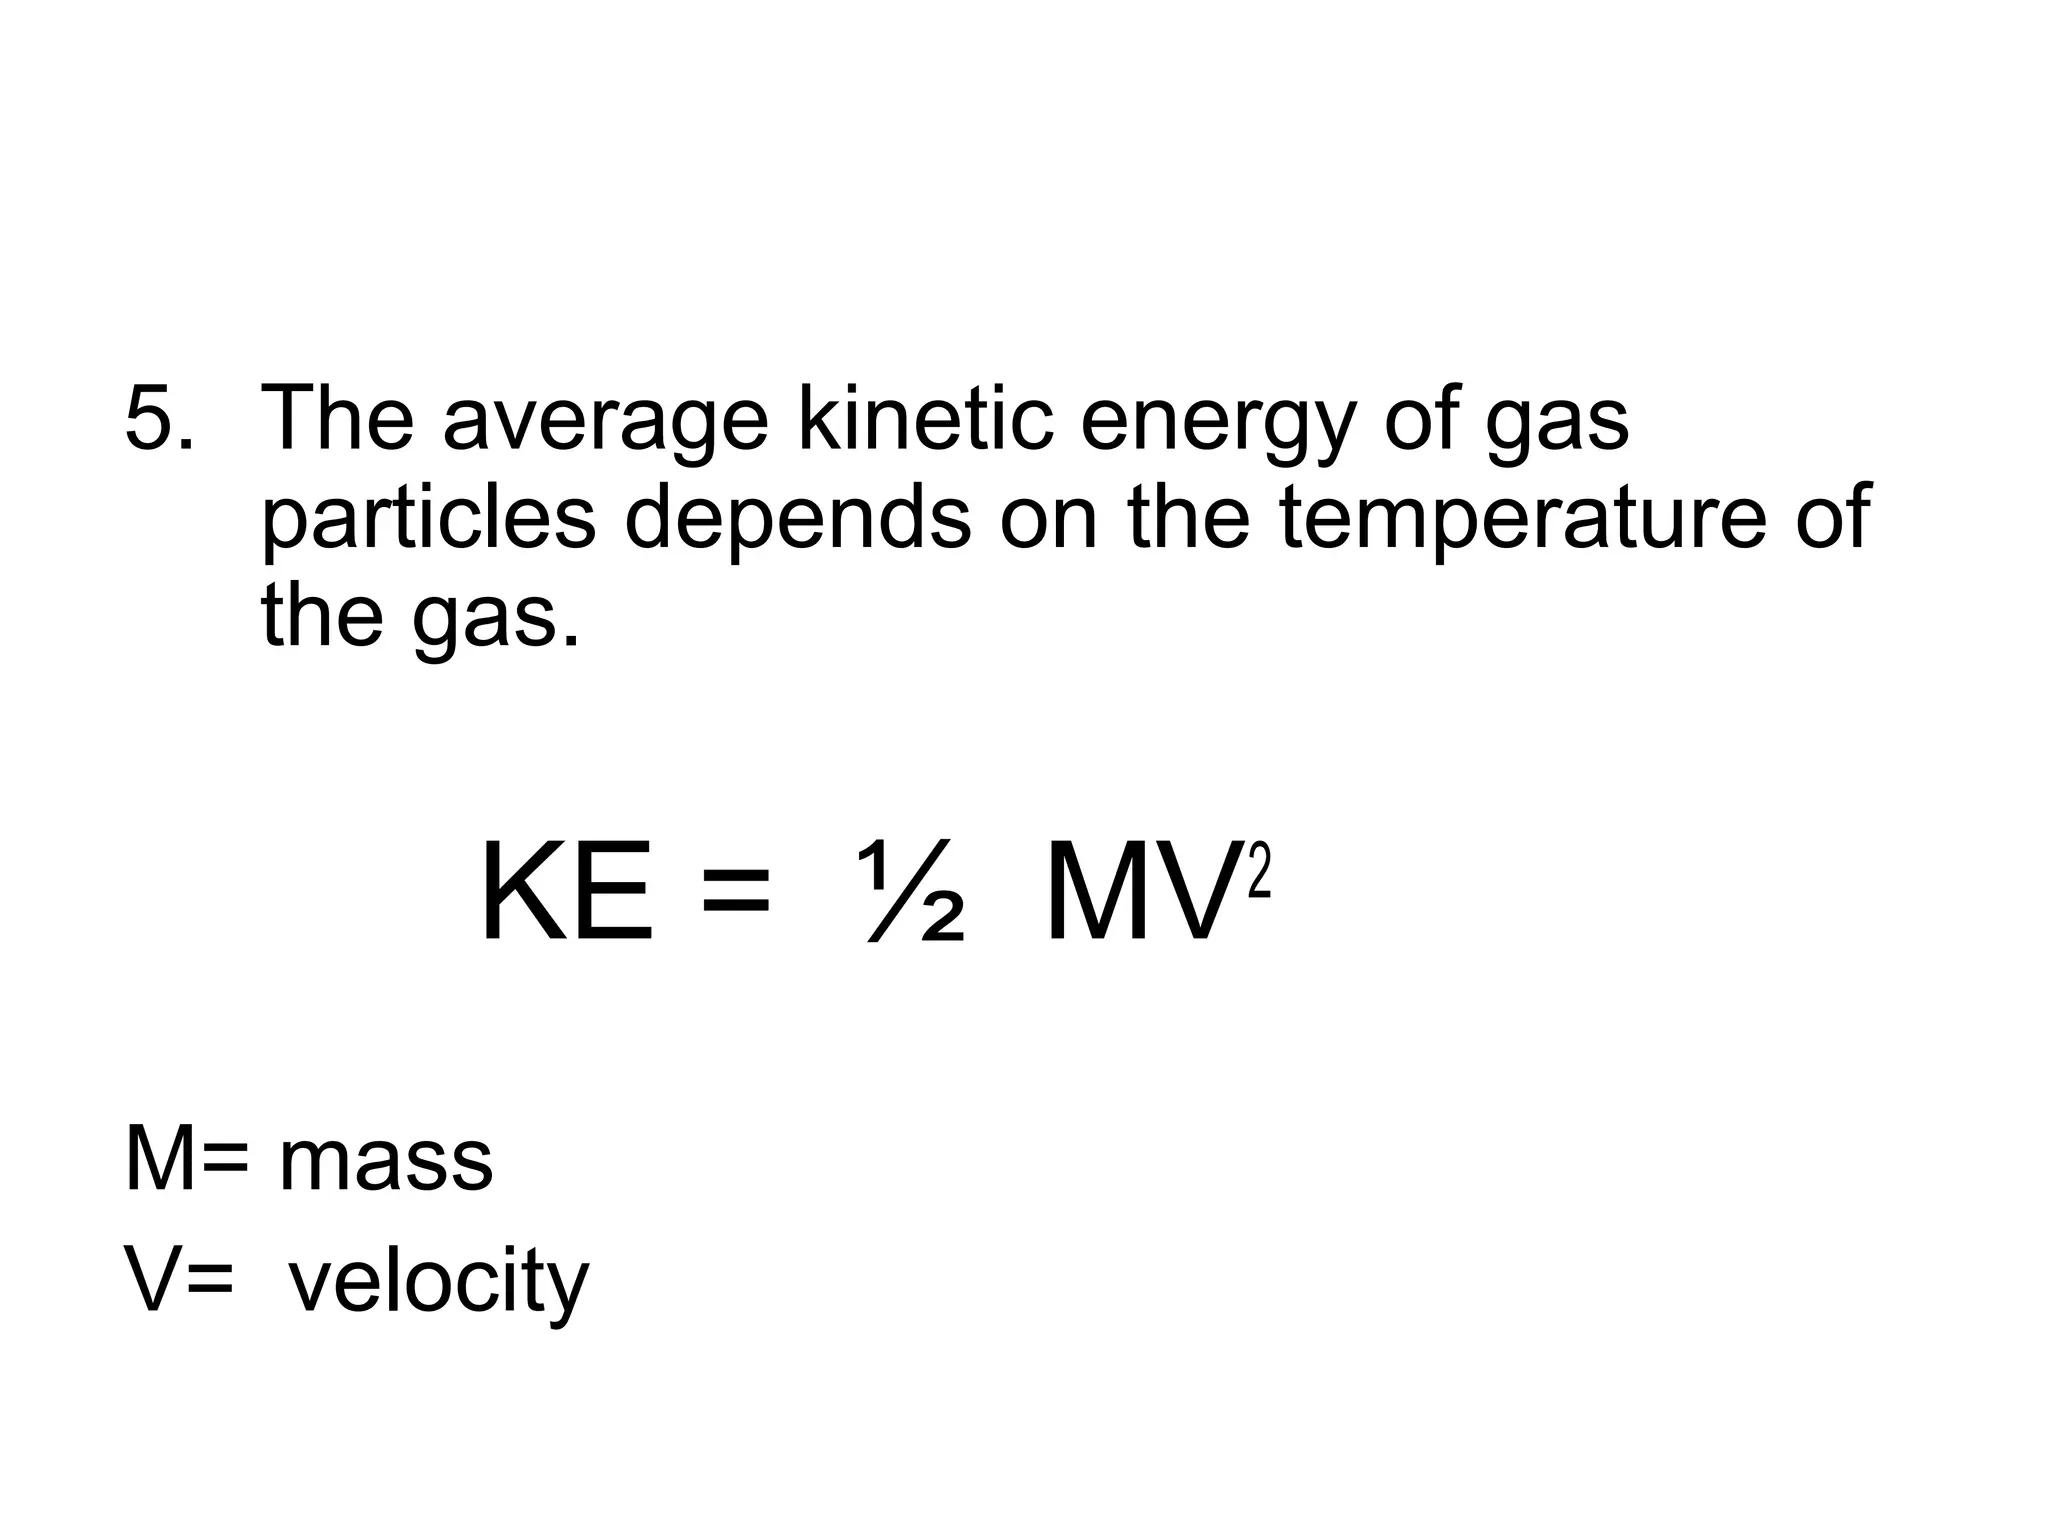
The kinetic energy formula applied to molecular motion is fundamentally just KE = 1/2 mv², but that simplicity hides a lot of practical complexity when you actually need to work with it in chemistry. I've spent years watching students and even professionals fumble through thermodynamics and kinetics problems because they treated the molecular kinetic energy framework as purely theoretical rather than something that breaks in predictable ways under real conditions. When you connect kinetic energy to temperature, you're looking at the average translational kinetic energy per molecule, which gives you KE_avg = 3/2 kT where k is Boltzmann's constant and T is absolute temperature. At room temperature, that works out to roughly 6.17 x 10^-21 joules per molecule for any gas. The mass term drops out entirely, which trips people up constantly. They expect heavier molecules to have more kinetic energy at the same temperature, but they don't. Heavier molecules move slower to compensate, and that's where the root mean square velocity equation becomes essential.
The Kinetic Energy Formula Unlocking The Dynamics Of Molecular Motion For Chemistry
The RMS velocity equation v_rms = sqrt(3RT/M) is the practical tool here, where R is the gas constant in J/(mol·K), T is temperature in Kelvin, and M is molar mass in kg/mol. Notice the molar mass has to be in kilograms, not grams. I see this mistake repeatedly in lab reports and exam work, and it produces answers that are exactly 1000 times too large because the square root of 1000 is about 31.6, throwing every subsequent calculation off by a wide margin. Here's where things get practically interesting. I once had a student running gas diffusion experiments in an undergraduate lab, trying to validate Graham's law by measuring the rate at which ammonia and hydrogen chloride gases met in a glass tube. The theoretical prediction based on RMS velocities came out about 12 percent higher than what we actually observed. We spent two weeks troubleshooting before realizing the issue wasn't the formula but the experimental setup. The glass tube had microscopic surface irregularities that created turbulent flow regimes at the concentrations we were using, and the diffusion wasn't purely molecular. Once we switched to longer tubes with larger diameters and lower pressures, the measurements aligned much closer to prediction. The kinetic theory assumptions of elastic collisions and negligible intermolecular forces are idealizations that break down in specific conditions, and you need to know which ones. Another thing that doesn't get enough attention is the distinction between translational, rotational, and vibrational kinetic energy. The 3/2 kT result applies strictly to translational motion in three dimensions. For diatomic and polyatomic molecules, you have additional degrees of freedom. Nitrogen at room temperature has rotational kinetic energy contributing another kT to the total per molecule, bringing it to 5/2 kT. That's why the molar heat capacity at constant volume for N is about 5/2 R rather than 3/2 R. If you're doing any thermochemistry work with real gases, ignoring these contributions introduces systematic errors that grow larger as you move to more complex molecules. Methane, for instance, has three translational and six rotational degrees of freedom at moderate temperatures, plus vibrational modes that start activating around 300 to 400 K and above.
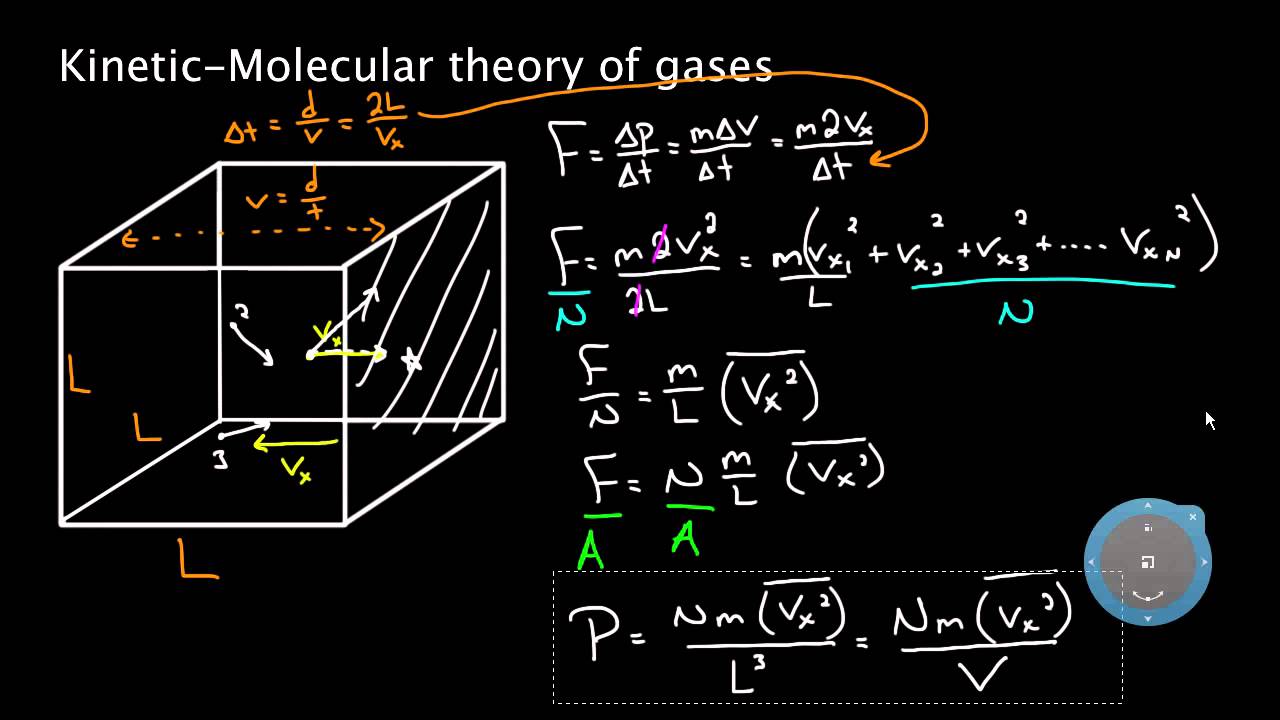
The Maxwell-Boltzmann distribution is what you actually need when the question isn't about average kinetic energy but about how many molecules exceed a certain energy threshold. This matters enormously for reaction kinetics. The Arrhenius equation's exponential term essentially comes from integrating the Maxwell-Boltzmann distribution above the activation energy barrier. I've used this repeatedly when troubleshooting why a reaction I expected to proceed rapidly at a given temperature was barely moving. Often the issue isn't the catalyst or concentration but that the activation energy is high enough that even at elevated temperature, only a tiny fraction of molecules in the high-energy tail of the distribution can actually react. Doubling the temperature doesn't double the reaction rate. It can increase it tenfold or more depending on Ea. There are real limitations to this framework that you should acknowledge upfront. The kinetic molecular theory assumes point particles with no volume and no intermolecular forces, which means it fails for dense gases and liquids. Near condensation points or in high-pressure systems, the ideal gas approximation embedded in these equations introduces significant error. Van der Waals corrections or more sophisticated equations of state become necessary, and the simple RMS velocity formula no longer gives you the full picture. For liquid-phase chemistry, which is where most synthesis actually happens, the concept of molecular kinetic energy still applies but the interpretation changes. Molecules in liquids are constantly colliding and exchanging energy on femtosecond timescales, and their motion isn't free-flight between collisions the way it is in gases. Molecular dynamics simulations handle this by numerically integrating Newton's equations of motion for each atom, but those require force fields and computational resources that most people don't have access to. If you're working with real-world data and need to convert between kinetic energy units and more practical chemistry units like electronvolts or kilojoules per mole, the conversion factors matter. One electronvolt per molecule equals about 96.485 kJ/mol. At 298 K, the average translational kinetic energy of a gas molecule is approximately 2.48 kJ/mol or 0.0257 eV. These numbers give you a quick mental reference for whether an activation energy or binding energy is large or small relative to thermal energy. If an energy barrier is below about 3RT, roughly 7.4 kJ/mol at room temperature, the process will be fast and essentially barrierless on a human timescale. Above 50 kJ/mol, you're looking at reactions that need significant heating or catalysis to proceed at reasonable rates.
Get the Full Details
The practical workflow I recommend is straightforward. Calculate the RMS velocity for your molecule at your temperature of interest using the molar mass in kg/mol. Check whether the ideal gas assumption is reasonable for your pressure range using the compressibility factor or by comparing your conditions to the critical point. If you're dealing with liquids or high pressures, switch to simulation or empirical data rather than relying on the analytic formulas. For reaction rate questions, use the Maxwell-Boltzmann distribution to estimate the fraction of molecules above your activation energy, then compare that to your experimental rate to validate or refine your model. I keep a reference sheet with the key equations, the common molar masses in kg/mol, and the conversion factors taped next to my monitor. It saves maybe fifteen minutes per problem set, but those minutes add up over a semester of work and more importantly they prevent the kind of systematic errors that show up as weird outliers in data you've already collected. Getting the units right from the start is the single highest-impact habit you can build when working with kinetic energy and molecular motion calculations.