So you need to actually account for London dispersion forces in your work and most textbooks aren't helping you.
I spent years watching people in computational chemistry and materials science completely misjudge van der Waals interactions, then wonder why their DFT calculations were off by 15 to 30 percent. The core problem is simple: London dispersion forces are universally attractive, they weaken rapidly with distance following an inverse sixth-power relationship, and they are the reason nonpolar molecules stick together at all. Without them, methane wouldn't be a liquid at low temperatures. Without them, graphite layers would fly apart instead of stacking neatly. But here is what nobody tells you until you have burned through weeks of debugging time — standard functionals in density functional theory, the ones you probably learned to use first, do not account for these forces correctly. Or at all, depending on which functional you pick. Let me walk through how I actually approached this, not the textbook version. I was modeling the adsorption energy of benzene on a graphene surface. My initial calculation using PBE, a very common generalized gradient approximation functional, showed almost no binding. Zero point five kilojoules per mole roughly. That was physically wrong. Benzene absolutely binds to graphene, and the experimental value sits around forty to fifty kilojouues per mole. I had just wasted three days because I did not include dispersion corrections from the start.
The Invisible Forces At Play Understanding London Dispersion Forces
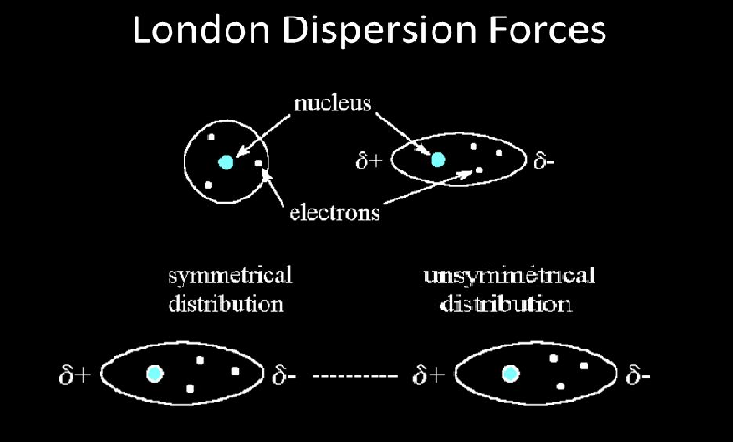
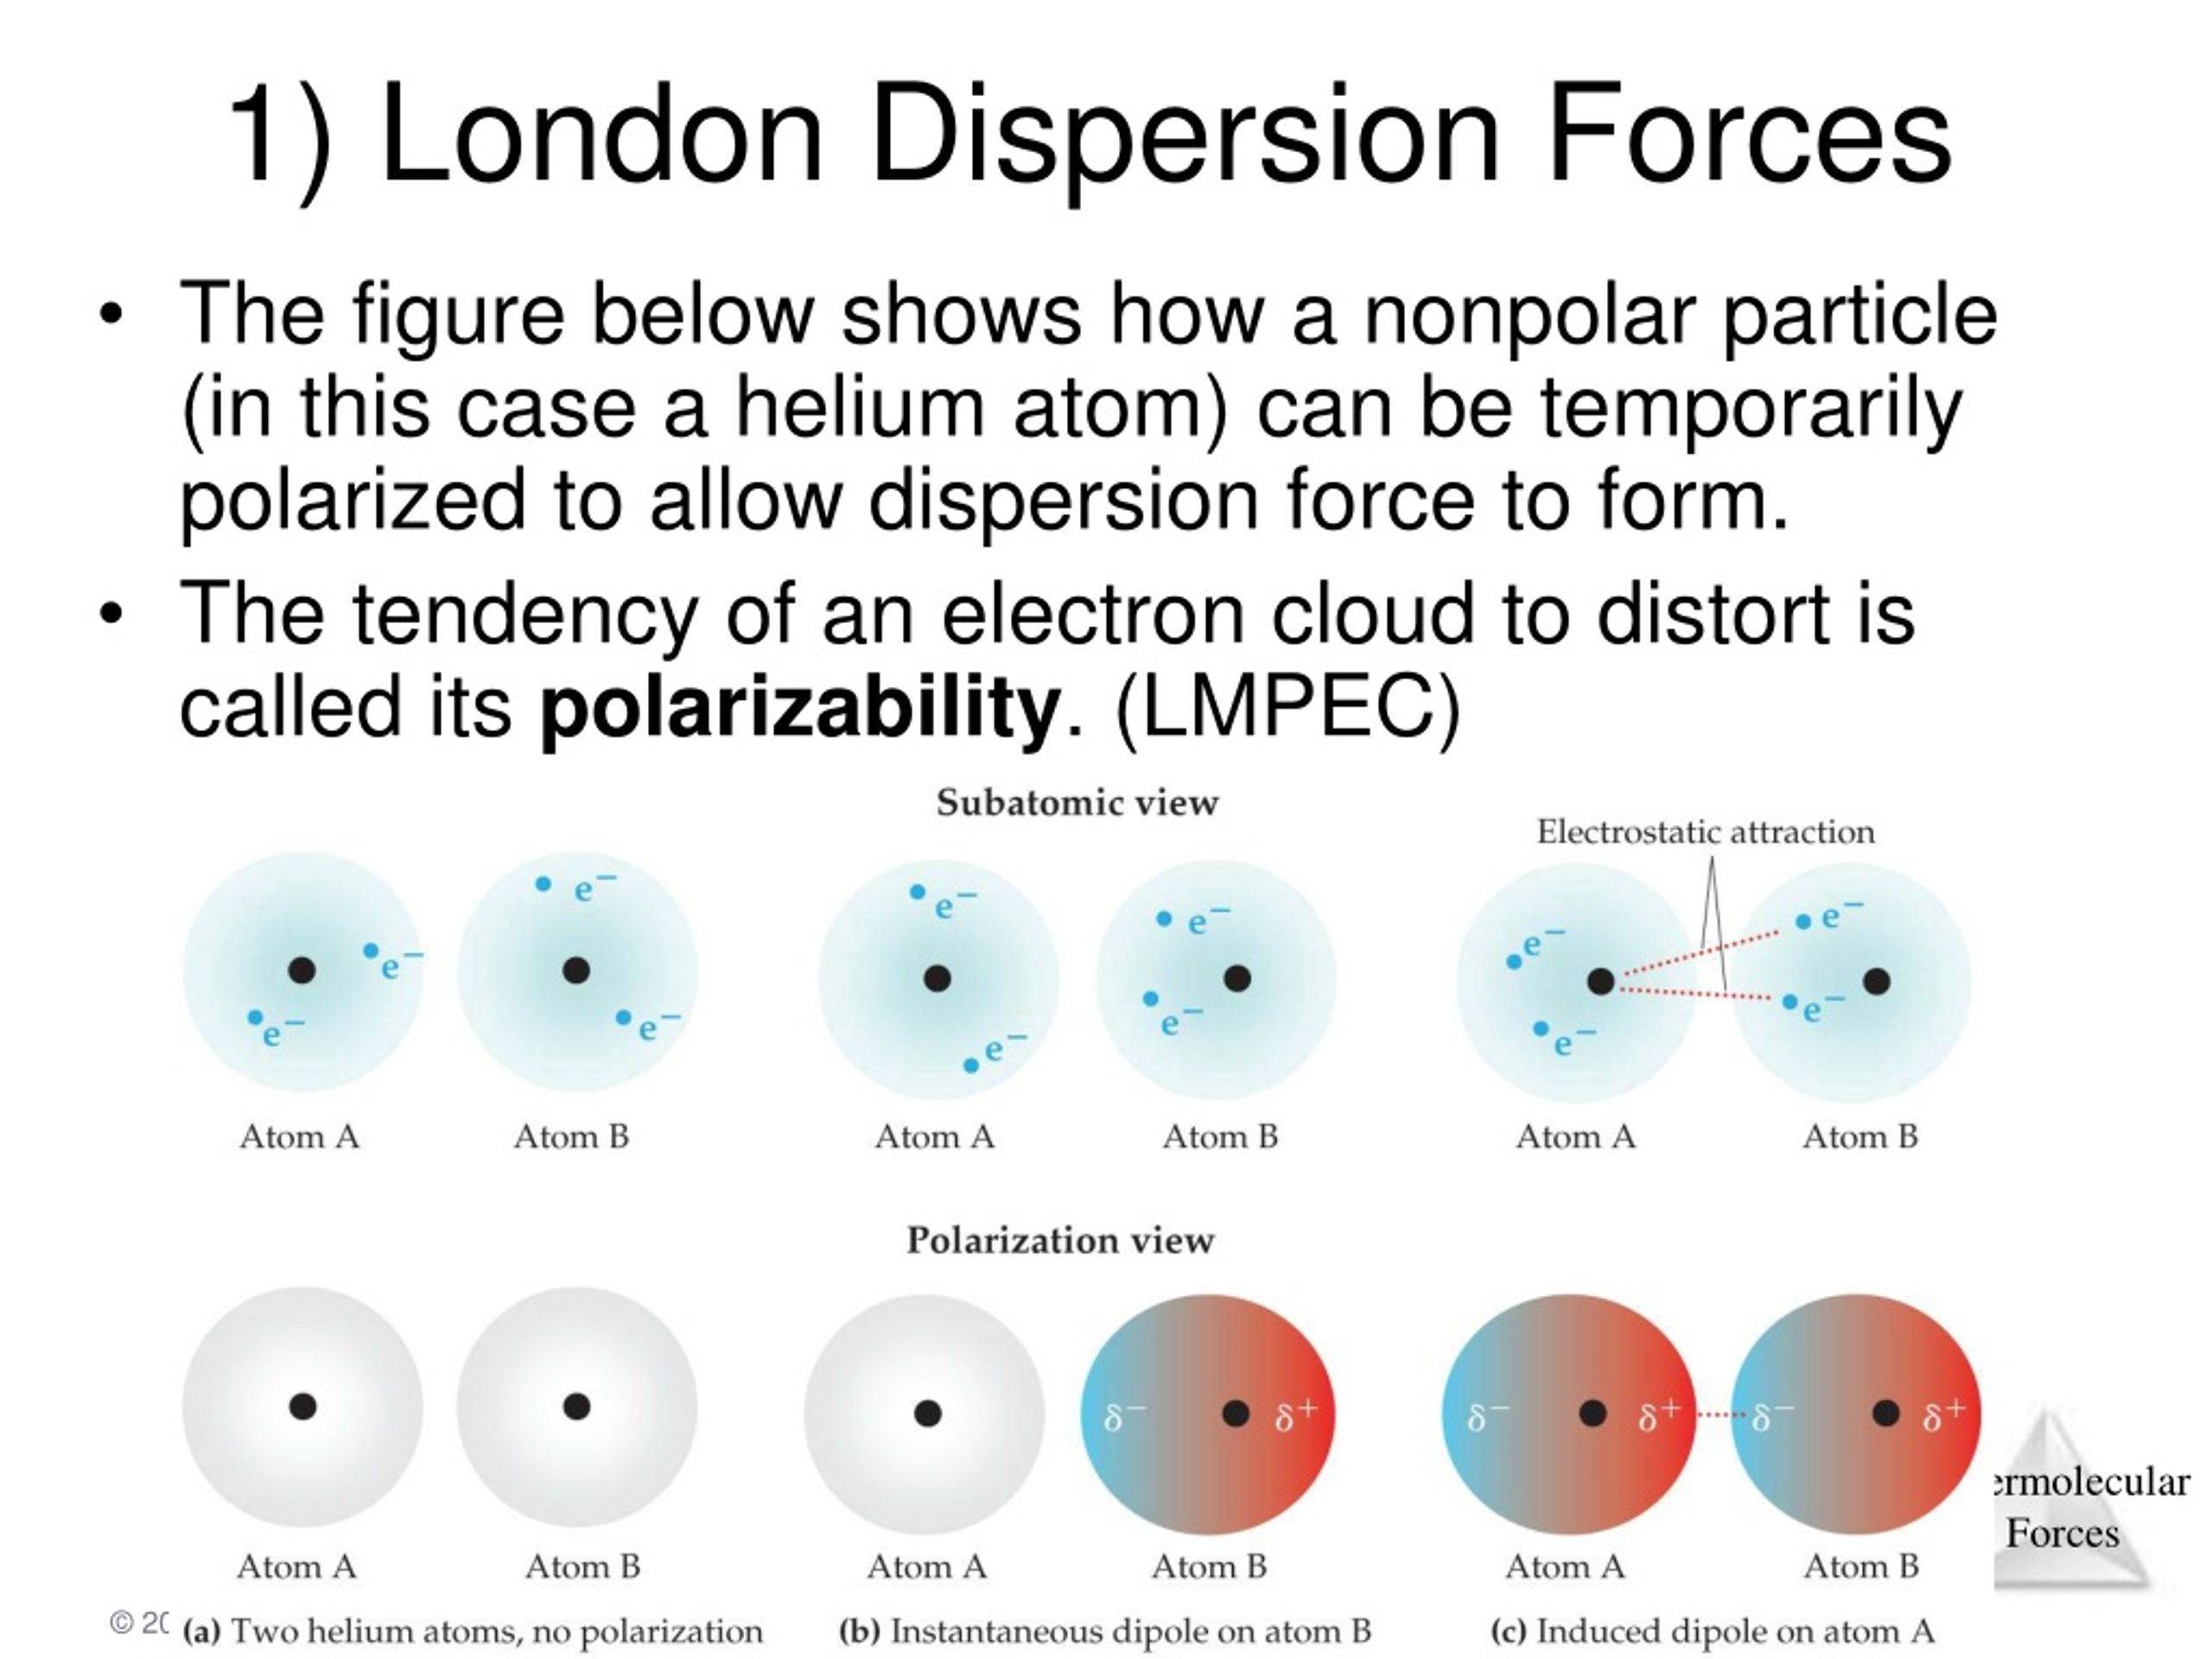
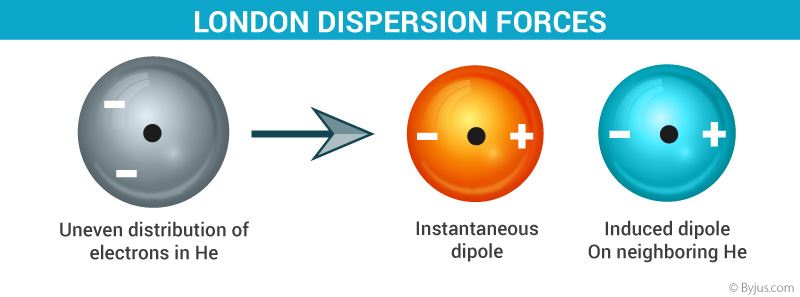
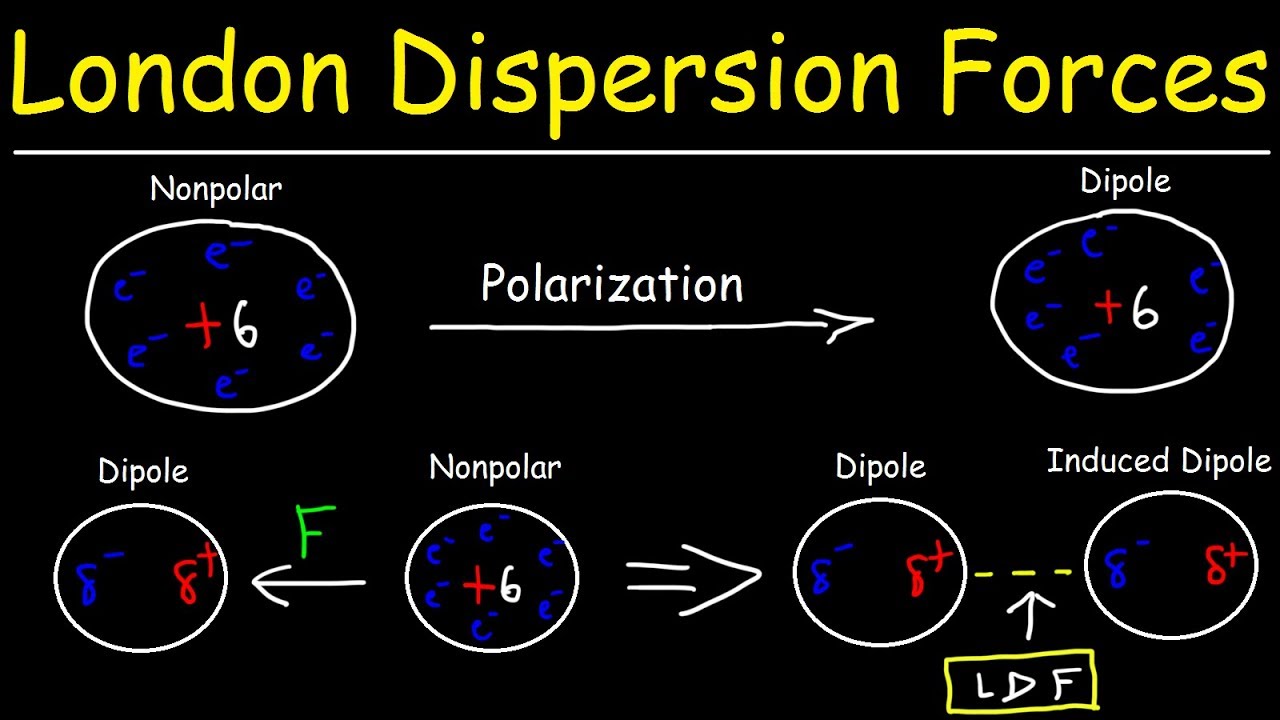
Here is the practical breakdown of what is happening at the quantum mechanical level. Electrons are constantly moving. At any given instant, there will be a random fluctuation in electron density on one molecule, creating a temporary dipole. This temporary dipole induces a complementary dipole in a neighboring molecule almost instantaneously, since electromagnetic interactions travel fast enough that you can treat this as simultaneous for most practical purposes. The result is an attractive interaction. It works even between atoms with zero permanent dipole moment, like argon or nitrogen. The strength depends heavily on polarizability, which scales with the size and softness of the electron cloud. Larger atoms and molecules with more diffuse electron clouds exhibit significantly stronger dispersion interactions. A good rule of thumb is that polarizability roughly scales with molecular volume, so going from CH4 to C8H18 is not a small increase in dispersion contribution. The mathematical description comes from the London equation, which relates the interaction energy to the ionization potentials and polarizabilities of the two interacting species, divided by the sixth power of the separation distance. You do not need to derive this by hand in most cases, but you should know that the R^-6 dependence means dispersion forces become negligible past about three to four angstroms in typical organic systems. They dominate at medium range, but they get crushed by electrostatic interactions and covalent bonding at short range, and they get overwhelmed by entropy at high temperature. Now for the part that matters if you are actually doing calculations or designing materials. There are several ways to incorporate dispersion into your workflows, and choosing the wrong one is where most people get stuck.
The DFT-D approach developed by Grimme adds an empirical correction term to standard DFT calculations. You pick DFT-D2, DFT-D3, or DFT-D3(BJ), and the program adds a C6*R^-6 term with parameters fitted to reference data. This is the most widely used method and it works well for the vast majority of organic and organometallic systems. The BJ damping function in D3(BJ) tends to be more reliable than the original zero-damping version, especially for systems where atoms get very close. I use D3(BJ) almost exclusively now. It adds maybe ten percent to your computational cost and fixes the biggest problems. Then there are dispersion-corrected functionals like B97-D and M06-2X, which have some dispersion built into their parameterization. M06-2X is a hybrid meta-GGA functional that tends to perform well for noncovalent interactions without additional corrections, but it is expensive and not always the best choice for transition metal systems. B97-D includes an empirical dispersion term. These are fine but less flexible than adding DFT-D corrections post hoc. For wavefunction-based methods, coupled cluster with perturbative triples, CCSD(T), is the gold standard and it captures dispersion naturally. The problem is that it scales as N^7 with system size, so you are looking at maybe twenty to fifty atoms max on a reasonable cluster, and even then it will take hours or days. For anything larger, you are out of luck unless you use local correlation methods, which are faster but require more setup knowledge.
Get the Full Details
Force fields like OPLS-AA or CHARMM handle dispersion through Lennard-Jones 12-6 potentials, where the attractive term is the C/R^6 component and the repulsive term is the B/R^12 component. This is standard for molecular dynamics simulations of proteins, polymers, and large supramolecular assemblies. The caveat here is that force field parameters for dispersion coefficients are fitted to reproduce bulk properties, not electronic structure. If you are studying something unusual, like a novel metal-organic framework or an exotic adsorbate, the default parameters might not transfer well and you should validate against higher-level calculations or experimental data. I ran into a specific edge case recently that illustrates why this matters. I was modeling a halogen-bonding interaction between an iodine-containing organic molecule and a -system. Halogen bonds are directionally specific and involve a sigma-hole on the halogen atom. Standard dispersion corrections treated the iodine atom fine, but the directional electrostatic component of the halogen bond was poorly described by my chosen functional. I ended up needing a combination of a range-separated hybrid functional with D3(BJ) corrections plus careful analysis of the electron density topology using quantum theory of atoms in molecules. The total interaction energy shifted by about eight kilojoules per mole when I switched from PBE-D3 to B97X-D. That is a massive difference when you are trying to predict binding selectivity. Another common pitfall involves solid-state systems. Periodic DFT calculations of layered materials, MOFs, and porous frameworks are extremely sensitive to the treatment of dispersion. If you model a MOF without dispersion corrections, your predicted pore volumes and lattice parameters can be wildly off. I once saw a published structure where the unit cell volume was off by twelve percent because the authors used a functional without any dispersion treatment. The framework looked plausible visually, but the distances between aromatic rings in the pores were completely wrong. Always verify your lattice parameters against experimental X-ray data when you can.
For practical guidance on getting started, if you are using Gaussian or ORCA for small molecule calculations, add the appropriate dispersion keyword. In Gaussian that is usually EmpiricalDispersion=GD3BJ. In ORCA you add %methodDisp dftd3 true end. The difference in computational time is negligible for systems under two hundred atoms. If you are doing periodic calculations with VASP or Quantum ESPRESSO, make sure your pseudopotentials and functional choice are compatible with dispersion corrections. VASP has the DFT-D2 and DFT-D3 options built in via the IVDW parameter. Quantum ESPRESSO can use the TS (Tkatchenko-Scheffler) method through the vdW-DF functionals or external dispersion packages. The limitations are real and you should know them before you invest time. Empirical dispersion corrections like DFT-D3 are fitted parameters. They work remarkably well across diverse chemical spaces, but they are not universal. There are known failures in systems with strong charge transfer character, highly polarizable heavy metal complexes, and situations where many-body dispersion effects become important. Many-body dispersion, where the polarizability of one atom is affected by the presence of a third atom, is not captured by pairwise additive corrections like DFT-D3. For large van der Waals complexes and condensed-phase systems, methods like the many-body expansion or DFT with van der Waals density functionals such as revDOD or r2SCAN-3c can be more accurate, though they are computationally more demanding. If you are working with very large systems where even DFT-D3 is too slow, consider semi-empirical methods with dispersion corrections like GFN2-xTB, which include an analytical dispersion term and can handle systems with thousands of atoms in reasonable time. The accuracy is lower than DFT, but for conformational searches and initial geometry screening, it is often sufficient and orders of magnitude faster.
The bottom line is that London dispersion forces are not optional. They are a fundamental component of intermolecular interactions that affects everything from boiling points to crystal packing to biomolecular recognition. Ignoring them is a mistake you will notice immediately in your results. Including them with the right method is straightforward if you know which tools to use and where the approximations break down.