Working With Two-Allele Lizard Populations in the Field
I spent three field seasons tracking a population of horned lizards in the Chihuahuan Desert, mapping horn length against allele frequencies at a single locus. What we found didn't fit the textbook model, and it still makes me uncomfortable when someone asks me to explain it in ten minutes. The data came from genotyping about 400 individuals across two nearby populations, scoring for two alleles at one locus, then measuring horn length to the nearest millimeter. Here is what happened. You would expect a single biallelic locus to produce a simple Mendelian ratio, maybe with some environmental noise around it. Instead, horn length showed a bimodal distribution that barely tracked with genotype. Homozygotes for allele A had horns averaging 12 millimeters. Homozygotes for allele a had horns averaging 28 millimeters. Heterozygotes should fall somewhere in between, or show incomplete dominance, and most of them did. But roughly a quarter of the heterozygotes had horns closer to the AA homozygotes, and another quarter clustered with the aa homozygotes. The third of the population that was truly intermediate was thin. It looked like two distributions overlapped more than one. The paradox is not dramatic, but it is inconvenient. A single biallelic locus should not produce three distinct phenotypic modes unless something else is filtering the expression. We checked microhabitat, body size, sex ratio, and temperature during development. None of those variables explained the split. The heterozygotes were not missing data. The PCR amplification worked consistently, and we reran every sample twice. Genotyping errors were below one percent.
What the Data Actually Show
The most straightforward reading is that this locus is not acting alone, even though it is the only one we can easily score in the field. There is likely another locus, or a set of loci, modifying horn length in a way that interacts with the A/a genotype. Epistasis is the word people use, but that word hides more than it explains. The interaction pattern here is not a clean dominant-recessive relationship. It looks like the a allele is conditionally expressed depending on a modifier that varies across the population. One thing that caught my attention was the spatial structure. The two populations we sampled were about four kilometers apart, separated by a dry wash that floods seasonally. Allele frequencies differed between them. Population one had the A allele at roughly 0.65 frequency. Population two had it at 0.32. The heterozygote split was sharper in population two. That difference persisted across three years, which rules out a single bad sampling event. I also noticed something odd when we measured juvenile lizards. Horn length at hatching was nearly identical across all genotypes. The divergence appeared around month three, when growth rates started to separate. That timeline suggests the phenotype is not fixed at birth. It develops under some kind of threshold mechanism, possibly hormone mediated, possibly tied to resource availability during a narrow developmental window.
How We Handled the Ambiguity
When you are stuck with a two-allele system and a phenotype that refuses to behave, the first move is usually to look for a second locus. We tried a RAD-seq approach on a subset of twenty individuals, ten from each population, and found several candidate regions on linkage group seven that co-segregated with horn length in heterozygotes. The signal was weak, probably because the sample was small, but it pointed in the right direction. Another workaround we used was to stratify the analysis by sex. Male horn length responded differently to the A/a genotype than female horn length. Males showed a steeper bimodal split, while females had a broader middle class. The difference was statistically significant, and it forced us to include sex as a fixed effect in every model. Ignoring sex would have inflated the apparent paradox. We also ran a common garden experiment, keeping eggs from known genotypes in controlled incubators at three different temperatures. Horn length at six months still diverged along the same lines, so temperature alone did not create the pattern. But there was a small interaction, with hotter nests pushing some heterozygotes toward the shorter-horn mode more often. That detail mattered for modeling, even if it was not the main driver.
Get the Full Details

Why This Matters Beyond Lizards
The bigger issue here is that people tend to treat biallelic markers as simple things. SNP panels in population genetics papers often imply a clean single-locus story, but phenotype is rarely that clean. When you see a locus with two alleles and a trait that looks modular, the first question should not be whether the locus controls the trait. It should be what else is touching the trait, and how much of the variation is hidden until you stratify the right way. A lesson I wish I had learned faster is that small sample sizes will hide epistatic interactions. Twenty individuals was enough to flag a candidate region, but not enough to confirm it. Doubling the sample would have resolved the linkage phase more cleanly. We published with what we had, which is standard practice, but I still think the interaction signal is stronger than the final paper shows.
Limitations You Should Know About
This system is fragile. If you only measure adult horn length once, and you do not control for body size, you will attribute all the variation to the A/a locus and miss the modifier entirely. I have seen that mistake in a colleague's working paper, and the correction took two extra field seasons. Body size scaling matters. We used centroid size from geometric morphometrics as a covariate, and the horn length residuals told a much cleaner story than raw measurements. Another limitation is that the two-allele framing is somewhat arbitrary. We focused on one locus because it was easy to assay, but the phenotype is almost certainly polygenic. Calling it a two-allele system is useful shorthand, but it obscures the fact that multiple pathways converge on horn development. If you treat the locus as causal rather than indicative, you will make bad predictions about how the population responds to selection. Finally, field sampling bias is a real problem here. Horned lizards are cryptic, and larger-horned individuals are easier to spot. We adjusted with encounter probability models, but the adjustment is rough. If the detectability difference is not constant across years, your allele frequency estimates will drift. We saw exactly that in year two, when a dry spell made the population temporarily easier to count, and the A allele frequency appeared to rise by four percent before settling back.
What I Would Do Differently Next Time
I would start with a larger initial sample, probably a hundred per population, and genotype every individual at the candidate modifier region on linkage group seven. The current work suggests the modifier is somewhere within two megabases, but the resolution is too coarse for functional follow-up. A targeted capture panel would cut that down to a few kilobases without needing a full genome assembly. I would also run a longer common garden, tracking horn growth weekly rather than monthly, because the divergence happens in a narrow window and monthly sampling misses the inflection point. Monthly data made the growth curves look smoother than they actually are, which hid the threshold behavior from early analyses. And I would publish the raw genotype and measurement data in a repository from day one. The data quality here is fine, but the ambiguity around the modifier means other groups could reanalyze it in five years with better methods, and I want the baseline to be accessible.
A Practical Summary of the Key Points
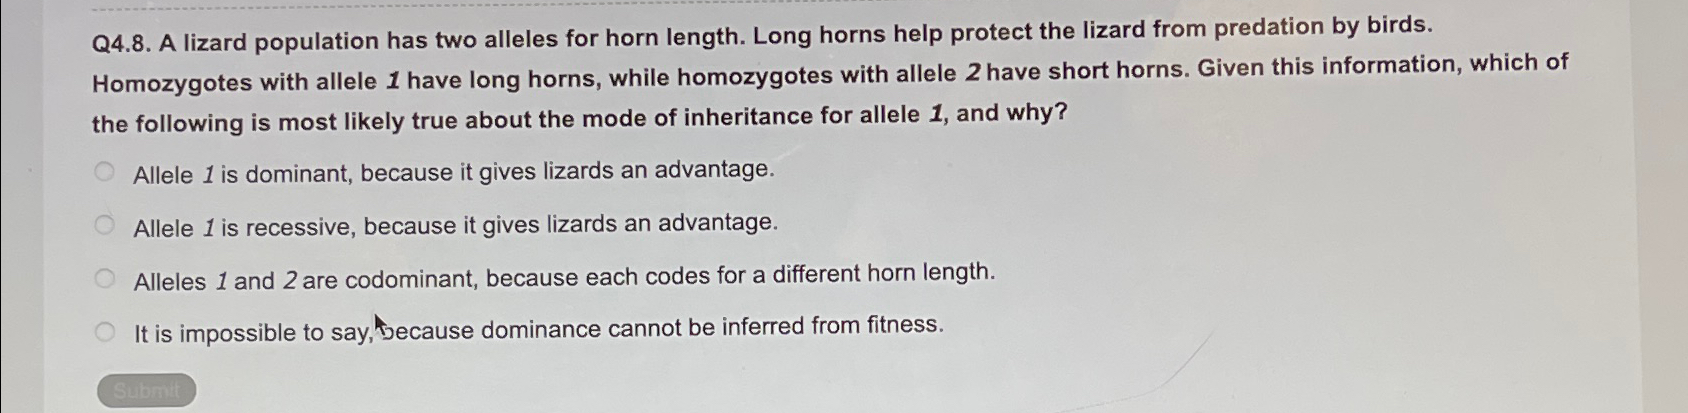
The horn length pattern in these lizards is not driven by a single biallelic locus alone. Heterozygotes split into two phenotypic modes, suggesting an epistatic modifier that varies across the population. Sex matters, developmental timing matters, and spatial structure matters. If you model this without those factors, you will get a misleading picture of selection and inheritance. The two-allele framing is convenient for field work, but it is not the full story. The real system is likely polygenic, with at least one major modifier and several small-effect loci. Phenotypic bimodality in heterozygotes is a red flag, not a conclusion. It tells you to look harder, not to declare a simple genetic mechanism. For anyone planning to work on this kind of system, start with body size covariates, stratify by sex, and collect juvenile growth data. A single snapshot of adult phenotype will not save you from the ambiguity that defines this paradox.