Understanding Molecular Geometry From The Ground Up
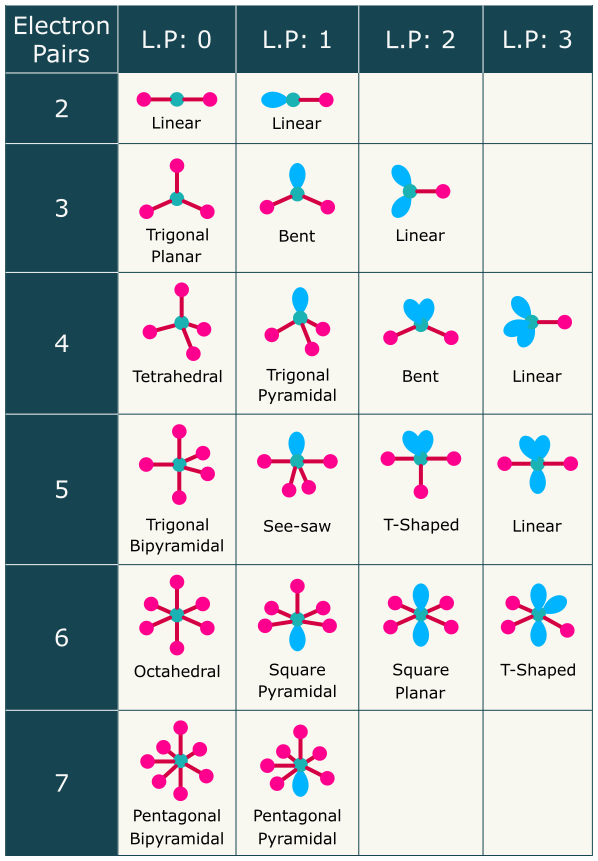
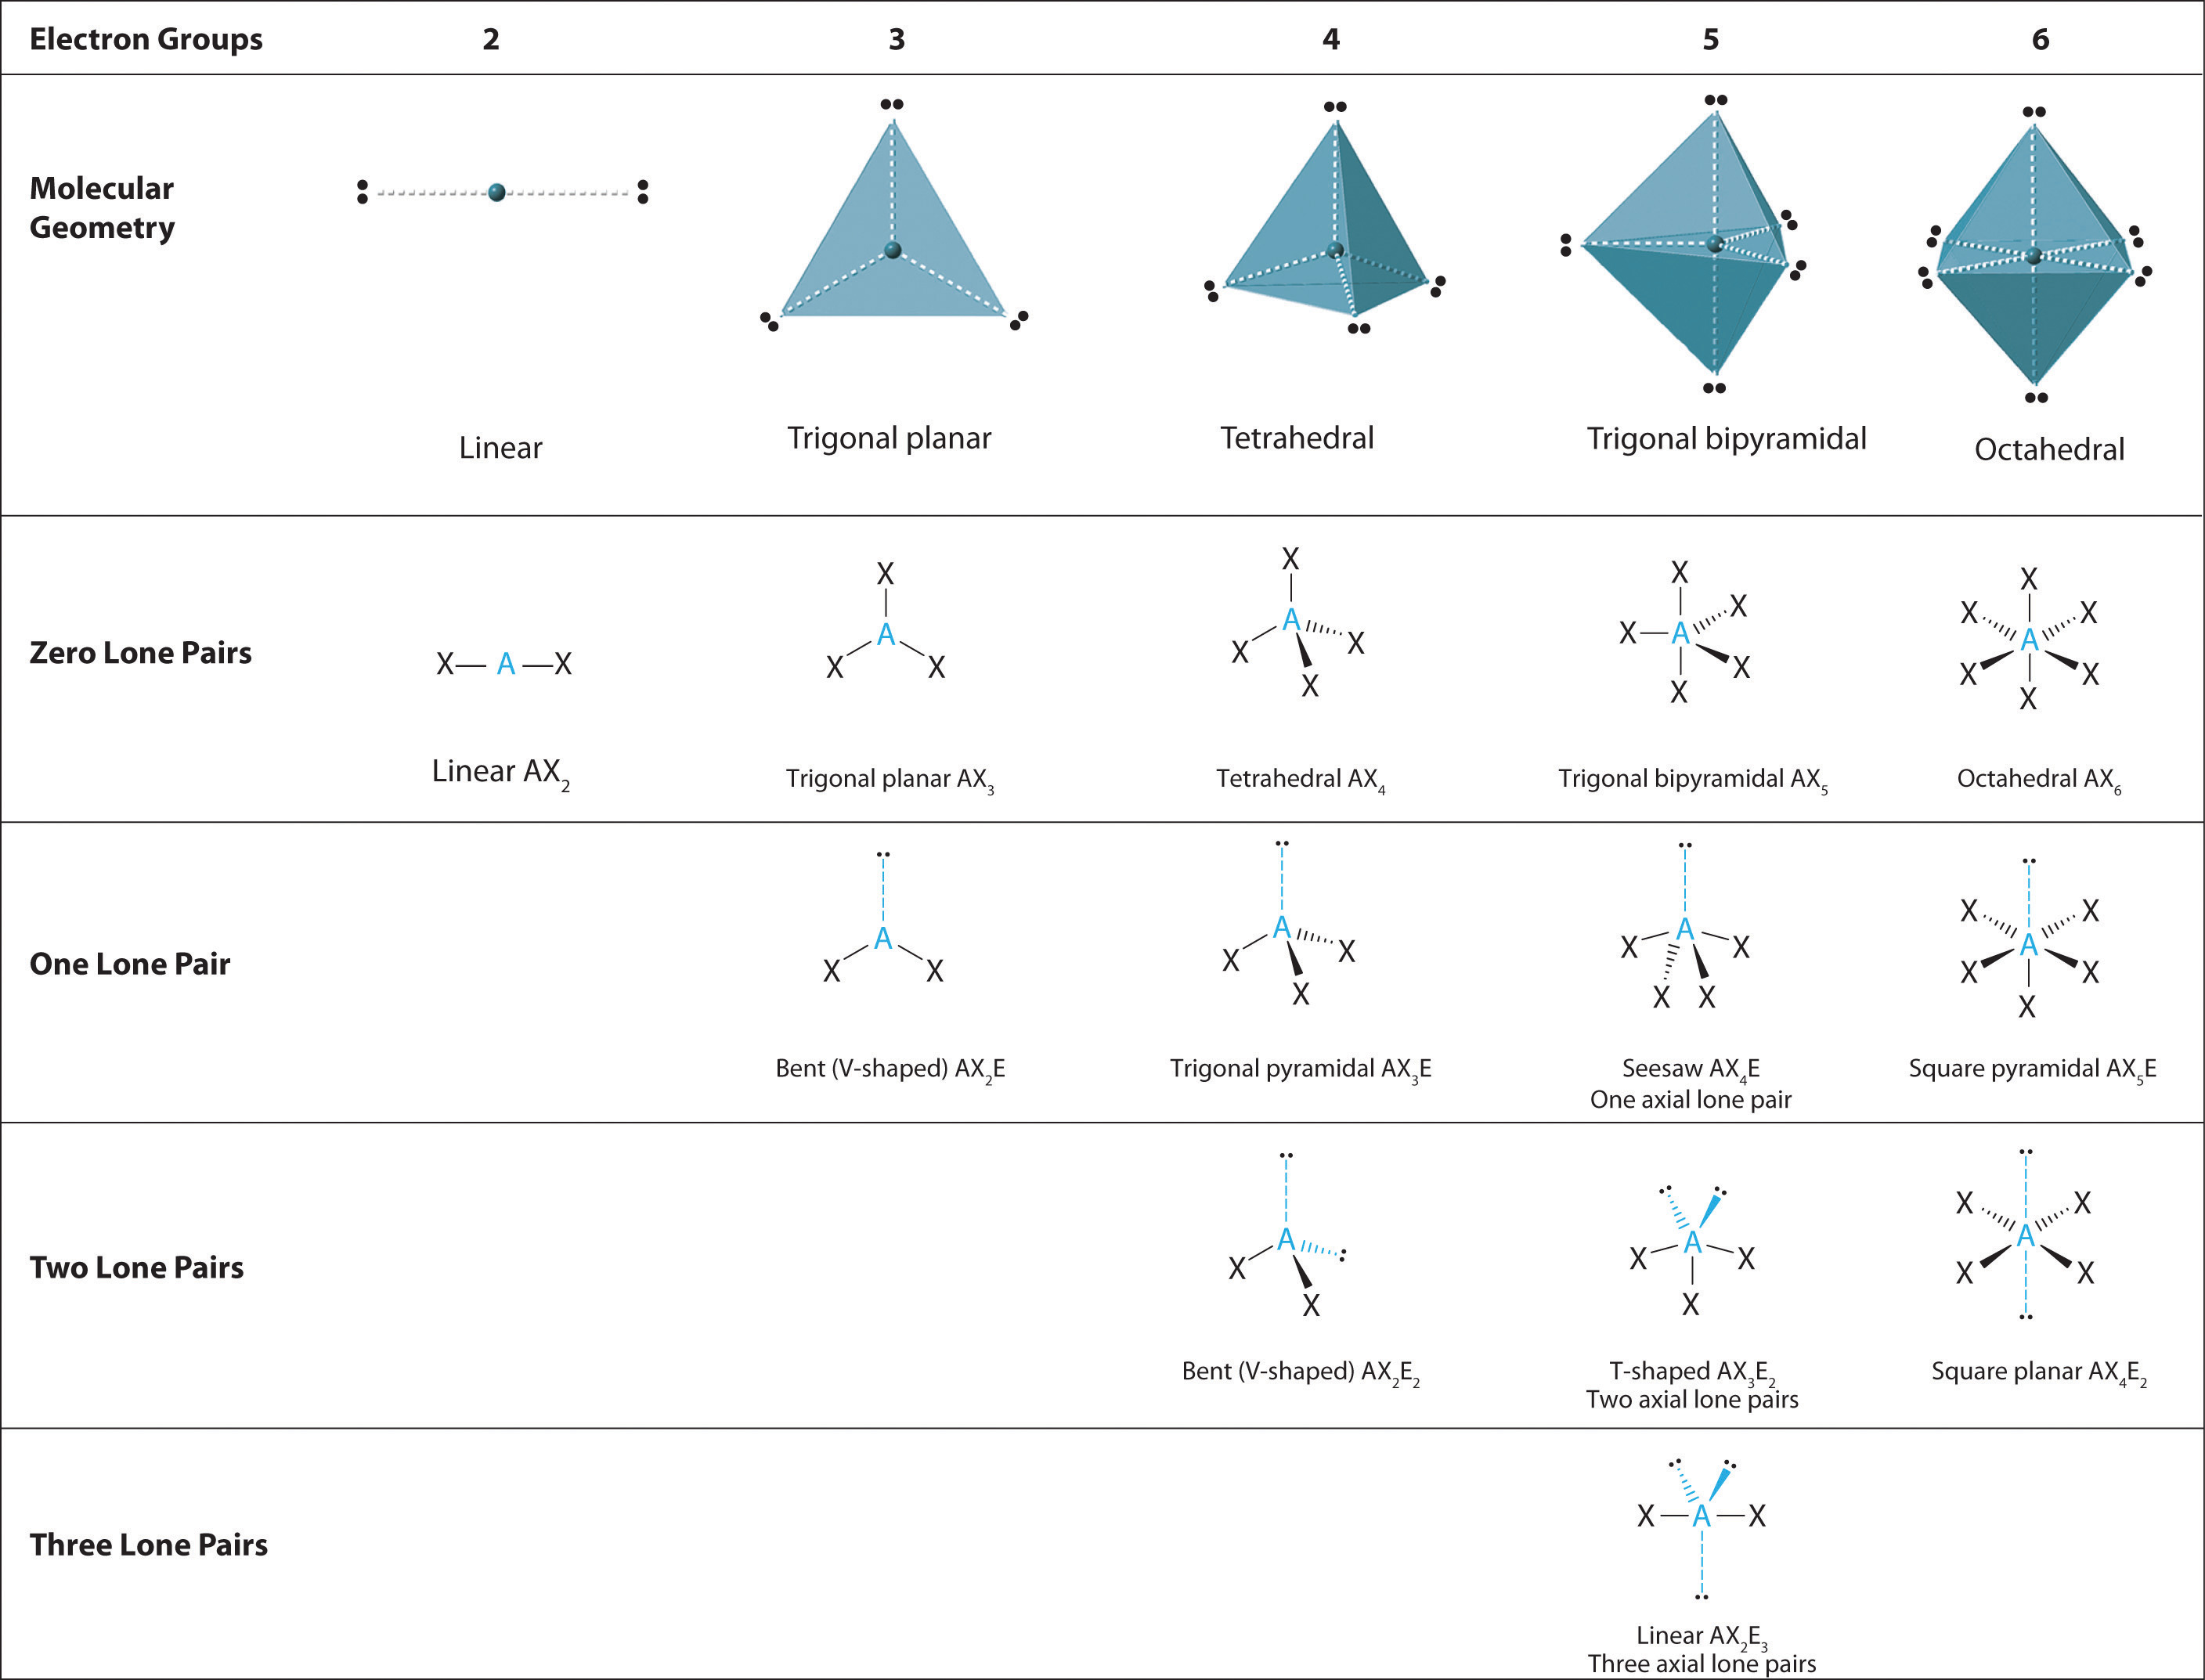
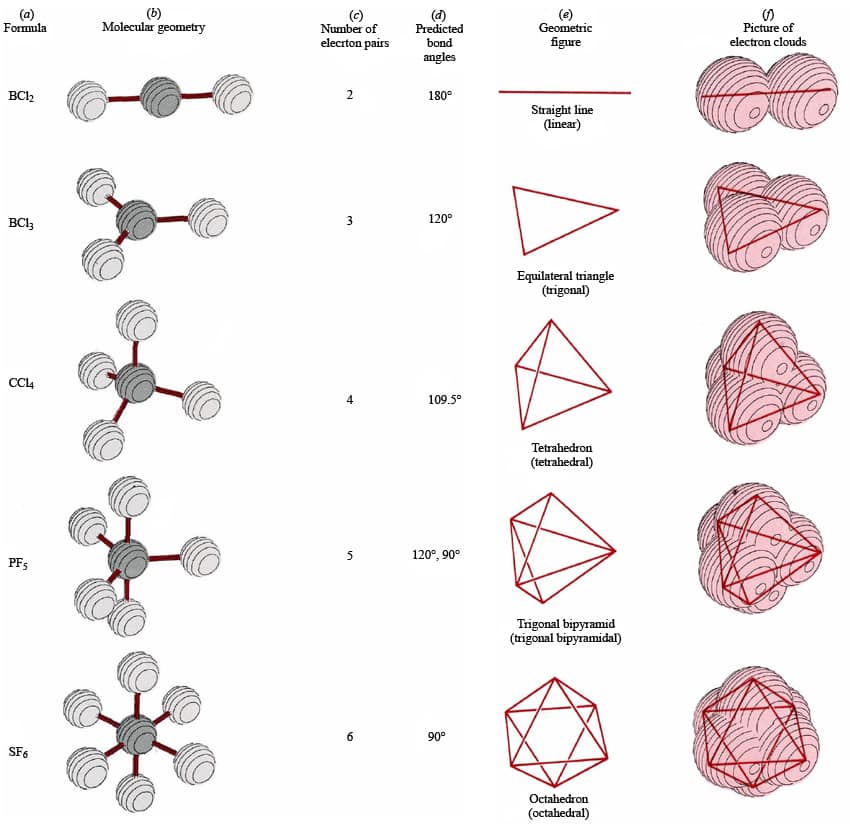
Most chemistry students learn VSEPR theory as a memorization exercise. Draw dots, count bonds, look up a table. That approach gets you through a few exams and then falls apart the moment you encounter something that doesn't fit the standard model. The actual geometry of a molecule is determined by the behavior of electron domains around a central atom. What I mean by that is simpler than the textbooks make it, but also harder to apply when you are working with real compounds. The core idea is that electron pairs repel each other. Not metaphorically — there is an electrostatic force at work. When you have four regions of electron density around an atom, they will arrange themselves as far apart as possible. That gives you a tetrahedral arrangement. Three regions give you trigonal planar. Two regions give you linear. The shape you observe depends on whether those electron domains are bonding pairs or lone pairs.
The Hidden Geometry Of Molecules How Electron Shapes Dictate The World
This phrase comes up often in literature, and it refers to the fact that molecular shape determines everything about how a substance behaves. Intermolecular forces. Solubility. Reactivity. The reason water is a liquid at room temperature and hydrogen sulfide is a gas comes down to bent molecular geometry creating a dipole moment. Without that bent shape, there is no polarity, and the whole chemistry changes. This is not theoretical. It is why your body functions the way it does. Here is where people usually get tripped up. Lone pairs occupy more space than bonding pairs. This is not a minor detail. A lone pair will compress the bond angles around it. In ammonia, the ideal tetrahedral angle is 109.5 degrees, but the H-N-H angle is actually 107 degrees because the lone pair pushes the bonding pairs closer together. In water, with two lone pairs, the angle drops further to about 104.5 degrees. If you ignore the lone pair effect, your predictions are wrong every time. I learned this the hard way during a computational chemistry project a few years back. I was modeling a phosphorus-containing compound and kept getting bond angles that did not match the experimental data. The structure looked right on paper but the calculations were off by several degrees. What I had missed was that the phosphorus atom had a stereochemically active lone pair that was not obvious from the Lewis structure alone. The workaround was to run a geometry optimization with a higher level of theory and include explicit treatment of the lone pair rather than relying on the standard VSEPR template. The optimized angle came within one degree of the reported value after that change. That single correction took me from a day and a half of troubleshooting down to about three hours of re-running calculations.
Advanced Considerations Beyond The Basics
Not every molecule follows the simple VSEPR pattern. Transition metals are the first place this breaks down. d-orbital participation changes everything. An octahedral complex might distort into a square planar or tetragonal geometry depending on the electronic configuration of the metal and the ligands attached to it. Crystal field theory explains this, but VSEPR cannot handle it at all. Beginners who try to force VSEPR onto transition metal complexes end up with nonsense structures. Another area where things get messy is hypervalent molecules. Phosphorus pentachloride exists as PCl5 in the gas phase with trigonal bipyramidal geometry. But in the solid state, it dissociates into PCl4+ and PCl6- ions. The geometry you predict depends entirely on the phase you are working in. If you are doing computational work, you need to specify the conditions or you will be comparing apples to oranges. Resonance structures also complicate geometry prediction. Ozone is a classic example. You might draw two resonance forms and assume the molecule has two distinct bond lengths. It does not. Both O-O bonds are identical at about 127.8 picometers, intermediate between a single and a double bond. The actual geometry is bent with a bond angle of roughly 116.8 degrees. The electron density is delocalized across all three atoms, and that delocalization affects the shape in ways a simple Lewis structure cannot capture.
Get the Full Details
If you want to move beyond qualitative predictions, you need to understand molecular orbital theory at least well enough to know when VSEPR fails. Hybridization is useful as a shorthand, but sp3d hybridization for trigonal bipyramidal geometry is essentially a fiction. The bonding in SF4 is better described using three-center four-electron bonds rather than invoking d-orbital hybridization, which computational chemistry shows contributes negligibly to the bonding. This distinction matters when you are trying to explain results to someone who actually knows the subject.
Practical Approaches For Prediction And Verification
For quick manual predictions, the steric number method is still the most efficient tool available. Count the number of atoms bonded to the central atom, add the number of lone pairs, and you get the steric number. Steric number 4 with zero lone pairs is tetrahedral. Steric number 4 with one lone pair is trigonal pyramidal. Steric number 4 with two lone pairs is bent. The pattern holds for steric numbers 2 through 6 in most main-group compounds. When you need accuracy, computation is the only reliable route. Gaussian, ORCA, and Q-Chem are standard packages. For a routine organic molecule at the DFT level with a medium basis set, a geometry optimization typically takes anywhere from 15 minutes to an hour on a modern workstation. That is fast enough for iterative work but slow enough that you cannot just throw thousands of structures at it without thinking about your workflow. Setting up a proper job requires selecting the right functional, basis set, and convergence criteria. Using B3LYP with a 6-31G* basis set is a reasonable default for organic molecules, but it is not universal. Some systems require dispersion corrections. Some require range-separated functionals. Testing your method against a known reference compound before committing to it saves weeks of debugging later. X-ray crystallography remains the gold standard for experimental verification, but it has limitations. You need a good crystal. That is not always possible. Neutron diffraction gives better information about hydrogen positions, but it requires access to a neutron source, which is rare and expensive. Electron diffraction works for gas-phase molecules and gives you bond lengths and angles directly, but interpretation is more complex. For most practical purposes, a combination of IR spectroscopy and NMR will tell you enough about molecular geometry to proceed with your work without needing high-end structural data.
Where The Method Breaks Down
VSEPR and even basic quantum chemistry calculations struggle with molecules that have significant electron correlation effects. Heavy elements with relativistic effects are another problem. Gold clusters, for instance, do not follow the patterns you would expect from simple electron counting. The geometry of Au25(SR)18 is dictated by relativistic contraction of the s-orbitals and expansion of the d-orbitals, which VSEPR cannot account for at all. DFT with relativistic effective potentials is the practical solution, but it requires specialized knowledge and computational resources that most people do not have access to. Molecules with near-degenerate electronic states are also problematic. Strontium monoxide in its ground state has a geometry that standard single-reference methods describe poorly because the electronic structure has significant multireference character. In these cases, you need CASSCF or similar multiconfigurational methods. They are computationally expensive and technically demanding. If your molecule falls into this category, the simple answer is to consult someone who already knows how to run these calculations rather than trying to learn it on the job. The biggest practical limitation I encounter is the assumption that a single static geometry represents the molecule. Molecules vibrate. At room temperature, the atoms are constantly moving. What we call a bond length is actually an average over many vibrational states. For most purposes this average is good enough. But if you are studying reaction dynamics or spectroscopic properties, you need to account for vibrational averaging, and that adds significant complexity to the calculation. Zero-point energy corrections are typically a few kilojoules per mole, which sounds small until you are trying to resolve energy differences of that magnitude between competing reaction pathways.
The geometry of molecules is not just an academic curiosity. It is the reason certain drugs bind to certain proteins, why catalysts work the way they do, and why some materials conduct electricity while others do not. Understanding electron shapes gives you a framework for predicting behavior rather than memorizing outcomes. The framework has real limitations, and knowing where those limitations are is as important as knowing how to use it. Most people stop learning once they can draw a water molecule correctly. The ones who get things done keep going past that point.


