Why Your Drug Molecule Doesn't Fit The Receptor
When I was first working through lead optimization for a kinase inhibitor series, our compound had great in vitro potency but absolutely terrible cell permeability. We spent three weeks trying different solubilizing groups before I actually looked at the dihedral angles around the central amide bond. The molecule wasn't just flexible - it was sampling conformations that placed the hydrogen bond acceptor roughly 180 degrees away from where the receptor expected it. That's not a solubility problem. That's a geometry problem. Bond angles aren't decorative. They determine whether a methyl group points into a hydrophobic pocket or directly at a water molecule that's competing for that space. They determine whether your ester can even reach the catalytic serine in an enzyme active site. Most people learn VSEPR theory in their second semester of chemistry and then never think about it again until they're staring at a crystallization hit that makes no thermodynamic sense.
The Hidden Geometry Of Life How Bond Angles Dictate Molecular Destiny
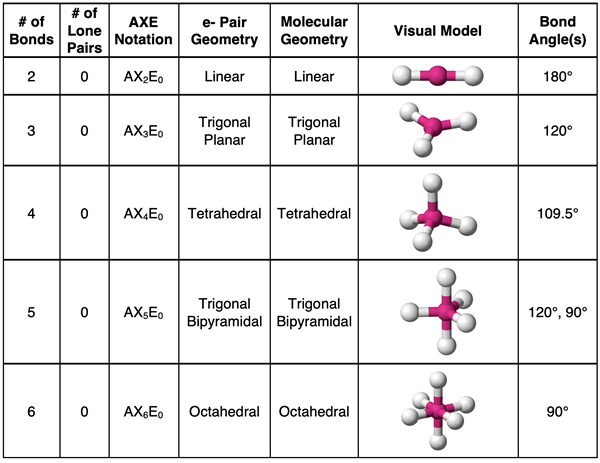
Let me explain how this actually works when you're trying to design something that binds to a biological target. Start with the central atom and count the electron domains around it. Two domains means 180 degree separation - linear geometry. Three domains gives you roughly 120 degrees in a trigonal planar arrangement. Four domains is approximately 109.5 degrees for tetrahedral, though you'll see deviations when lone pairs are involved. This is standard textbook material, but the practical implications are where things get interesting. Here's what the textbooks don't emphasize enough: bond angles are not fixed constants. They flex. A C-C-N angle in a protein backbone can shift by several degrees depending on the steric environment around it. When I model a ligand into a binding site and the geometry looks right on paper but the docking score is garbage, I usually check whether the bond angles forced by the target pocket are creating strain energy that outweighs any favorable interactions. Strain penalties of 2 to 4 kcal/mol are common in rigidified analogs, and that translates to a 10 to 100-fold difference in binding affinity depending on your conditions. I ran into a specific case last year where we were optimizing a GPCR agonist. The SAR data was completely flat across a series of analogs - adding or removing a methylene in the linker didn't seem to change binding at all. I built out the 3D models and looked at the bond angles more carefully. What I found was that the linker was long enough to allow the pharmacophore to reach the binding site through multiple conformations, but short enough that the optimal bond angle was being compromised in every single one. The molecule was paying an angular strain penalty of roughly 3 kcal/mol across the entire series, which masked any subtle electronic effects we were trying to measure. I proposed shortening the linker by one carbon and increasing the sp2 character of the central atom to lock in a more favorable geometry. The new analog showed a 25-fold improvement in Ki. The binding pocket hadn't changed. The geometry had.
The counter-intuitive part that trips up most people new to this is that more rigidity doesn't automatically mean better binding. Rigidifying a flexible bond to lock it into what looks like the bioactive conformation sounds like the right move, and it often is, but you're also eliminating the entropic cost of binding only if the locked conformation is actually the one the receptor selects. If you rigidify into the wrong angle, you've made things worse, not better. I've seen molecules go from nanomolar to micromolar potency because someone took a flexible rotatable bond and froze it into a conformation that was 15 degrees off from what the crystal structure showed. Another thing beginners consistently miss: hybridization state changes during binding. A nitrogen that's sp2 in solution can rehybridize toward sp3 when it forms a hydrogen bond in a protein pocket, shifting its bond angles by roughly 10 to 15 degrees. If you're doing molecular mechanics calculations with fixed hybridization states, your geometry will be off, and your energy estimates will be unreliable. Use force fields that allow for rehybridization or run short QM/MM relaxations on the binding site before trusting the numbers. There are tools for this. Open Babel handles basic geometry optimization and is free. Avogadro is good for visual inspection of bond angles in real time. For anything involving actual binding energy predictions, you want something like Gaussian or ORCA for QM calculations, though those require computational resources and patience. The workaround I use when I need quick geometry checks without setting up a full QM job is to build the molecule in Avogadro, run a MMFF94 minimization, then export the coordinates to a short DFT geometry optimization at the B3LYP/6-31G* level. That usually takes about 15 minutes on a decent workstation and catches the kind of angular strain issues that would take hours to diagnose by trial and error.
Get the Full Details
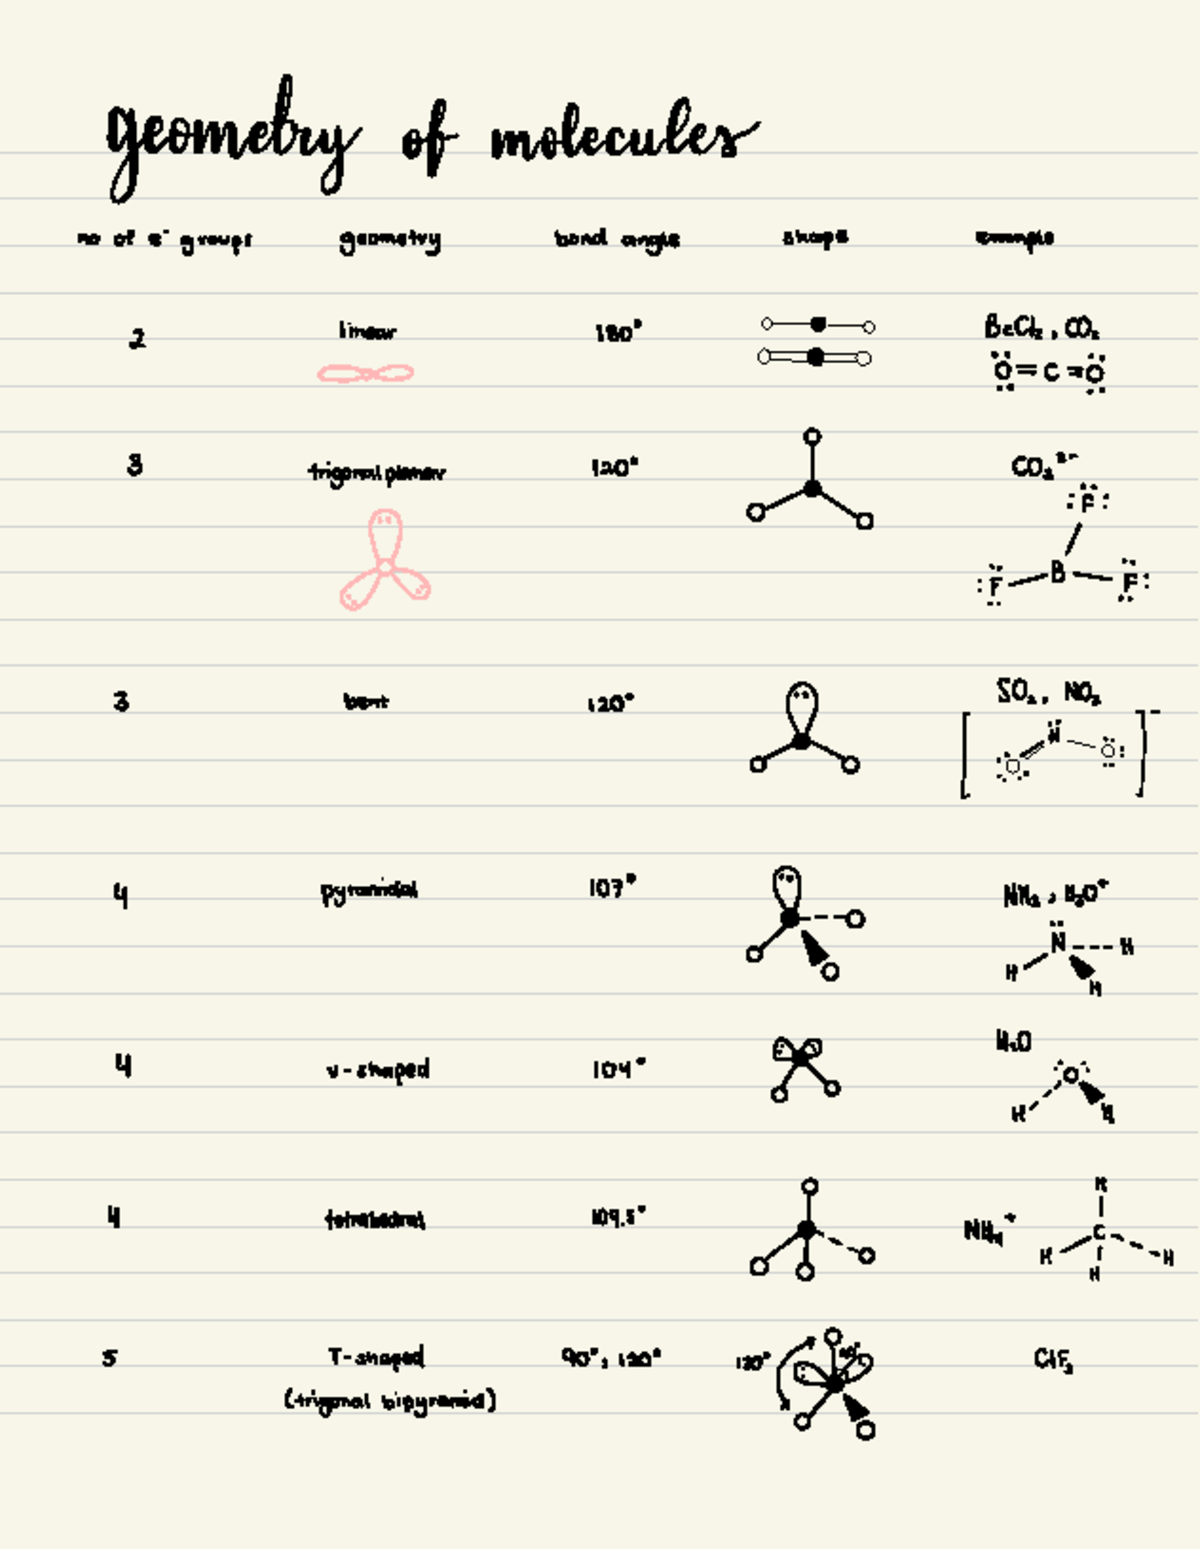
The limitation you need to accept is that bond angle analysis alone won't save a poorly designed molecule. If your hydrogen bond donors and acceptors aren't positioned correctly, no amount of angular optimization will fix that. Bond angles modulate binding affinity - they don't create it from nothing. You also can't rely on gas-phase geometry calculations to predict solution behavior without accounting for solvent effects. Implicit solvent models help, but explicit water molecules in the binding site can shift angles significantly, and ignoring them is a mistake I see repeated in computational chemistry papers regularly. When the geometry gets complicated - say you're dealing with a macrocyclic compound or a peptide with constrained ring systems - the standard VSEPR approach breaks down and you need to think in terms of conformational energy landscapes rather than individual bond angles. In those cases, I recommend using something like MacroModel or even just running extensive molecular dynamics simulations to see which angles are actually sampled under physiological conditions. The average bond angle from an MD trajectory will tell you more about reality than a single minimized structure ever will. The bottom line is that bond angles are a fundamental constraint on molecular behavior that most people gloss over because they seem simple. They're not simple. They're the difference between a molecule that fits and one that doesn't, and catching angular issues early in the design process saves a lot of wasted synthesis and assay time.