What You Actually Need To Know About Water Before You Start Simulating It
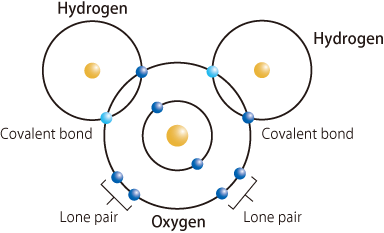
Water looks simple on paper. Two hydrogens, one oxygen, a bond angle of 104.5 degrees. You probably learned that in high school chemistry and never looked back at it. But if you have ever tried to model water in any computational setting, whether it is molecular dynamics, CFD, or quantum chemistry, you quickly learn that H2O is one of the most annoying molecules in existence. Not because it is hard to define, but because it refuses to behave like anything else. Here is the thing most people miss: water is not a rigid molecule. The bonds stretch. The angle bends. And in any realistic simulation, you have to decide whether to treat those degrees of freedom as fixed or flexible. That single choice cascades into everything else. Rigid water models like SPC/E or TIP4P run orders of magnitude faster than flexible ones like TIP4P/Flex, but they completely miss certain infrared spectral features and fail to capture the coupling between rotational and translational motion at high temperatures. I spent about six months debugging a simulation that kept producing anomalous diffusion coefficients for water near a charged surface. The system looked fine on paper. The force field was standard TIP3P. The box size was adequate. The temperature coupling was set correctly. The problem turned out to be that I was using a 2 femtosecond timestep with constrained bonds, which is apparently fine for bulk water but completely breaks down when water is sitting within two angstroms of a silica surface under an electric field. The constraints were allowing energy to leak into unphysical vibrational modes, and the integrator was quietly drifting. Switching to a 1 femtosecond timestep fixed it, but cut my throughput in half. I ended up running the production phase on a different machine with more nodes rather than trying to optimize further.
The deeper issue is that water has no single correct description. Every force field makes a different compromise. TIP3P overestimates diffusion by roughly 40 percent compared to experiment. SPC/E corrects that somewhat but underestimates the dielectric constant. TIP4P/2005 gets the density and diffusion closer to real values but requires a longer cutoff and more computation per step. Then there is the polarizable models like SWM4-NDP or AMOEBA, which handle things like dielectric screening and ion hydration much better, but they are roughly five to ten times more expensive per timestep and notoriously finicky to converge in certain boundary conditions. You pick your poison based on what property actually matters for your work. Another thing nobody warns you about is the sensitivity to long-range electrostatics treatment. Water is highly polar, so the way you handle Coulombic interactions matters enormously. Particle Mesh Ewald is the standard, but the parameters you choose for it can shift your results more than you might expect. I had a case where changing the grid spacing from 1.0 to 0.8 Angstroms in the PME calculation altered the radial distribution function gOO significantly around the first peak, even though the total energy difference was negligible. If you are publishing simulation work on water, you should report your PME parameters explicitly. Most people do not, and it makes replication painful. There is also the question of quantum nuclear effects, which become relevant when you start looking at isotope substitution or low-temperature behavior. Protons are light. They delocalize. Path integral molecular dynamics can capture this, but it multiplies your computational cost by the number of beads you need, typically somewhere between 8 and 32 at room temperature. For most practical applications you do not need it, but if you are studying proton transfer mechanisms or the subtle differences between H2O and D2O, classical mechanics simply will not give you the right answer. There is no middle ground here.
If you are just starting out and want to experiment without building everything from scratch, GROMACS has decent built-in water models and the documentation covers the common setups adequately. OpenMM is another solid option if you prefer a Python interface and want to prototype something quickly. For quantum mechanical treatment, CP2K handles water reasonably well with its mixed Gaussian and plane-wave approach, though the setup is less straightforward than the classical packages. None of these are particularly difficult to install anymore, but each one has its own quirks when it comes to output formats and energy reporting conventions, so plan to spend a day just getting comfortable with the basics before you run anything production-scale. The real takeaway is that water is deceptively complicated, and the complexity is mostly hidden in the assumptions you make without thinking about them. Rigid or flexible. Which force field. How you treat electrostatics. Whether nuclear quantum effects matter for your specific question. Getting the answers right depends on recognizing which of those choices actually impacts what you are measuring, and being willing to redo a simulation when you realize you picked wrong.
Get the Full Details