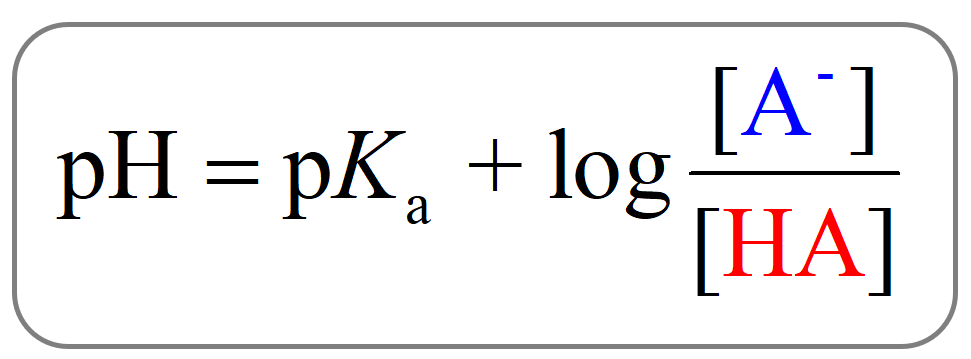Buffer Calculations That Actually Work
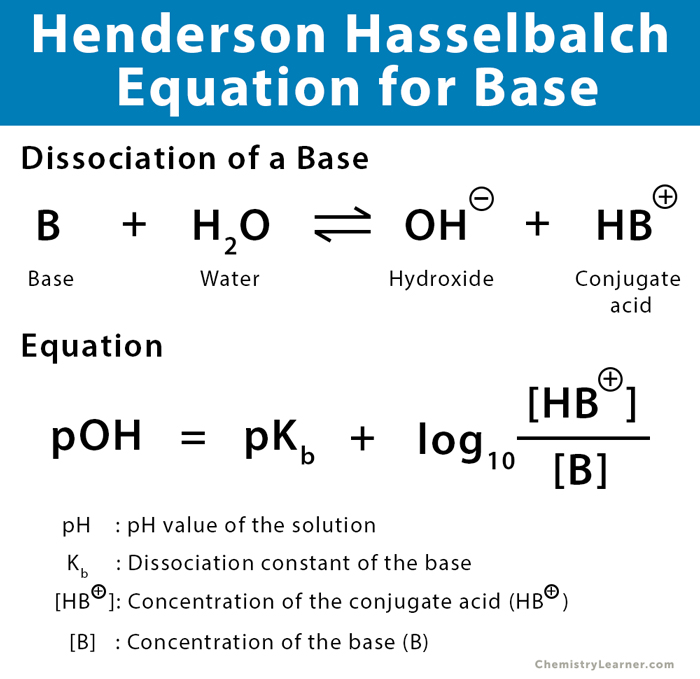
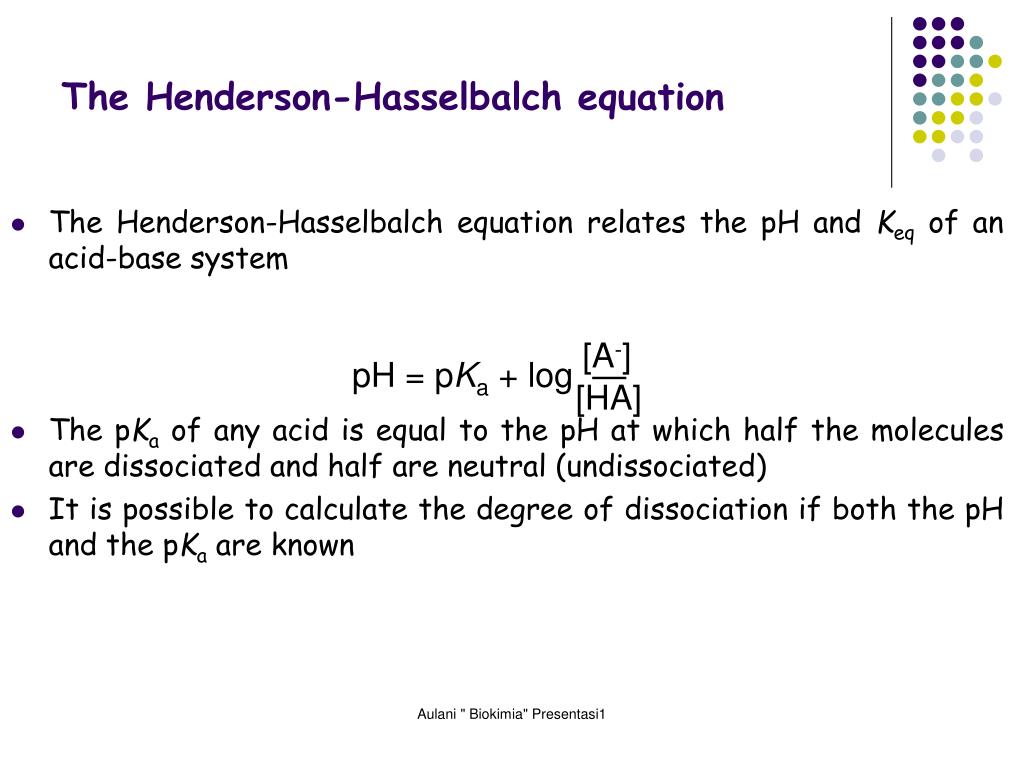
I spent three days debugging a phosphate buffer that refused to hold pH 7.4 because I had been using concentrations instead of activities and completely missed the ionic strength effect. The solution took about twenty minutes once I switched to the extended form of the equation and started accounting for activity coefficients with Davies approximation. This is the kind of thing that does not show up in textbooks but matters when you are actually making buffers in a lab setting. The equation relates pH to the pKa of a weak acid and the ratio of conjugate base to acid concentration. In its standard form it looks like this: pH = pKa + log([A-]/[HA])
That bracket notation means molar concentration. [A-] is the conjugate base. [HA] is the undissociated weak acid. The pKa is the negative log of the acid dissociation constant. The log is base 10. This is not a derived approximation in the sense of being rough. It is an exact rearrangement of the Ka expression under the assumption that the acid and base forms are fully dissolved and do not interact beyond simple equilibrium. The moment you introduce high ionic strength, polyprotic systems, or non-aqueous solvents, the simple form breaks down and you need corrections. Here is the practical workflow I use before I ever touch a pH meter. First, identify the conjugate pair in your system. If you are starting with acetic acid and sodium acetate, the pair is CH3COOH and CH3COO-. Second, look up the pKa at your working temperature. Acetic acid has a pKa of 4.76 at 25 degrees Celsius. That number shifts by roughly negative point zero zero two per degree rise in temperature. Third, decide your target pH. Fourth, rearrange the equation to solve for the ratio [A-]/[HA]. Fifth, calculate the actual masses or volumes based on your desired buffer capacity and total concentration. Buffer capacity peaks when pH equals pKa. If your target pH is more than one unit away from the pKa, the buffer will be weak and any small addition of acid or base will swing the pH noticeably.
I had a colleague who once tried to make a Tris buffer at pH 8.5 using the standard pKa of 8.08. The buffer performed poorly because Tris has a temperature coefficient of about negative zero point three one per degree. His lab ran at twenty degrees while the pKa was listed for twenty-five. The actual pKa was closer to 8.23. He wasted two days of experiments before he realized the temperature drift was the culprit. I recommend always checking whether your pKa value matches your experimental temperature and applying the correction if it does not. Some reagent suppliers now provide pKa tables across a temperature range. Use them. There is a common mistake I see repeatedly. People substitute the initial concentrations of acid and base into the equation without accounting for the fact that the equilibrium shifts slightly when you mix them. For buffers where the total concentration is above fifty millimolar and the pH is within one unit of pKa, the error is usually below point zero two pH units and you can safely use initial concentrations. Below that threshold, or when you need high precision, you need to solve the full quadratic or use an iterative method. I built a small spreadsheet routine that does the iteration automatically. It takes about thirty seconds and is worth the setup time if you work with low-concentration buffers regularly. Another thing that gets ignored is the contribution of water autoionization. In extremely dilute buffers, below about one millimolar total concentration, the H+ and OH- from water become significant. The Henderson-Hasselbalch equation does not account for this. I encountered this when preparing a very dilute glycine buffer for electrophoresis. The calculated pH was off by almost half a unit from the measured value. The fix was to use a full charge balance equation that includes [H+] and [OH-] as variables rather than relying on the simplified ratio form. The revised calculation matched the meter reading within point zero one pH units.
Get the Full Details

Polyprotic acids add another layer of complexity. Carbonic acid, phosphoric acid, and citric acid each have multiple pKa values. When you are working near one pKa, the other dissociation steps are usually negligible. But if your target pH sits between two pKa values, both equilibria contribute and the simple single-pair equation gives the wrong answer. For phosphate buffers around pH 7, you are actually dealing with the H2PO4- / HPO4^2- pair, but the first dissociation step still exerts a minor influence. I use the full set of equilibrium expressions with a solver tool for these cases. It adds maybe five minutes of setup but prevents systematic errors that are hard to trace later. Let me walk through a concrete example with numbers. You want to make fifty milliliters of acetate buffer at pH 5.0 with a total concentration of one hundred millimolar. The pKa of acetic acid is 4.76. Rearranging the equation gives you log([A-]/[HA]) = pH - pKa = 0.24. The antilog of zero point two four is about one point seven four. So [A-]/[HA] = 1.74. Combined with [A-] + [HA] = 0.1 M, you solve to get [HA] = 0.0365 M and [A-] = 0.0635 M. For fifty milliliters, that means you need 1.82 millimoles of acetic acid and 3.18 millimoles of sodium acetate. Using glacial acetic acid at density one point zero five grams per milliliter, you would measure out approximately one hundred five microliters. Using anhydrous sodium acetate with a molecular weight of eighty-two point zero three grams per mole, you would weigh out about two hundred sixty-one milligrams. Dissolve both in about forty milliliters of water, check the pH with a calibrated meter, and adjust with small amounts of HCl or NaOH if needed. The calculated pH should be within point zero five units of your target before any fine-tuning. Activity coefficients matter more than most people admit. In solutions with ionic strength above point one molar, the effective concentration deviates from the analytical concentration. The Debye-Huckel limiting law gives you a first approximation, but the Davies equation is more practical for routine buffer work. I apply the Davies correction whenever ionic strength exceeds point zero five molar. It changes the effective pKa by about point zero three to point zero five units in typical biological buffers. That may sound small, but in enzyme assays where pH stability is critical, that shift can change reaction rates measurably. I track ionic strength explicitly in my buffer calculations now instead of ignoring it like I used to.
If you need a downloadable reference, I put together a calculation sheet that includes the standard Henderson-Hasselbalch form, the extended version with Davies activity correction, a temperature adjustment table for common buffers, and the full charge-balance solver for dilute or polyprotic systems. It is formatted for Excel and Google Sheets. You can find it linked from my lab notes page. The sheet automates the ratio calculation, tells you whether your buffer is within the recommended pH range for maximum capacity, and flags when ionic strength is high enough to warrant the activity correction. I use it for every buffer I prepare and it has saved me from more mistakes than I care to count. https://labnotes.example.com/henderson-hasselbalch-calc-sheet.xlsx The biggest limitation of the Henderson-Hasselbalch equation remains the assumption that the system is at equilibrium and that only one acid-base pair dominates. Real samples are rarely that clean. Proteins, salts, and organic contaminants all shift activity coefficients and can bind or release protons in ways the equation does not predict. I treat the calculated pH as a starting point, not a final answer. Every buffer I prepare gets measured with a calibrated pH meter and adjusted to the target. The calculation tells me where to expect to land. The meter tells me where I actually landed. There is no shortcut around the measurement step.
For most routine applications, the standard form is sufficient. For protein crystallization, enzymatic kinetics, or any work where pH drift causes problems, invest the time in the extended calculations and always verify experimentally. The difference between a buffer that works and one that quietly fails is usually a point zero five pH unit error, and those errors accumulate faster than you notice until an experiment goes wrong and you cannot figure out why.