
Why bonding theory still trips people up in practice
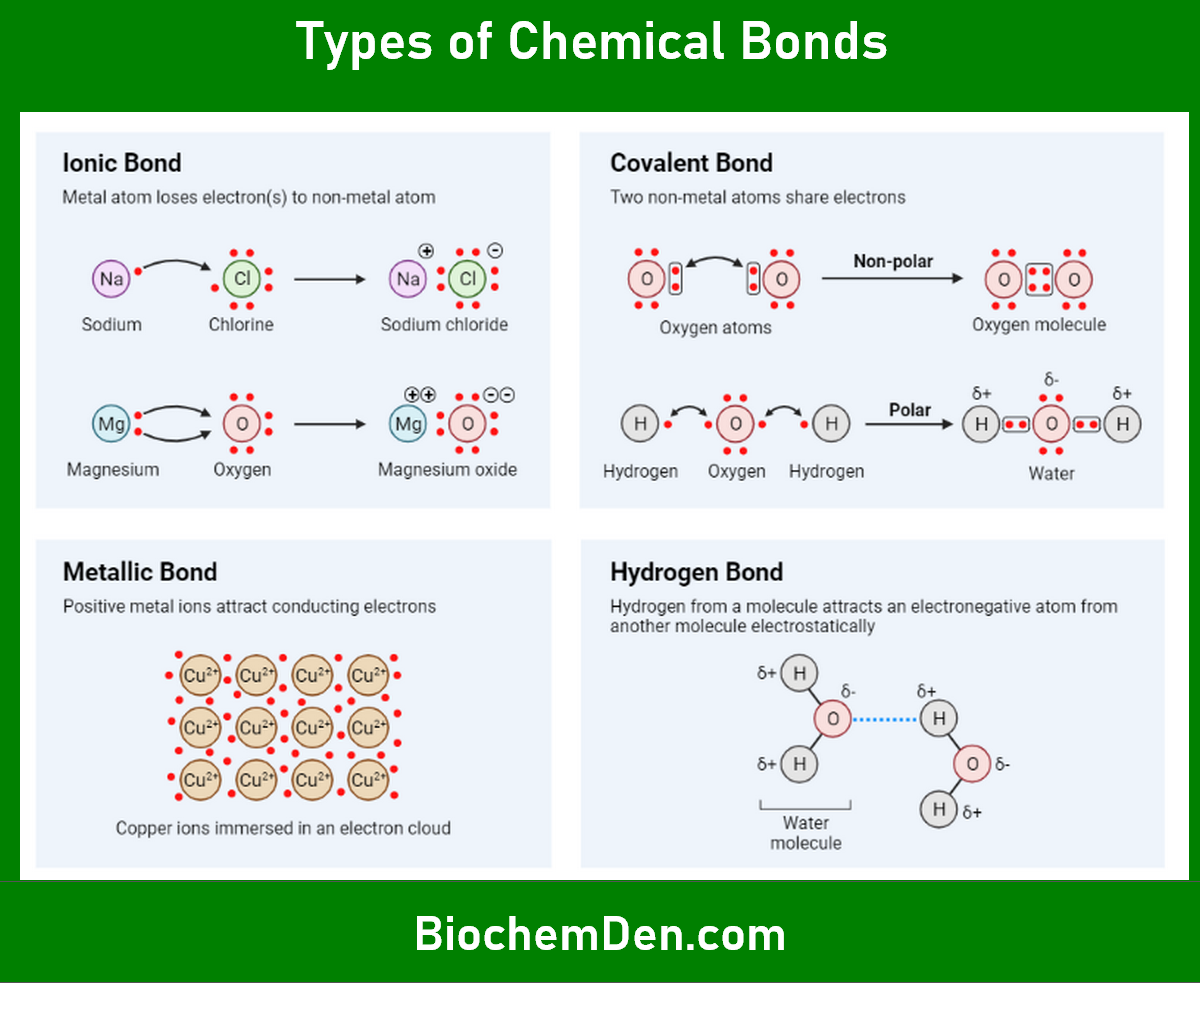
Chemical bonding isn't as clean as the textbooks make it look. The standard model covers ionic, covalent, and metallic bonds. But in the lab or when working with real molecules, things rarely fall neatly into those categories. You get partial charges, borderline cases, and interactions that shift depending on your computational method or measurement technique. At its core, bonding is about electrons redistributing themselves to lower the total energy of a system. Nuclei attract electrons. Electrons repel each other. Nuclei repel each other. The equilibrium distance between atoms is where those competing forces balance out. That's it. Everything else is interpretation. I spent years working with organometallic complexes where the textbook definitions kept breaking down. We had a palladium compound with a bridging ligand that showed up as covalent by X-ray diffraction but behaved like an ionic interaction in solution. The bond lengths were ambiguous. The computational results varied wildly depending on whether we used DFT or wavefunction-based methods. What actually helped was checking the electron density topology using QTAIM analysis, which gives you a more objective picture of where bonds actually exist versus where they're just assumed to exist.
How to approach bonding without getting lost in theory
Start with what you can measure. Bond lengths from crystallography or spectroscopy. Bond angles from geometry optimization. Ionization energies from photoelectron spectroscopy. These are facts. The interpretation layer comes after. When predicting bond character, use electronegativity differences as a first filter, but don't stop there. Pauling scales are useful approximations. They fail when d-orbitals are involved, when back-bonding occurs, or when the system is highly polarized. A bond between carbon and chlorine has an electronegativity difference of about 0.5 on the Pauling scale. That suggests covalent. But the bond is clearly polar. The dipole moment doesn't lie. Here's something beginners consistently miss: bond strength doesn't always correlate with bond order. A single bond can be stronger than a double bond if the atoms are small and the orbitals overlap well. Silicon-silicon single bonds are weaker than expected because silicon's 3p orbitals are diffuse. That's why disilanes are reactive even though they're technically saturated.
Common pitfalls that waste time
Assuming VSEPR predicts actual molecular geometry in transition metal complexes. It doesn't. Crystal field effects and ligand field stabilization dominate. A tetrahedral complex might distort to square planar depending on the metal's d-electron count and the ligand field strength. I've seen people spend days trying to rationalize geometries with VSEPR before realizing the ligand field model was the right tool. Another trap: treating resonance structures as real. They're not. They're a drawing convention. The actual molecule is a hybrid. When you're calculating properties, you're working with one electronic state, not a mixture of your drawings. This matters when you're interpreting UV-Vis spectra or computational output. The oscillator strength comes from the actual electron density distribution, not from whichever resonance contributor looks most reasonable on paper. Orbital hybridization is similarly a model, not a physical reality. sp3 hybridization explains methane's tetrahedral geometry well. But it's computationally expensive to justify in larger systems, and modern methods often skip it entirely. Natural bond orbital analysis gives you a different view that sometimes aligns better with what's actually happening electronically. Neither is wrong. Both are approximations.
Get the Full Details
When standard bonding models completely fail
Three-center two-electron bonds are the classic example. Diborane B2H6 has bridging hydrogens that don't fit any simple bonding category. The terminal B-H bonds are normal two-center covalent bonds. The bridging ones involve two boron atoms sharing two electrons across three centers. Lewis structures can't represent this without drawing impossible formal charges or violating the octet rule in ways that feel wrong. Aromaticity in non-planar systems also breaks conventional models. Twistaranes and helicenes show aromatic character despite significant geometric distortion. The Hückel rule works for planar rings. It becomes questionable when the pi system is forced out of plane. Computational magnetic criteria like NICS values help, but even those have limitations depending on where you place the probe point. Agostic interactions in organometallic chemistry are another edge case. A C-H bond interacts with an empty metal orbital without fully breaking. The hydrogen sits between carbon and metal. Is it a bond? Is it an interaction? The answer depends on your definition. X-ray structures show elongated C-H distances. NMR shows characteristic shifts. Computational studies show bonding orbitals with mixed character. All of it is consistent. None of it fits neatly into ionic or covalent.
Practical workflow for analyzing bonding in unfamiliar systems
Get the geometry first. Optimized structure from computation or experimental data. Check for unreasonable bond lengths or angles. If something looks off, the geometry optimization might have converged to a local minimum instead of the global one. Run a frequency calculation to confirm it's a true minimum. Then calculate the electron density. QTAIM gives you bond critical points and bond paths. These are topological features of the electron density, not assumptions. If a bond path exists between two atoms, there's a bonding interaction. If it doesn't, the interaction is weak or negligible. This is more objective than judging by bond length alone, though bond length correlates reasonably well in most cases. Population analysis helps too. Mulliken populations are basis-set dependent and unreliable. NBO or Atoms in Molecules charges are better. Look at the charge distribution. If atoms are nearly neutral, the bond is likely covalent. If there's significant charge separation, ionic character is present. The percentage ionic character can be estimated from the dipole moment relative to the theoretical maximum for a full charge transfer.
For metals, check the d-orbital occupancy and ligand field splitting. The 18-electron rule works for many organometallics but fails for early transition metals and lanthanides. Don't force it. Use crystal field theory or ligand field theory instead. The color of the complex often tells you the splitting energy directly. A blue complex has a different d-d transition energy than a red one. That energy difference maps to the ligand field strength.
The hard truth about bonding theory
Every bonding model is an approximation. Lewis structures ignore electron correlation. Molecular orbital theory assumes a mean-field approximation. Density functional theory depends on the functional you choose. No single model captures everything. The best approach is to use multiple methods and look for convergence in your conclusions. When different approaches agree, you're probably onto something real. When they disagree, dig deeper instead of picking a favorite. We don't actually know what a bond "is" in the fundamental sense. We know how electrons behave under certain conditions. We know what observables we can measure. The rest is model-building. That's not a weakness. It's just honest science.


