Understanding Activation Energy and the Arrhenius Equation
The Arrhenius equation is k = A × e^(-Ea/RT). That's it. It relates reaction rate constants to temperature and activation energy. Most people memorize the formula, plug numbers into a calculator, and move on. That's fine for homework. It falls apart the moment you're actually running reactions and things don't go as predicted. I spent years working in process chemistry before moving into consulting. The first time this equation bit me, I was scaling up a nitration reaction. The lab batch went smoothly at 0°C. The plant scale-up required cooling to -10°C for safety reasons. According to the Arrhenius parameters I'd calculated, the rate should have been about 40% slower. It turned out the rate was roughly 10% of what I expected. I'd missed something fundamental about how pre-exponential factors behave when you change solvent conditions at scale. It took me three weeks and a lot of ego damage to figure out that the A factor isn't actually constant across different temperature ranges and solvent environments. It shifts. People don't tell you that in undergrad.
The Energy Of Activation Equation Unlocking The Secrets Of Chemical Reactions
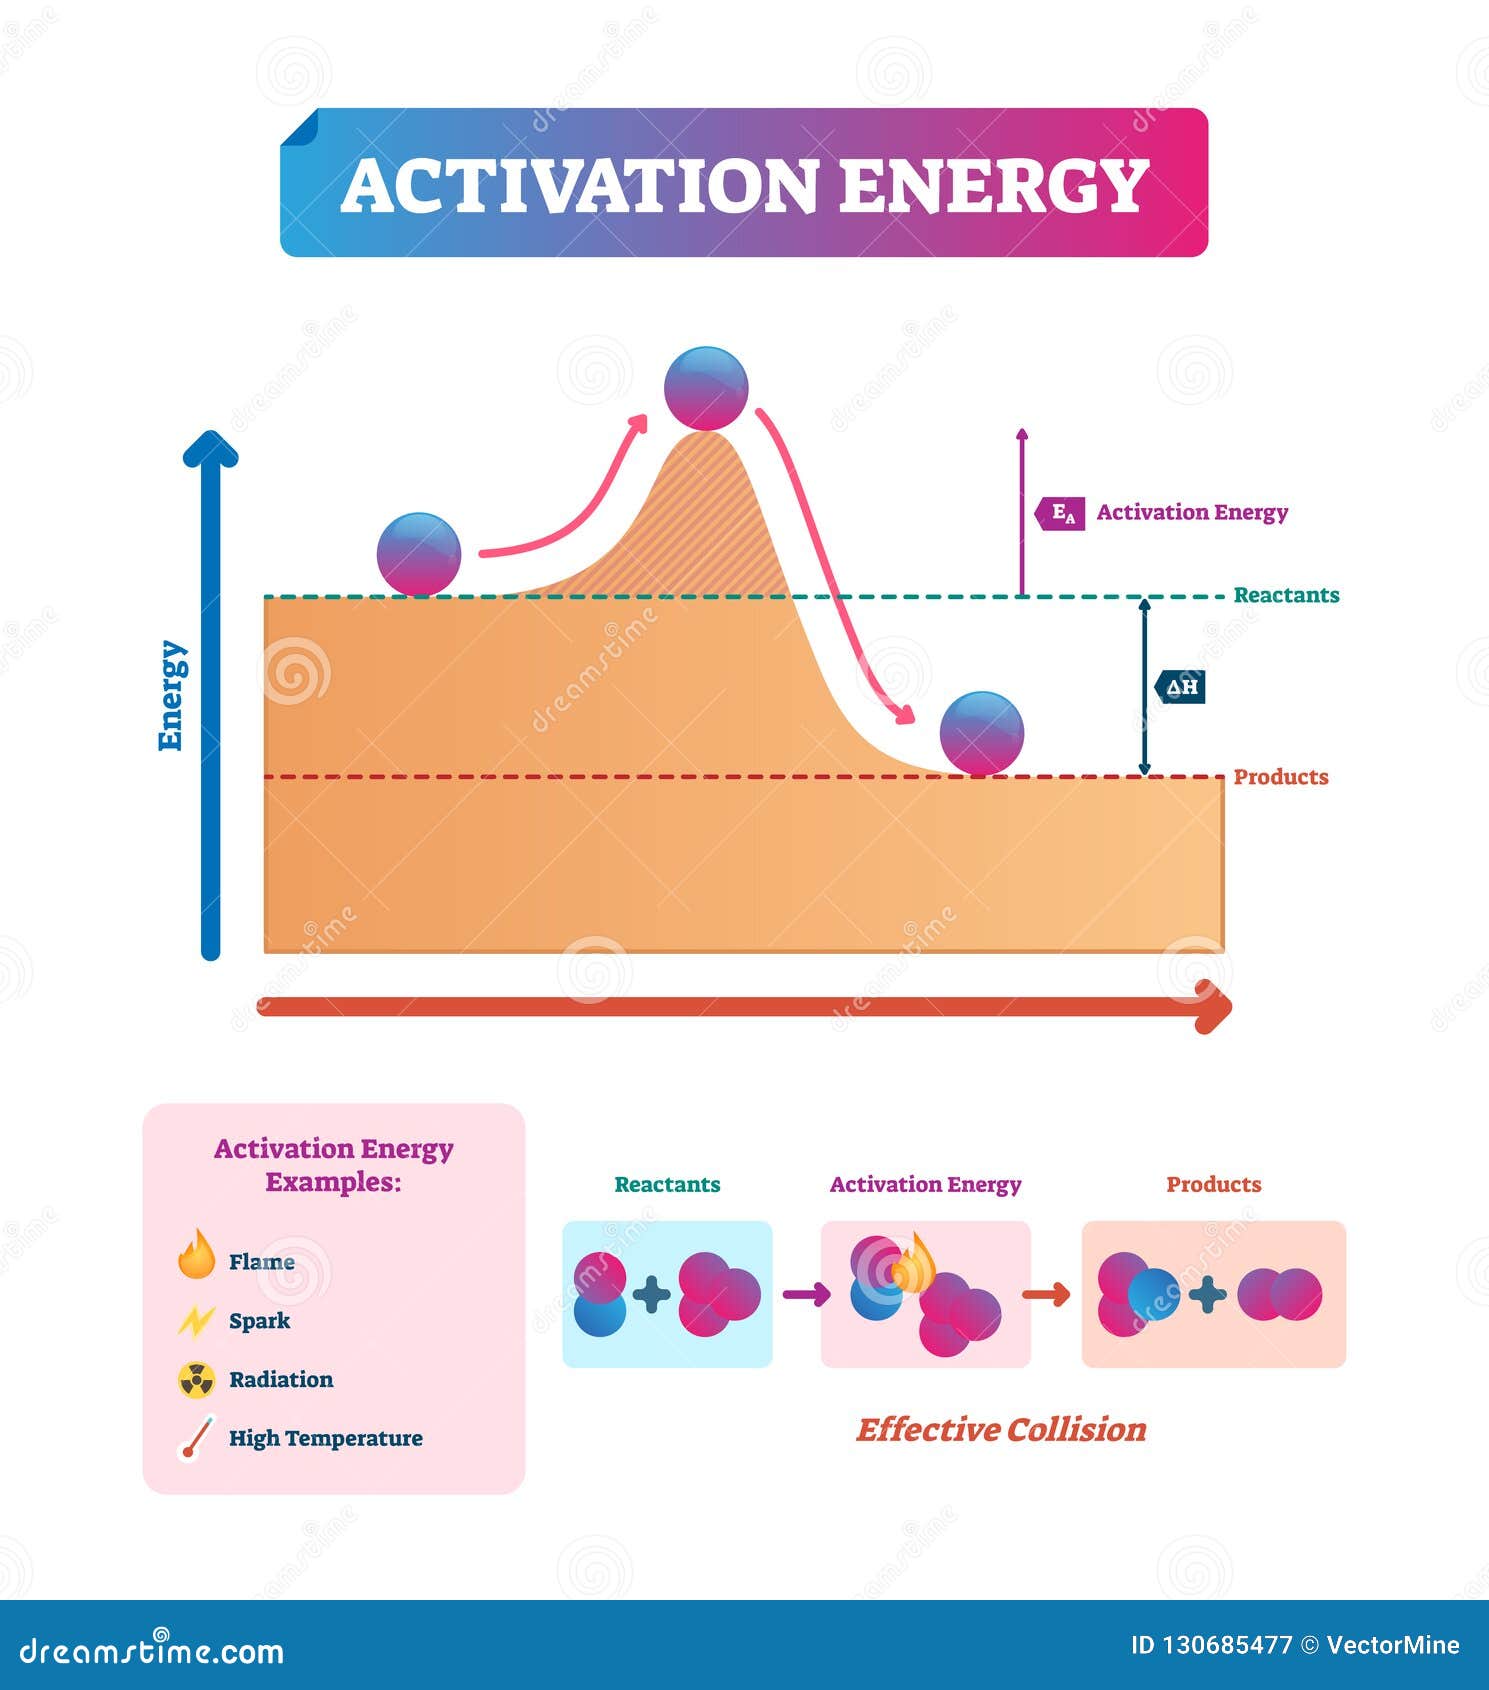
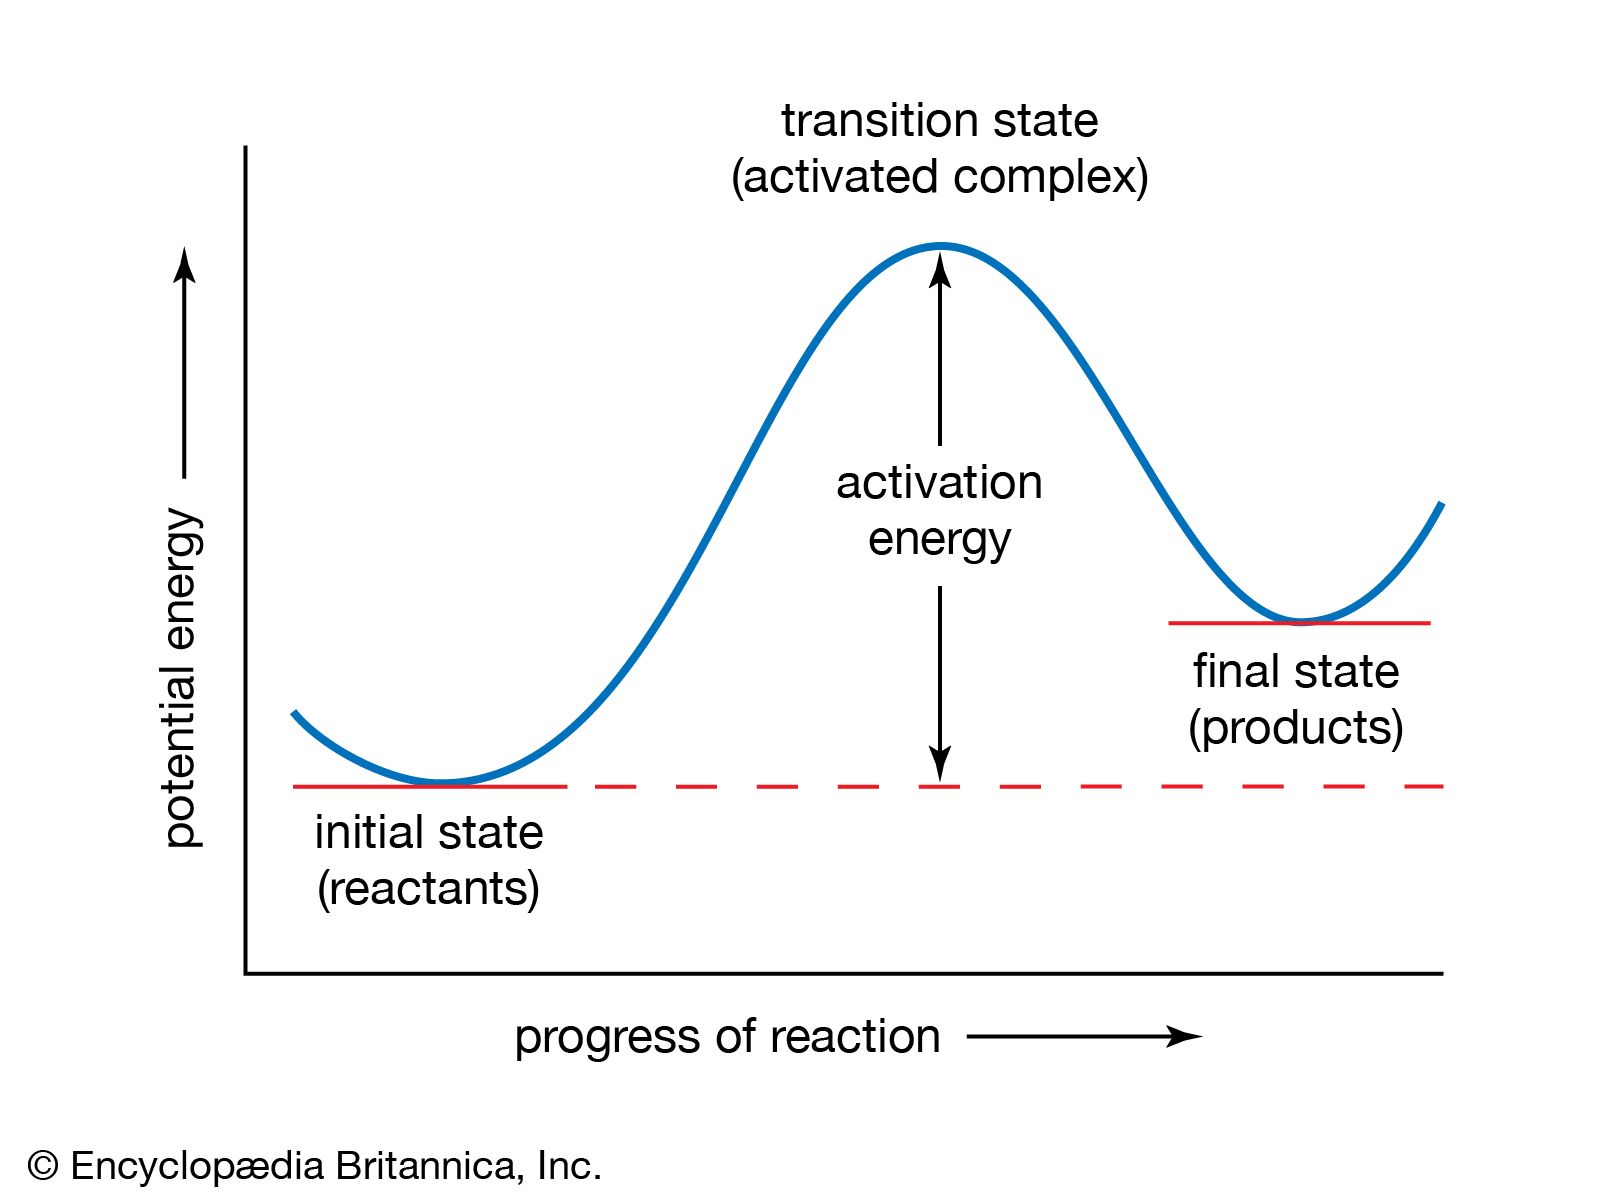
Activation energy, Ea, is the minimum energy barrier reactants must overcome to form products. The pre-exponential factor A represents collision frequency and orientation. R is the gas constant, 8.314 J/(mol·K). T is temperature in Kelvin. The exponential term e^(-Ea/RT) gives the fraction of molecules with enough energy to surmount the barrier at a given temperature. Here's the part most guides skip. To find Ea experimentally, you run the reaction at multiple temperatures and plot ln(k) versus 1/T. The slope equals -Ea/R. You need at least four data points spanning a reasonable temperature range. Three points won't cut it because one bad measurement throws everything off. I typically run reactions at five temperatures, usually covering a 30-40 degree range. Anything less and the uncertainty in Ea becomes unusable. The linearized form of the equation is ln(k) = ln(A) - Ea/(RT). This is your working equation for data analysis. When you plot it, the y-intercept gives you ln(A). From there you exponentiate to get A. Simple on paper. In practice, experimental error in k values propagates through the exponential calculation, and small temperature measurement errors become significant when you're dealing with 1/T plots.
A counter-intuitive thing about activation energy: a higher Ea doesn't always mean a slower reaction. If the pre-exponential factor A is sufficiently large, a reaction with high activation energy can still proceed rapidly at elevated temperatures. I've seen this in catalytic processes where the catalyst dramatically increases A even though Ea remains substantial. Beginners fixate on Ea alone and miss the full picture. Another common pitfall. The Arrhenius equation assumes a single, well-defined activation energy barrier. Real reactions often involve multiple steps with competing pathways. At different temperatures, the dominant pathway can shift. You'll get an apparently curved Arrhenius plot, which means your "Ea" is actually an effective value that changes with temperature. When this happens, the simple model breaks down and you need to consider mechanisms involving parallel or consecutive elementary steps. Practical note on units. Ea is typically reported in kJ/mol. Make sure your R value matches. If Ea is in J/mol, use R = 8.314. If Ea is in kJ/mol, convert to J/mol first or use R = 0.008314 kJ/(mol·K). Mixing these up is the most common arithmetic error I see. It produces results that are off by a factor of 1000, which is easy to miss if you're just glancing at the final number.
Get the Full Details

Temperature sensitivity is where this equation matters most in practice. A rule of thumb many people cite is that reaction rates double for every 10°C increase. That's roughly correct for reactions with Ea around 50 kJ/mol near room temperature. But for reactions with Ea of 100 kJ/mol, the rate can quadruple over the same temperature range. For Ea of 20 kJ/mol, it might only increase by 30%. The rule of thumb is useful for quick estimates but dangerous for anything requiring precision. When I'm designing a process and need to predict behavior at temperatures outside my experimental range, I always flag the uncertainty. Extrapolating Arrhenius parameters beyond your measured range is risky. The equation works well within the temperature window you tested. Beyond that, you're guessing. I typically limit extrapolation to within 20-30 degrees of my highest measured temperature. Beyond that, I run additional experiments rather than trust the model blindly. The equation also has limitations with enzymatic and diffusion-controlled reactions. Enzymes denature at high temperatures, creating a bell-shaped rate curve that the Arrhenius equation cannot describe. Diffusion-controlled reactions show much weaker temperature dependence because the rate is limited by molecular mobility, not by overcoming an energy barrier. In those cases, the Stokes-Einstein relationship or transition state theory provides better frameworks.
For solid-phase reactions and heterogeneous catalysis, the concept of effective activation energy becomes complicated. Mass transfer limitations, surface coverage effects, and pore diffusion all introduce temperature dependencies that aren't captured by the simple Arrhenius form. I've seen engineers apply the equation to fixed-bed reactor data without accounting for external mass transfer resistance, then wonder why their calculated Ea values were negative. A negative apparent activation energy is usually a red flag that mass transfer, not surface kinetics, is controlling the observed rate. If you're working with this equation and want to verify your calculations, most computational chemistry packages can generate rate constants from transition state theory calculations. These provide a theoretical Ea and A factor that you can compare against experimental data. The agreement is usually within a factor of 2-5 for well-behaved systems. Larger discrepancies suggest that your assumed mechanism or the transition state structure needs revision. I keep an Excel template for Arrhenius analysis that handles the ln(k) versus 1/T regression automatically, including uncertainty bounds on Ea and A. It's straightforward to set up and saves time when you're processing multiple reaction datasets. The built-in solver functions in most chemistry software packages can do the same regression, but they don't always give you easy access to the confidence intervals on your parameters, which matters when you're reporting results or making scale-up decisions.
The bottom line is that the Arrhenius equation is a tool, not a law. It describes a large class of reactions well enough for most practical purposes. It fails when the assumptions behind it break down, and recognizing those failure modes is what separates someone who can use the equation from someone who just knows how to use a calculator.