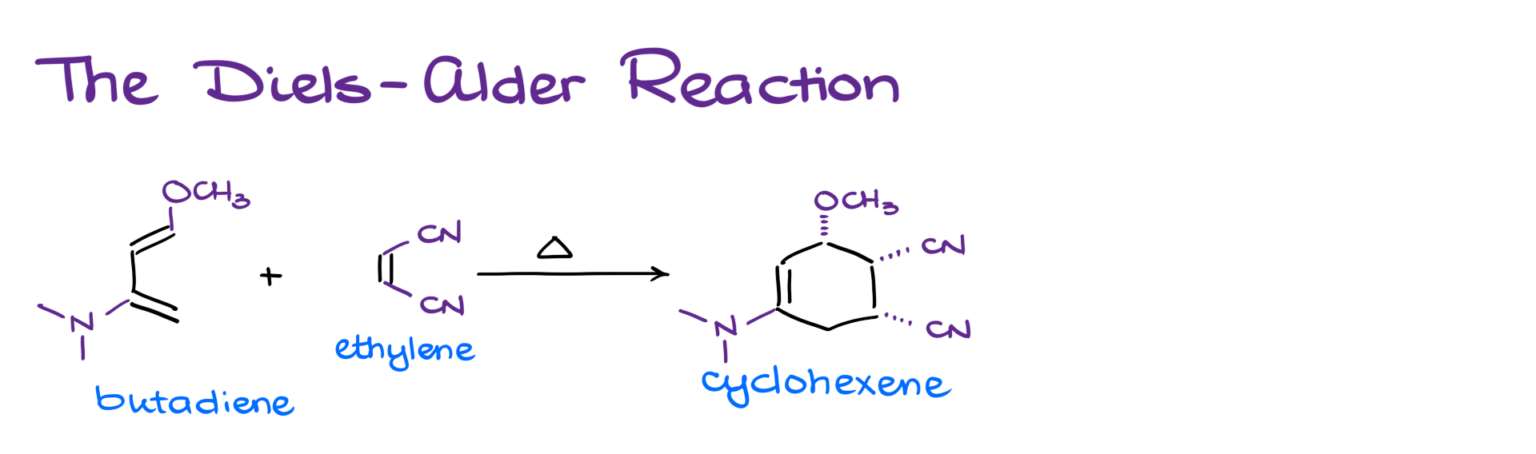
What actually happens when you run a Diels-Alder
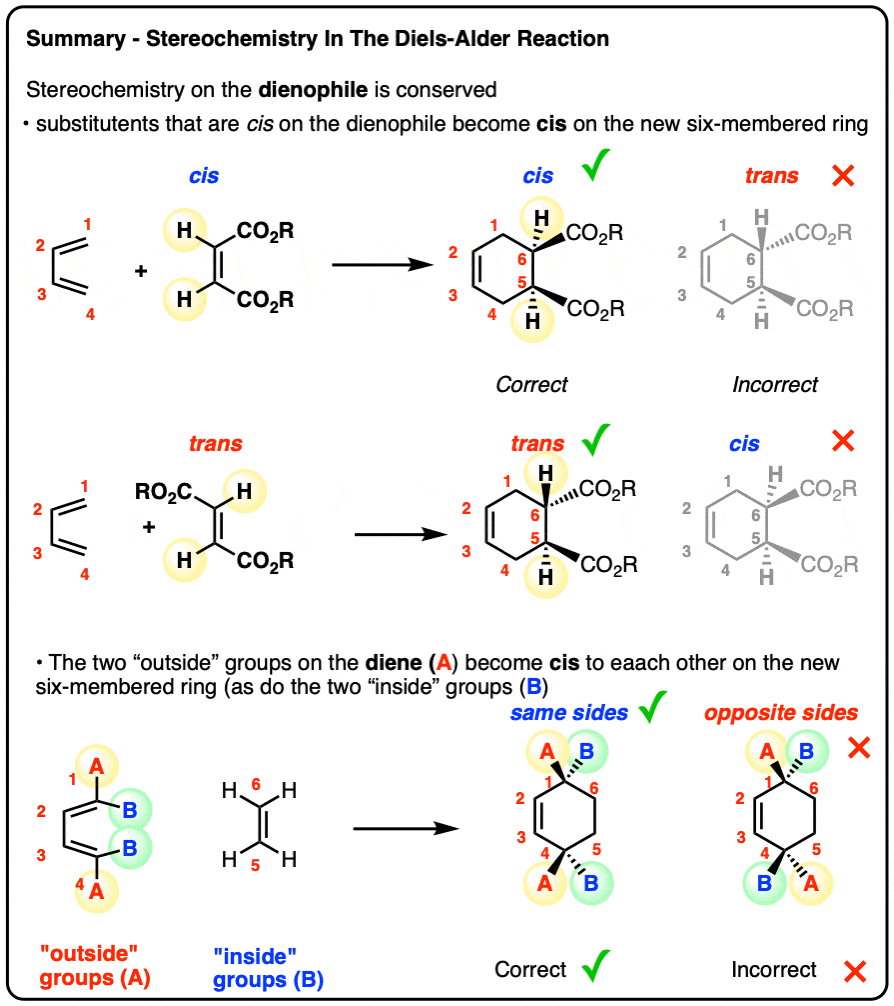
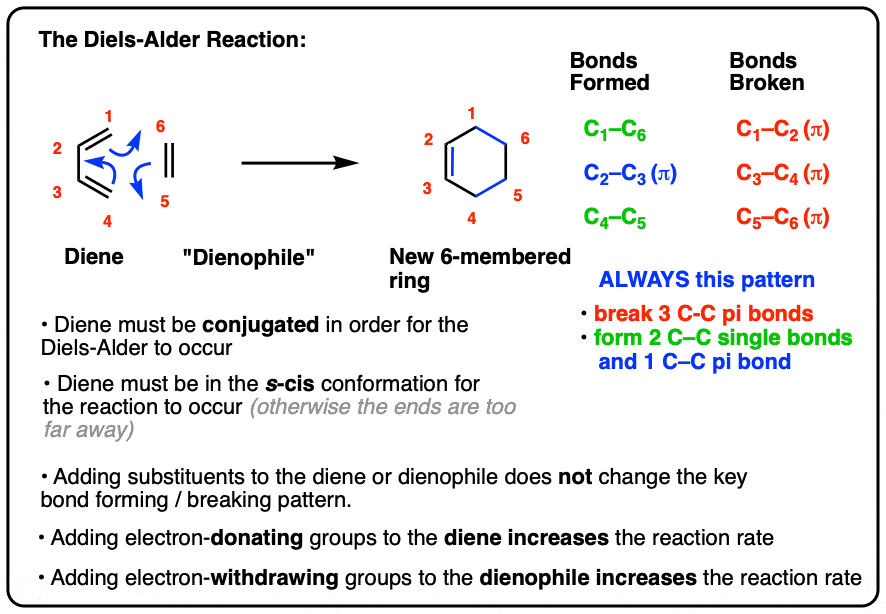
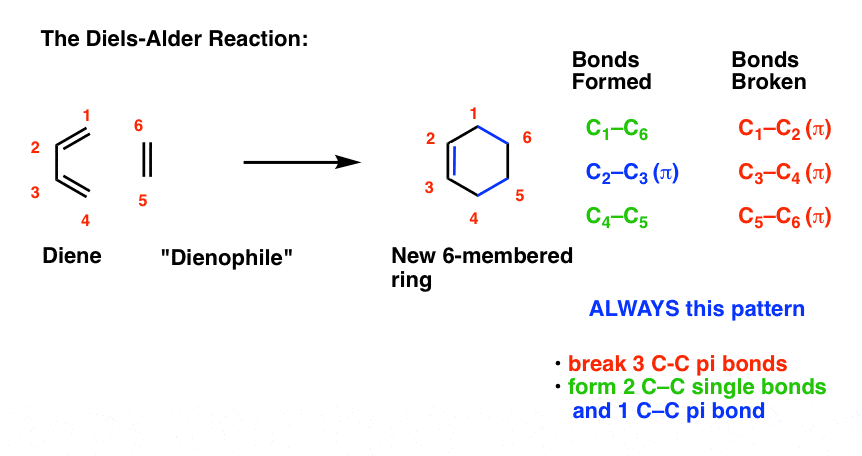
It is a [extract_itex][4+2][/extract_itex] cycloaddition. The diene contributes four pi electrons, the dienophile contributes two, and a six-membered ring forms in a single step without any intermediates. You mix them together, apply heat or pressure, and a cyclohexene appears. That is the entire scope of it at its most basic level. The reaction is stereospecific, which means the geometry of the dienophile is preserved in the product. If you start with a cis-disubstituted alkene, the substituents stay cis in the ring. Trans stays trans. The diene also has to be in the s-cis conformation to react, and that is why 1,3-cyclopentadiene is such a common partner—it is locked in the right shape already.
The Dielsalder Reaction A Concerted Reaction Expediting Organic Synthesis
I learned this reaction in undergraduate organic chemistry and then forgot most of it until I needed to use it three years later on a total synthesis project. That gap between knowing it exists and actually running it successfully is wider than most students realize. The Diels-Alder reaction builds complexity fast. In one step you can install up to four stereocenters, two new carbon-carbon bonds, and a ring all at once. That is why it shows up in so many synthesis routes, including industrial processes for pharmaceutical intermediates and materials. The efficiency is real, but the conditions can be finicky depending on what you are working with.
Setting up the reaction in practice
You need a conjugated diene and an electron-poor dienophile for the standard version. Electron-withdrawing groups on the dienophile like esters, ketones, nitriles, or amides accelerate the reaction significantly. The normal electron demand Diels-Alder is by far the most common scenario you will encounter in the lab. Solvent choice matters less than people think. This reaction often proceeds fine in toluene, xylene, or even neat conditions. The diene and dienophile are usually nonpolar enough that a polar solvent adds complexity without much benefit. I have run successful reactions in toluene at 110 degrees Celsius under reflux, and I have also run identical reactions in chlorobenzene at 130 degrees. Both worked. The difference was mostly boiling point and how hot you could push it. When the substrates are unreactive, Lewis acid catalysis becomes useful. Diethyl aluminum chloride, boron trifluoride etherate, or indium triflate can lower the activation energy and allow the reaction to proceed at lower temperatures. Lower temperatures often mean better endo selectivity as well.
Get the Full Details
The endo rule and selectivity problems
The endo product is usually the kinetic product and forms faster because of secondary orbital interactions in the transition state. But that is not a guarantee. I ran a Diels-Alder reaction a few years back using cyclopentadiene and methyl acrylate with AlCl3 as a Lewis acid catalyst, and I got a roughly 60:40 endo-to-exo ratio instead of the 90:10 ratio the literature predicted. The issue turned out to be the stoichiometry of the Lewis acid. Using a full equivalent rather than the typical 10 to 20 mol percent shifted the selectivity back toward the expected range. It was a subtle thing and not something I caught from reading a textbook. The inverse electron demand Diels-Alder works differently. There the diene is electron-poor and the dienophile is electron-rich. Tetrazines reacting with strained alkenes like trans-cyclooctene fall into this category and react incredibly fast, sometimes at room temperature or below. These are the reactions that have taken over bioorthogonal chemistry in the last decade because they are fast and selective in complex biological environments.
A real problem I ran into
I was working on a synthesis that required a bulky aryl-substituted dienophile with a sensitive functional group. The standard thermal conditions degraded the product. Switching to a Lewis acid at low temperature created a new problem: the AlCl3 complexed too strongly with the ester and essentially shut down reactivity. I ended up using a milder Lewis acid, scandium triflate, at minus 78 degrees Celsius in dichloromethane. The reaction completed in about 45 minutes instead of the 12 hours it would have taken under thermal conditions, and the yield was around 82 percent after purification. The scandium system was more expensive per gram, but the time savings and cleaner reaction profile justified it. The regiochemistry of unsymmetrical dienes and dienophiles can also be unpredictable if you do not think through the electronic effects. The major product generally places the electron-donating group on the diene and the electron-withdrawing group on the dienophile in a 1,2 or 1,4 relationship depending on the substituents. You can predict it by looking at the largest coefficients in the frontier molecular orbitals, but doing that calculation by hand is error-prone. Running a quick DFT calculation or consulting a precomputed table of common diene-dienophile pairs saves time compared to letting the reaction run and then analyzing a messy product mixture by NMR.
Limitations you should know about
Not everything works with this reaction. Steric bulk around the reactive centers can shut it down entirely. A diene that cannot adopt the s-cis conformation will not react under normal conditions. (E,E)-1,4-diphenyl-1,3-butadiene reacts sluggishly because the phenyl groups prefer the s-trans geometry, and forcing it into s-cis costs conformational energy. Some dienes decompose before they react. Cyclopentadiene dimerizes over time through a retro Diels-Alder followed by re-polymerization. If you are using old stock, you need to crack the dimer by distillation before running your intended reaction. I once skipped that step because the bottle looked fresh, and my reaction gave me almost nothing but starting material recovered. The cyclopentadiene had been sitting in the drawer for two months. The retro Diels-Alder is also a real possibility at elevated temperatures. If your product is thermally unstable or if the forward reaction is reversible, you can lose yield simply by heating too long or too hot. Monitoring the reaction by thin-layer chromatography or in situ IR prevents overcooking, which is something I wish I had learned earlier in my career.
If the substrate is extremely unreactive and no amount of Lewis acid or temperature optimization helps, alternative approaches like stepwise annulation or radical cyclization might be more practical. The Diels-Alder is powerful, but it is not universal.
Workup and purification notes
Purifying Diels-Alder products depends entirely on what you used. Simple cases with volatile dienes like cyclopentadiene just need a rotary evaporation and a flash column. More complex substrates with multiple stereocenters and similar polarity to side products require careful chromatography or sometimes recrystallization. I typically run a small scale test reaction first to check the TLC pattern before committing a full gram of material. The test tells you whether the product is a single spot or a streak, and whether any unreacted starting material remains stubbornly on the plate. NMR analysis of the product is where the real information lives. The vinyl protons in the cyclohexene product show characteristic coupling patterns that confirm the regiochemistry and stereochemistry. Vicinal coupling constants of around 10 Hz indicate axial-axial relationships and help assign the relative stereochemistry without needing X-ray crystallography, though X-ray is always definitive when you can get a good crystal.