Concerted Doesn't Mean Everything Happens At The Same Instant
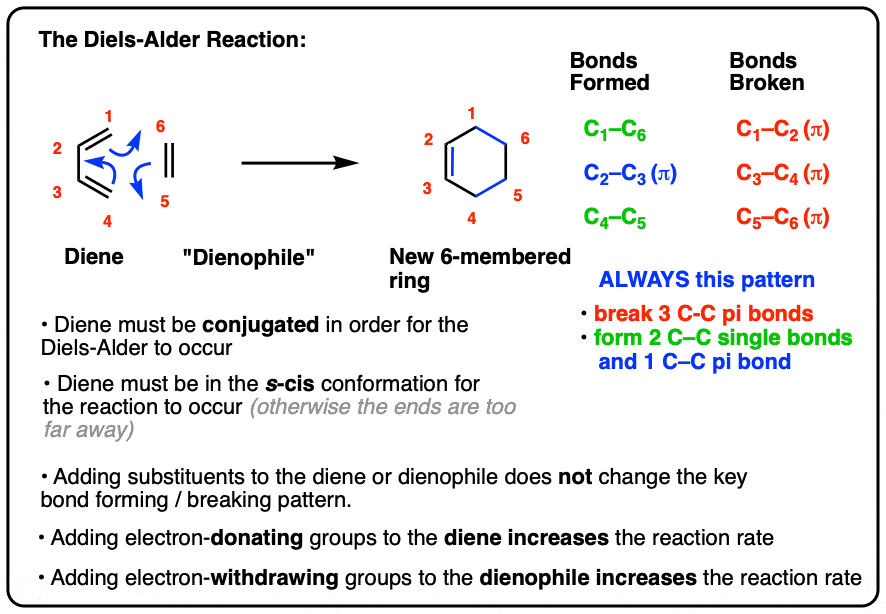
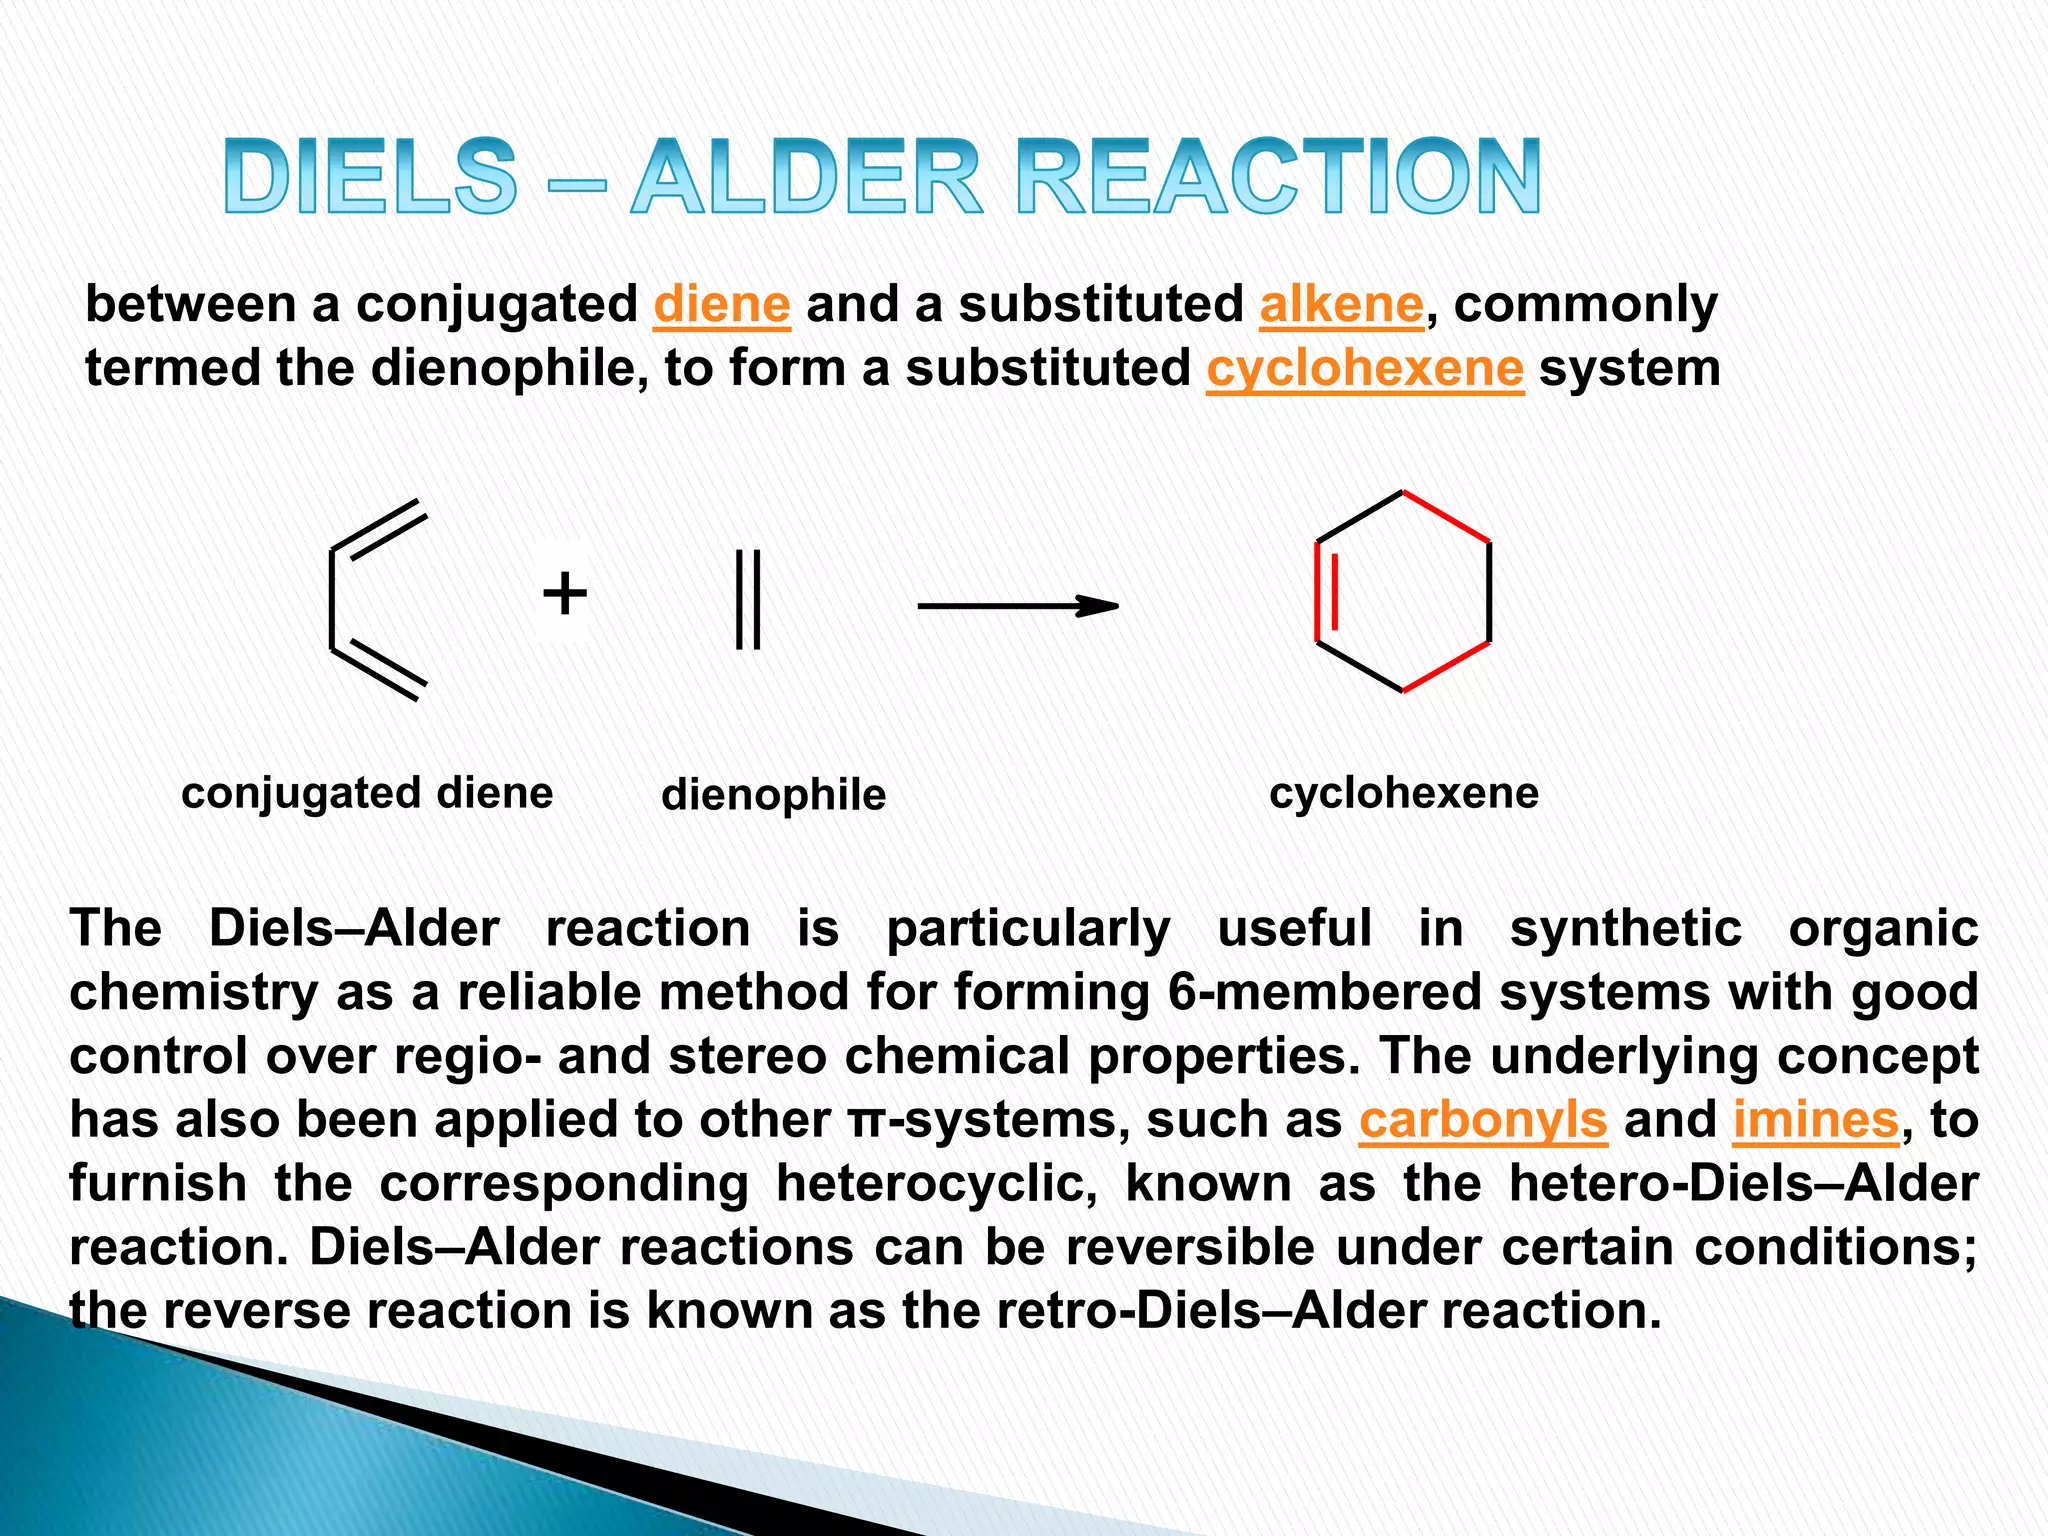
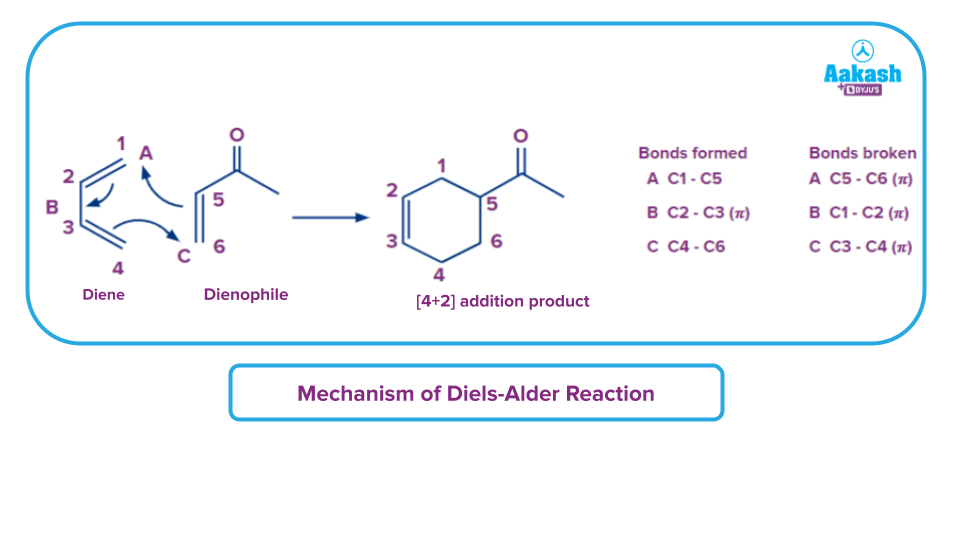
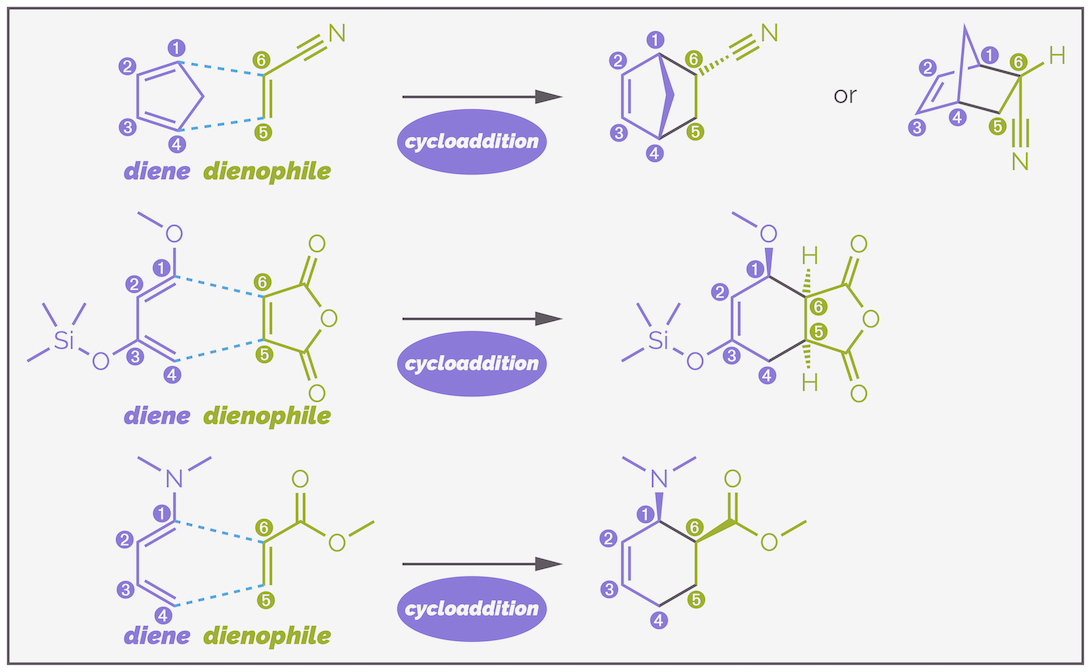
A lot of textbooks make it sound like every bond in a Diels-Alder is forming and breaking simultaneously, like a perfectly choreographed routine. It's not. Concerted just means there are no intermediates. No carbocations, no radicals, no discrete stepwise stage where one bond is done and the other is still waiting in the wings. The electrons flow through a single transition state and you get your product. That's the definition. How the electron density actually redistributes is messier than the diagrams suggest. In a standard intermolecular Diels-Alder, the diene contributes four pi electrons and the dienophile contributes two. These six electrons move through a cyclic six-membered transition state to form two new sigma bonds and one new pi bond inside the ring. That's it. Thermal conditions do the trick. You don't need light, you don't need a radical initiator, you just heat it or sometimes leave it at room temperature if the partners are reactive enough. Here's what most people miss: the new bonds don't form equally. With unsymmetrical dienes and dienophiles, one bond is further along in the transition state than the other. I spent a whole day trying to figure out why my cross-coupling between a 1-substituted butadiene and methyl acrylate gave a regioisomer ratio that didn't match the frontier molecular orbital prediction on paper. The issue wasn't that the reaction wasn't concerted. It was that the asynchronicity was high enough that the transition state looked more like a late-stage diradical to computational chemistry software. Once I switched to a more electron-deficient dienophile and ran it with a Lewis acid catalyst like trimethylaluminum in dichloromethane at 0 degrees Celsius, the selectivity snapped into place and I got the expected major isomer in about 87 percent yield after workup.
The term synchronous is used when both new bonds are forming at roughly the same rate. Asynchronous means one is ahead. Both are still concerted. Both are still pericyclic. The distinction matters because it affects your stereochemical outcomes and your regiochemistry, not whether the reaction works at all.
Orbital Symmetry Is The Real Constraint
The Woodward-Hoffmann rules govern whether a pericyclic pathway is allowed under thermal or photochemical conditions. For the Diels-Alder, you're looking at a suprafacial-suprafacial [4+2] process. Both the diene and dienophile interact on the same face of their pi systems. This is thermally allowed because the HOMO of the diene overlaps constructively with the LUMO of the dienophile in a continuous cyclic fashion. No node issues. No symmetry mismatch. If you tried the same reaction photochemically, you'd be promoting an electron and flipping the orbital symmetry, which would make the [4+2] pathway forbidden while opening up a [2+2] route instead. You don't need to memorize the full correlation diagram to use this. Just know that normal Diels-Alder runs on heat, not light. I once watched a grad student try to force a Diels-Alder at room temperature with a very unreactive ethylene dienophile and a plain 1,3-butadiene. Nothing happened for three days. He nearly scrapped the whole project. The reaction wasn't broken. The partners were just too. He switched to a dimethyl acetylenedicarboxylate dienophile and added a pinch of aluminum chloride, and the reaction completed in under two hours at the same temperature. The lesson is that while the mechanism is fixed, the kinetics are entirely dependent on your substituent choices.
Get the Full Details
Endo Versus Exo: Kinetic And Thermodynamic Control
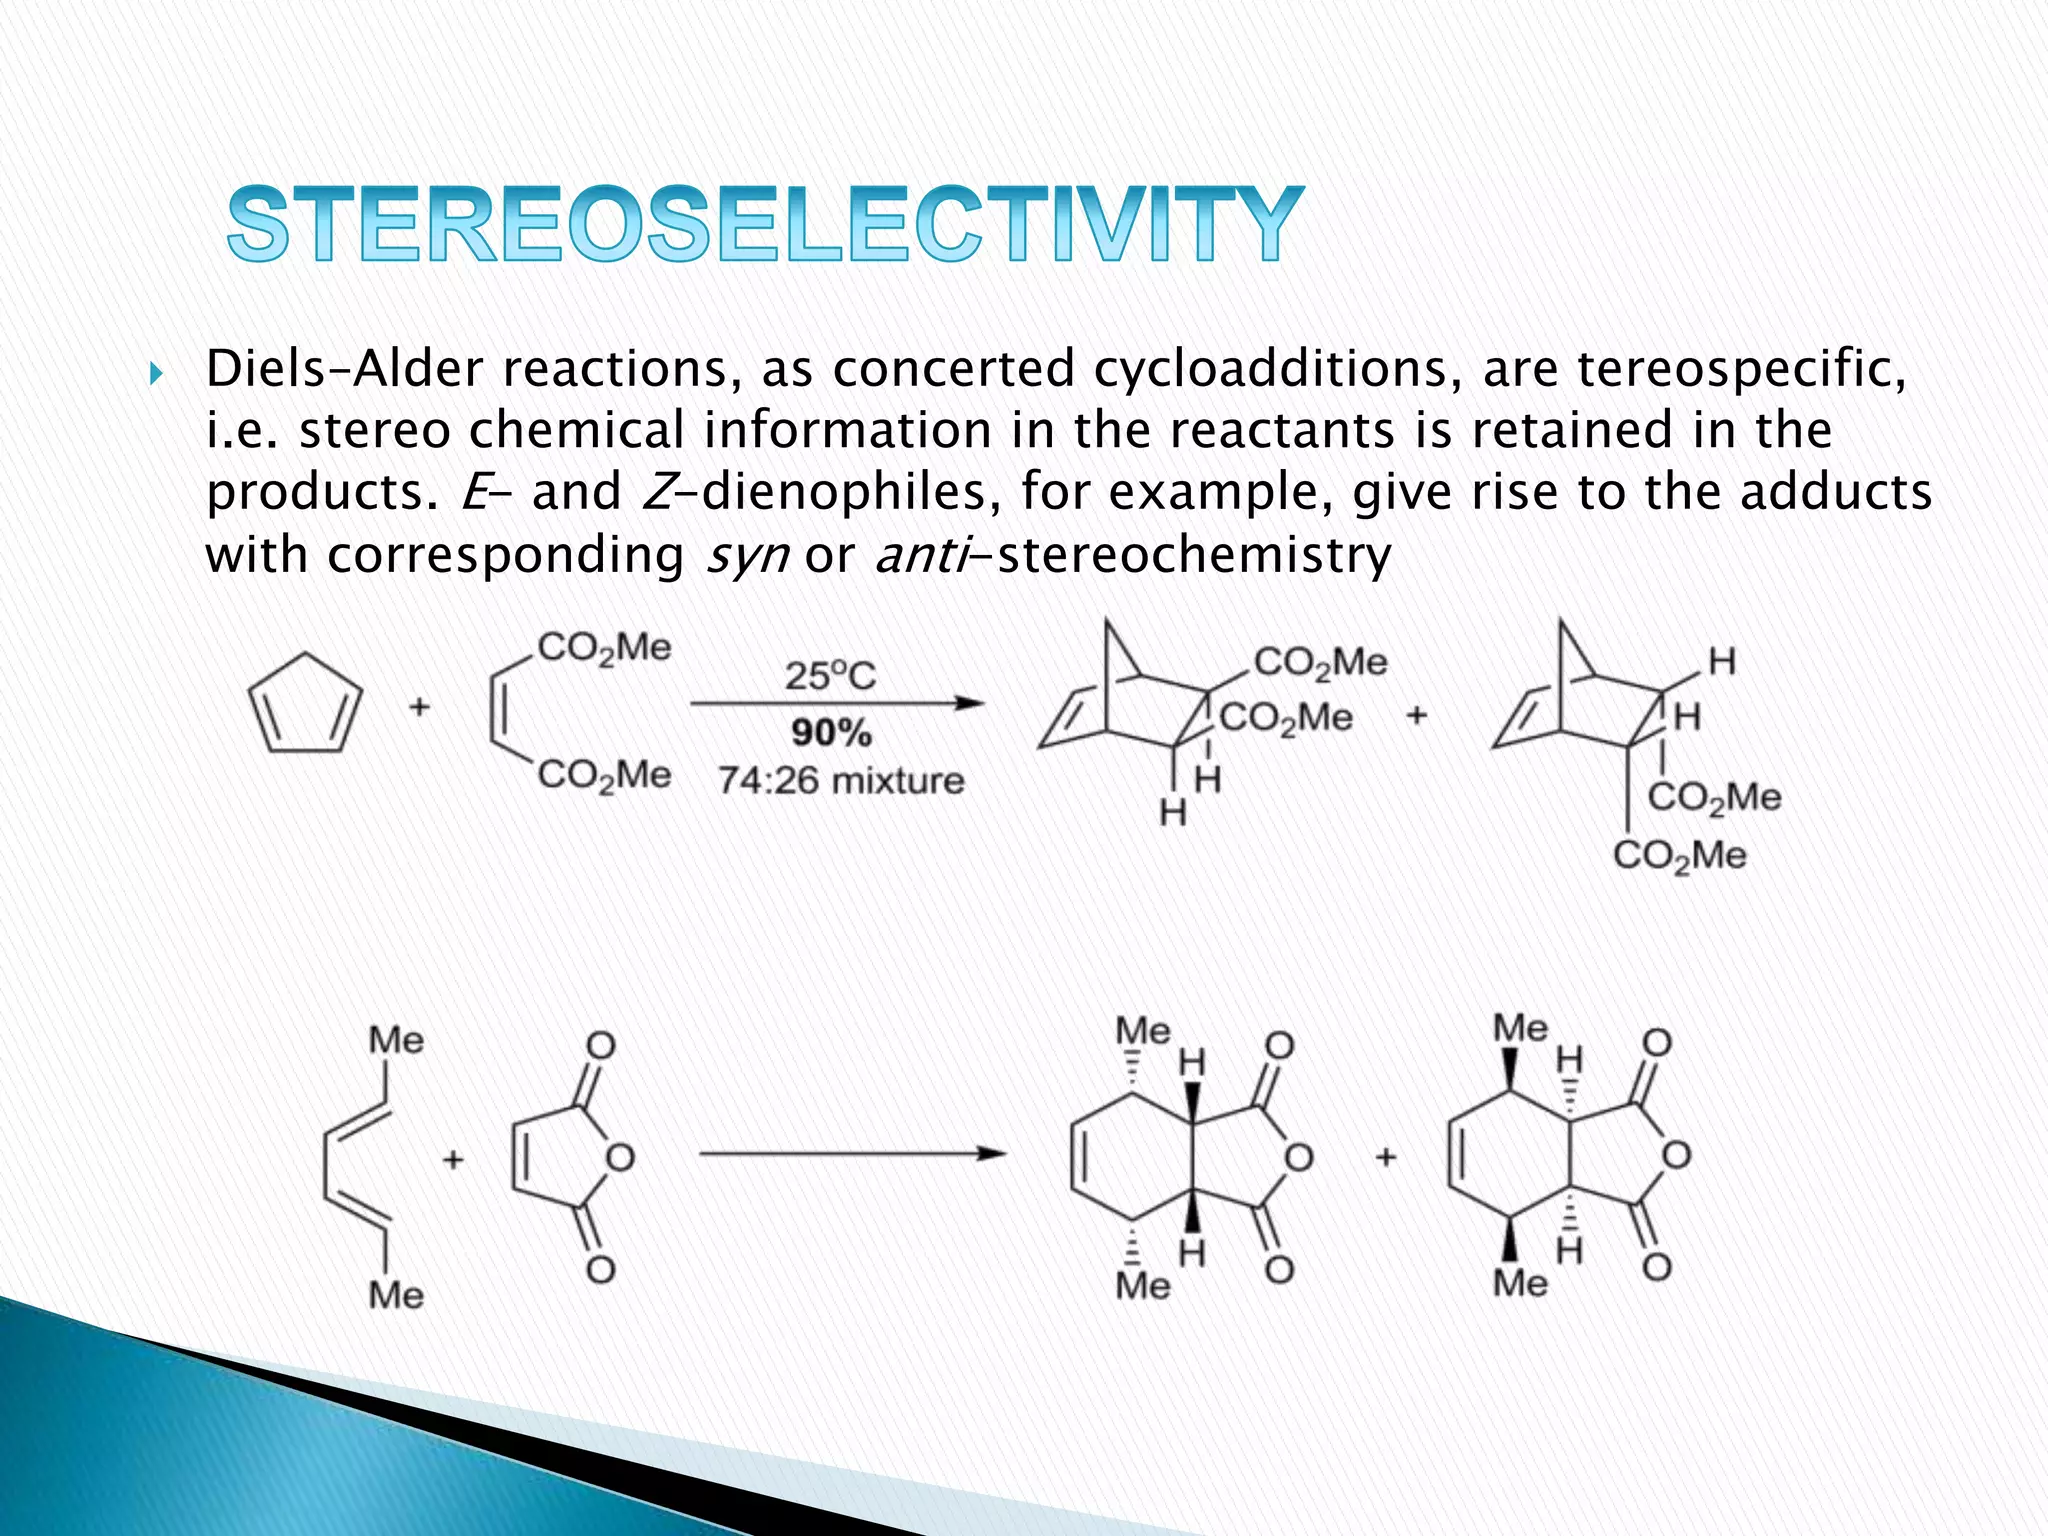
When the dienophile has a pi-bonded substituent like a carbonyl, you get two possible stereoisomers. The endo product places that substituent under the newly formed ring, while the exo places it away. The endo rule says you generally get the endo product under kinetic control because secondary orbital interactions between the substituent's pi system and the developing pi bond in the transition state lower the activation energy. This is real. I've seen endo selectivities above 95 to 5 with cyclic dienophiles like cyclopentadiene and maleic anhydride. But here's the catch that trips people up: the endo product isn't always the thermodynamic product. In some cases, especially with bulky substituents or when you heat the reaction long enough, the exo isomer becomes more stable and can dominate. I ran a reaction where I initially got 90 percent endo at 60 degrees Celsius, but after leaving the crude mixture at reflux for twelve hours, the ratio flipped to mostly exo. I had to monitor it by NMR to see the shift happen in real time. If you're running a synthesis where stereochemistry matters, you need to decide upfront whether you want kinetic or thermodynamic control and design your conditions around that.
Limitations And When The Diels-Alder Fails
The Diels-Alder isn't a universal solution. A few hard limitations to keep in mind: Geometric constraints on the diene: The diene must be able to adopt an s-cis conformation. If it's locked in s-trans like in some rigid bicyclic systems, the reaction either won't proceed or will require extreme conditions. Open-chain dienes can rotate, but if steric bulk around the central bond makes s-cis prohibitively high in energy, you're stuck. Dienophile reactivity: Electron-poor dienophiles work well. Electron-rich alkenes and alkynes are essentially unreactive toward standard dienes unless you activate them with strong Lewis acids or switch to inverse-electron-demand variants where the diene is electron-poor and the dienophile is electron-rich.
Steric hindrance: Bulky substituents at the reacting centers of either partner can suppress the reaction dramatically. I had a case where a tert-butyl group on the diene terminal carbon killed reactivity entirely. Moving it one position over made the reaction work fine. Position matters more than you'd expect. Ring strain: Forming highly strained bicyclic products is possible, but the strain can push the equilibrium backward. Removing the product as it forms, usually by precipitation or distillation, can help drive the reaction forward. Otherwise you're just sitting in an equilibrium with low conversion.

Practical Tips That Actually Matter
When running a Diels-Alder in practice, solvent choice matters more than people admit. Toluene at reflux is a reliable default for moderate-temperature reactions. Dichloromethane works fine for Lewis acid-catalyzed variants at low temperature. If you're doing an inverse-electron-demand Diels-Alder with a tetrazine and a strained alkene, the reaction can be fast enough to run at room temperature in acetonitrile, sometimes complete within minutes. Workup is usually simple. Dilute with an organic solvent, wash with brine or a mild aqueous solution depending on your reagents, dry over magnesium sulfate, and concentrate. Purification by column chromatography is standard unless the product crystallizes cleanly, in which case recrystallization can give you high purity without a column. The one thing I see people consistently get wrong is trusting the textbook drawing of a perfectly symmetrical transition state. Reality is almost always asymmetric in some direction. Check your regiochemistry with computational modeling or by running a small screening set of similar partners before committing to a full-scale reaction. It saves hours of purification work later.