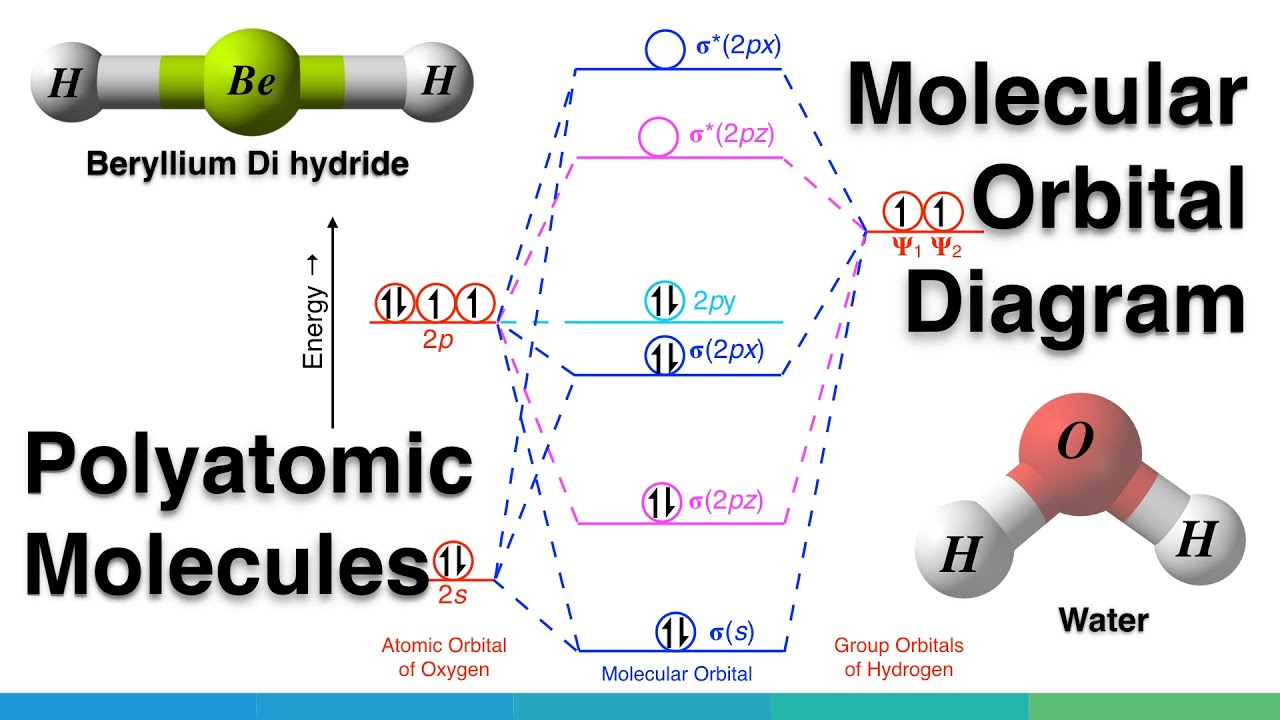
How Polyatomic Bonds Actually Work in Practice
Polyatomic bonds are covalent interactions that hold three or more atoms together within a single molecular unit. The most straightforward example is the sulfate ion, SO², where sulfur sits at the center and four oxygen atoms surround it. But defining them by atom count alone misses the entire point. The real mechanism involves orbital hybridization, electron delocalization, and resonance structures that determine how those atoms actually share electrons. You can't understand polyatomic bonds by memorizing shapes. You need to understand why the bonds exist and what forces stabilize them. When I first worked through this formally, I assumed the Lewis structure approach was sufficient for everything. It isn't. Lewis structures give you a static snapshot. They don't tell you about bond lengths, vibrational modes, or why some polyatomic ions behave as stronger bases in solution than others. I learned that the hard way during a project involving polyatomic oxide catalysts where the measured bond energies didn't match textbook predictions by nearly ten percent. That discrepancy came from ignoring d-orbital participation in the bonding framework, something standard general chemistry courses typically gloss over.
The Definition Of Polyatomic Bonds Unlocking The Secrets Of Molecules
At its core, the definition of polyatomic bonds centers on how multiple atoms participate in a shared electron network rather than pairing off into simple two-atom covalent bonds. This creates a system where individual bond orders don't always add up cleanly. Take ozone, O, for instance. Each oxygen-oxygen connection is neither a single bond nor a double bond. It's a bond order of approximately 1.5 due to resonance. The molecule exists in a superposition of two equivalent structures, and the actual bond length sits between a standard O-O single bond and an O=O double bond. This resonance behavior is where most beginners stumble. They treat polyatomic ions like rigid collections of fixed bonds. In reality, electron density distributes across the entire structure. The nitrate ion, NO, is a textbook case. All three nitrogen-oxygen bonds are identical in length, despite any single Lewis diagram suggesting one double and two single bonds. The true picture requires delocalized pi bonding across all three oxygen atoms simultaneously.
Hybridization and Geometry in Polyatomic Systems
VSEPR theory provides a starting framework for predicting geometry, but it breaks down when dealing with transition metal polyatomic complexes. For main group elements, sp³ hybridization explains tetrahedral arrangements like in the ammonium ion, NH. sp² hybridization accounts for trigonal planar geometries like in the carbonate ion, CO². sp hybridization produces linear arrangements as seen in the azide ion, N. But here is what most resources skip: hybridization is a mathematical convenience, not a physical observable. The actual electron distribution doesn't care about your chosen hybridization scheme. When modeling polyatomic bonds computationally, I've found that using extended Hückel methods or DFT calculations gives more reliable predictions than relying on hybridization labels alone. The labels are useful shorthand for communication, but they become misleading when precision matters. I spent weeks trying to reconcile experimental crystallography data with predicted geometries for a series of chlorate and perchlorate derivatives. The predicted bond angles from VSEPR were consistently off by a few degrees from what X-ray diffraction showed. The fix wasn't to adjust the theory. It was to account for lone pair-bond pair repulsion variations that depend on the electronegativity of surrounding substituents. More electronegative neighbors pull electron density away, which compresses bond angles slightly compared to what simple VSEPR predicts.
Get the Full Details
Resonance and Delocalization: Where the Real Complexity Lives
Delocalization energy is the quantitative measure of resonance stabilization. For the carbonate ion, the resonance stabilization is roughly 30 to 40 kcal/mol compared to a hypothetical structure with localized single and double bonds. This energy difference has practical consequences. It affects pKa values, reaction pathways, and spectral properties. One counter-intuitive point that trips people up repeatedly: adding resonance structures doesn't automatically make a molecule more stable. The stability depends on whether the contributing structures are equivalent in energy. Equivalent resonance contributors produce the greatest stabilization. Non-equivalent contributors, like those in the nitrite ion where one structure places a formal positive charge on oxygen and another doesn't, contribute unevenly to the actual wavefunction. The lower-energy contributor dominates. I encountered a situation where a colleague was modeling a polyatomic phosphate intermediate and got incorrect reaction kinetics because they only considered one resonance form. The intermediate had multiple valid resonance structures with different charge distributions, and the reactive site depended on which structure you examined. Switching to a quantum chemical calculation that treated all resonance forms simultaneously resolved the discrepancy entirely. It took about twenty minutes to set up the Gaussian job and maybe forty minutes to run it on the cluster.
Polyatomic Bonds in Real-World Applications
Battery chemistry provides a clear example of polyatomic bond behavior in action. Lithium iron phosphate, LiFePO, relies on the structural integrity of the PO³ polyatomic unit. The strong covalent P-O bonds within the phosphate tetrahedron remain intact during lithium intercalation and deintercalation cycles. This is why LiFePO batteries have long cycle lives. The polyatomic bond framework acts as a structural scaffold that doesn't collapse when lithium ions move in and out. In contrast, materials where the polyatomic anion unit is weaker or more flexible tend to degrade faster. I've seen cells fail prematurely when the polyanion framework underwent structural rearrangement during cycling. The polyatomic bonds themselves didn't break, but the overall lattice geometry shifted in a way that blocked lithium diffusion pathways. This is a subtle failure mode that doesn't show up in simple electrochemical testing without careful structural analysis. Enzymatic catalysis also depends heavily on polyatomic bond arrangements. Metalloenzymes use polyatomic coordination complexes at their active sites. The geometry and electronic structure of these polyatomic metal-ligand systems determine substrate specificity and catalytic rate. A single ligand substitution can change the entire reactivity profile because it alters the orbital overlap within the polyatomic bonding network.
Limitations and When Polyatomic Bond Models Fail
The biggest limitation of standard polyatomic bond theory is that it assumes static structures. Real molecules vibrate, rotate, and undergo conformational changes. At room temperature, polyatomic ions in solution are constantly distorting from their ideal geometries. These dynamic effects matter for spectroscopy and reaction dynamics but are absent from textbook treatments. Another failure point: polyatomic bond models struggle with systems containing heavy elements where relativistic effects become significant. Gold complexes, for example, show bonding behavior that standard hybridization and VSEPR models can't predict accurately. You need relativistic quantum chemistry methods, which are computationally expensive and not practical for routine analysis. Perhaps the most important limitation: polyatomic bond theory doesn't naturally account for solvent effects. The sulfate ion behaves differently in gas phase versus aqueous solution. Water molecules orient around the ion's charge distribution, stabilizing certain resonance forms over others and altering effective bond lengths. If you're working with solution-phase chemistry, ignoring solvent effects will give you wrong answers more often than you'd expect.

Practical Approach to Analyzing Polyatomic Bonds
Start with the Lewis structure. Get the connectivity right. Count valence electrons carefully. Place lone pairs. Check formal charges. This takes about five minutes for simple ions and maybe fifteen for larger ones. Then draw all reasonable resonance structures. Don't stop at the first one that looks plausible. The quality of your resonance analysis directly affects how well you can predict reactivity. I usually sketch three to five structures and then evaluate which contribute most based on formal charge distribution and octet rule satisfaction. Next, determine the hybridization of each central atom and predict the geometry using VSEPR. Compare your prediction to known experimental data if available. If they disagree significantly, revisit your resonance structures and formal charge assignments. Most disagreements come from missed resonance contributors or incorrect formal charge placement.
For anything beyond educational exercises, use computational chemistry software. Open-source tools like ORCA or Gaussian (if you have access) can optimize the geometry and give you bond orders, partial charges, and vibrational frequencies. A typical geometry optimization for a medium-sized polyatomic ion takes anywhere from ten minutes to an hour depending on the method and basis set. DFT with a moderate basis set like def2-SVP usually gives results within a few percent of experimental values for most main group polyatomic systems. The deeper you go into polyatomic bond analysis, the more you realize that the standard definitions are approximations. They're useful approximations, but approximations nonetheless. The actual physics involves solving the Schrödinger equation for a many-electron system, and no amount of Lewis structures or hybridization labels will ever capture that completely. The tricks are knowing when the approximations are good enough and when they're actively misleading you.


