Working With Limiting Reactants In The Lab
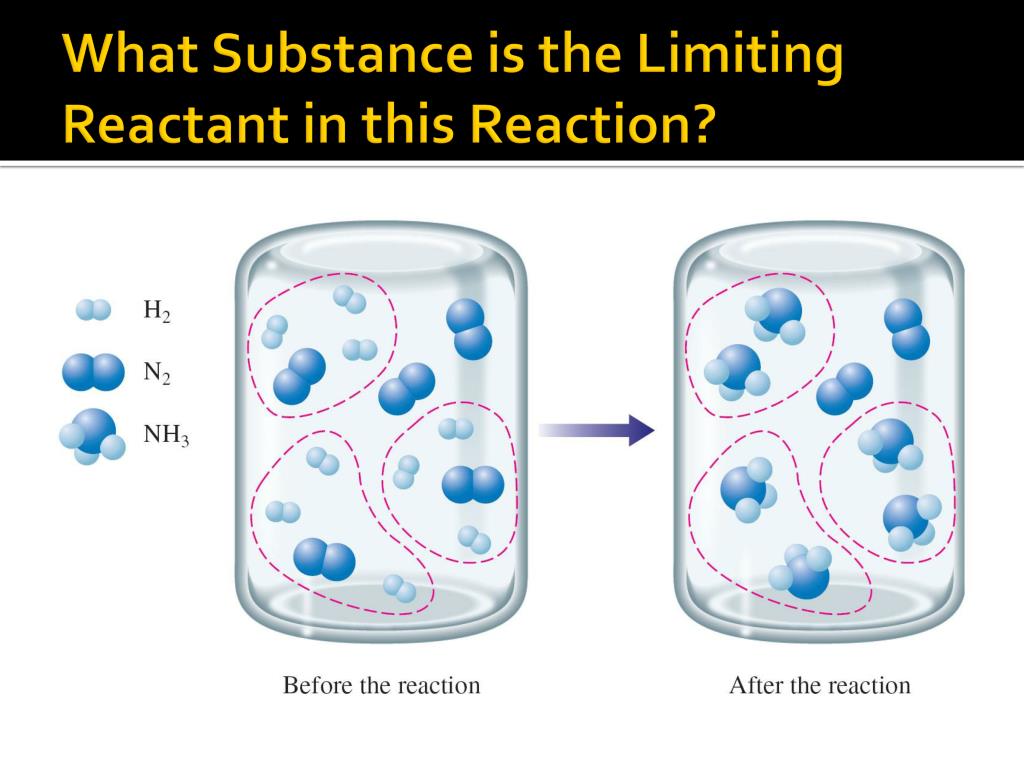
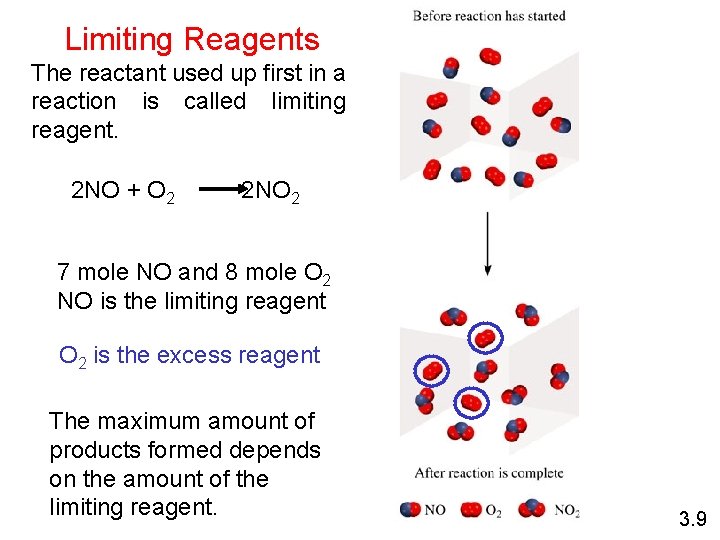
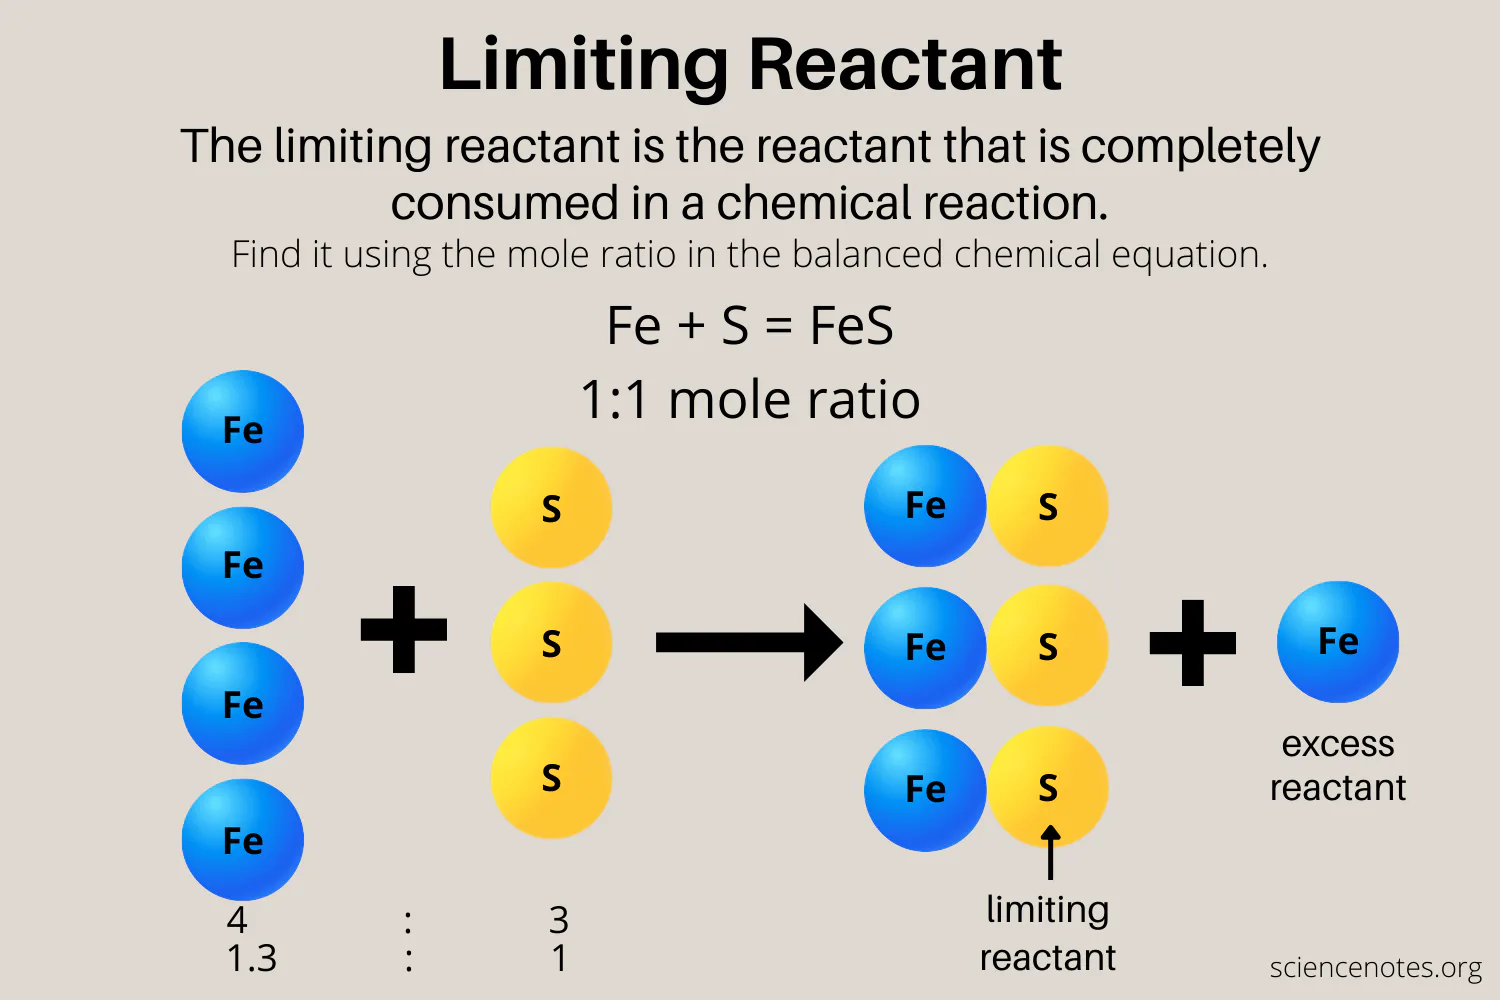
Most people learn limiting reactants from a textbook problem where you mix exactly 2.0 moles of hydrogen with 1.5 moles of oxygen and calculate water yield. That is not how it works outside a classroom. In practice, you are dealing with reagents that are impure, concentrations that drift over time, and reaction conditions you cannot control perfectly. The concept itself is straightforward, but the execution is where things fall apart. A limiting reactant is simply the substance that gets consumed first and therefore caps how much product you can form. Everything else is in excess. You determine it by converting your starting amounts into moles, dividing by the stoichiometric coefficient from the balanced equation, and comparing the results. The smallest ratio wins. That is the theory. Here is what the theory does not tell you. In a real batch synthesis, I once ran a Grignard reaction where the limiting reagent kept shifting between batches because the commercial-grade bromobenzene contained varying amounts of phenol as a stabilizer. The phenol quenches the Grignard reagent before it ever touches your intended substrate. Standard limiting reactant calculations assume your starting material is pure. That assumption cost me three wasted batches and roughly fourteen hours of chromatography time. The workaround was straightforward but tedious: I began titrating each new bottle of bromobenzene against a standard base to determine actual active halide content before committing to stoichiometric calculations. Once I had the true concentration, I could calculate the actual limiting reagent rather than trusting the label.
Another thing textbooks rarely emphasize is that the limiting reactant is not always the one you think it is when you have competing side reactions. If your desired pathway and a side pathway consume the same reagent but at different rates, the limiting reagent for your target product may be completely different from the limiting reagent for the overall consumption of that material. I learned this running a nitration where I was chasing a mono-nitrated product. The limiting reactant for the mono-substitution was the aromatic starting material by one calculation, but dinitration was consuming the nitric acid faster than I expected. I ended up running the reaction with the aromatic in slight excess relative to the mono-pathway because controlling the nitric acid addition rate mattered more than following the simple stoichiometric ratio. The yield of the desired product doubled when I stopped treating the limiting reactant as a single number and started treating it as a dynamic variable. Here is the practical method I use now when setting up any reaction that depends on identifying a limiting reagent: First, write out the balanced equation for the main reaction and note any known side reactions that consume the same reagents. Second, determine the actual moles of each reactant available, accounting for purity, hydration state, and concentration accuracy. Third, divide each value by its stoichiometric coefficient across all relevant reactions. Fourth, identify which pathway each reactant limits and decide which limitation matters for your desired outcome. Fifth, adjust your ratios accordingly. This usually takes about twenty minutes for a standard organic synthesis and prevents most of the yield loss I used to encounter.
There are situations where this approach breaks down entirely. If a reaction is diffusion-controlled or mass-transfer limited rather than kinetically limited, the concept of a limiting reactant becomes almost meaningless. Stirring speed, surface area, and phase transfer become the actual constraints. I ran a heterogeneous catalytic hydrogenation once where switching from a magnetic stir bar to a mechanical overhead stirrer increased the effective reaction rate by roughly threefold, even though the limiting reactant calculations were identical in both setups. In cases like that, your limiting reactant analysis is mathematically correct but chemically irrelevant. You are better off optimizing mixing or catalyst surface area instead of chasing a theoretical yield that the reaction never reaches anyway. Another honest limitation is that limiting reactant analysis assumes complete reaction. It tells you the maximum possible product, not the actual product. Real reactions have equilibrium constraints, incomplete conversions, and losses during workup. A typical organic reaction with clean stoichiometry and careful technique might reach sixty to eighty percent of the theoretical yield after purification. If your limiting reactant calculation gives you five grams of theoretical product, plan on getting three to four grams at best. Planning around the theoretical yield rather than an estimated practical yield is one of the fastest ways to undersupply downstream steps or waste reagents in excess trying to compensate for losses that the calculation never accounted for. When you need something more precise than standard stoichiometric calculations, consider running a small scale screening reaction at ten percent of your intended scale before committing to a full batch. This usually takes about an hour and reveals whether your limiting reactant assumptions hold up under actual conditions. You will catch issues like unexpected side reactions, reagent degradation, or solubility problems that no amount of calculation will predict. The screening step itself becomes a practical limiting reactant determination based on observation rather than theory.
Get the Full Details
For routine stoichiometric work, free tools like the limiting reactant calculators on PubChem or simple spreadsheet models do the arithmetic without issue. There is no reason to overcomplicate the math when the chemistry is straightforward. The value comes from knowing when the math stops being useful and your hands-on judgment needs to take over.