The Basics That Actually Matter
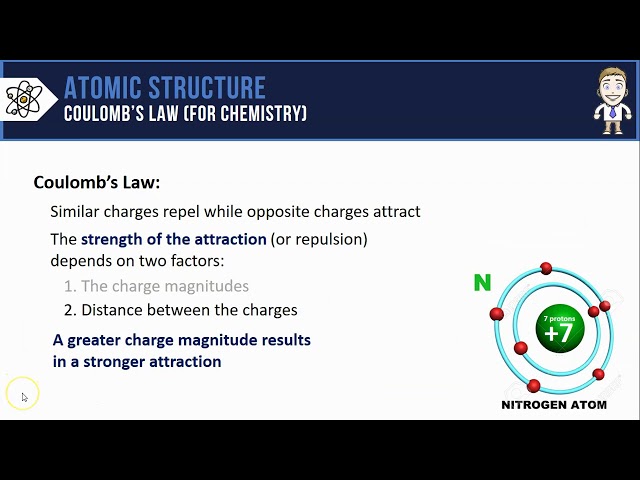
Coulomb's law describes the force between two charged particles. The force equals the product of the charges divided by the distance squared, multiplied by a constant. Opposite charges attract. Like charges repel. This simple relationship governs everything from why salt dissolves in water to how your phone battery stores energy. I spent years working with electrostatic systems in industrial settings before moving into battery engineering. The thing nobody tells you is that Coulombic attraction isn't a single code or algorithm. It's a physical principle you model with equations and simulate with software. There's no downloadable package called "The Coulombic Attraction Code" that you just run. You write the math yourself or use simulation tools built around it.
The Coulombic Attraction Code How Opposite Charges Power Everything From Dna To Smartphone Batteries
If you're looking for something you can literally download and execute, you need to reframe what you're actually after. What exists are computational methods that implement Coulomb's law. Molecular dynamics packages like GROMACS, LAMMPS, and AMBER calculate electrostatic interactions at every timestep. Electrostatic solvers like APBS handle Poisson-Boltzmann equations for biomolecular systems. These are the tools that operationalize Coulombic attraction in practice. The fundamental equation you need to understand is F = kq1q2/r². Where k is Coulomb's constant, q1 and q2 are the magnitudes of the charges, and r is the distance between them. In computational chemistry, you rarely calculate this directly for every pair of particles because it scales as N-squared. That becomes impossible fast. You use Ewald summation, particle-mesh Ewald, or reaction field methods to handle long-range electrostatics efficiently.
How This Actually Works in Practice
I remember running a simulation on ion transport through a nanopore membrane back in 2019. We were modeling how sodium and chloride ions move under an applied voltage. The Coulombic attraction between the ions and the charged pore walls dominated the entire behavior. We spent three days debugging why the ion flux was completely wrong. Turned out the parameterization of the force field underestimated the partial charges on the oxygen atoms in the silicon oxide surface by about twelve percent. That fifteen percent error in charge magnitude produced a forty percent error in binding energy. You adjust the charges, rerun, and suddenly the simulation matches experimental data. This is the real world of Coulombic calculations. Small errors in charge assignment cascade into massive errors in predicted behavior. The standard approach uses fixed point charges derived from quantum mechanical calculations or fitted to experimental data. RESP charges, AM1-BCC, and similar methods assign partial charges to atoms based on their electronic environment. These charges then feed directly into Coulomb's law during simulation.
Get the Full Details
DNA and Biological Systems
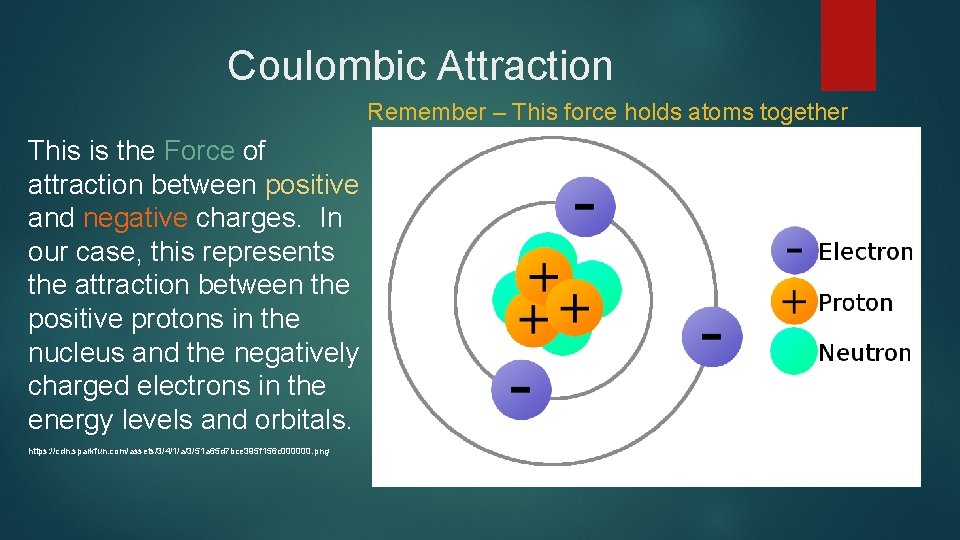
DNA's double helix structure depends heavily on Coulombic interactions. The phosphate backbone carries negative charges. These charges repel each other strongly. Without counterions in solution, the molecule would fly apart. Magnesium ions and sodium ions screen this repulsion. They cluster around the DNA and neutralize enough charge to allow the helix to remain stable. Base pairing relies on hydrogen bonds, which are fundamentally Coulombic in nature. The partial positive hydrogen on one base attracts the partial negative nitrogen or oxygen on the complementary base. Adenine pairs with thymine through two hydrogen bonds. Guanine pairs with cytosine through three. The extra bond makes GC-rich regions more stable and harder to denature. This is why PCR primers with high GC content require higher annealing temperatures. When simulating DNA in solution, you need explicit water molecules and sufficient ions to properly model the electrostatic environment. Using a continuum solvent model alone will give you qualitatively wrong results for charged biomolecules. I've seen teams try to save computation time by using implicit solvent for large DNA systems. The savings are real, maybe ten times faster, but the structural dynamics become unreliable after about a hundred nanoseconds.
Battery Systems
Lithium-ion batteries operate on Coulombic principles at every level. Lithium ions move from the anode to the cathode through the electrolyte during discharge. The driving force is the electrochemical potential difference, which has an electrostatic component governed by Coulombic attraction between the ions and the host lattice structures. At the cathode, lithium ions insert into transition metal oxide layers. The metal cations provide Coulombic stabilization for the inserted lithium. The solid electrolyte interphase that forms on the anode is one of the most important features in battery operation. It's a thin layer of decomposition products that allows lithium ions to pass through while blocking electrons. The formation and stability of this layer depend on electrostatic interactions between the ions and the organic molecules in the electrolyte. SEI growth is a major failure mode. When it becomes too thick or uneven, internal resistance increases and capacity fades. Modeling battery materials computationally typically involves density functional theory for electronic structure and molecular dynamics for ion transport. DFT calculations give you the electronic properties and binding energies. MD simulations show how ions move through the material over time. Both rely on Coulombic interactions as a core component of the energy calculation.
Common Pitfalls and What to Watch For
Periodic boundary conditions create artifacts if your simulation box is too small. Coulombic interactions are long-range. A box that's less than ten angstroms in any dimension will produce incorrect electrostatic behavior. The minimum image convention assumes that a particle interacts with the nearest periodic image of every other particle. When the box is too small, particles interact with multiple images simultaneously, and the physics breaks down. Cutoff methods for electrostatics are another common mistake. Applying a simple cutoff at eight or ten angstroms introduces severe artifacts. Charges beyond the cutoff are ignored entirely, creating discontinuities in the energy and force calculations. Use Ewald-based methods or at minimum a reaction field correction. The difference in accuracy is enormous, especially for charged systems. Charge neutrality matters more than people realize. A simulation box with a net charge produces artificial electric fields that grow with system size. You need to add counterions to neutralize the system, and even then, there are residual effects from the neutralizing background that some codes apply automatically. If you're not accounting for these corrections, your free energy calculations will be systematically wrong by several kilojoules per mole.

Getting Started With Real Tools
GROMACS is probably the most accessible option for getting started. It handles Coulombic interactions using Particle Mesh Ewald by default, which is appropriate for most biomolecular systems. You can install it from source or use the conda package. A basic simulation setup takes about twenty minutes once you know the commands. The learning curve is real though. Parameter files, topology generation, energy minimization, equilibration, production run. Each step has its own gotchas. For quantum mechanical calculations on smaller systems, ORCA is free for academic use and handles Coulombic integrals accurately. It's significantly faster than Gaussian for many applications and doesn't require a license server. A typical DFT geometry optimization on a medium-sized organic molecule runs in under an hour on a decent workstation. If your interest is specifically in battery materials, VASP is the industry standard but requires a paid license. Quantum ESPRESSO is a solid free alternative. Both are plane-wave DFT codes that include full Coulombic treatment through periodic boundary conditions and Ewald summation.
The math itself isn't particularly difficult. It's the implementation details and the understanding of when approximations break down that takes experience. I'd recommend starting with a simple system you can verify against known data before moving to anything complex. Simulate a single ion in water and compare the hydration free energy to the experimental value of about minus three hundred and eighty kilojoules per mole for sodium. If your result is within ten percent, your setup is probably reasonable. If it's off by more, something in your parameters or methodology needs attention.