Prokaryotic Gene Regulation Is Messier Than Textbooks Make It Look
I spent three weeks troubleshooting a lac operon construct that refused to repress even when the culture was sitting in high-lactose, low-glucose media. The plasmid had the right promoter, the right operator sequence, and the right lacI gene. Nothing worked. Turned out the copy number was twelve times what the supplier claimed, which means the Lac repressor was titrated out by the sheer volume of operator sites flooding the cell. Every student who has ever tried to use a pUC-based expression vector for repressor-dependent systems runs into this eventually. Gene expression in prokaryotes is not a simple on-off switch. It is a layered system of transcriptional control, translational control, and post-translational modification that operates simultaneously. Understanding how it works matters whether you are engineering a metabolic pathway, building a biosensor, or just trying to get a protein to express at the level you need without killing the cells.
The Complex Dance Of Gene Expression In Prokaryotes Unlocking The Secrets Of Control
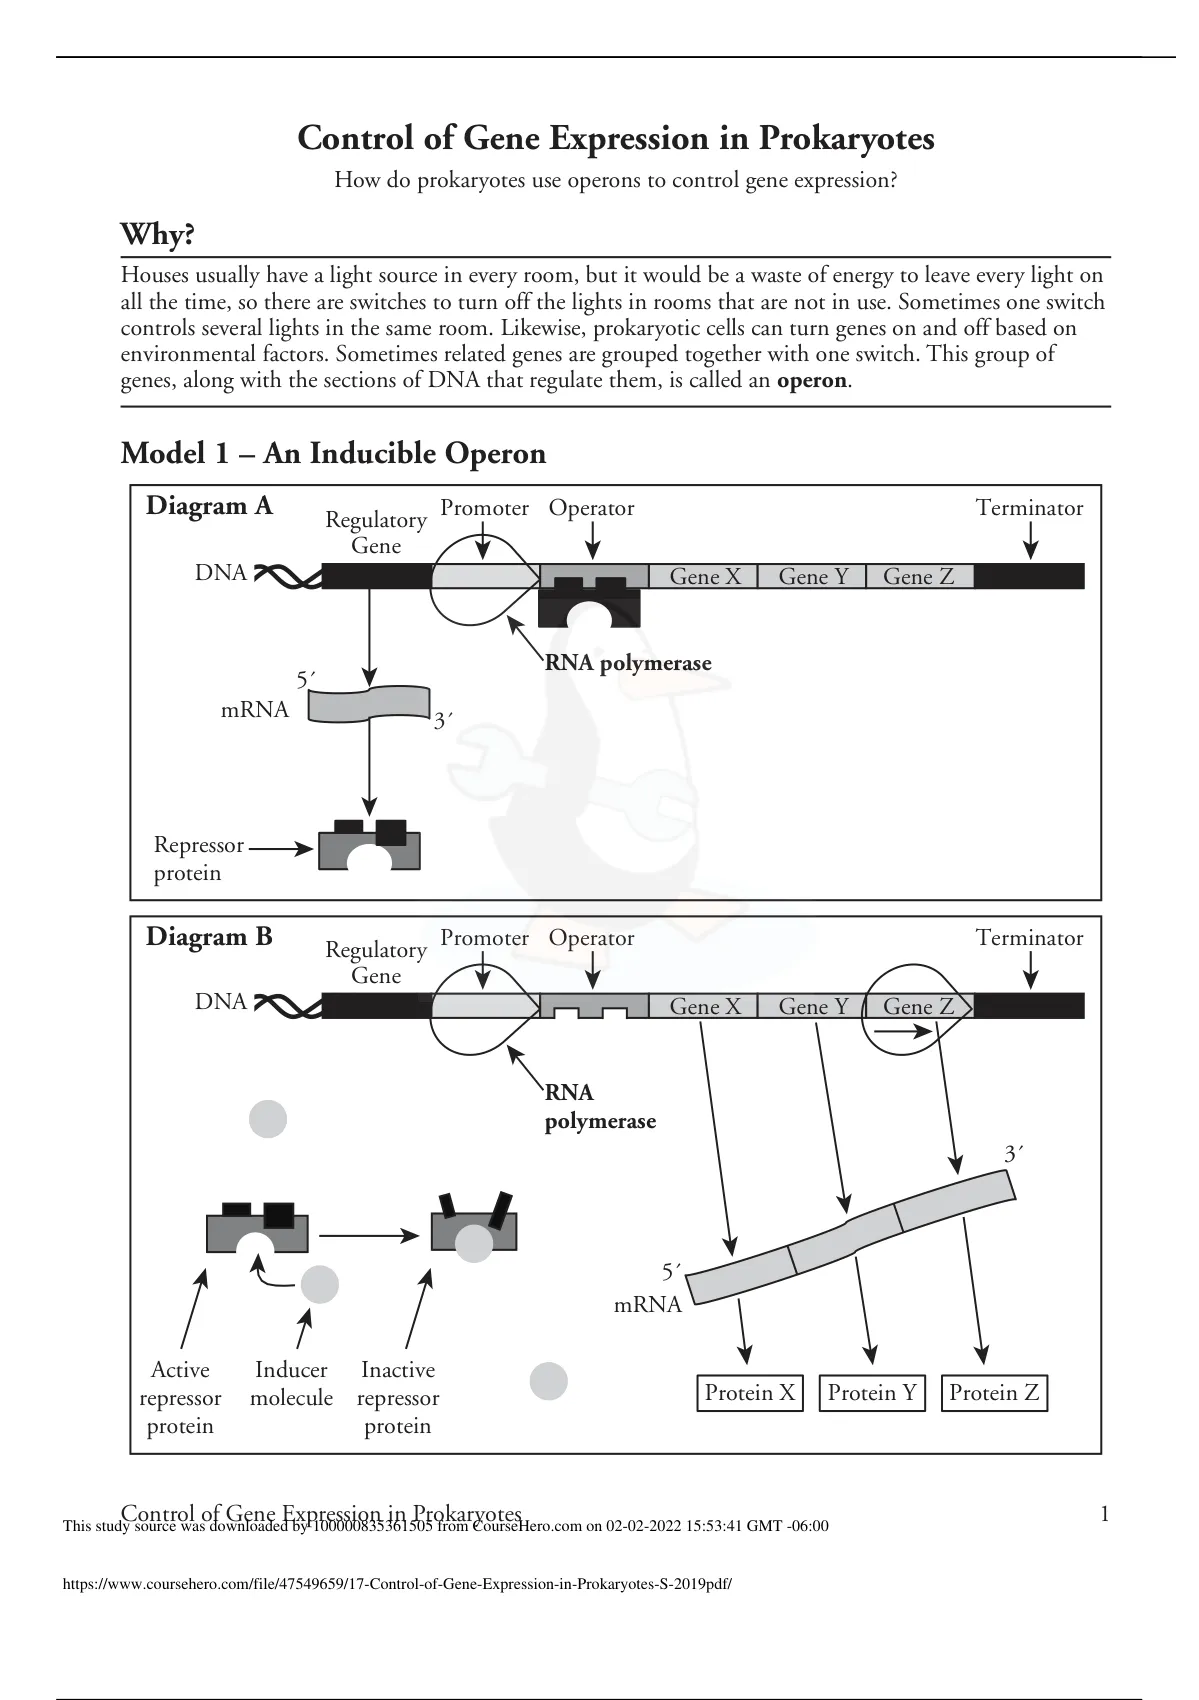
At the core is the operon model. DNA is transcribed into polycistronic mRNA, meaning one transcript carries multiple coding sequences. The lac operon, the trp operon, the araBAD operon — these are the canonical examples you will see in every textbook. But the real mechanism is more nuanced than repression and activation alone. Repressors bind operator sequences and physically block RNA polymerase from initiating transcription. The trp repressor needs tryptophan as a co-repressor to become active. Without tryptophan, the repressor falls off and transcription proceeds. With tryptophan, the repressor-tryptophan complex binds the operator and transcription stops. That part is straightforward. Activators work the other direction. CAP-cAMP binds upstream of the lac promoter when glucose is low, bending the DNA and recruiting RNA polymerase more efficiently. This is why the lac operon only fully activates when glucose is absent and lactose is present. Two signals, two inputs, integrated at the DNA level.
But here is where people get tripped up. Attenuation. The trp operon does not just get turned off by the repressor. It also uses a premature transcription termination mechanism that depends on ribosome positioning during translation. If tryptophan is abundant, the ribosome moves quickly through the Trp codons in the leader peptide, allowing the terminator hairpin to form and transcription to stop before the structural genes are reached. If tryptophan is scarce, the ribosome stalls, an anti-terminator hairpin forms instead, and transcription continues. This is translational-transcriptional coupling, and it means the cell is monitoring actual tRNA charging levels, not just metabolite concentration. I ran into this when I was trying to get consistent expression from a trp-driven promoter in a minimal medium. The repressor mechanism shut things down predictably, but attenuation added a second layer of control that was extremely sensitive to growth rate. Slow-growing cells showed much stronger repression than fast-growing cells, even at identical tryptophan concentrations. This is not something you would guess from the standard diagram. The ribosome speed is tied to the cell cycle, and the cell cycle is tied to everything else happening in the culture. Sigma factor switching is another critical control point that gets shortchanged in introductory courses. E. coli has seven major sigma factors, and each one directs RNA polymerase to a different set of promoters. RpoD sigma 70 handles housekeeping genes. RpoS sigma S takes over during stationary phase and stress. RpoH sigma 32 responds to heat shock. RpoN sigma 54 controls nitrogen regulation. The cell does not just turn individual genes on and off. It reprograms its entire transcriptional machinery to match environmental conditions.
Get the Full Details

This has direct practical consequences. If you are expressing a recombinant protein and your culture hits stationary phase early because of a contamination or a bad inoculum density, you are no longer transcribing from sigma 70 promoters with the same efficiency. Your construct may contain a T7 promoter, which requires T7 RNA polymerase, but if the strain carrying that polymerase is under stress and downregulating the lacUV5 promoter driving T7 pol expression, your yield drops to near zero. I learned this the hard way when a routine expression run produced nothing and it took me two days to realize the overnight culture had sat too long before inoculation. Small RNA regulators add another dimension. Antisense RNAs in bacteria can bind mRNA and either block ribosome access or recruit RNase E for degradation. The RyhB sRNA in E. coli represses non-essential iron-using proteins when iron is scarce, redirecting resources to essential pathways. These are not edge cases. They are fundamental control mechanisms, and they mean that even if your promoter is perfectly induced, the mRNA you produce might get degraded before translation ever begins. Translational control matters too. The Shine-Dalgarno sequence strength, mRNA secondary structure around the start codon, codon usage bias, and tRNA abundance all affect how efficiently a given mRNA gets translated. A strong promoter does not guarantee strong protein expression. I have seen constructs with robust T7 promoters produce barely detectable protein simply because the mRNA formed a stable hairpin that sequestered the ribosome binding site. A single point mutation four nucleotides upstream of the start codon — enough to disrupt the secondary structure — increased expression tenfold.
Post-translational control is the final layer. Protein stability, proteolysis, and allosteric regulation all determine how much functional protein actually accumulates. ClpXP and Lon proteases degrade misfolded or unnecessary proteins. Some operons produce regulatory proteins that inhibit their own synthesis through feedback loops. The lambda phage decision between lysis and lysogeny is a classic example of protein-level bistability driven by Cro and cI repressors competing for overlapping operator sites. If you are designing a synthetic circuit or optimizing expression, you need to think about all these layers. Here is a practical checklist based on what I have learned from experience: First, check plasmid copy number. High-copy vectors amplify the titration problem for repressor-based systems. If you are using a repressor, switch to a low-copy backbone like p15A or pSC101 derivatives. This alone resolved my lac operon issue.
Second, measure growth rate, not just optical density. Attenuation and sigma factor switching respond to growth conditions, not cell density. A culture at OD600 of 0.8 could be in exponential phase or early stationary depending on the medium and temperature. Plate counts or live-dead staining give you a clearer picture. Third, verify your inducer concentration against the actual Kd of your regulator. IPTG is not a perfect analogue of allolactose. It does not get metabolized, so the intracellular concentration is different from what you would calculate from the media supplement. For fine-tuned expression, dose-response curves under your specific conditions are necessary. Standard protocols calling for 0.1 or 1 mM IPTG are starting points, not recommendations. Fourth, watch for metabolic burden. Strong expression drains resources. Cells slow down, protein quality drops, and unintended stress responses kick in. I have seen T7-based expression systems trigger the stringent response because the cell could not keep up with rRNA and tRNA demand. Adding a weaker promoter or using a tunable system like the rhamnose promoter can reduce this.

Fifth, consider codon optimization carefully. It is not a universal fix. Over-optimization can cause ribosome traffic jams when rare codons are removed entirely, and it can create unintended secondary structures or cryptic splice sites in some contexts. A balanced approach that preserves natural codon usage patterns in the middle of the gene while optimizing the N-terminal region usually works better. The bottom line is that prokaryotic gene expression is a system of integrated controls, not a collection of independent switches. Every component affects every other component. You cannot isolate transcription from translation, or translation from growth rate, or growth rate from plasmid burden. The organisms that taught us the operon model were doing elegant basic science, but they were also working with wild-type strains in controlled conditions. When you take those systems out of the textbook and put them into a bioreactor or a cloning strain, the complexity shows up immediately. I still run into unexpected behavior. Last month I was working with an arabinose-inducible system and saw variable expression between replicates even though the induction protocol was identical. It turned out to be a carryover effect from the previous growth medium — residual glucose was affecting AraC conformation through catabolite repression, even though the arabinose concentration was sufficient to induce. The lesson was mundane but expensive in terms of time: always wash cells before resuspending in induction medium, and never assume that a clean transfer protocol means a clean physiological state.